

Computing the (1)H NMR spectrum of a bulk ionic liquid from snapshots of car-parrinello molecular dynamics
Alessandro Bagno1, Fabio D'Amico, Giacomo Saielli
1Dipartimento di Scienze Chimiche, Università di Padova via Marzolo, 135131 Padova, Italy. alessandro.bagno@unipd.it

Abstract:
We have investigated the performance of several computational protocols in predicting the NMR spectrum of a molecular ion in a complex liquid phase such as an ionic liquid. To do this, we computed the proton NMR chemical shifts of the 1-ethyl-3-methylimidazolium cation [emim](+) in [emim][Cl]. Environmental effects on the imidazolium ring proton chemical shifts are quite significant and must be taken into account explicitly. Calculations performed on the isolated imidazolium cation as well as on the [emim][Cl] ion pair grossly fail to reproduce the correct spacing between proton signals. In contrast, calculations performed on clusters extracted from the trajectory of a Car-Parrinello molecular dynamics simulation yield very good results.

Related Concept Videos
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:19
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:26
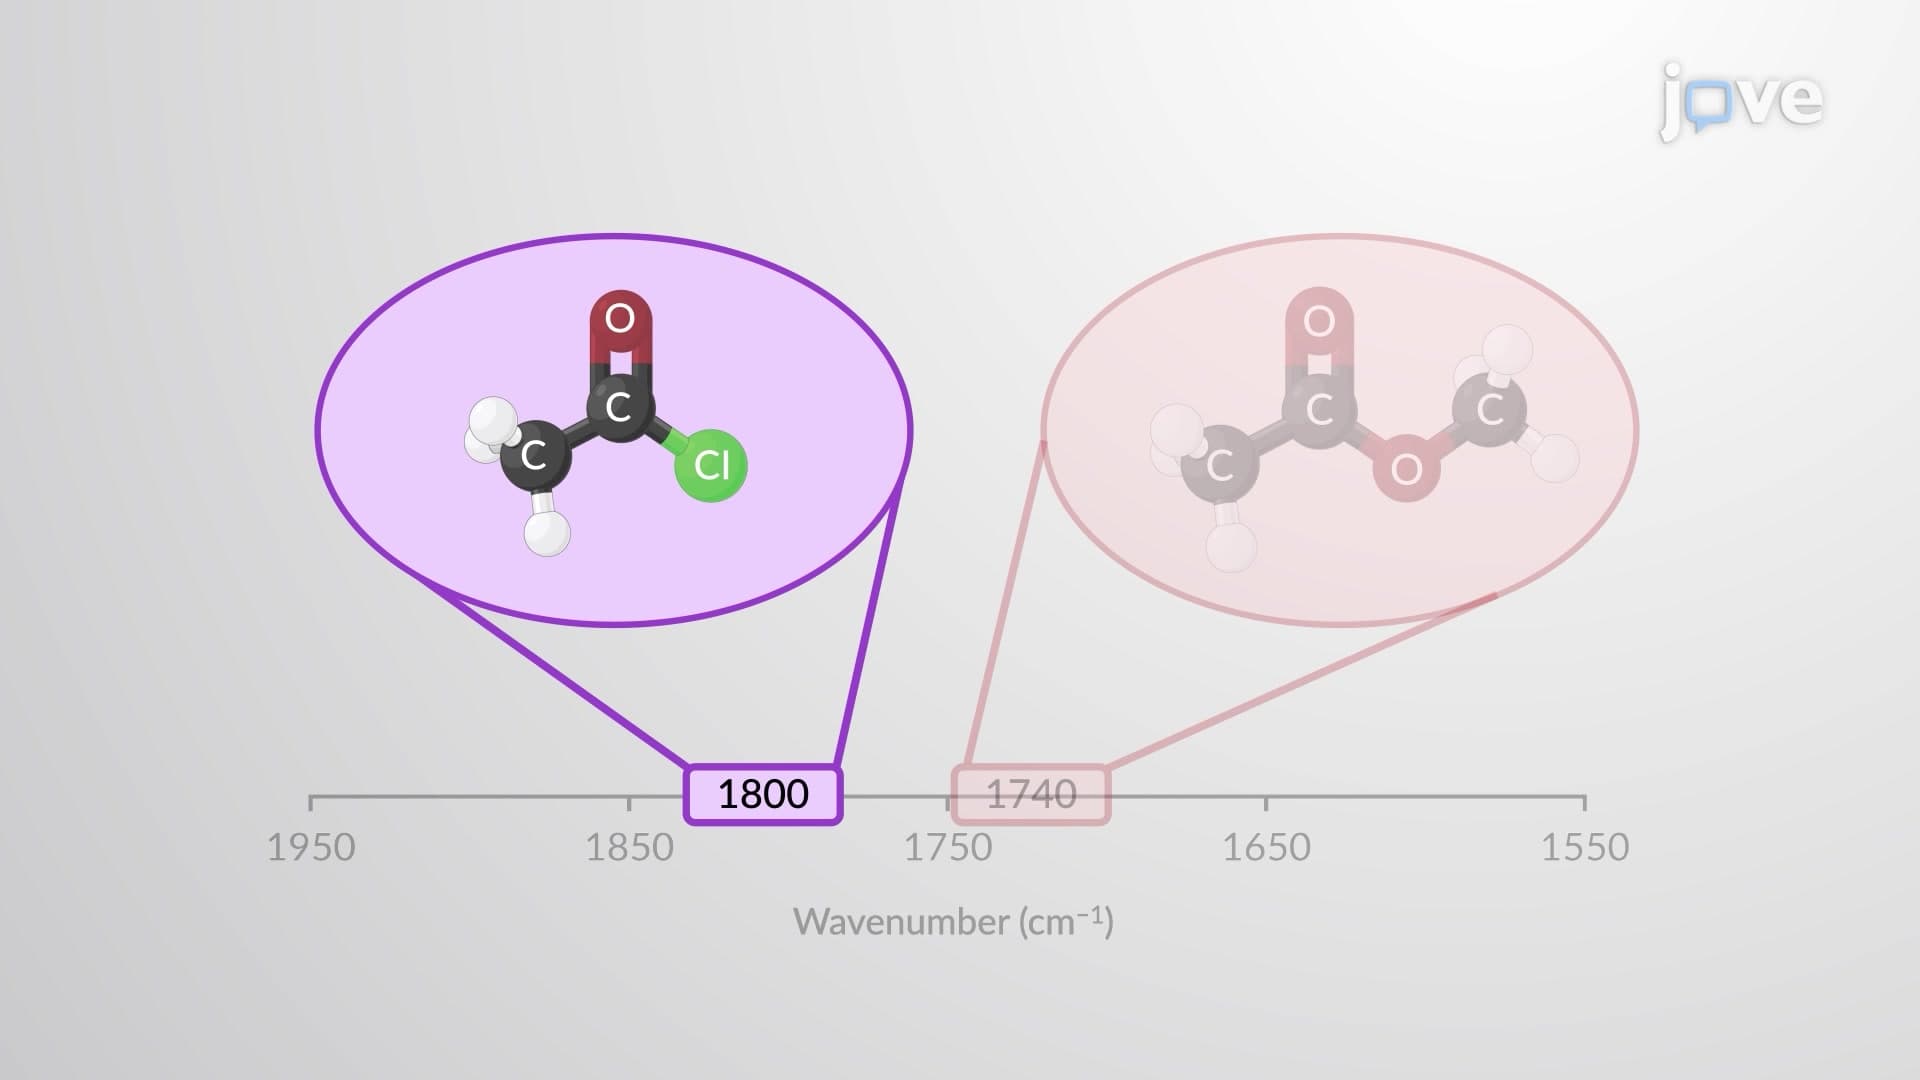
Spectroscopy of Carboxylic Acid Derivatives
In the...
 01:28
IR and UV–Vis Spectroscopy of Carboxylic Acids
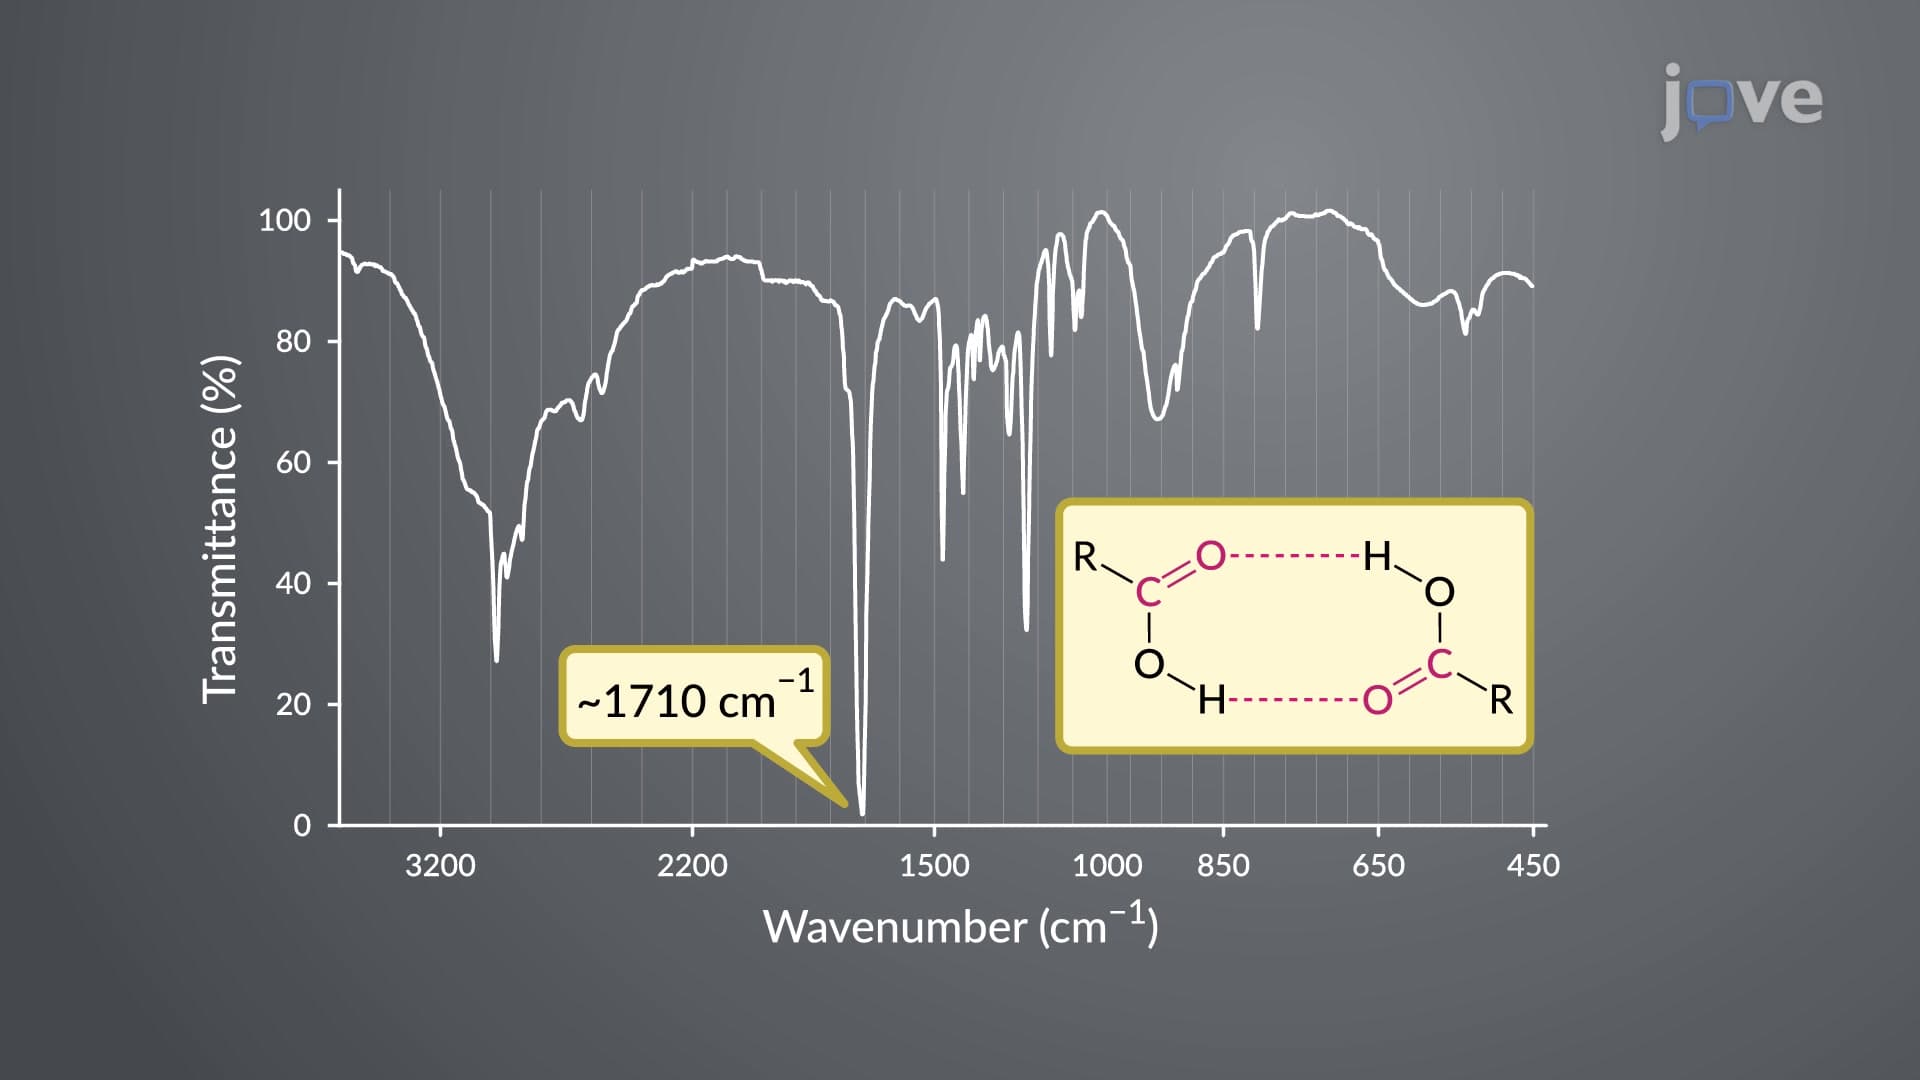
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency, 1710 cm−1. The C=O bond of the...