Related Experiment Video
Updated: Jul 16, 2026
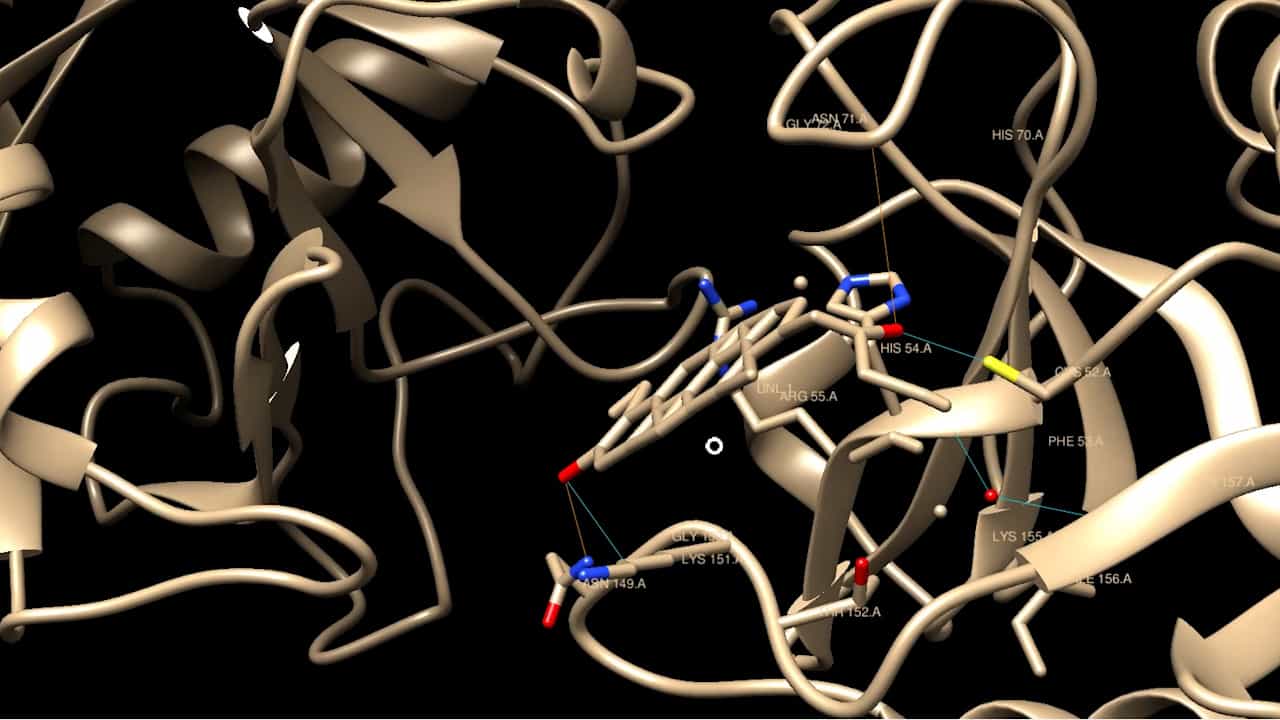

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Exploration of enzyme-ligand interactions in CYP2D6 & 3A4 homology models and crystal structures using a novel
Britta Kjellander1, Collen M Masimirembwa, Ismael Zamora
1Discovery DMPK & Bioanalytical Chemisty Department, AstraZeneca R&D Mölndal, SE-431 81, Sweden. Britta.kjellander@astrazeneca.com

Abstract:
New crystal structures of human CYP2D6 and CYP3A4 have recently been reported, and in this study, we wanted to compare them with previously used homology models with respect to predictions of site of metabolism and ligand-enzyme interactions. The data set consisted of a family of synthetic opioid analgesics with the aim to cover both CYP2D6 and CYP3A4, as most of these compounds are metabolized by both isoforms. The program MetaSite was used for the site of metabolism predictions, and the results were validated by experimental assessment of the major metabolites formed with recombinant CYP450s. This was made on a selection of 14 compounds in the data set. The prediction rates for MetaSite were 79-100% except for the CYP3A4 homology model, which picked the correct site in half of the cases. Despite differences in orientation of some important amino acids in the active sites, the MetaSite-predicted sites were the same for the different structures, with the exception of the CYP3A4 homology model. Further exploration of interactions with ligands was done by docking substrates/inhibitors in the different structures with the docking program GLUE. To address the challenge in interpreting patterns of enzyme-ligand interactions for the large number of different docking poses, a new computational tool to handle the results from the dockings was developed, in which the output highlights the relative importance of amino acids in CYP450-substrate/inhibitor interactions. The method is based on calculations of the interaction energies for each pose with the surrounding amino acids. For the CYP3A4 structures, this method was compared with consensus principal component analysis (CPCA), a commonly used method for structural comparison to evaluate the usefulness of the new method. The results from the two methods were comparable with each other, and the highlighted amino acids resemble those that were identified to have a different orientation in the compared structures. The new method has clear advantages over CPCA in that it is far simpler to interpret and there is no need for protein alignment. The methodology enables structural comparison but also gives insights on important amino acid substrate/inhibitor interactions and can therefore be very useful when suggesting modifications of new chemical entities to improve their metabolic profiles.
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 00:49
Ligand Binding and Linkage
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes