Related Experiment Video
Updated: Jul 16, 2026
Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
In vitro prediction of stop-codon suppression by intravenous gentamicin in patients with cystic fibrosis: a pilot
Isabelle Sermet-Gaudelus1, Michel Renouil, Anne Fajac
1Centre de Ressources et de Compétence en Mucoviscidose, Hôpital Necker-Enfants Malades, AP-HP, Paris, France. isabelle.sermet@nck.aphp.fr

Background:
Cystic fibrosis (CF) is caused by mutations in the gene encoding the cystic fibrosis transmembrane conductance regulator (CFTR) protein, which acts as a chloride channel activated by cyclic AMP (cAMP). The most frequent mutation found in 70% of CF patients is F508del, while premature stop mutations are found in about 10% of patients. In vitro aminoglycoside antibiotics (e.g. gentamicin) suppress nonsense mutations located in CFTR permitting translation to continue to the natural termination codon. Pharmacologic suppression of stop mutations within the CFTR may be of benefit to a significant number of patients. Our pilot study was conducted to determine whether intravenous gentamicin suppresses stop codons in CF patients and whether it has clinical benefits.
Methods:
A dual gene reporter system was used to determine the gentamicin-induced readthrough level of the most frequent stop mutations within the CFTR in the French population. We investigated readthrough efficiency in response to 10 mg/kg once-daily intravenous gentamicin perfusions in patients with and without stop mutations. Respiratory function, sweat chloride concentration, nasal potential difference (NPD) and CFTR expression in nasal epithelial cells were measured at baseline and after 15 days of treatment.
Results:
After in vitro gentamicin incubation, the readthrough efficiency for the Y122X mutation was at least five times higher than that for G542X, R1162X, and W1282X. In six of the nine patients with the Y122X mutation, CFTR immunodetection showed protein at the membrane of the nasal epithelial cells and the CFTR-dependent Cl- secretion in NPD measurements increased significantly. Respiratory status also improved in these patients, irrespective of the gentamicin sensitivity of the bacteria present in the sputum. Mean sweat chloride concentration decreased significantly and normalised in two patients. Clinical status, NPD and sweat Cl- values did not change in the Y122X patients with no protein expression, in patients with the other stop mutations investigated in vitro and those without stop mutations.
Conclusion:
Suppression of stop mutations in the CFTR gene with parenteral gentamicin can be predicted in vitro and is associated with clinical benefit and significant modification of the CFTR-mediated Cl- transport in nasal and sweat gland epithelium.
Insights
Intravenous gentamicin can suppress cystic fibrosis transmembrane conductance regulator (CFTR) stop mutations, improving patient respiratory function and chloride transport. This treatment shows clinical benefit for specific CFTR mutations, particularly Y122X.
Area of Science:
- Genetics and Molecular Biology
- Pulmonology
- Pharmacology
Background:
- Cystic fibrosis (CF) results from mutations in the CFTR gene, with F508del being most common and stop mutations affecting ~10% of patients.
- Aminoglycoside antibiotics like gentamicin can suppress premature stop codons in vitro, offering a potential therapeutic strategy for CF.
Purpose of the Study:
- To evaluate if intravenous gentamicin can suppress CFTR stop mutations in CF patients.
- To assess the clinical benefits of gentamicin-induced stop codon suppression in CF patients.
Main Methods:
- A dual gene reporter system assessed gentamicin's readthrough efficiency for CFTR stop mutations in vitro.
- Intravenous gentamicin (10 mg/kg/day) was administered to CF patients with and without stop mutations.
- Key parameters measured included respiratory function, sweat chloride, nasal potential difference (NPD), and CFTR expression.
Main Results:
- Gentamicin demonstrated higher in vitro readthrough for Y122X than other common CFTR stop mutations.
- Six of nine Y122X patients showed CFTR protein at the cell membrane, increased CFTR-dependent Cl- secretion, improved respiratory status, and reduced sweat chloride.
- No significant changes were observed in patients lacking protein expression, those with other stop mutations, or those without stop mutations.
Conclusions:
- Parenteral gentamicin can suppress CFTR stop mutations, as predicted by in vitro studies.
- This suppression is linked to clinical benefits and improved CFTR-mediated chloride transport in CF patients.
More Related Videos
 07:04
Forskolin-induced Swelling in Intestinal Organoids: An In Vitro Assay for Assessing Drug Response in Cystic Fibrosis Patients
Published on: February 11, 2017
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
Related Concept Videos
 01:15
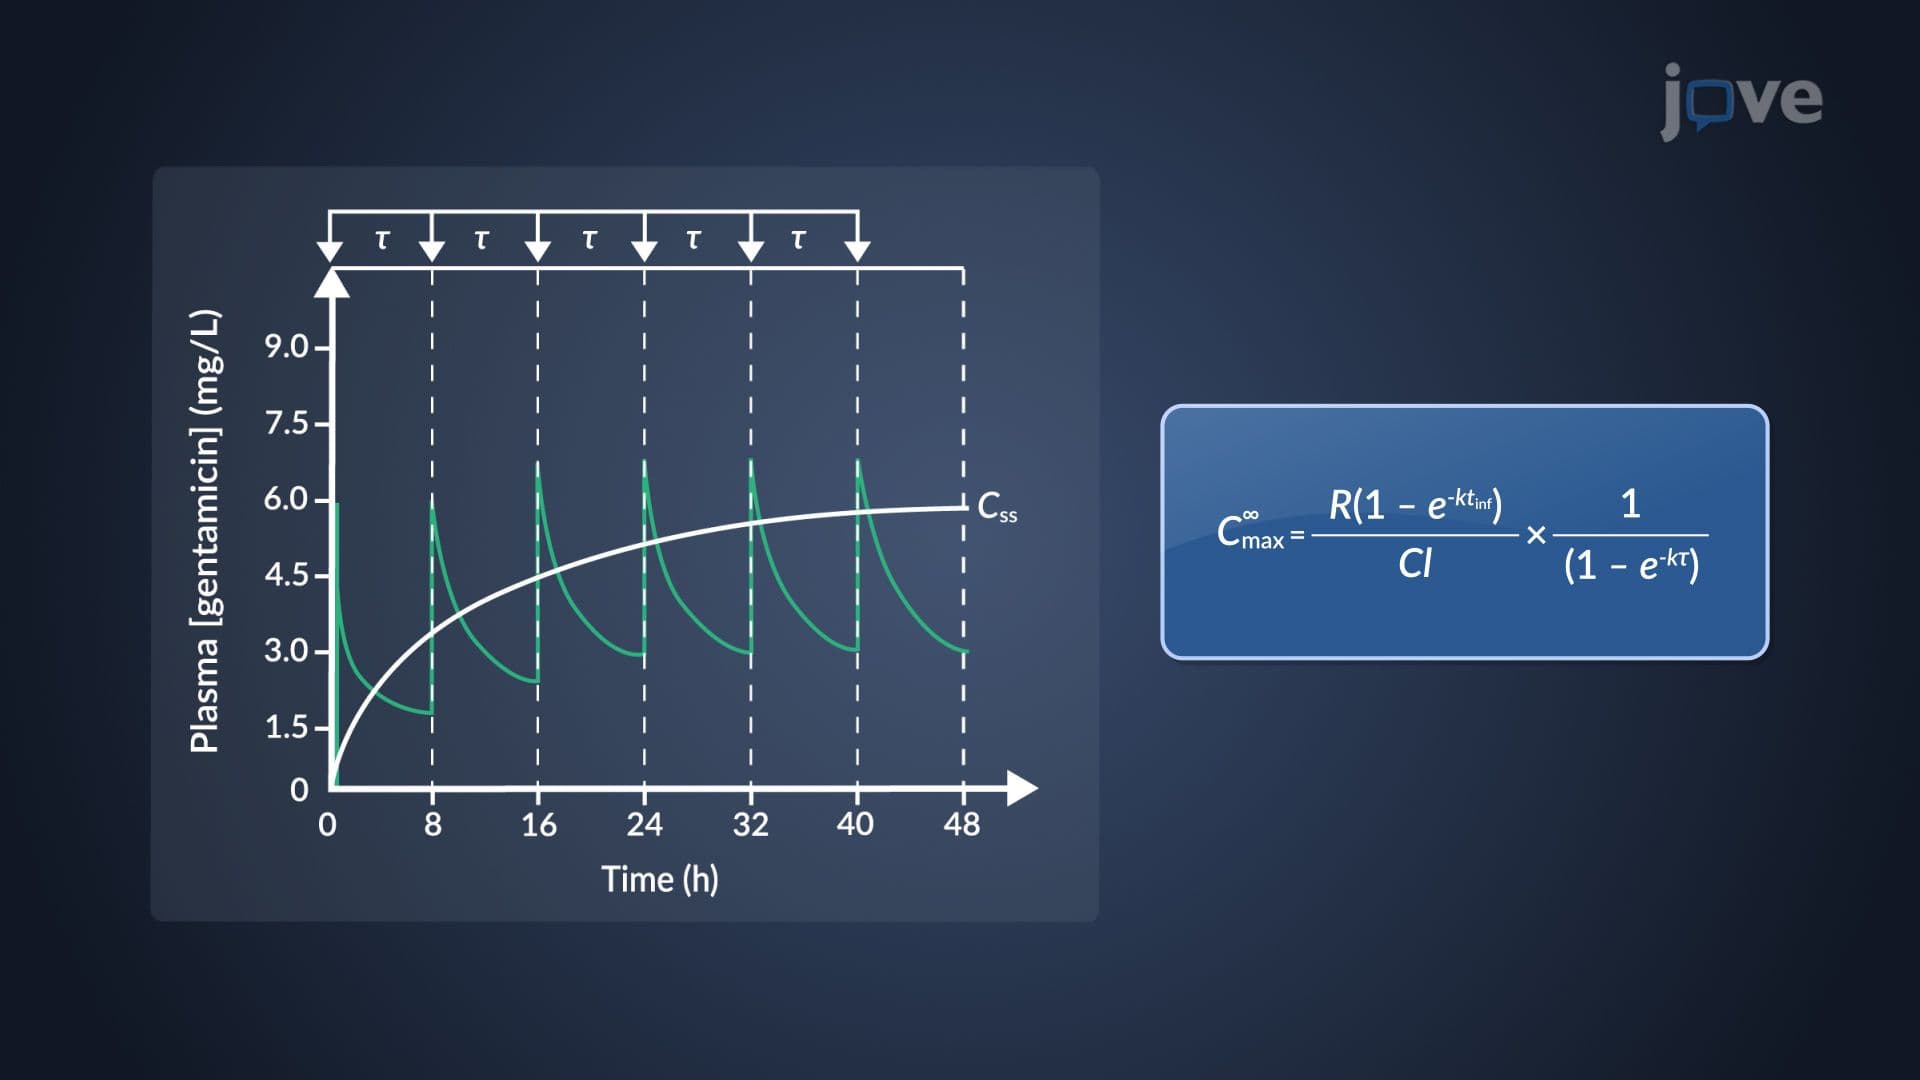
Determination of Multiple Dosing Parameters: Steady-State, Minimum and Maximum Concentrations
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...