Related Experiment Video
Updated: Jul 15, 2026
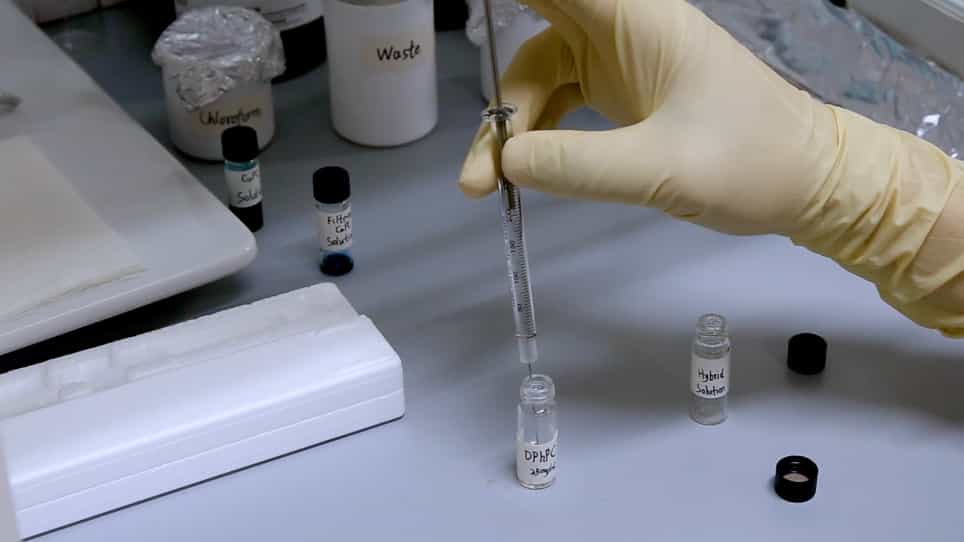
Self-Assembly of Hybrid Lipid Membranes Doped with Hydrophobic Organic Molecules at the Water/Air Interface
Published on: May 1, 2020
Electronic structure of copper phthalocyanine: an experimental and theoretical study of occupied and unoccupied
Fabrizio Evangelista1, Vincenzo Carravetta, Giovanni Stefani
1Dipartimento di Fisica, Università Roma Tre, via della Vasca Navale 84, I-00146 Roma, Italy.

Abstract:
An experimental and theoretical study of the electronic structure of copper phthalocyanine (CuPc) molecule is presented. We performed x-ray photoemission spectroscopy (XPS) and photoabsorption [x-ray absorption near-edge structure (XANES)] gas phase experiments and we compared the results with self-consistent field, density functional theory (DFT), and static-exchange theoretical calculations. In addition, ultraviolet photoelectron spectra (UPS) allowed disentangling several outer molecular orbitals. A detailed study of the two highest occupied orbitals (having a(1u) and b(1g) symmetries) is presented: the high energy resolution available for UPS measurements allowed resolving an extra feature assigned to vibrational stretching in the pyrrole rings. This observation, together with the computed DFT electron density distributions of the outer valence orbitals, suggests that the a(1u) orbital (the highest occupied molecular orbital) is mainly localized on the carbon atoms of pyrrole rings and it is doubly occupied, while the b(1g) orbital, singly occupied, is mainly localized on the Cu atom. Ab initio calculations of XPS and XANES spectra at carbon K edge of CuPc are also presented. The comparison between experiment and theory revealed that, in spite of being formally not equivalent, carbon atoms of the benzene rings experience a similar electronic environment. Carbon K-edge absorption spectra were interpreted in terms of different contributions coming from chemically shifted C 1s orbitals of the nonequivalent carbon atoms on the inner ring of the molecule formed by the sequence of CN bonds and on the benzene rings, respectively, and also in terms of different electronic distributions of the excited lowest unoccupied molecular orbital (LUMO) and LUMO+1. In particular, the degenerate LUMO appears to be mostly localized on the inner pyrrole ring.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...