Related Experiment Video
Updated: Jul 15, 2026

Interactive Molecular Model Assembly with 3D Printing
Published on: August 13, 2020
Correlated ab initio quantum chemical calculations of di- and trisaccharide conformations
Ian R Gould1, Hoda Abdel-Aal Bettley, Richard A Bryce
1Department of Chemistry, Imperial College London, London SW7 2AZ, United Kingdom. ian.gould@ic.ac.uk

Abstract:
High level correlated quantum chemical calculations, using MP2 and local MP2 theory, have been performed for conformations of the disaccharide, beta-maltose, and the trisaccharide, 3,6-di-O-(alpha-D-mannopyranosyl)-alpha-D-mannopyranose. For beta-maltose, MP2 and local MP2 calculations using the 6-311++G** basis set are in good agreement, predicting a global minimum gas-phase conformation with a counterclockwise hydrogen bond network and the experimentally-observed intersaccharide hydrogen bonding arrangement. For conformations of 3,6-di-O-(alpha-D-mannopyranosyl)-alpha-D-mannopyranose, MP2/6-311++G**, and local MP2/6-311++G** calculations do not provide a consensus prediction of relative energetics, with the MP2 method finding large differences in stability between extended and folded trisaccharide conformations. Local MP2 calculations, less susceptible to intramolecular basis set superposition errors, predict a narrower range of trisaccharide energetics, in line with estimates from Hartree-Fock theory and B3LYP and BP86 density functionals. All levels of theory predict compact, highly hydrogen-bonded conformations as lowest in energy on the in vacuo potential energy surface of the trisaccharide. These high level, correlated local MP2/6-311++G** calculations of di- and trisaccharide energetics constitute potential reference data in the development and testing of improved empirical and semiempirical potentials for modeling of carbohydrates in the condensed phase.
More Related Videos

Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 02:18
Fischer Projections
 02:27
Predicting Molecular Geometry
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:18
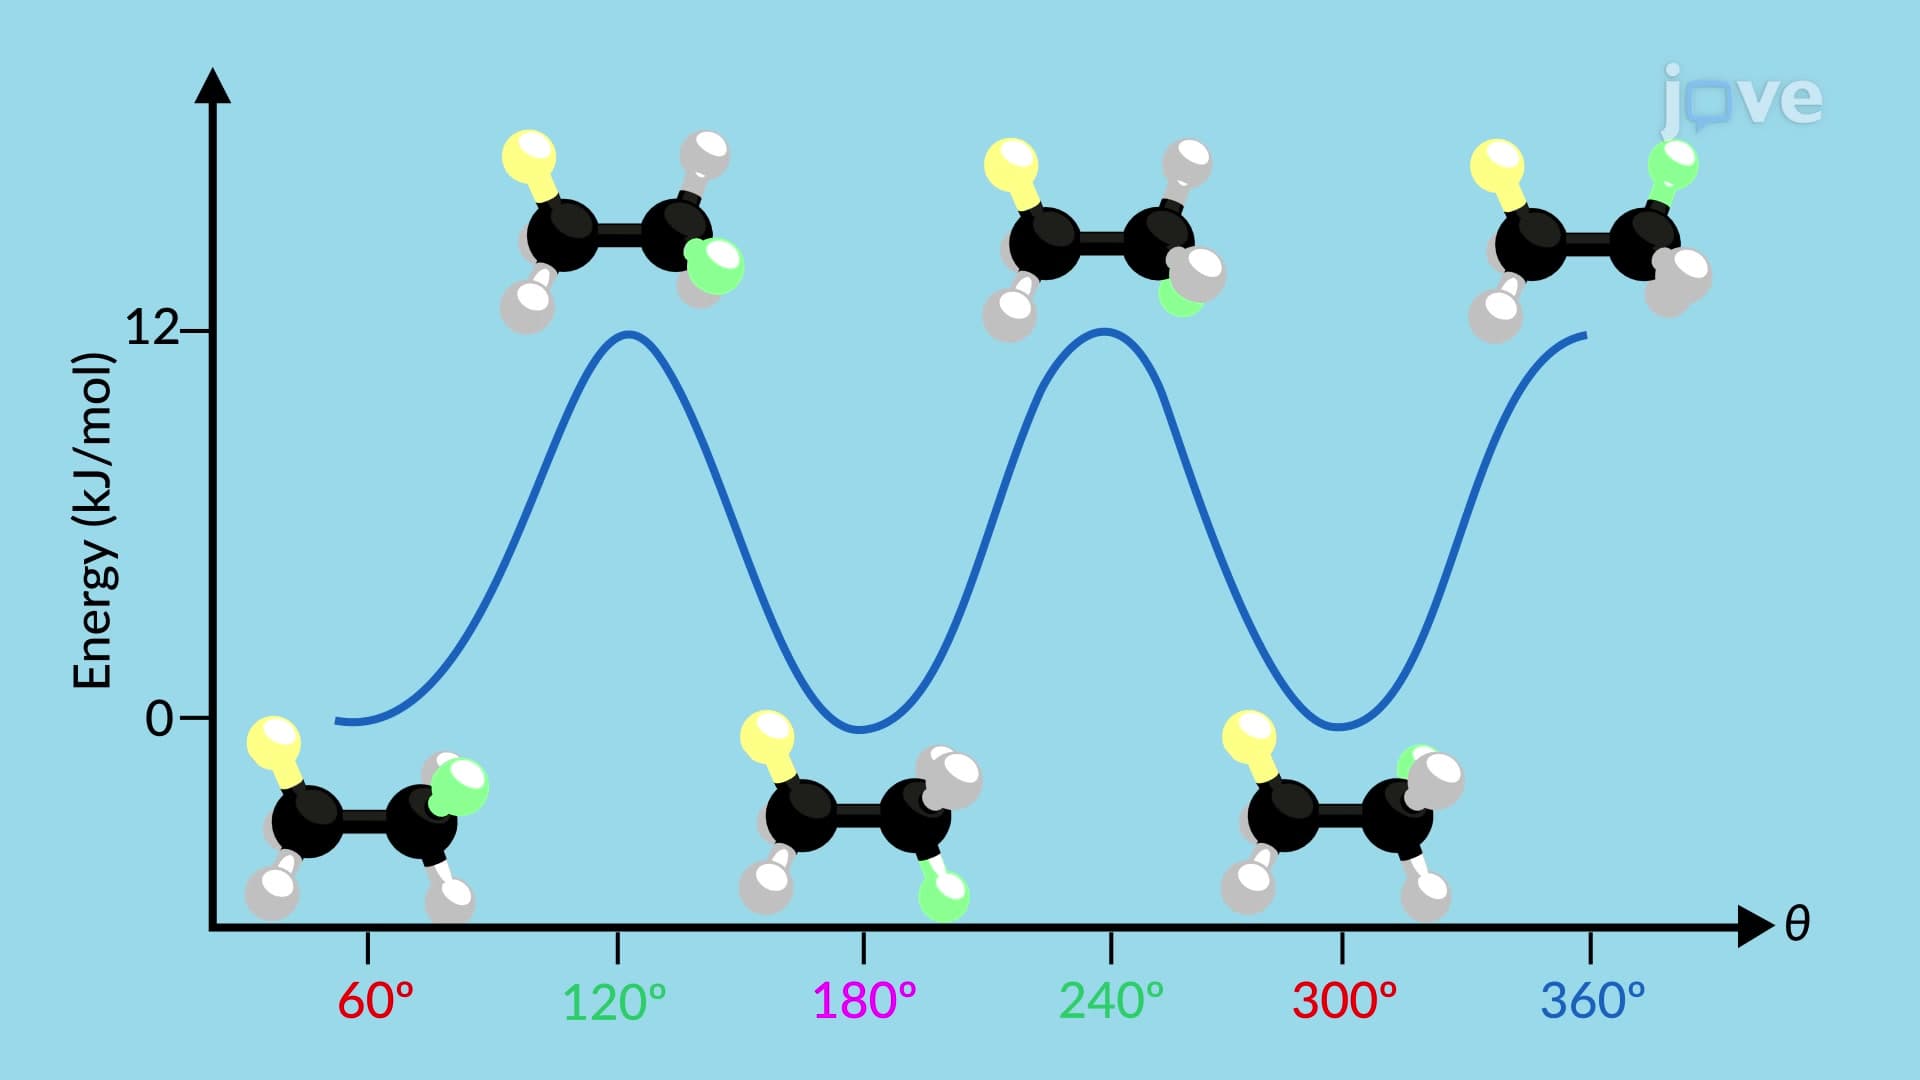
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...