Description of core excitations by time-dependent density functional theory with local density approximation, generalized gradient approximation, meta-generalized gradient approximation, and hybrid functionals
Related Experiment Video
Updated: Jul 15, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Yutaka Imamura1, Takao Otsuka, Hiromi Nakai
1Department of Chemistry, School of Science and Engineering, Waseda University, Tokyo 169-8555, Japan.

Abstract:
Time-dependent density functional theory (TDDFT) is employed to investigate exchange-correlation-functional dependence of the vertical core-excitation energies of several molecules including H, C, N, O, and F atoms. For the local density approximation (LDA), generalized gradient approximation (GGA), and meta-GGA, the calculated X1s-->pi* excitation energies (X = C, N, O, and F) are severely underestimated by more than 13 eV. On the other hand, time-dependent Hartree-Fock (TDHF) overestimates the excitation energies by more than 6 eV. The hybrid functionals perform better than pure TDDFT because HF exchange remedies the underestimation of pure TDDFT. Among these hybrid functionals, the Becke-Half-and-Half-Lee-Yang-Parr (BHHLYP) functional including 50% HF exchange provides the smallest error for core excitations. We have also discovered the systematic trend that the deviations of TDHF and TDDFT with the LDA, GGA, and meta-GGA functionals show a strong atom-dependence. Namely, their deviations become larger for heavier atoms, while the hybrid functionals are significantly less atom-dependent.

Related Concept Videos
 02:38
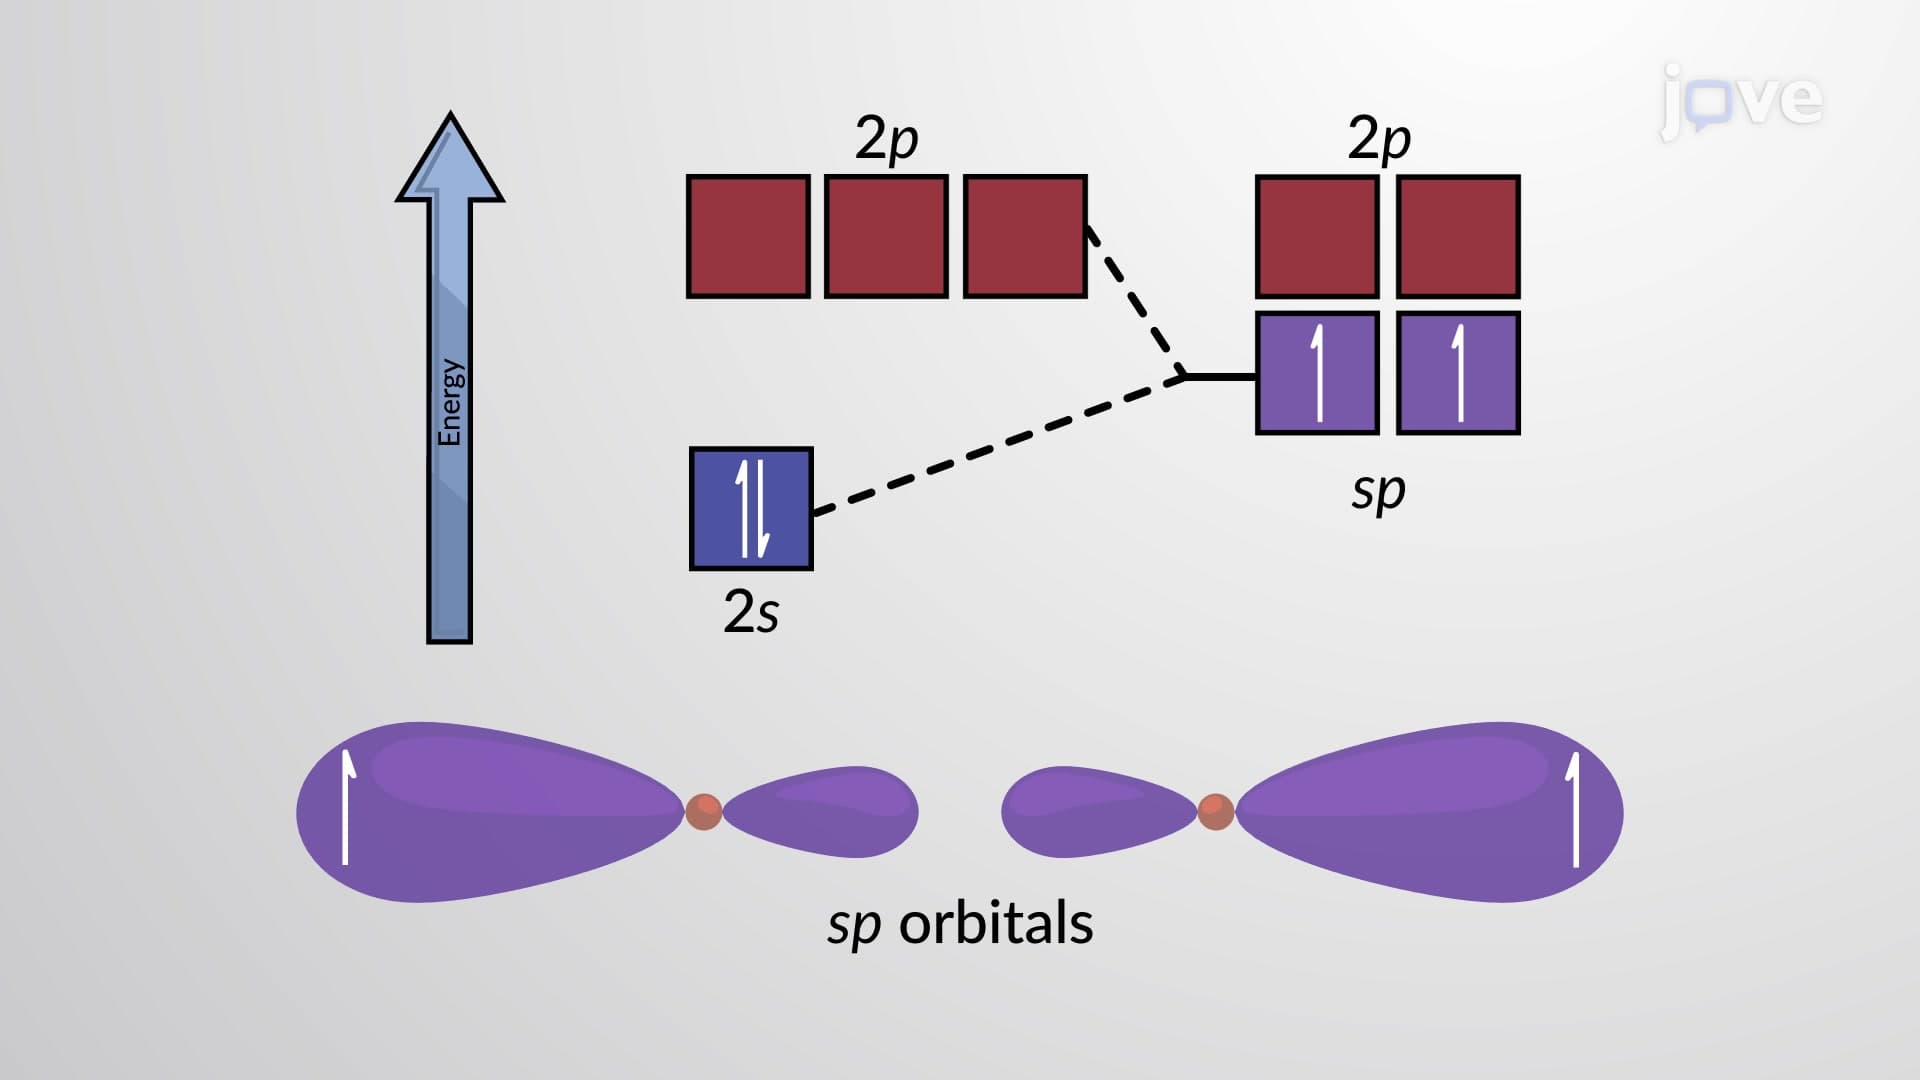
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:21
Energy Associated With a Charge Distribution
 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 03:24
Hybridization of Atomic Orbitals I
 02:35
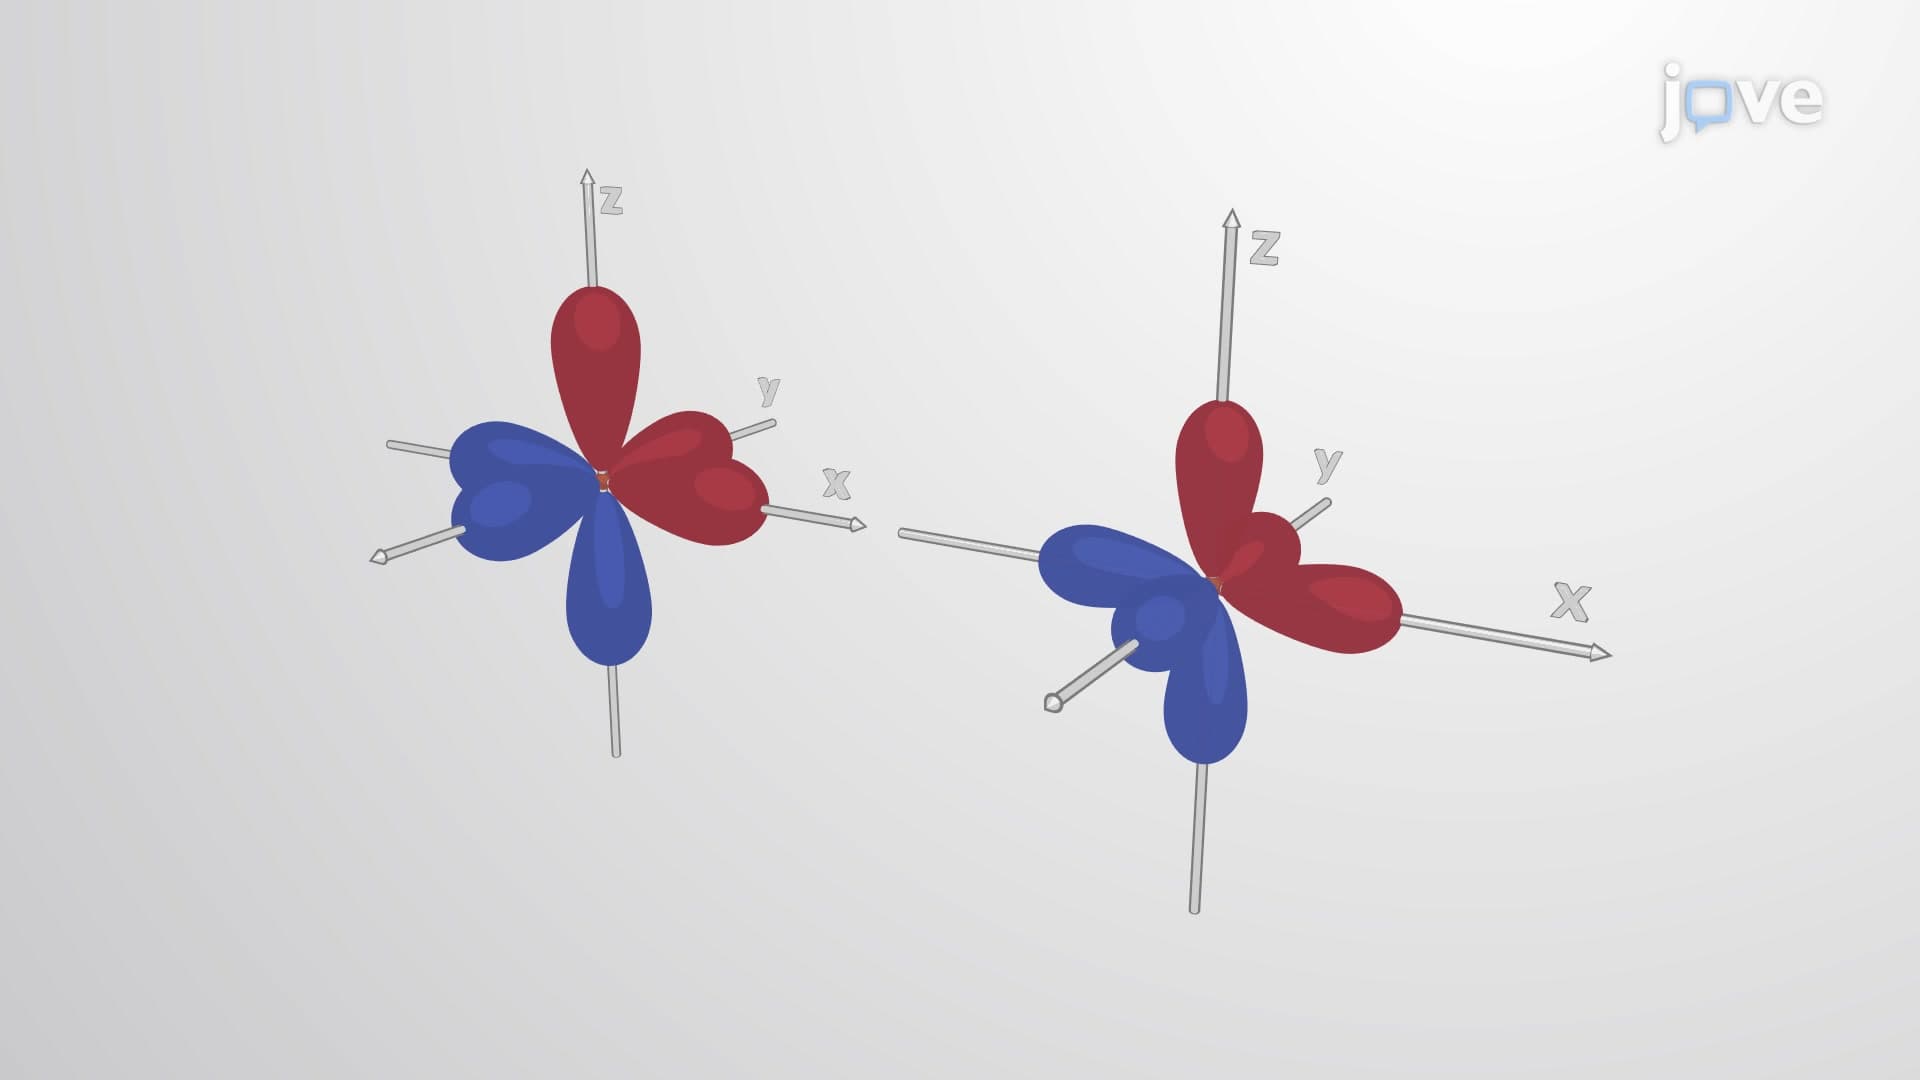
Molecular Orbital Theory I
 03:35
Hybridization of Atomic Orbitals II