Related Experiment Video
Updated: Jul 15, 2026
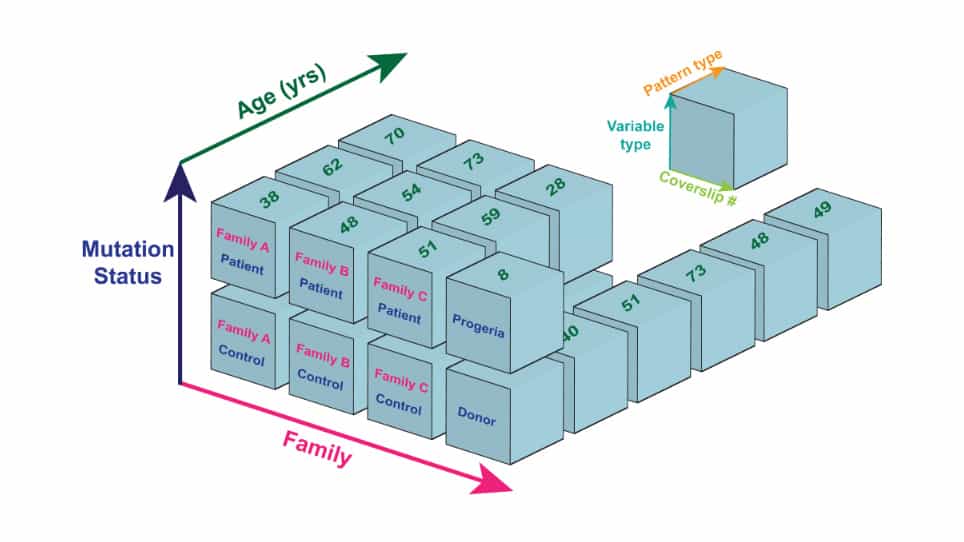

Databases to Efficiently Manage Medium Sized, Low Velocity, Multidimensional Data in Tissue Engineering
Published on: November 22, 2019
Hutchinson-Gilford progeria syndrome with severe skin calcinosis
S Nakamura1, Y Makita, A Takagi
1Department of Dermatology, Asahikawa Medical College, Asahikawa, and Takagi Dermatological Clinic, Nishi, Hokkaido, Japan. namu@asahikawa-med.ac.jp

Insights
This case study details Hutchinson-Gilford progeria syndrome (HGPS) in a young girl, highlighting extensive skin calcinosis and effective insulin-like growth factor treatment for chronic ulceration.
Area of Science:
- Genetics and Molecular Biology
- Dermatology
- Pediatric Endocrinology
Background:
- Hutchinson-Gilford progeria syndrome (HGPS) is a rare genetic disorder characterized by premature aging.
- The LMNA gene mutation is the primary cause of HGPS, affecting nuclear lamin A/C protein structure.
- Long-term follow-up studies are crucial for understanding the multifaceted clinical manifestations of HGPS.
Observation:
- A pediatric patient presented with sclerodermatous skin changes at one month of age, progressing to typical HGPS symptoms.
- Extensive, body-wide skin calcinosis was a notable observation, appearing to be underreported in existing literature.
- The patient developed a chronic ulceration following surgery for a skull fracture and subdural hematoma at age 16.
Findings:
- The patient harbored the G608G LMNA gene mutation, leading to a cryptic splice site and a truncated lamin A/C protein.
- Wet dressing with insulin-like growth factor demonstrated significant efficacy in treating the chronic ulceration.
- The case underscores the diverse dermatological and systemic complications associated with HGPS.
Implications:
- This case expands the understanding of HGPS clinical variability, particularly regarding extensive skin calcinosis.
- The successful application of insulin-like growth factor highlights a potential therapeutic avenue for HGPS-related complications.
- Further research into LMNA gene mutations and their impact on protein function is warranted for developing targeted HGPS therapies.
Abstract:
We describe a case of Hutchinson-Gilford progeria syndrome (HGPS) with long-term follow-up. A 1-month-old girl with marked sclerodermatous skin changes developed various symptoms of HGPS during follow-up. These included sclerotic skin, pigmentation, skin atrophy with translucent veins, wispy hair and alopecia, nail dystrophy and decreased sweating. Marked skin calcinosis was observed over almost the entire body, a symptom that has apparently been ignored in the literature. At 16 years old, the girl underwent surgery for a skull fracture and subdural haematoma, which was followed by chronic ulceration. Wet dressing with insulin-like growth factor was used with considerable effect. Mutation of the lamin A/C (LMNA) gene mutation, which encodes nuclear lamin A and C, has been reported to be the cause of HGPS. Our case showed the mutation G608G (GGC-->GGT), which resulted in a cryptic splice site and consequently in a truncated lamin A/C protein.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 01:33
Pleiotropy
 01:23
Skin Diseases and Disorders
Gram-positive Staphylococcus spp. and Streptococcus spp. are responsible for many of the most common skin infections. However, many...
 01:20
Inborn Errors of Metabolism
 01:14
Changes in Skin Color: Clinical Perspectives
Albinism
Albinism is a genetic disorder that affects (completely or partially) the coloring of skin, hair, and eyes. The defect is primarily...