A self-contained and portable density functional theory library for use in Ab Initio quantum chemistry programs
Related Experiment Video
Updated: Jul 15, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Paweł Sałek1, Andreas Hesselmann
1Laboratory of Theoretical Chemistry, The Royal Institute of Technology, Stockholm, Sweden. pawsa@theochem.kth.se

Abstract:
A major unresolved problem of density functional theory is the yet unknown exchange-correlation functional, which leads to a proliferation of its less or more successful approximations. A practical implementation of these numerous functionals can present a substantial challenge particularly if the higher order functional derivatives are required. We present a systematic method of functional implementation. The method allows a clean handling of a large number of functionals in a mutually independent way. We developed an extensive set of automatic test routines to facilitate functional and derivative testing with respect to the implementation correctness and numerical stability. An integral part of the presented solution is a program for automatic code generation from analytical formulas that uses only freely available tools. Code for evaluation of functionals and their first, second, third, and fourth derivatives can be generated, which accelerates the development, implementation, and testing of new functionals.

Related Concept Videos
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:40
MO Theory and Covalent Bonding
 02:35
Molecular Orbital Theory I
 01:26
Thermodynamic Potentials
 02:38
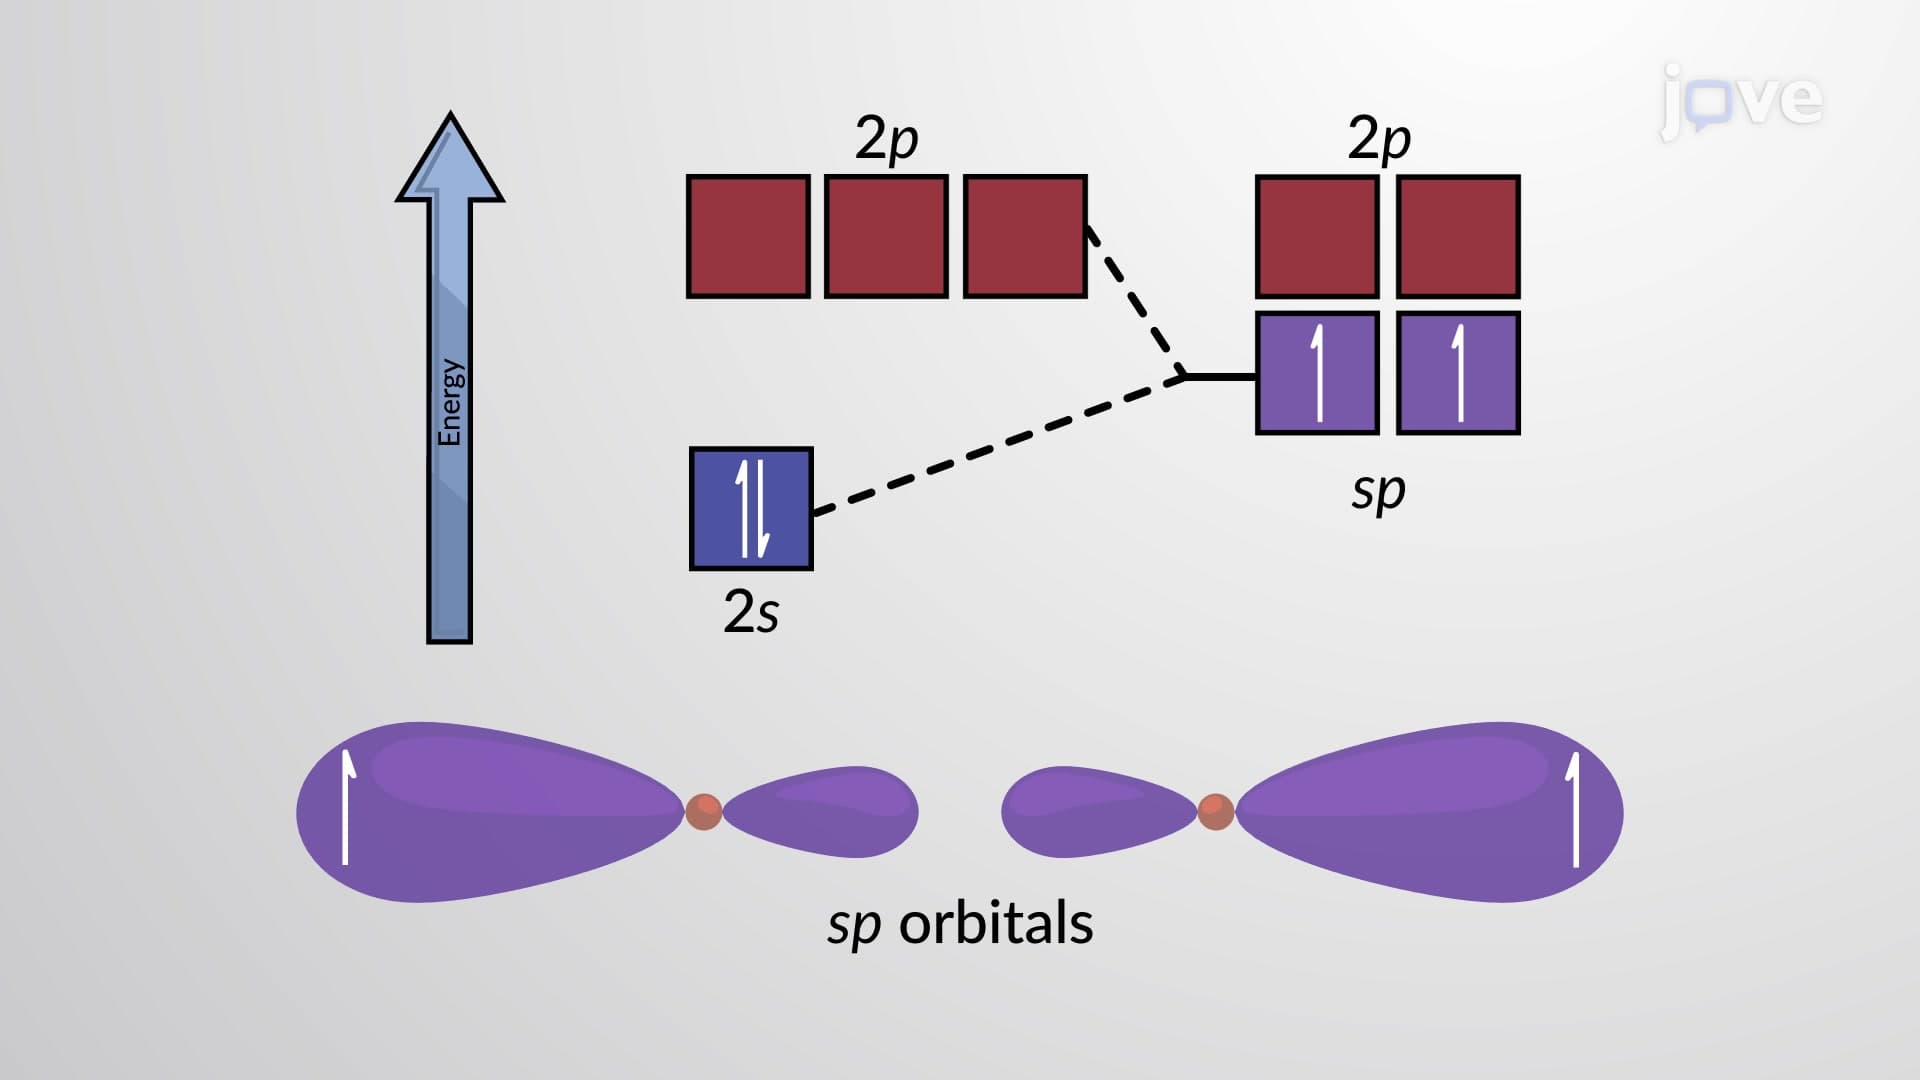
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...