Related Experiment Video
Updated: Jul 14, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Five-dimensional ab initio potential energy surface and predicted infrared spectra of H2-CO2 van der Waals complexes
Hong Ran1, Yanzi Zhou, Daiqian Xie
1Institute of Theoretical and Computational Chemistry, Key Laboratory of Mesoscopic Chemistry, School of Chemistry and Chemical Engineering, Nanjing University, Nanjing 210093, China.

Abstract:
The authors present a new five-dimensional potential energy surface for H2-CO2 including the Q3 normal mode for the nu3 antisymmetric stretching vibration of the CO2 molecule. The potential energies were calculated using the supermolecular approach with the full counterpoise correction at the CCSD(T) level with an aug-cc-pVTZ basis set supplemented with bond functions. The global minimum is at two equivalent T-shaped coplanar configurations with a well depth of 219.68 cm-1. The rovibrational energy levels for four species of H2-CO2 (paraH2-, orthoH2-, paraD2-, and orthoD2-CO2) were calculated employing the discrete variable representation (DVR) for radial variables and finite basis representation (FBR) for angular variables and the Lanczos algorithm. Our calculations showed that the off-diagonal intra- and intermolecular vibrational coupling could be neglected, and separation of the intramolecular vibration by averaging the total Hamiltonian with the wave function of a specific vibrational state of CO2 should be a good approximation with high accuracy. The calculated band origin shift in the infrared spectra in the nu3 region of CO2 is -0.113 cm-1 for paraH2-CO2 and -0.099 cm-1 for orthoH2-CO2, which agrees well with the observed values of -0.198 and -0.096 cm-1. The calculated rovibrational spectra for H2-CO2 are consistent with the available experimental spectra. For D2-CO2, it is predicted that only a-type transitions occur for paraD2-CO2, while both a-type and b-type transitions are significant for orthoD2-CO2.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 01:24
Van der Waals Interactions
 01:10
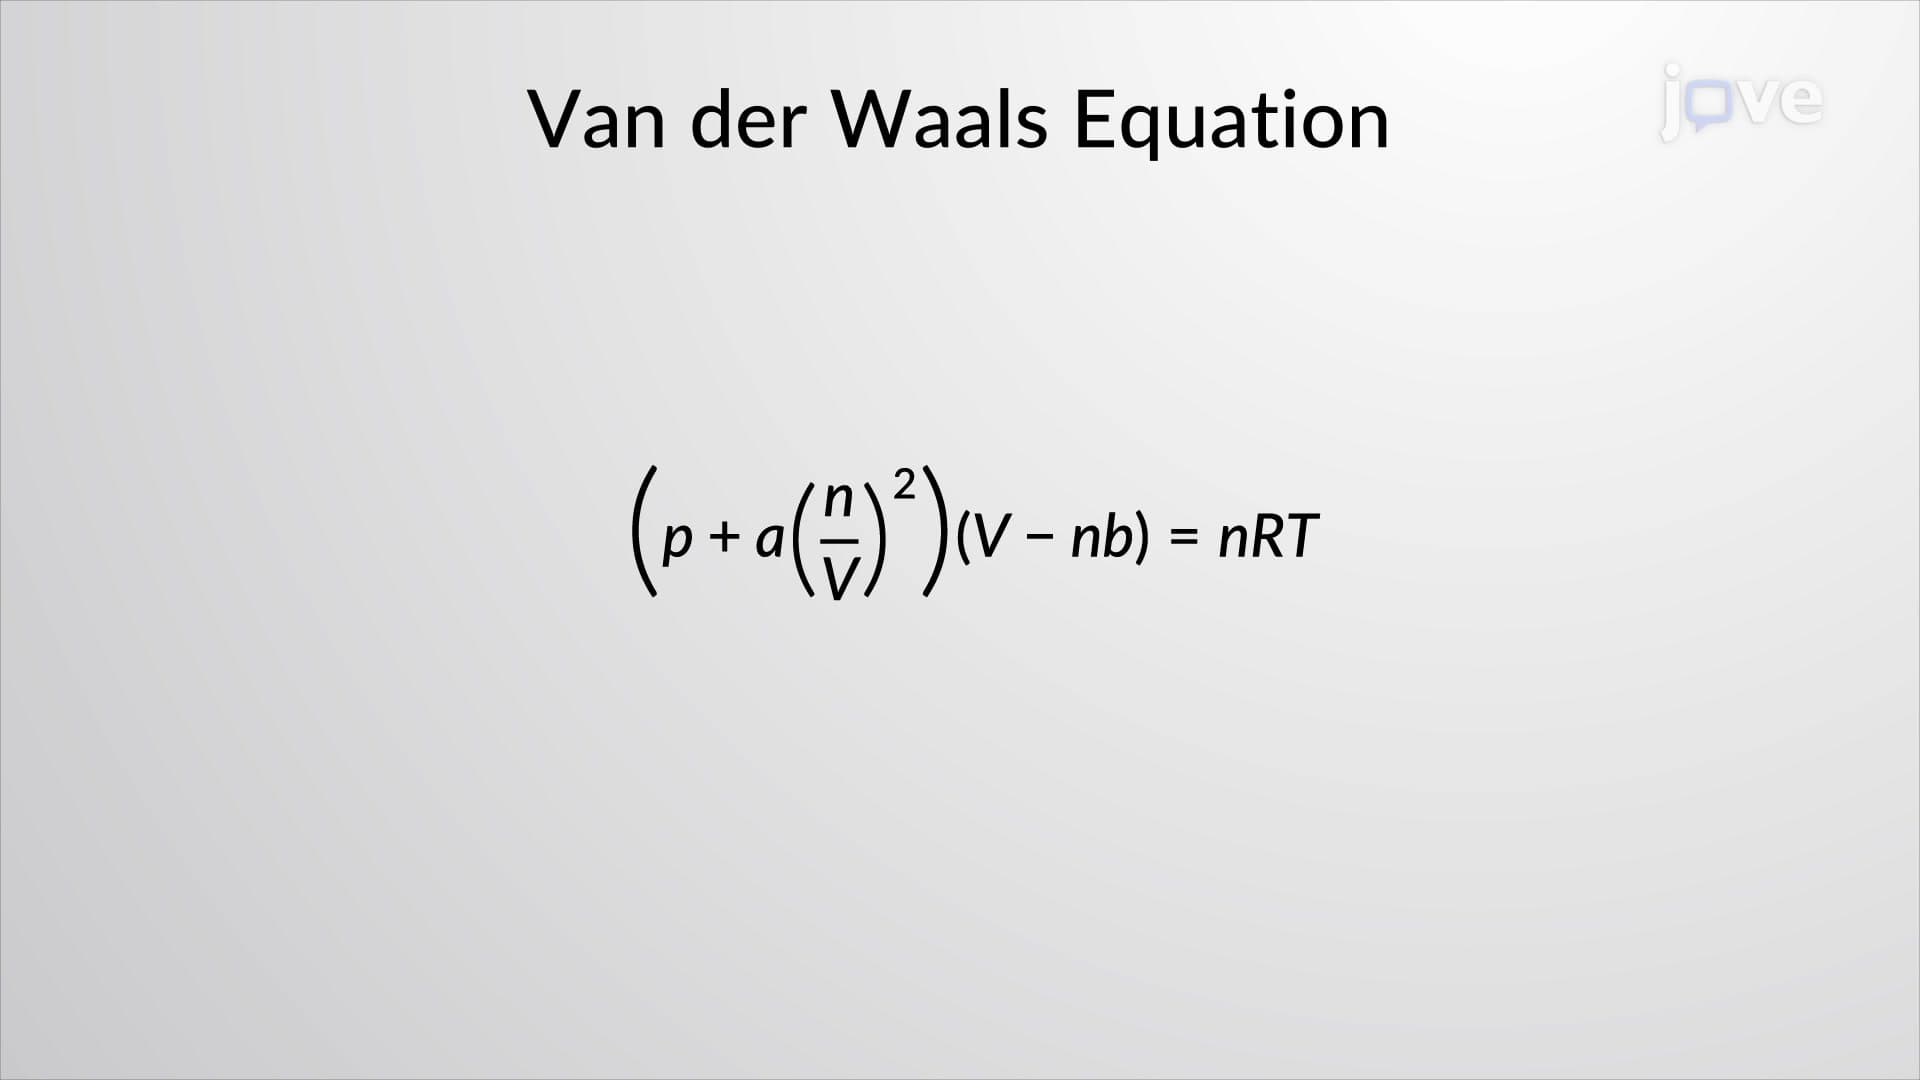
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:26
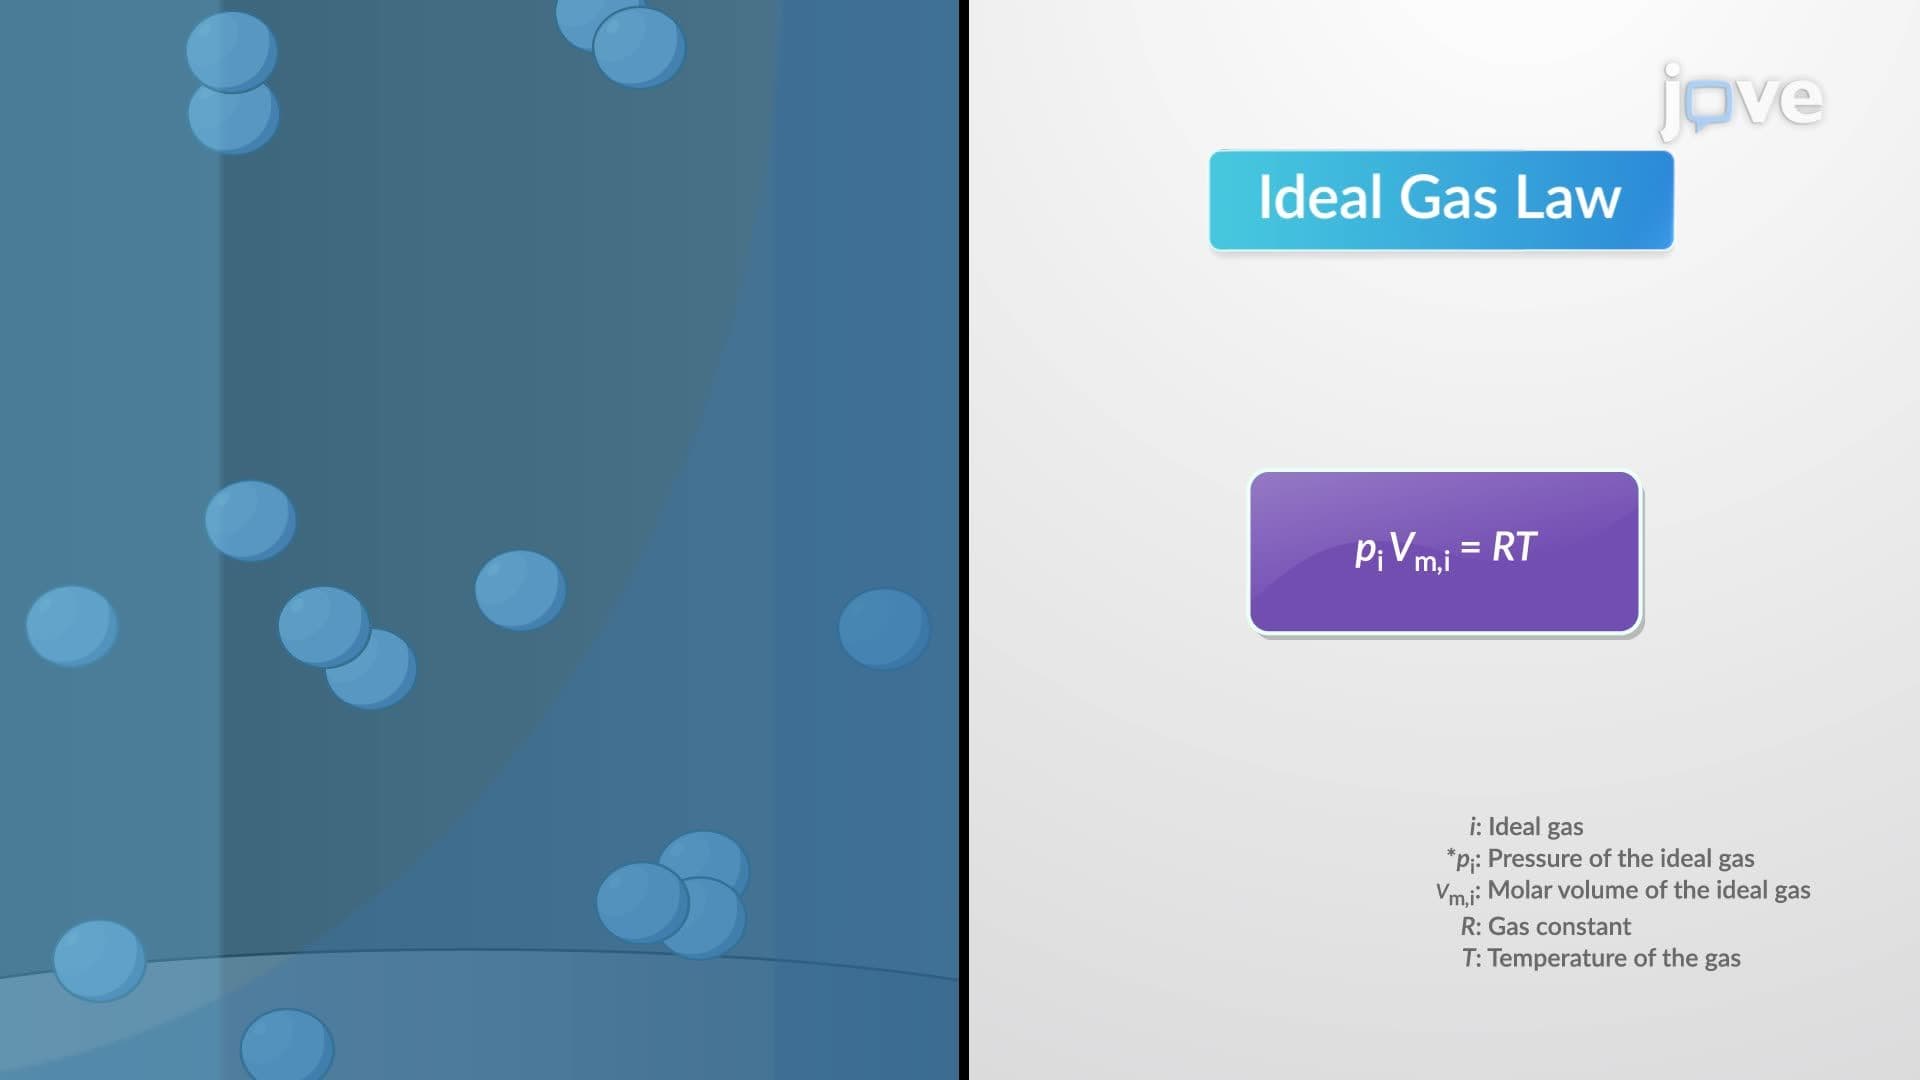
The Van der Waals Equation
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation