Related Experiment Video
Updated: Jul 14, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Efficiency of numerical basis sets for predicting the binding energies of hydrogen bonded complexes: evidence of
1Accelrys K K, Nishishinbashi TS Building, 3-3-1 Nishishinbashi, Minato-ku, Tokyo 105-0003, Japan. yinada@accelrys.com

Abstract:
Binding energies of selected hydrogen bonded complexes have been calculated within the framework of density functional theory (DFT) method to discuss the efficiency of numerical basis sets implemented in the DFT code DMol3 in comparison with Gaussian basis sets. The corrections of basis set superposition error (BSSE) are evaluated by means of counterpoise method. Two kinds of different numerical basis sets in size are examined; the size of the one is comparable to Gaussian double zeta plus polarization function basis set (DNP), and that of the other is comparable to triple zeta plus double polarization functions basis set (TNDP). We have confirmed that the magnitudes of BSSE in these numerical basis sets are comparative to or smaller than those in Gaussian basis sets whose sizes are much larger than the corresponding numerical basis sets; the BSSE corrections in DNP are less than those in the Gaussian 6-311+G(3df,2pd) basis set, and those in TNDP are comparable to those in the substantially large scale Gaussian basis set aug-cc-pVTZ. The differences in counterpoise corrected binding energies between calculated using DNP and calculated using aug-cc-pVTZ are less than 9 kJ/mol for all of the complexes studied in the present work. The present results have shown that the cost effectiveness in the numerical basis sets in DMol3 is superior to that in Gaussian basis sets in terms of accuracy per computational cost.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 02:38
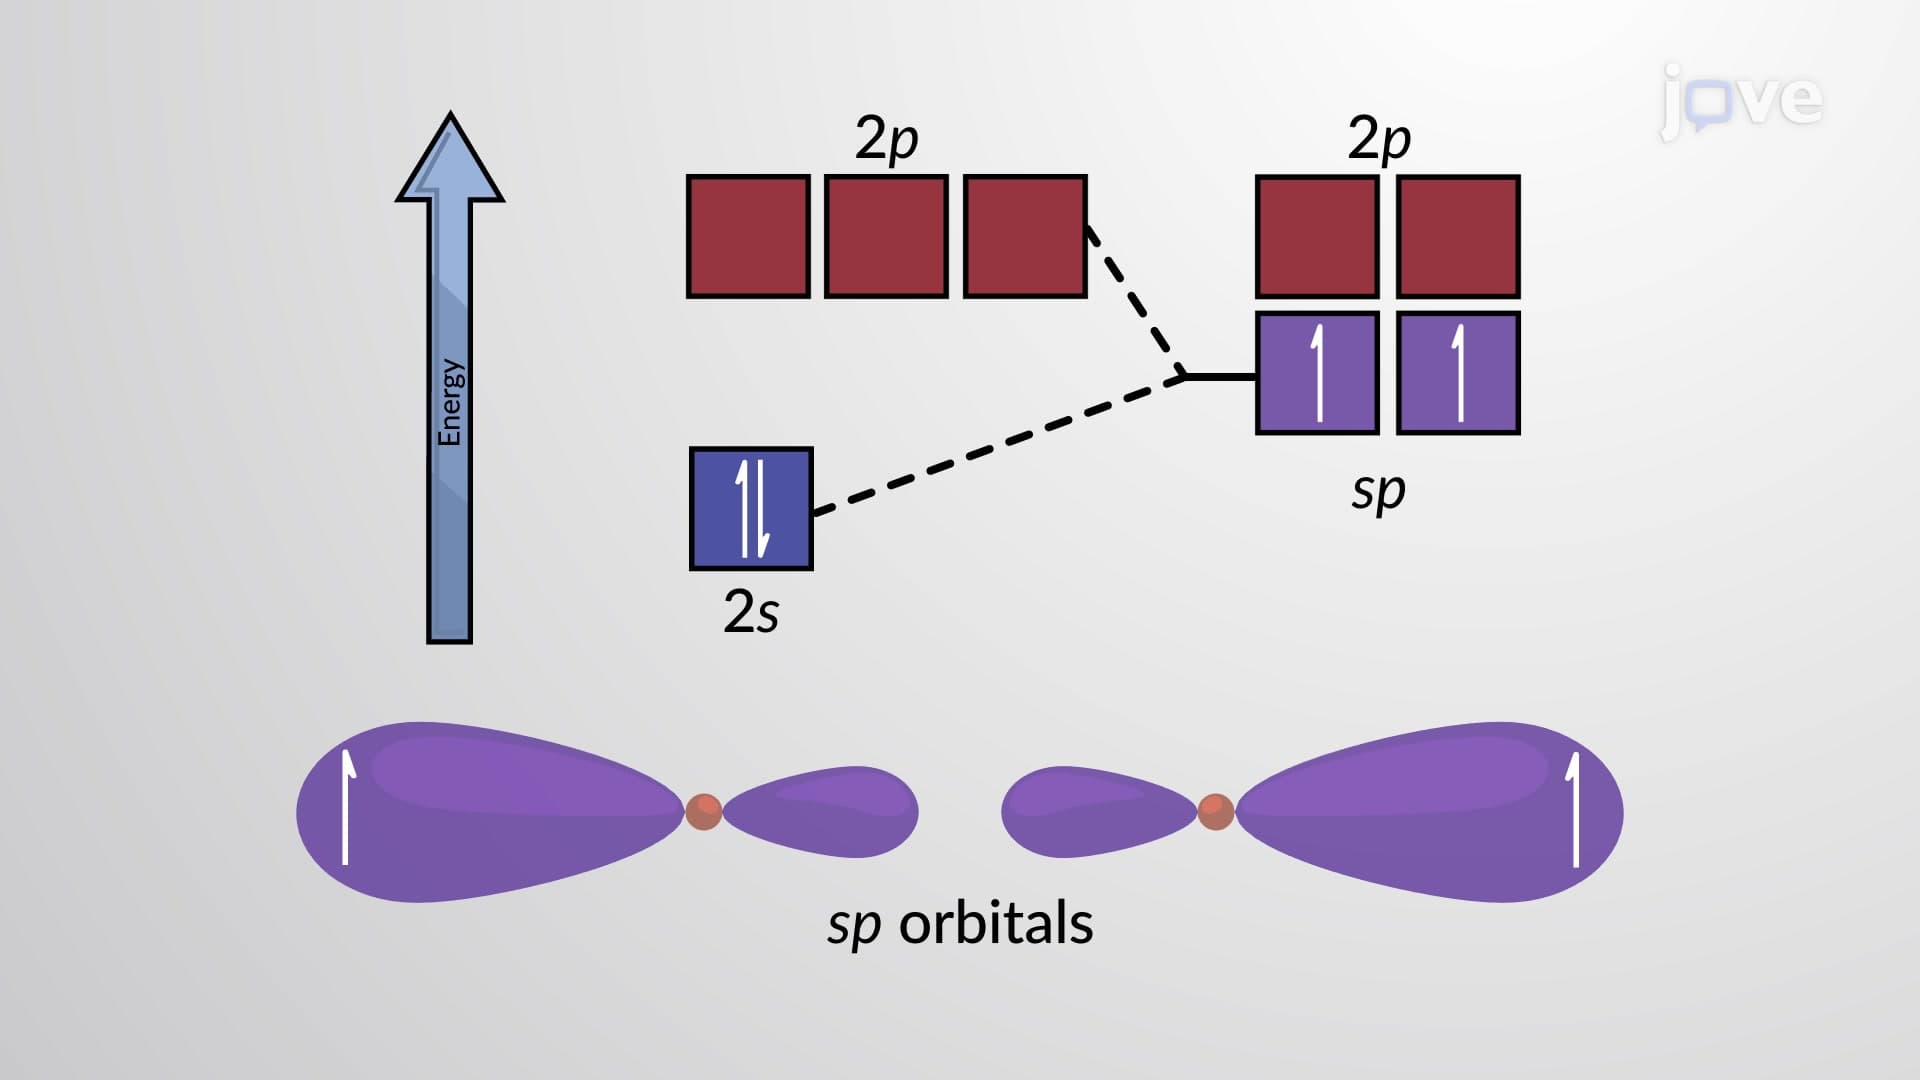
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:45
Valence Bond Theory
 02:42
Valence Bond Theory
 03:24
Hybridization of Atomic Orbitals I