Related Experiment Video
Updated: Jul 14, 2026

Preparation of SNS Cobalt(II) Pincer Model Complexes of Liver Alcohol Dehydrogenase
Published on: March 19, 2020
Ligand versus metal protonation of an iron hydrogenase active site mimic
Gerriet Eilers1, Lennart Schwartz, Matthias Stein
1Department of Photochemistry and Molecular Science, The Angström Laboratories, Uppsala University, Box 523, 75120 Uppsala, Sweden.

Abstract:
The protonation behavior of the iron hydrogenase active-site mimic [Fe2(mu-adt)(CO)4(PMe3)2] (1; adt=N-benzyl-azadithiolate) has been investigated by spectroscopic, electrochemical, and computational methods. The combination of an adt bridge and electron-donating phosphine ligands allows protonation of either the adt nitrogen to give [Fe2(mu-Hadt)(CO)4(PMe3)2]+ ([1 H]+), the Fe-Fe bond to give [Fe2(mu-adt)(mu-H)(CO)4(PMe3)2]+ ([1 Hy]+), or both sites simultaneously to give [Fe2(mu-Hadt)(mu-H)(CO)4(PMe3)2]2+ ([1 HHy]2 +). Complex 1 and its protonation products have been characterized in acetonitrile solution by IR, (1)H, and (31)P NMR spectroscopy. The solution structures of all protonation states feature a basal/basal orientation of the phosphine ligands, which contrasts with the basal/apical structure of 1 in the solid state. Density functional calculations have been performed on all protonation states and a comparison between calculated and experimental spectra confirms the structural assignments. The ligand protonated complex [1 H]+ (pKa=12) is the initial, metastable protonation product while the hydride [1 Hy]+ (pKa=15) is the thermodynamically stable singly protonated form. Tautomerization of cation [1 H]+ to [1 Hy]+ does not occur spontaneously. However, it can be catalyzed by HCl (k=2.2 m(-1) s(-1)), which results in the selective formation of cation [1 Hy]+. The protonations of the two basic sites have strong mutual effects on their basicities such that the hydride (pK(a)=8) and the ammonium proton (pK(a)=5) of the doubly protonated cationic complex [1 HHy]2+ are considerably more acidic than in the singly protonated analogues. The formation of dication [1 HHy]2+ from cation [1 H]+ is exceptionally slow with perchloric or trifluoromethanesulfonic acid (k=0.15 m(-1) s(-1)), while the dication is formed substantially faster (k>10(2) m(-1) s(-1)) with hydrobromic acid. Electrochemically, 1 undergoes irreversible reduction at -2.2 V versus ferrocene, and this potential shifts to -1.6, -1.1, and -1.0 V for the cationic complexes [1 H]+, [1 Hy]+, and [1 HHy]2+, respectively, upon protonation. The doubly protonated form [1 HHy]2+ is reduced at less negative potential than all previously reported hydrogenase models, although catalytic proton reduction at this potential is characterized by slow turnover.
More Related Videos
 10:01
Protein Film Infrared Electrochemistry Demonstrated for Study of H2 Oxidation by a [NiFe] Hydrogenase
Published on: December 4, 2017
 08:02
Benchtop Immobilized Metal Affinity Chromatography, Reconstitution and Assay of a Polyhistidine Tagged Metalloenzyme for the Undergraduate Laboratory
Published on: August 23, 2018
Related Concept Videos
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:19
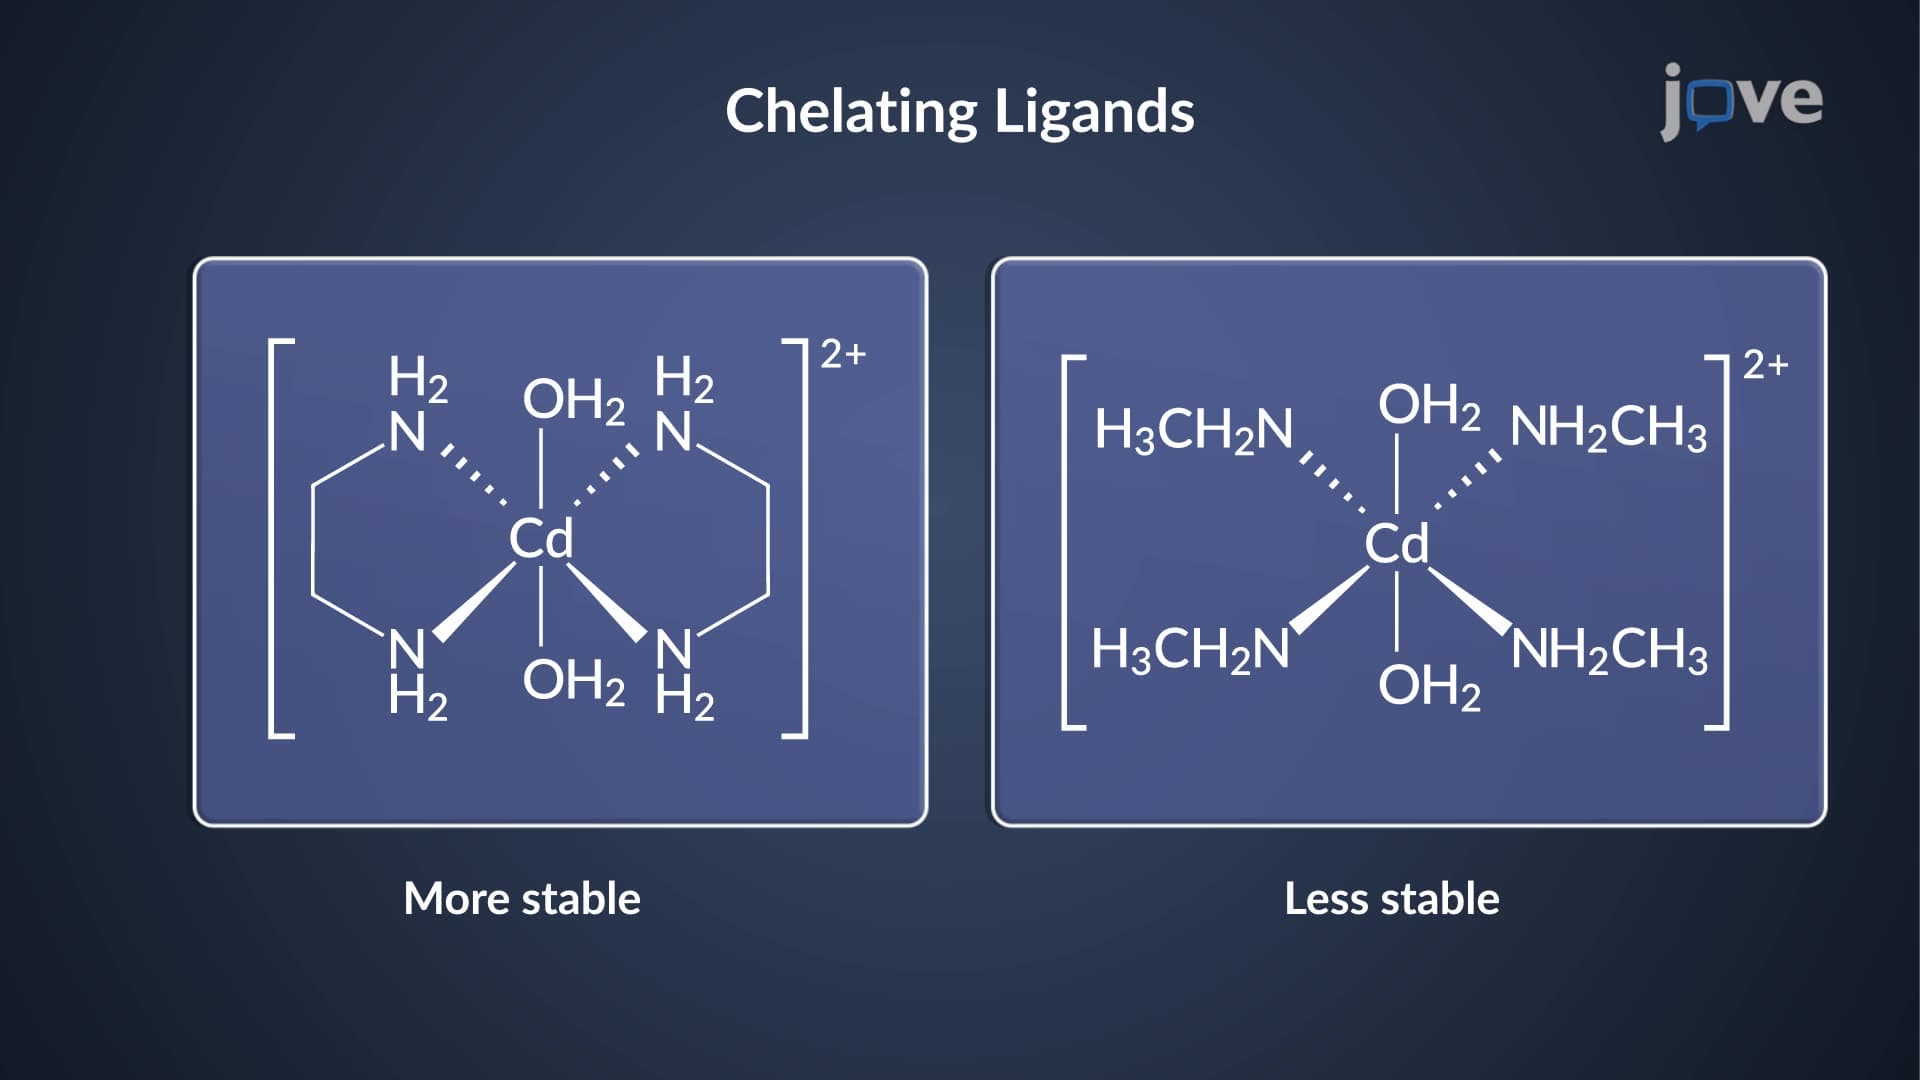
Complexation Equilibria: The Chelate Effect
 01:09
Complexation Equilibria: Factors Influencing Stability of Complexes
 00:49
Ligand Binding and Linkage