Related Experiment Video
Updated: Jul 14, 2026
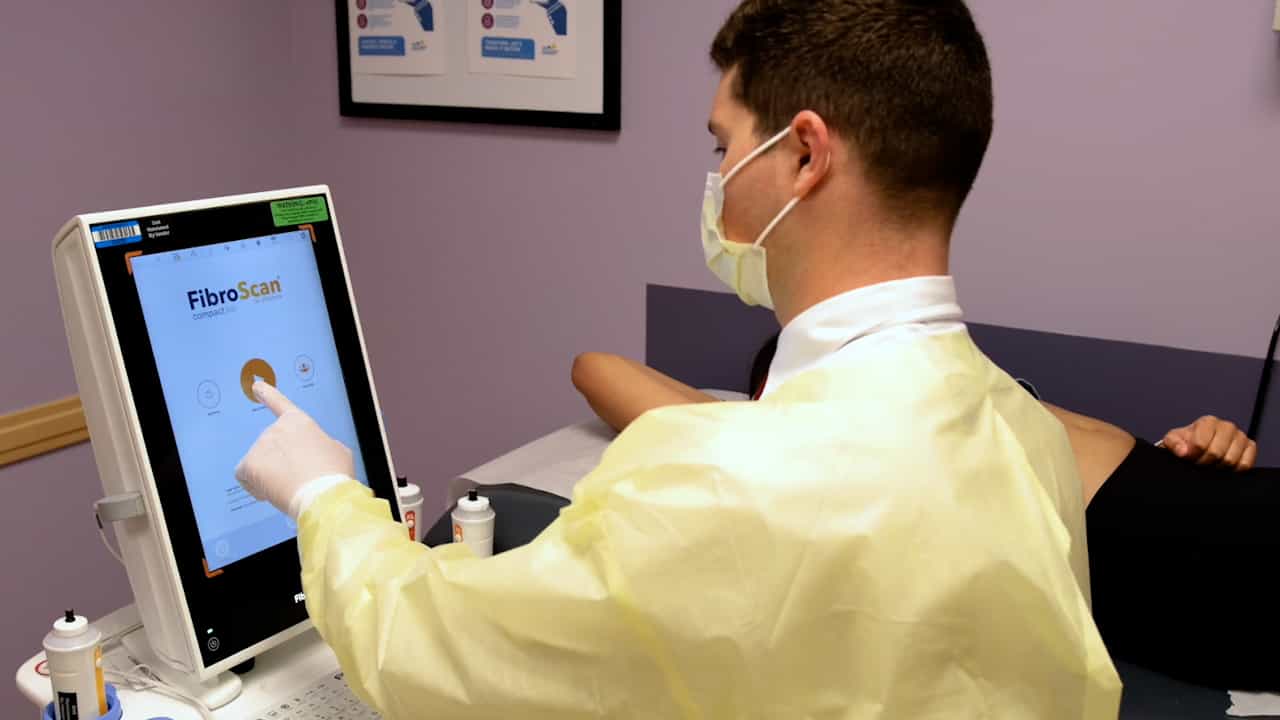

Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Does cystic fibrosis neonatal screening detect atypical CF forms? Extended genetic characterization and 4-year
L Narzi1, G Ferraguti, A Stamato
1Department of Paediatrics, Cystic Fibrosis Centre, University of Rome La Sapienza, Rome, Italy.

Abstract:
The neonatal screening protocol for cystic fibrosis (CF) is based on a first determination of blood immunoreactive trypsin (IRT1), followed by a first level genetic test that includes the 31 worldwide most common mutations of the cystic fibrosis transmembrane conductance regulator (CFTR) gene (DNA31), and a second determination of blood immunoreactive trypsin (IRT2). This approach identifies, in addition to affected subjects, a high proportion of newborns with hypertrypsinaemia at birth, in whom only one mutation is identified and who have a negative or borderline sweat test and pancreatic sufficiency. Although it has been suggested that hypertrypsinaemia may be caused by a single CFTR mutation, whether such neonates should be merely considered as healthy carriers remains a matter of debate as hypertrypsinaemia at birth may be a biochemical marker of a CFTR malfunction because of a second mild mutation. We analyzed, by means of an extended sequencing protocol, 32 newborns who tested positive at an IRT1/DNA31/IRT2 screening protocol and in whom only one CFTR mutation was found. The results obtained demonstrate that 62.5% of these newborns were also carrying a second mild CFTR mutation. The high proportion of compound heterozygous subjects, combined with the results of a 4-year follow-up in nine of these subjects all of whom displaying initial CF clinical symptoms, suggest that it may be possible to use the IRT1/DNA31/IRT2 protocol of neonatal screening to identify newborns with atypical forms of CF. In view of these findings, an extended genetic search for subjects with compound heterozygosity and a periodic clinical assessment should be considered.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 02:46
Genetic Screens
Forward genetic screens
Forward or “classical” genetic screens involve creating random mutations in an organism’s DNA using radiation, mutagens, or insertion of additional bases, which result in visible changes...
 01:29
Pharmacogenomics: Identification of New Drug Targets