Related Experiment Video
Updated: Jul 14, 2026

Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
An ab initio study of the CH3I photodissociation. I. Potential energy surfaces
Aleksey B Alekseyev1, Heinz-Peter Liebermann, Robert J Buenker
1Fachbereich C-Theoretische Chemie, Bergische Universität Wuppertal, Gaussstr. 20, D-42097 Wuppertal, Germany. alexeev@uni-wuppertal.de

Abstract:
The multireference spin-orbit (SO) configuration interaction (CI) method in its Lambda-S contracted SO-CI version is employed to calculate two-dimensional potential energy surfaces for the ground and low-lying excited states of CH3I relevant to the photodissociation process in its A absorption band. The computed equilibrium geometry for the X A1 ground state, as well as vibrational frequencies for the nu2 umbrella and nu3 symmetric stretch modes, are found to be in good agreement with available experimental data. The 3Q0+ state converging to the excited I(2P1/2o) limit is found to possess a shallow minimum of 850 cm(-1) strongly shifted to larger internuclear distances (RC-I approximately 6.5a0) relative to the ground state. This makes a commonly employed single-exponent approximation for analysis of the CH3I fragmentation dynamics unsuitable. The 4E(3A1) state dissociating to the same atomic limit is calculated to lie too high in the Franck-Condon region to have any significant impact on the A-band absorption. The computed vertical excitation energies for the 3Q1, 3Q0+, and 1Q states indicate that the A-band spectrum must lie approximately between 33,000 and 44,300 cm(-1), i.e., between 225 and 300 nm. This result is in very good agreement with the experimental findings. The lowest Rydberg states are computed to lie at >or=49,000 cm(-1) and correspond to the ...a(1)2n3a1(6sI) leading configuration. They are responsible for the vacuum ultraviolet absorption lines found experimentally beyond the A-band spectrum at 201.1 nm (49,722 cm(-1)) and higher.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 01:21
Chemical Ionization (CI) Mass Spectrometry
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:13
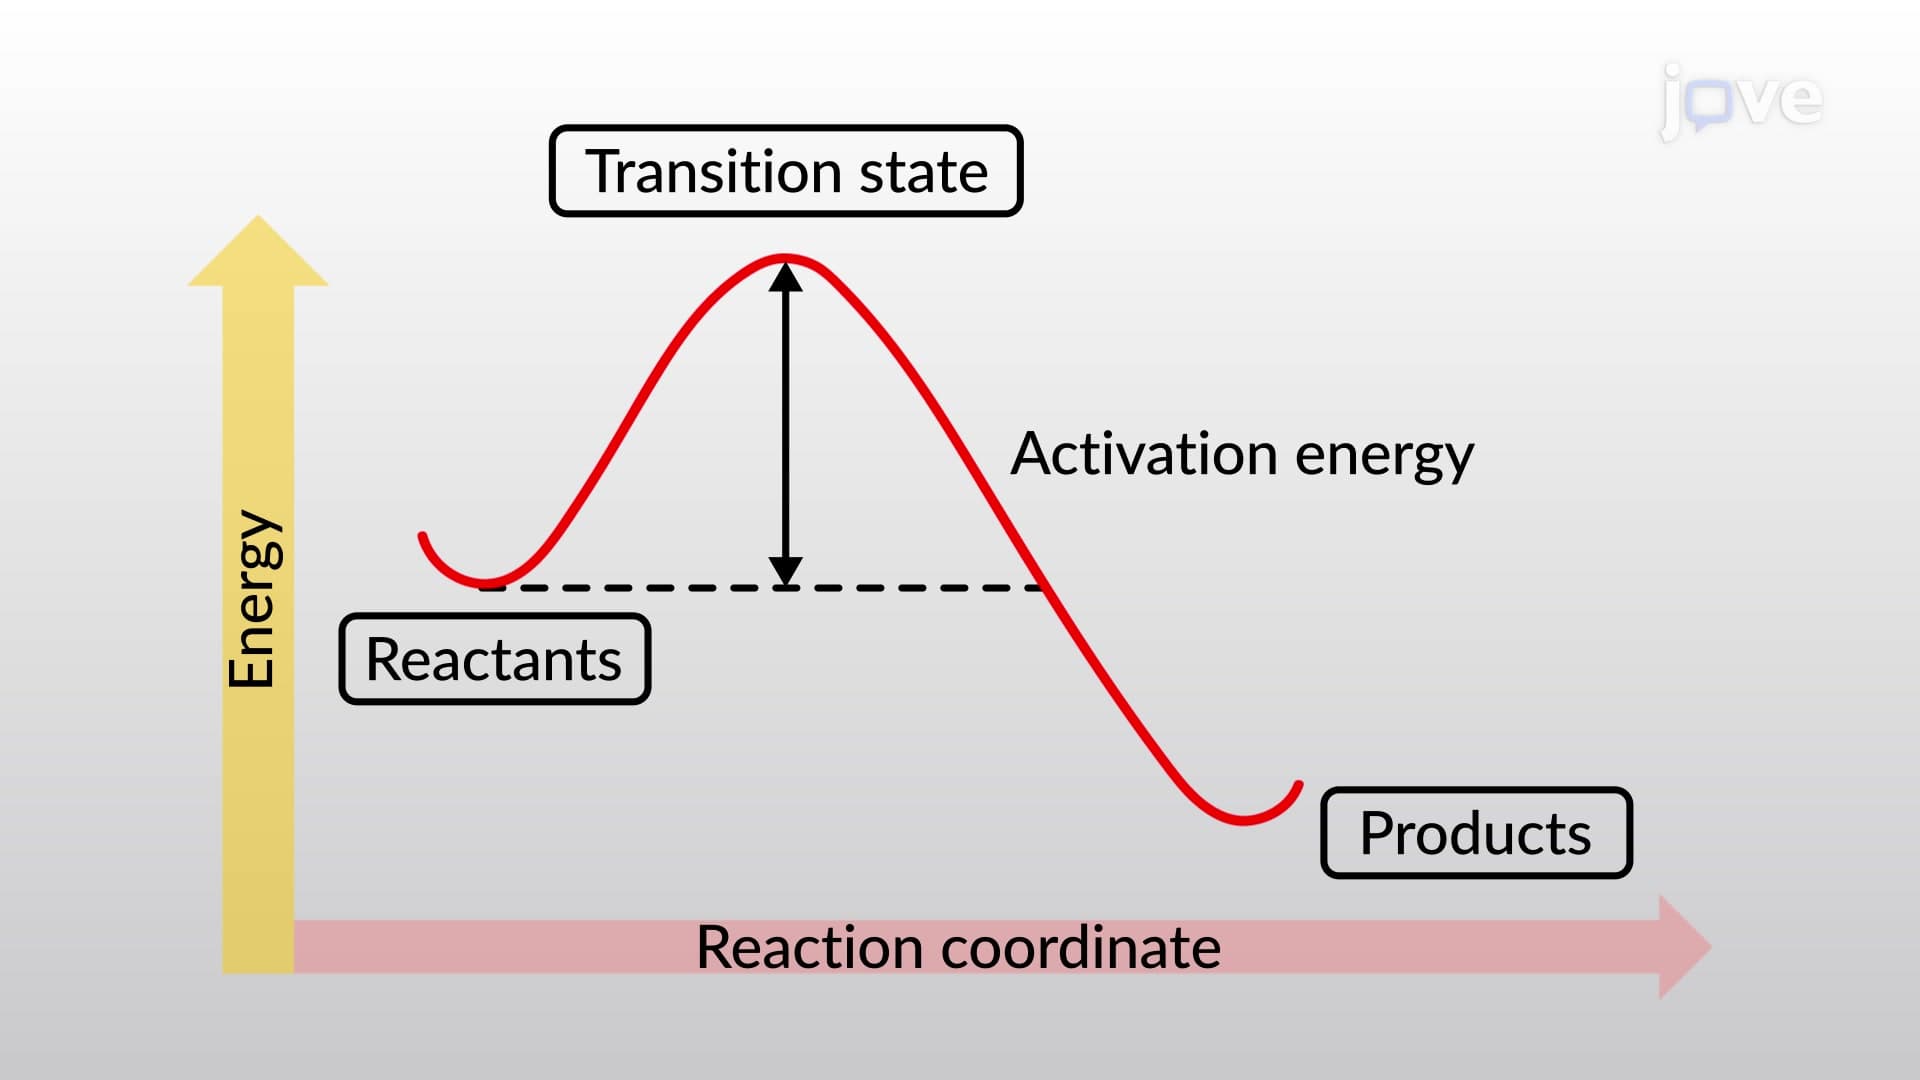
Bond Dissociation Energy and Activation Energy
 01:13
Force and Potential Energy in One Dimension
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:36
Molecular Geometry and Dipole Moments