Time-dependent density functional theoretical study of the absorption properties of BN-substituted C60 fullerenes

C N Ramachandran1, N Sathyamurthy

  • 1Department of Chemistry, Indian Institute of Technology-Kanpur, Kanpur 208016, India.

Related Concept Videos

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution00:52

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution

At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
Stability of Substituted Cyclohexanes02:30

Stability of Substituted Cyclohexanes

This lesson discusses the stability of substituted cyclohexanes with a focus on energies of various conformers and the effect of 1,3-diaxial interactions.
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

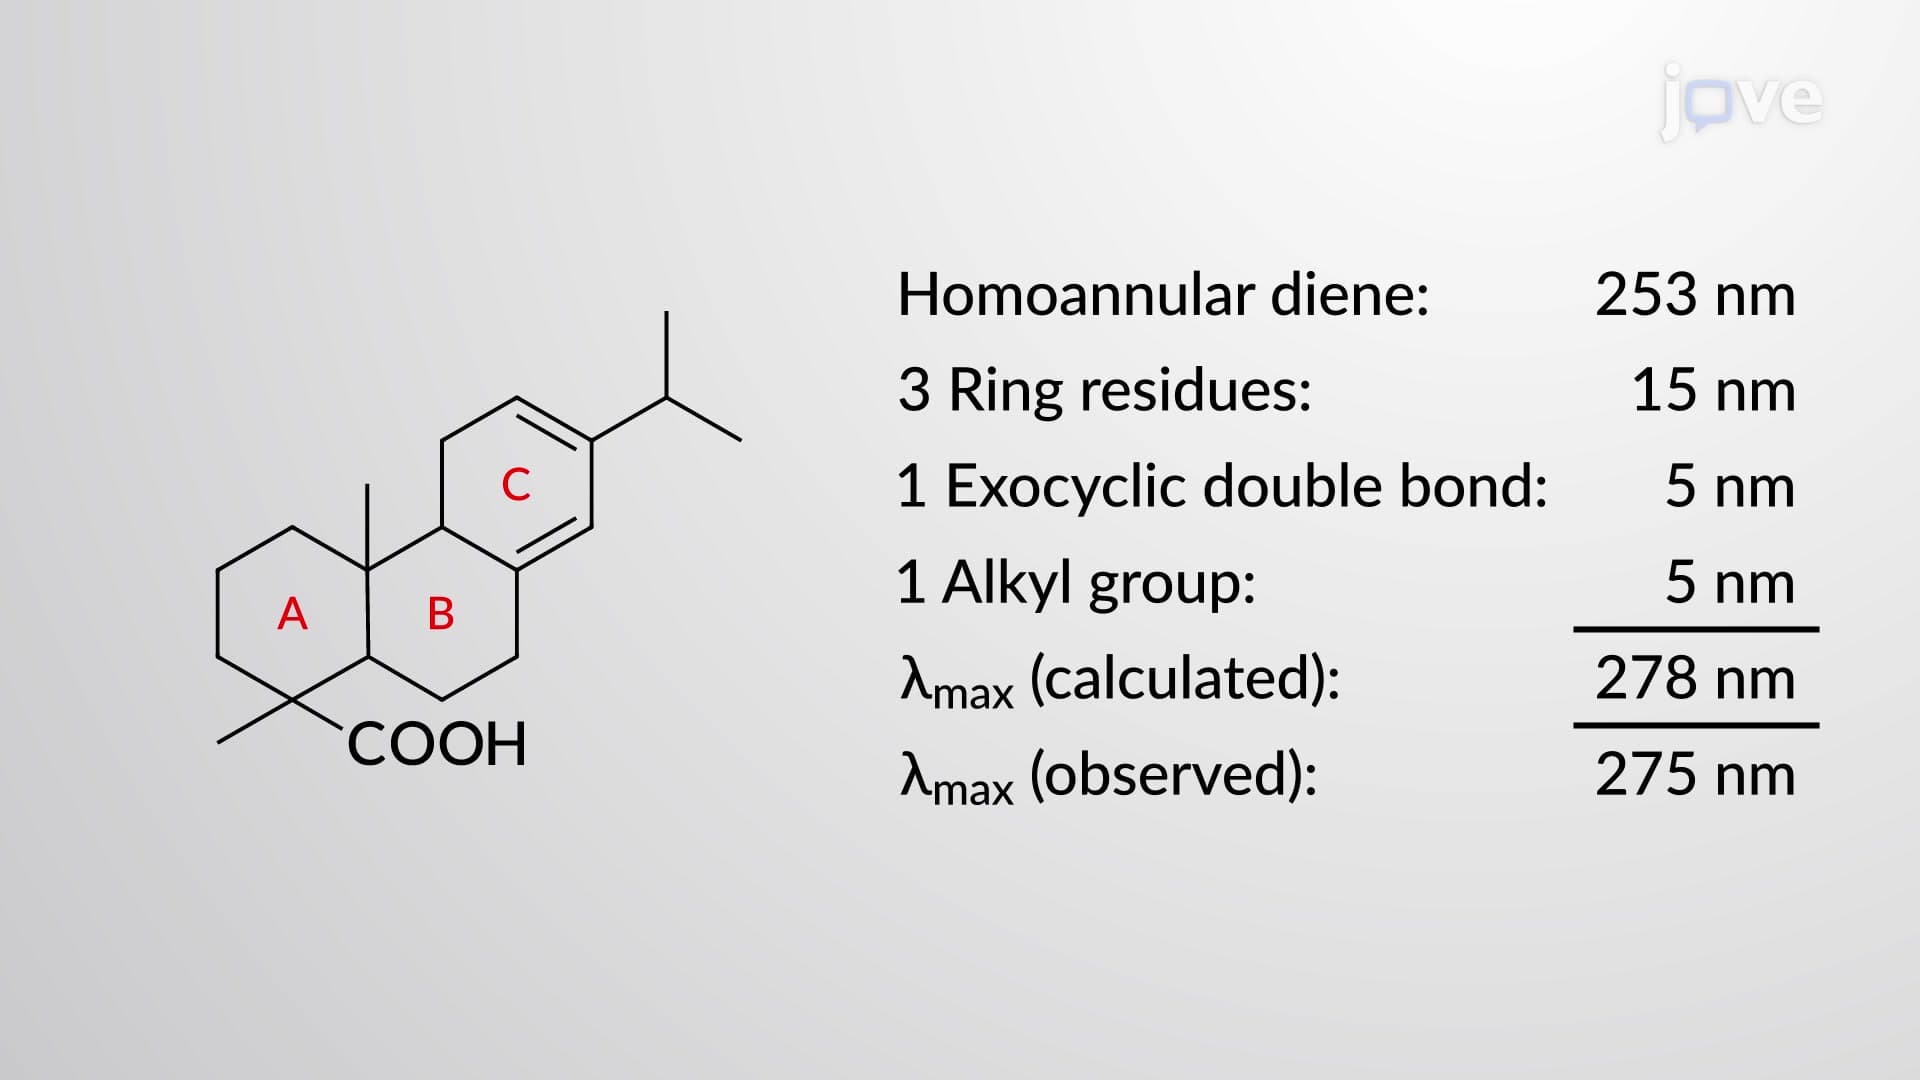
UV–Vis Spectroscopy: Woodward–Fieser Rules

UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
Structure of Benzene: Molecular Orbital Model01:18

Structure of Benzene: Molecular Orbital Model

According to the molecular orbital (MO) model, benzene has a planar structure with a regular hexagon of six sp2 hybridized carbons. As shown in Figure 1, each carbon is bonded to three other atoms with C–C–C and H–C–C bond angles of 120°. The C–H bond length is 109 pm, and the C–C bond length is 139 pm which is midway between the single bond length of sp3 hybridized carbons (154 pm) and sp2 hybridized carbons (133 pm).
π Molecular Orbitals of 1,3-Butadiene01:24

π Molecular Orbitals of 1,3-Butadiene

Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR01:15

¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR

The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.