Related Experiment Video
Updated: Jul 14, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Evaluation of molecular mechanics calculated binding energies for isolated and monolayer organic molecules on
Thomas R Rybolt1, Christina E Wells, Charles R Sisson
1Department of Chemistry, University of Tennessee at Chattanooga, Chattanooga, TN 37403, USA.

Abstract:
The calculated molecule-surface binding energy, E(cal)( *), for physical adsorption was determined using molecular mechanics MM2 parameters for a model graphite surface and various organic molecules. The results for E(cal)( *) were compared to published experimental binding energy values, E( *), from gas chromatography (GC) or thermal desorption (TD). The binding energies from GC were for isolated molecules in the Henry's law region of adsorption, and the binding energies from TD were for molecules in monolayer coverage on a highly oriented pyrolytic graphite (HOPG). A simple desorption model was used to allow the calculation of monolayer coverage to include both molecule-surface and molecule-molecule interactions and then the results were compared to experimental values. For the 14 TD organic adsorbates (polyaromatic hydrocarbons, alcohols, benzene, substituted benzenes, methane, chloroalkanes, N,N-dimethylformamide, and C(60) Buckyball), the experimental versus calculated binding energies were E( *)=1.1193E(cal)( *) and r(2)=0.967. The GC E( *) values were also well correlated by calculated E(cal)( *) values for a set of 11 benzene and methyl substituted benzenes and for another set of 10 alkanes and haloalkanes. The TD E(cal)( *) mechanics computation provides a useful comparison to the one for GC data since adsorbate-adsorbate interactions as well as adsorbate-surface must be considered.
More Related Videos
 10:01
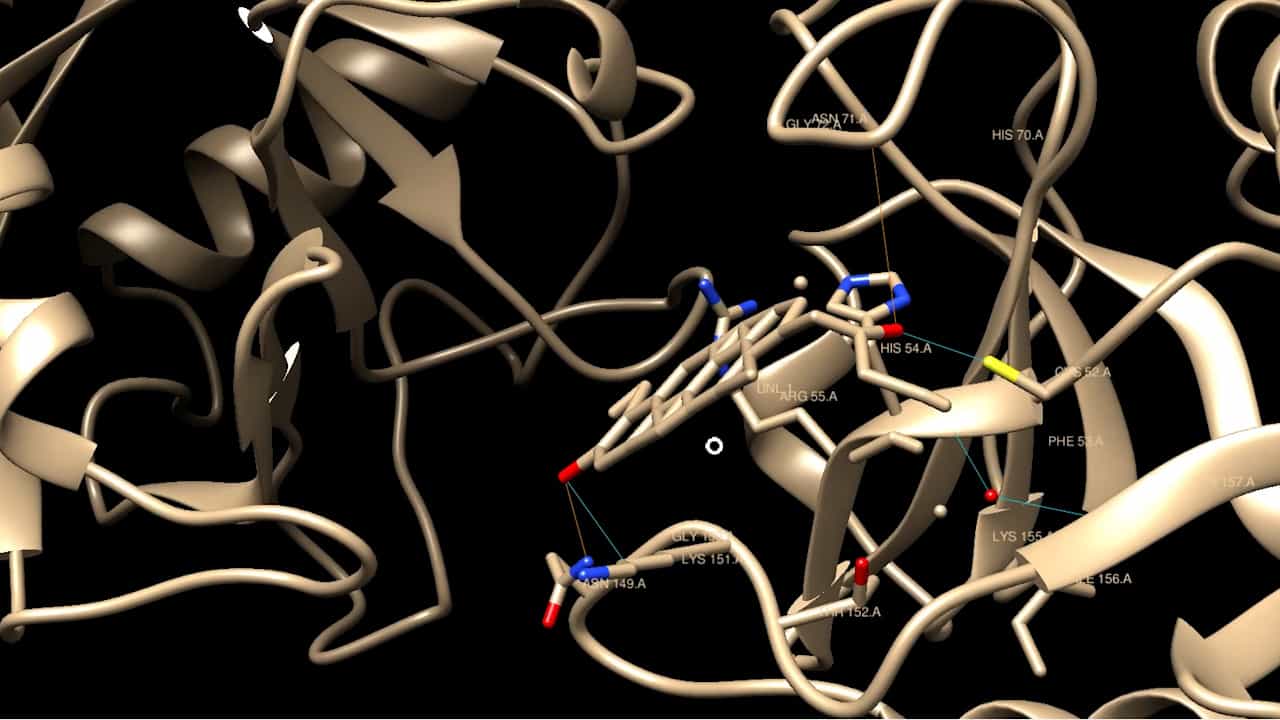
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:13
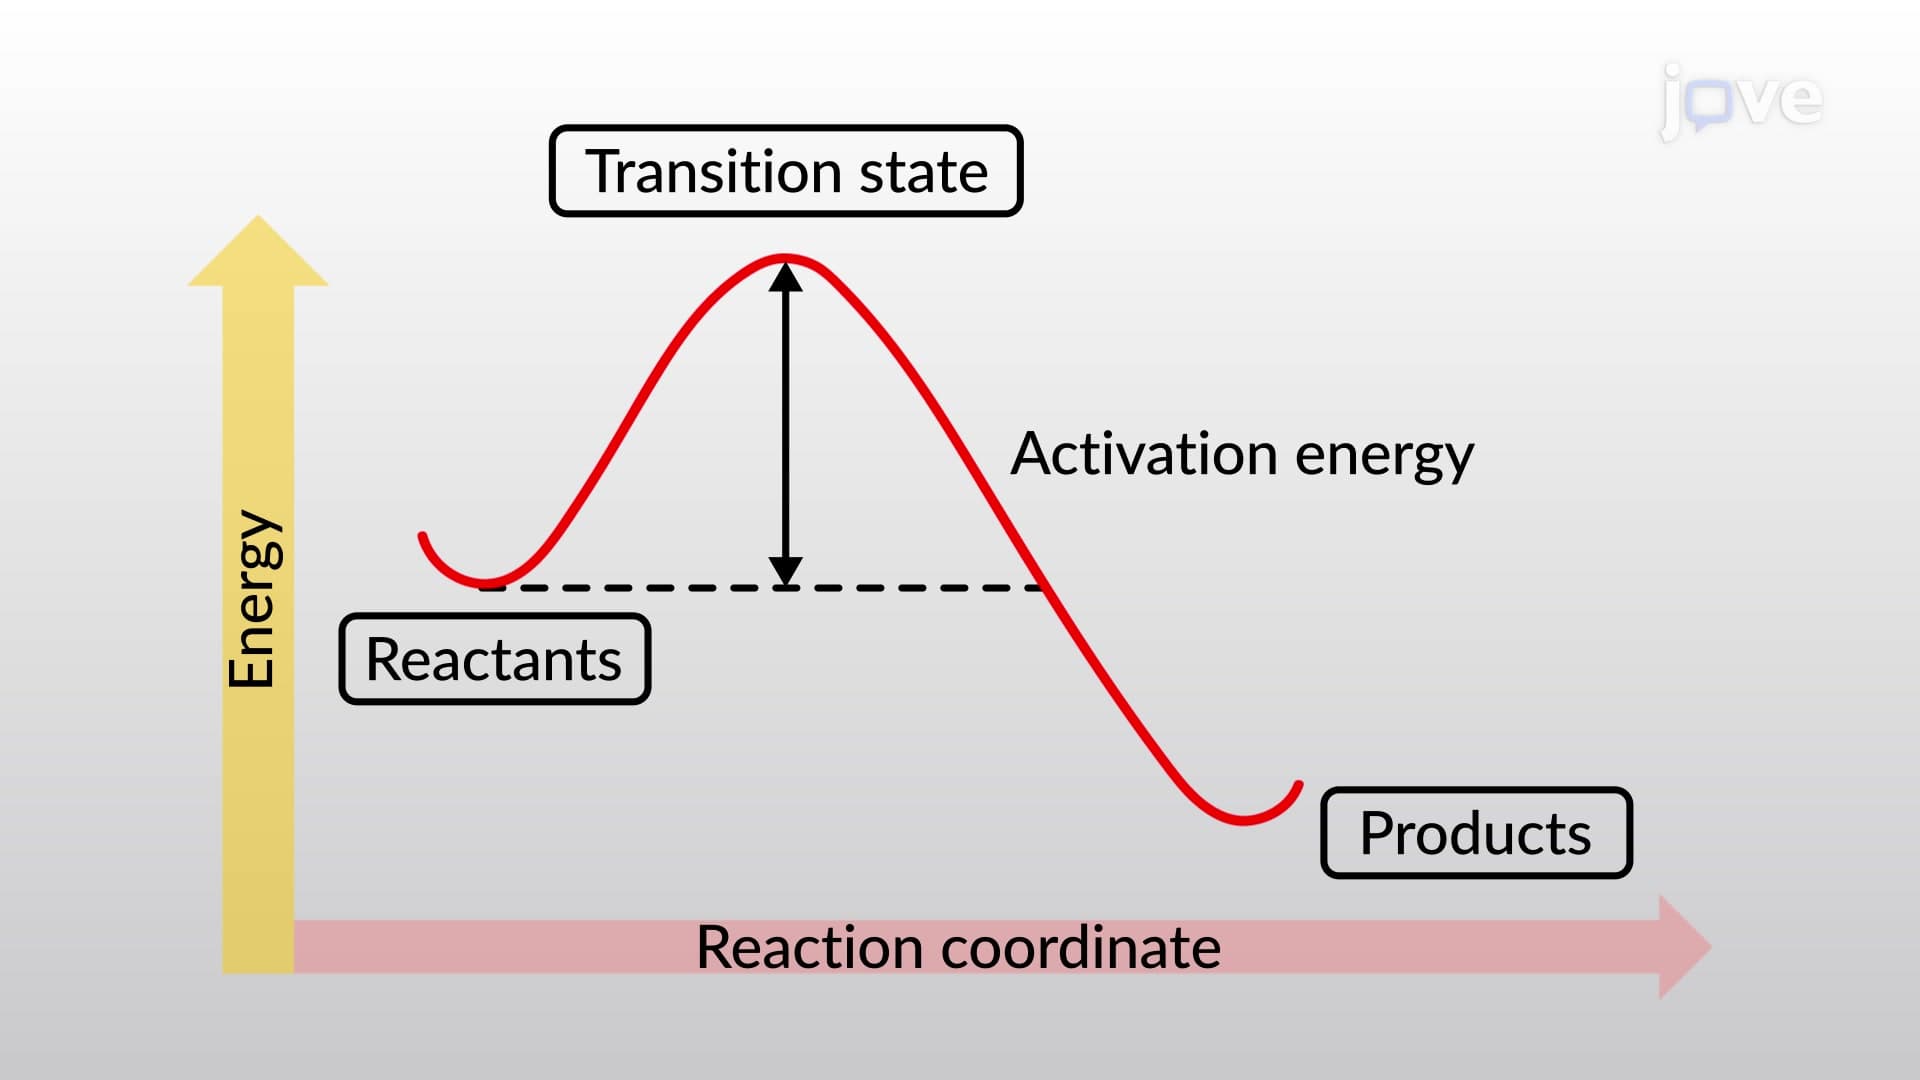
Bond Dissociation Energy and Activation Energy
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...