On reduced amino acid alphabets for phylogenetic inference
Related Experiment Video
Updated: Jul 13, 2026

An Integrated Approach for Microprotein Identification and Sequence Analysis
Published on: July 12, 2022
1Department of Mathematics and Statistics, Dalhousie University, Halifax, Nova Scotia, Canada. susko@mathstat.dal.ca

Abstract:
We investigate the use of Markov models of evolution for reduced amino acid alphabets or bins of amino acids. The use of reduced amino acid alphabets can ameliorate effects of model misspecification and saturation. We present algorithms for 2 different ways of automating the construction of bins: minimizing criteria based on properties of rate matrices and minimizing criteria based on properties of alignments. By simulation, we show that in the absence of model misspecification, the loss of information due to binning is found to be insubstantial, and the use of Markov models at the binned level is found to be almost as effective as the more appropriate missing data approach. By applying these approaches to real data sets where compositional heterogeneity and/or saturation appear to be causing biased tree estimation, we find that binning can improve topological estimation in practice.
Related Concept Videos
 03:21
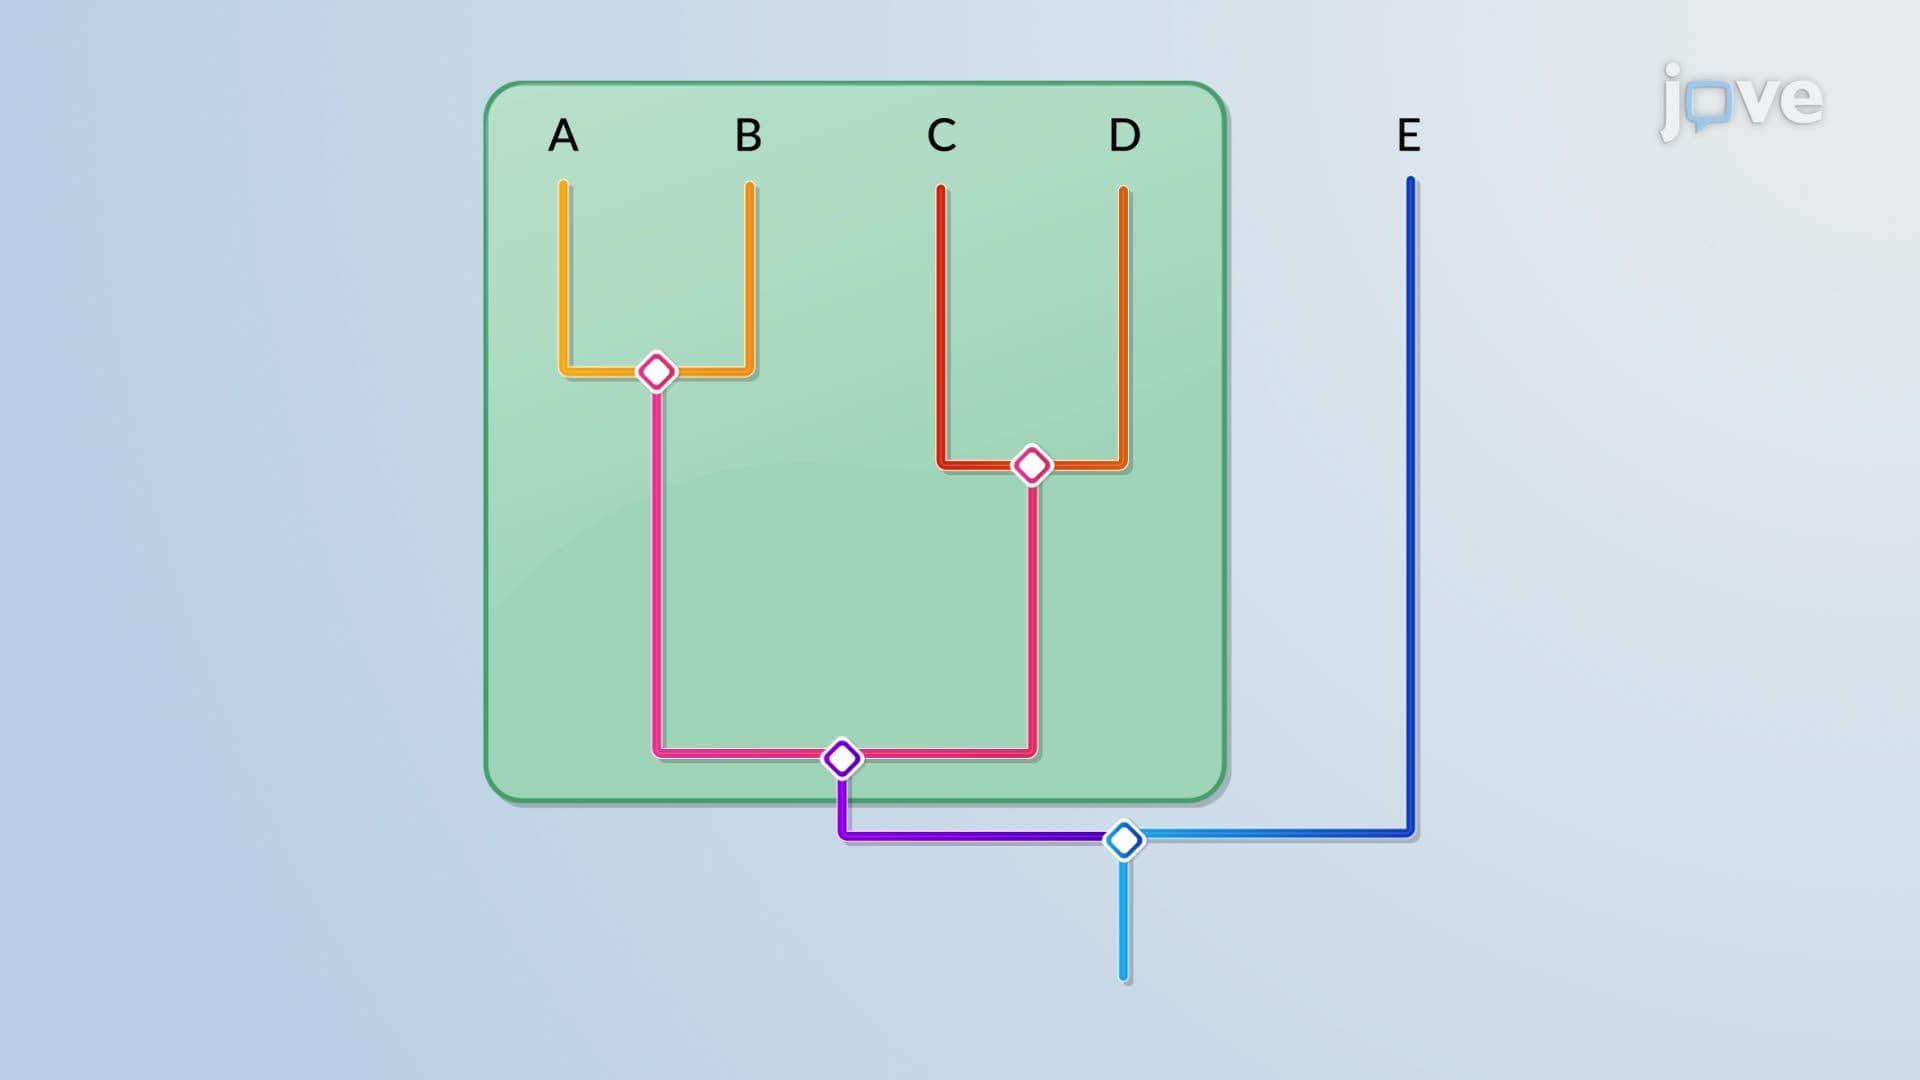
Phylogenetic Trees
03:21Phylogenetic Trees
 03:42
Amino acids
 02:26
Conservation of Protein Domains
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
02:26Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:54
Evolutionary Relationships through Genome Comparisons