Related Experiment Video
Updated: Jul 13, 2026


Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Quasiclassical trajectory calculations of acetaldehyde dissociation on a global potential energy surface indicate

Abstract:
A recent experimental study [Houston, P. L.; Kable, S. H. Proc. Natl. Acad. Sci. U.S.A. 2006, 103, 16079] of the photodissociation of acetaldehyde (CH(3)CHO) has suggested two distinct mechanisms for the production of the molecular products CH(4) + CO. One corresponds to the traditional transition state mechanism and the other to a transition state-skirting path similar to the roaming channel previously reported in formaldehyde. To investigate this theoretically, a full-dimensional potential energy surface (PES) has been constructed. The PES was fit with permutationally invariant polynomials to 135,000 points calculated using coupled cluster theory with single and double excitations and a perturbative treatment of triple excitations [CCSD(T)] and correlation consistent basis sets of double- and triple-zeta quality. To test the accuracy of the PES additional CCSD(T) and multireference configuration interaction calculations were carried out. Quasiclassical trajectory calculations were run on the PES starting at the acetaldehyde equilibrium geometry and also at the conventional transition state (TS) for the molecular products CH(4) + CO. The former calculations agree well with the experimental results of Houston and Kable; however, those from the TS do not. The implications for a non-transition state, roaming mechanism in this molecule are discussed.


Related Concept Videos
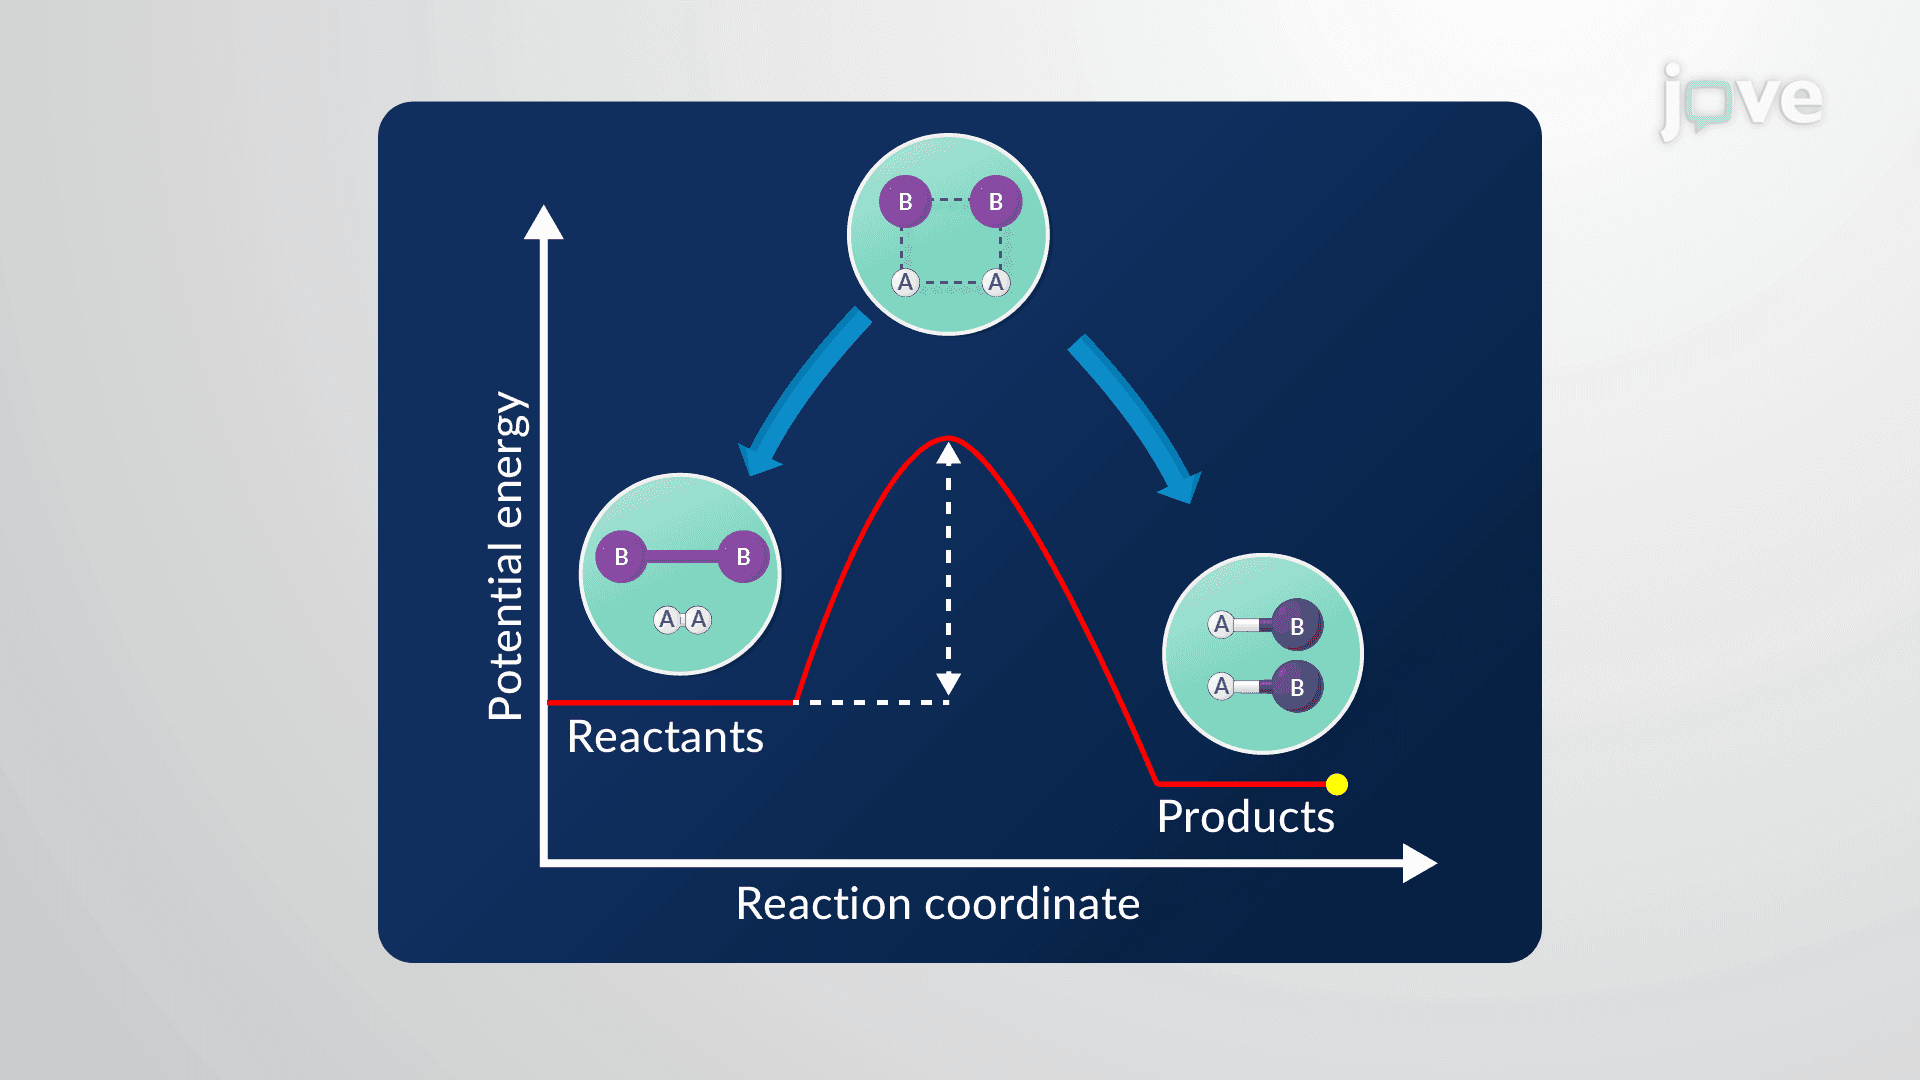 01:25
Transition State Theory
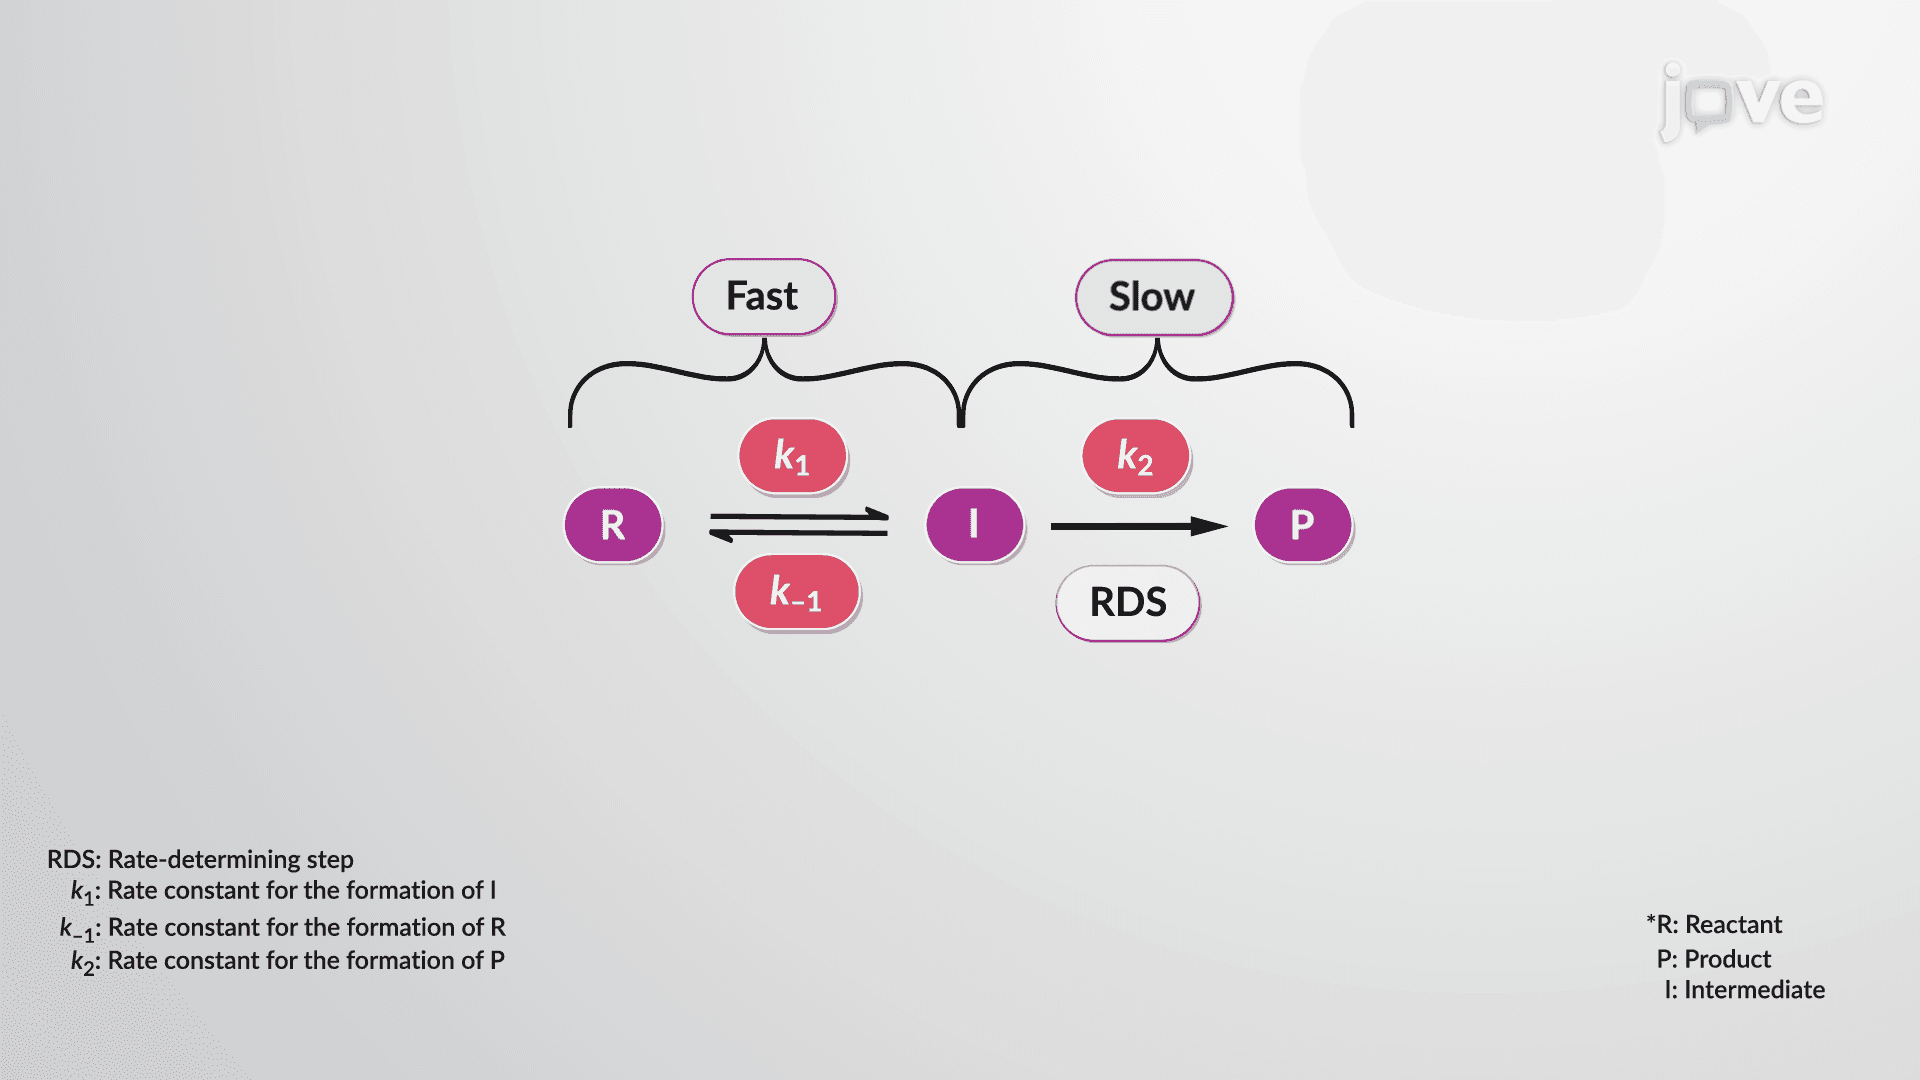 01:26
Reaction Mechanisms: The Steady-State Approximation
 02:13
Energy Diagrams, Transition States, and Intermediates
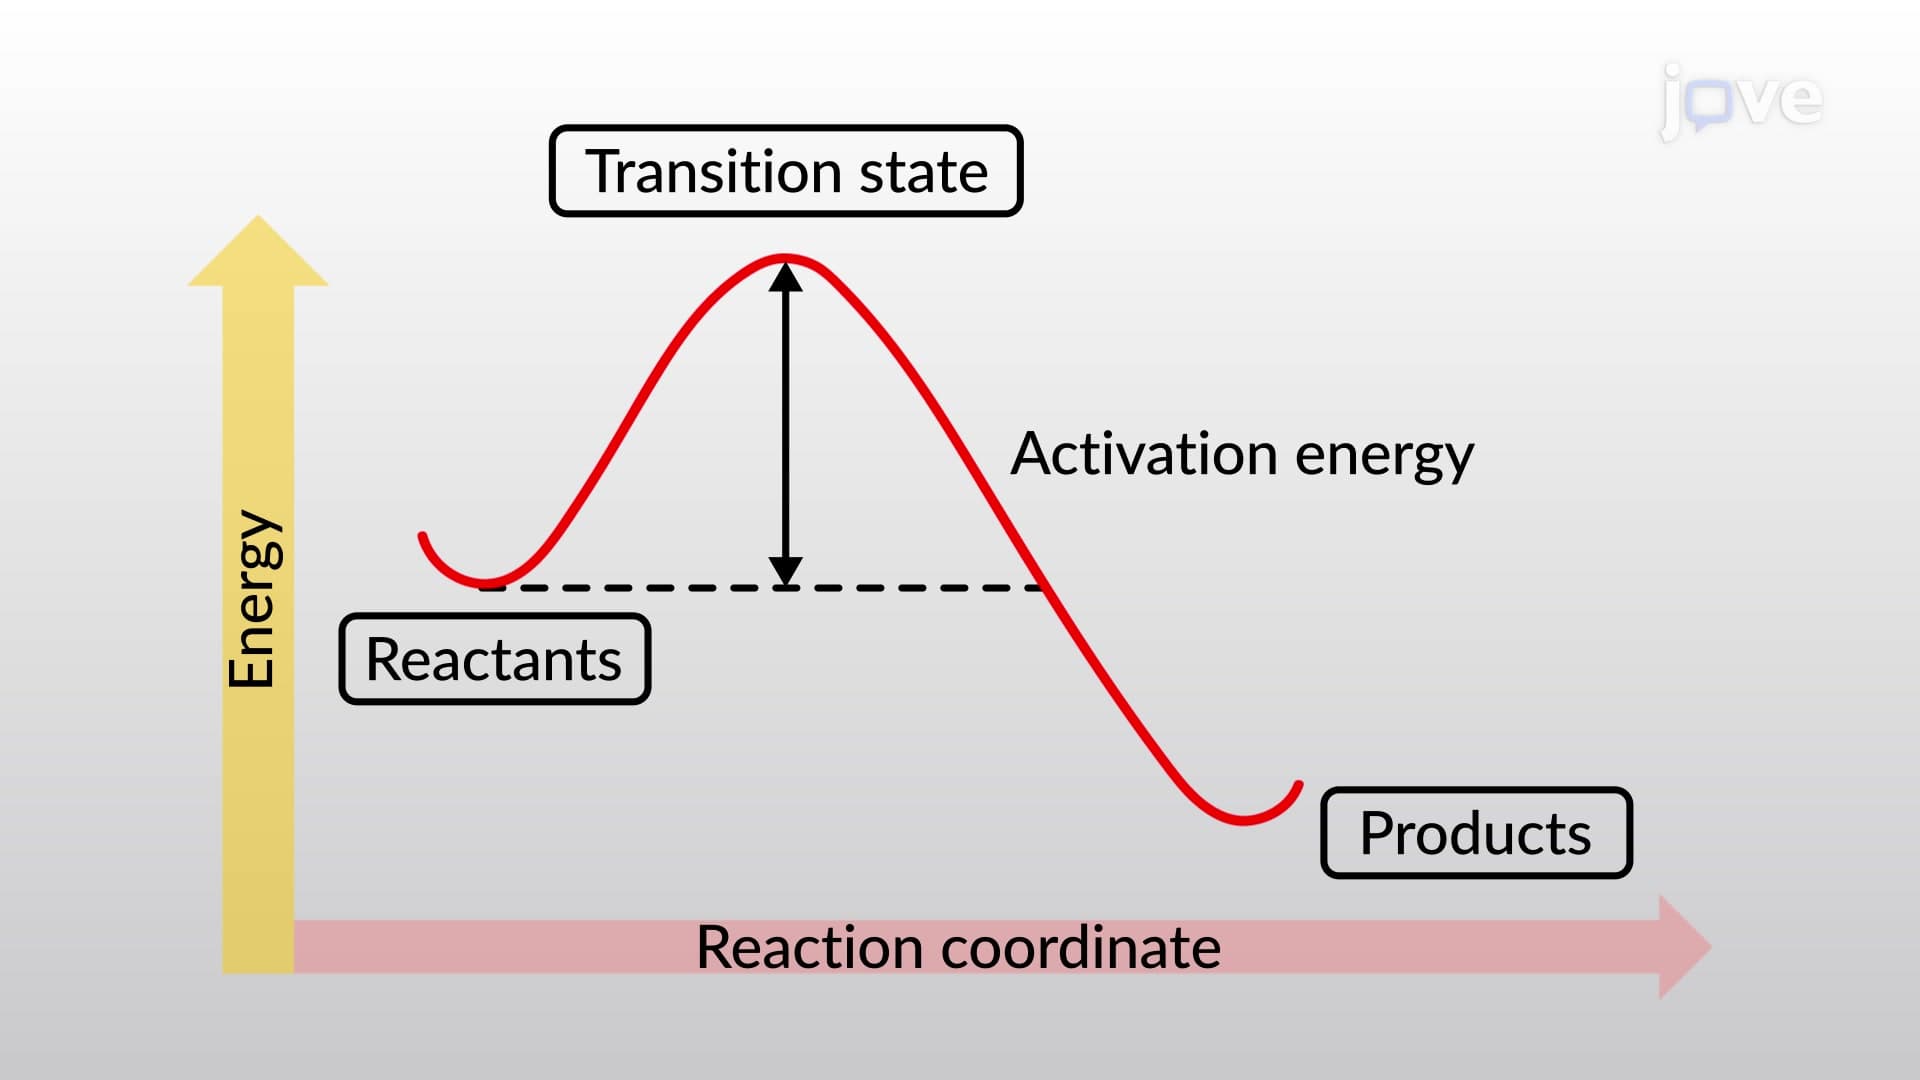 02:13
Bond Dissociation Energy and Activation Energy
 02:45
Acid-Catalyzed Hydration of Alkenes
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration