Related Experiment Video
Updated: Jul 13, 2026
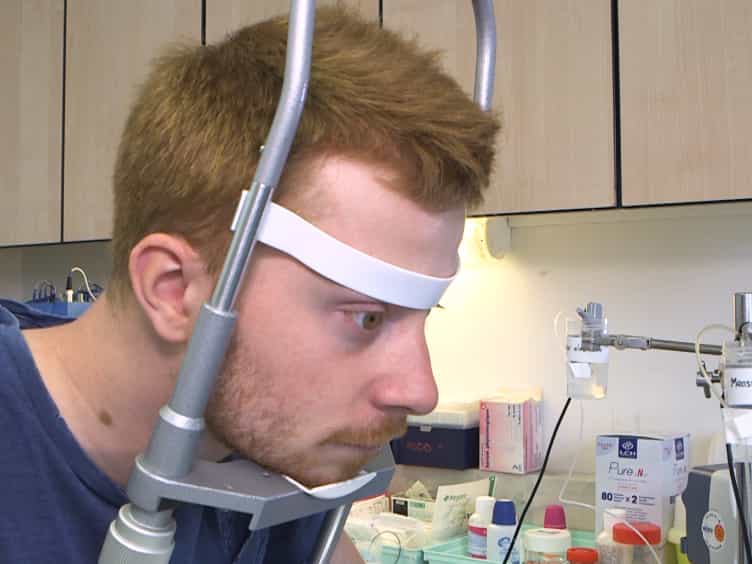
Standardized Measurement of Nasal Membrane Transepithelial Potential Difference (NPD)
Published on: September 13, 2018
Repeated aerosolized AAV-CFTR for treatment of cystic fibrosis: a randomized placebo-controlled phase 2B trial
Richard B Moss1, Carlos Milla, John Colombo
1Department of Pediatrics, Stanford University, Stanford, CA 94305, and Division of Respiratory Diseases, Children's Hospital Boston, MA 02115, USA. rmoss@stanford.edu

Abstract:
Previous studies have demonstrated that delivery of a recombinant adeno-associated virus (AAV) vector encoding the complete human cystic fibrosis transmembrane regulator (CFTR) cDNA (tgAAVCF) to the nose, sinus, and lungs of subjects with cystic fibrosis (CF) was safe and well tolerated. In a small randomized, double-blind study of three doses of aerosolized tgAAVCF or placebo at 30-day intervals, encouraging but non-significant trends in pulmonary function and induced sputum interleukin 8 (IL-8) levels were seen at early time points. This larger study was conducted to verify these trends. One hundred and two subjects aged 12 years and older with mild-to-moderate cystic fibrosis (forced expiratory flow in 1 sec [FEV1]:60% predicted) were randomized to two aerosolized doses of 1x10(13)DNase-resistant particles of tgAAVCF (n=51) or matching placebo (n=51) administered 30 days apart. Although tgAAVCF was well tolerated, the study did not meet its primary efficacy end point of statistically significant improvement in FEV1 30 days after initial administration of tgAAVCF compared with placebo. There were no significant differences in spirometric lung function over time, induced sputum biologic markers, or days of antibiotic use in either treatment group. Thus repeated doses of aerosolized tgAAVCF were safe and well tolerated, but did not result in significant improvement in lung function over time. Because gene transfer is the simplest, most basic way to correct the underlying genetic defect that leads to disease in CF, further research is warranted to develop an effective gene transfer agent for the treatment of CF.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History