Related Experiment Video
Updated: Jul 11, 2026
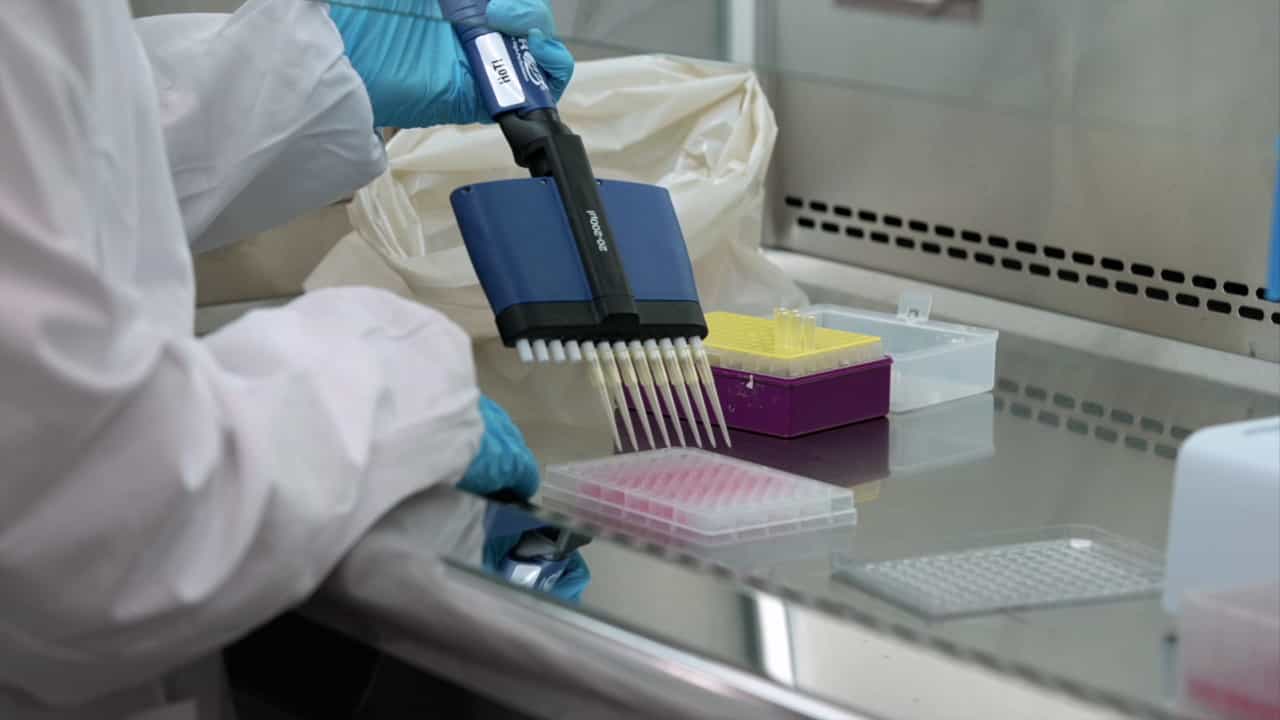

Assay Development for High-Throughput Drug Screening Against Mycobacteria
Published on: October 25, 2024
[Development of a large-scale comparative genome system and its application to the analysis of mycobacteria genomes]
Yasubumi Sakakibara1, Yasunori Osana, Kris Popendorf
1Department of Biosciences and Informatics, Keio University, Japan. yasu@bio.keio.ac.jp

Abstract:
As the number of whole genome sequences available continues to increase rapidly, the raw scale of the sequence data being used in analysis is the first hurdle for comparative genome analysis. When performing whole genome alignments, large-scale rearrangements make it necessary to first find out roughly which short well-conserved segments correspond to what other segments (termed anchors). Successful results have been achieved by adapting tools like BLAT and BLASTZ on a problem-to-problem basis, but the work required to perform a single alignment is considerable. Recently, new programs such as Mauve and Pattern-Hunter can handle slightly larger inputs, but the memory/time requirements for sequences like Human and Chimp X chromosomes are prohibitive for most computational environments. Our novel algorithm, which we have implemented in a program called Murasaki (available at http://murasaki.dna.bio.keio.ac.jp), makes it possible to identify anchors of multiple large sequences on the scale of several hundred megabases (e.g. three mammal chromosomes) in a matter of minutes. We also demonstrate an application of Murasaki to the comparative analysis of multiple mycobacteria genomes.
More Related Videos
Related Concept Videos
 01:29
Modern Molecular Taxonomy
 01:20
Applications of Molecular Taxonomy
 02:54
Evolutionary Relationships through Genome Comparisons
 02:02
Genomics
 01:08
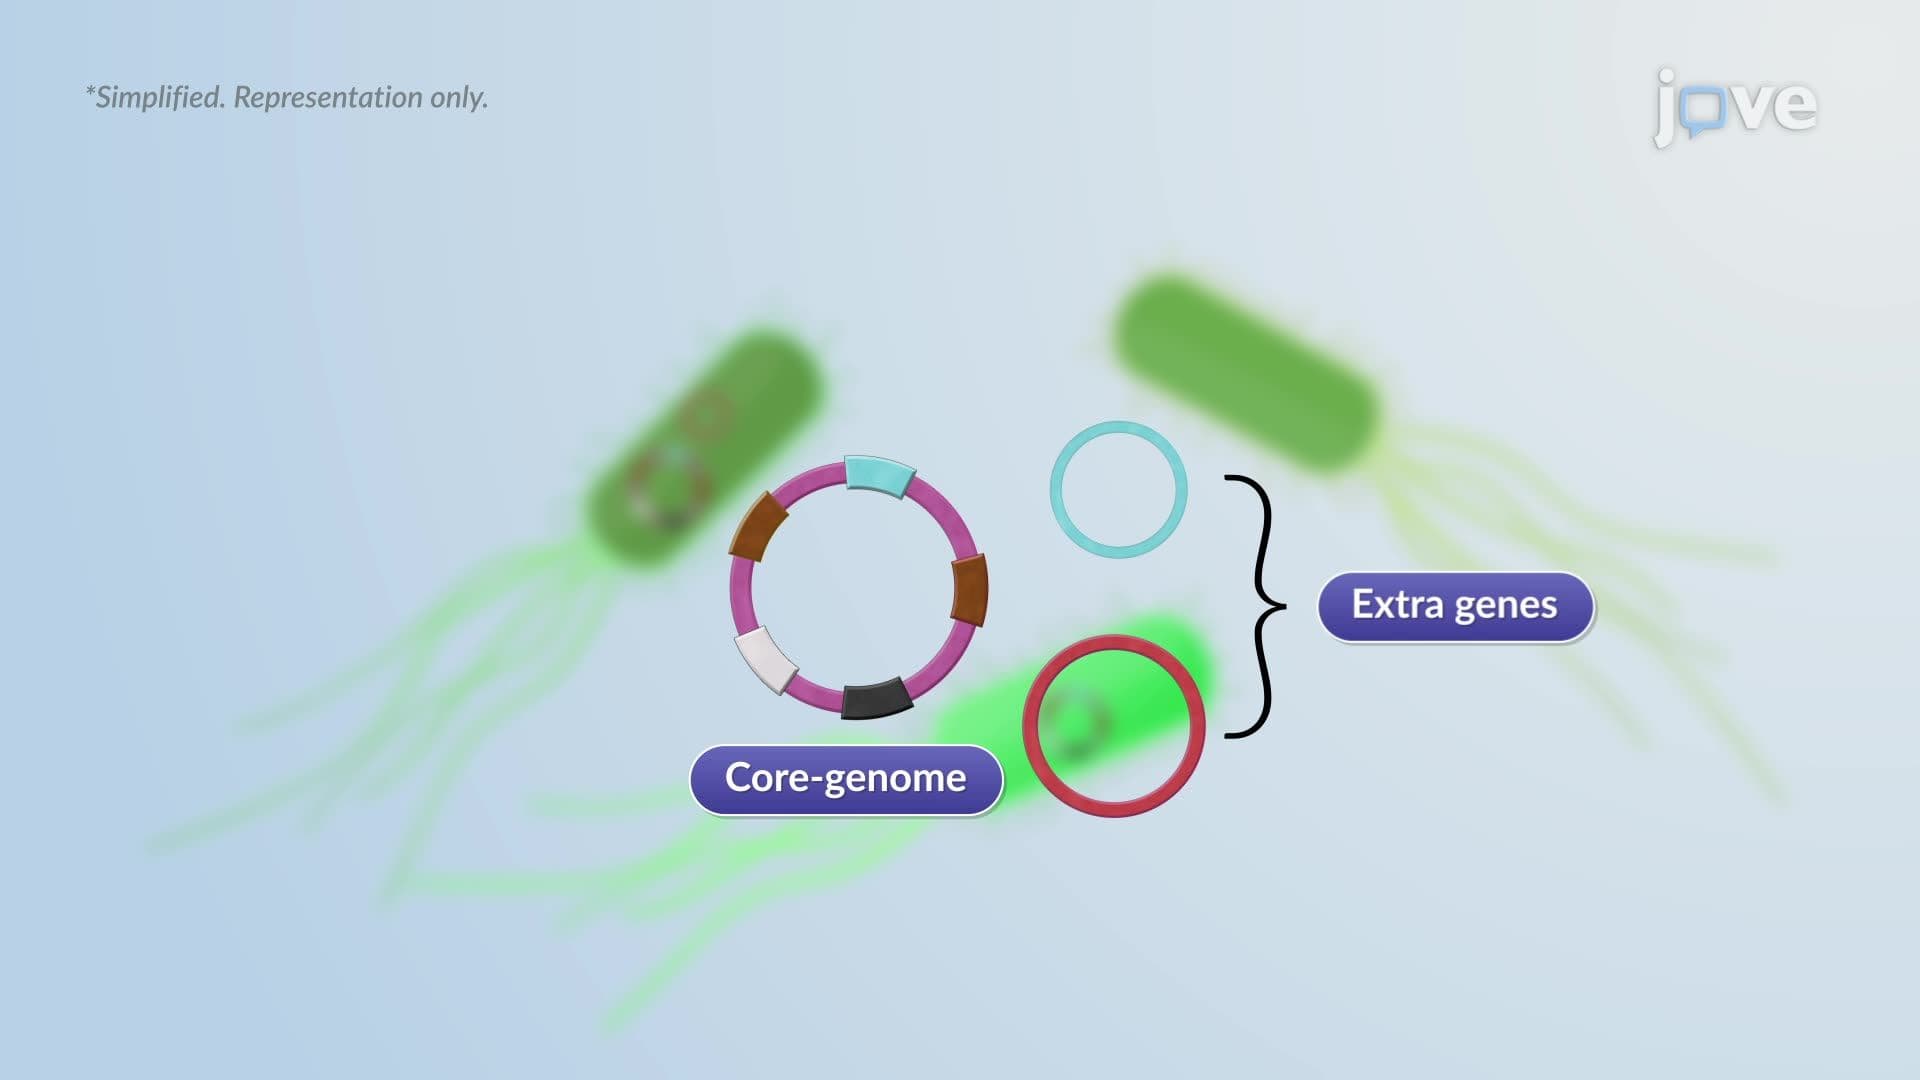
Evolution of Microbial Genome
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes