Related Experiment Video
Updated: Jul 10, 2026

Preparation and Pathogen Inactivation of Double Dose Buffy Coat Platelet Products using the INTERCEPT Blood System
Published on: December 7, 2012
Pharmacokinetics of Beriplex P/N prothrombin complex concentrate in healthy volunteers
Helmut Ostermann1, Sabine Haertel, Sigurd Knaub
1University Hospital Munich - Grosshadern, Medical Clinic III, Department of Haematology and Oncology, Ludwig Maximilian University, Munich, Germany.

Abstract:
Prothrombin complex concentrates (PCCs) are widely administered for emergency oral anticoagulation reversal and for coagulation defects in liver disease. Pharmacokinetic data may help to optimize treatment. The objective of this study was to characterize the pharmacokinetics of a PCC (Beriplex P/N) containing coagulation factors II (FII), VII (FVII), IX (FIX) and X (FX) and anticoagulant proteins C and S. Fifteen healthy volunteers received a single rapid 50 IU/kg infusion of PCC and underwent frequent blood sampling until 144 hours (h) after infusion. Coagulation factors and anticoagulant protein pharmacokinetic parameters were estimated by non-linear regression. The mean infusion rate of PCC was 7.9 ml/min, equivalent to 196.4 IU/min. By the earliest post-infusion sampling point at 5 minutes (min), plasma FIX concentration increased by a median of 73%. Median increases in FII, FVII and FX at 5 min were 122%, 62% and 158%, respectively. Proteins C and S also increased rapidly. The median terminal half-life of FIX was 16.7 h, FII 59.7 h, FVII 4.2 h and FX 30.7 h. The median in-vivo recovery of FIX was 1.57 %/IU/kg and that of the other three coagulation factors > 2 %/IU/kg. Plasma concentration of thrombogenicity marker D-dimer did not increase, and there was no clinical evidence of thrombosis. Through up to 12 weeks follow-up there were no laboratory findings indicating PCC-related viral exposure. Rapid PCC infusion produced prompt sustained increases in coagulation factors and anticoagulant proteins with no clinical evidence of thrombosis or viral transmission.
Related Concept Videos
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:09
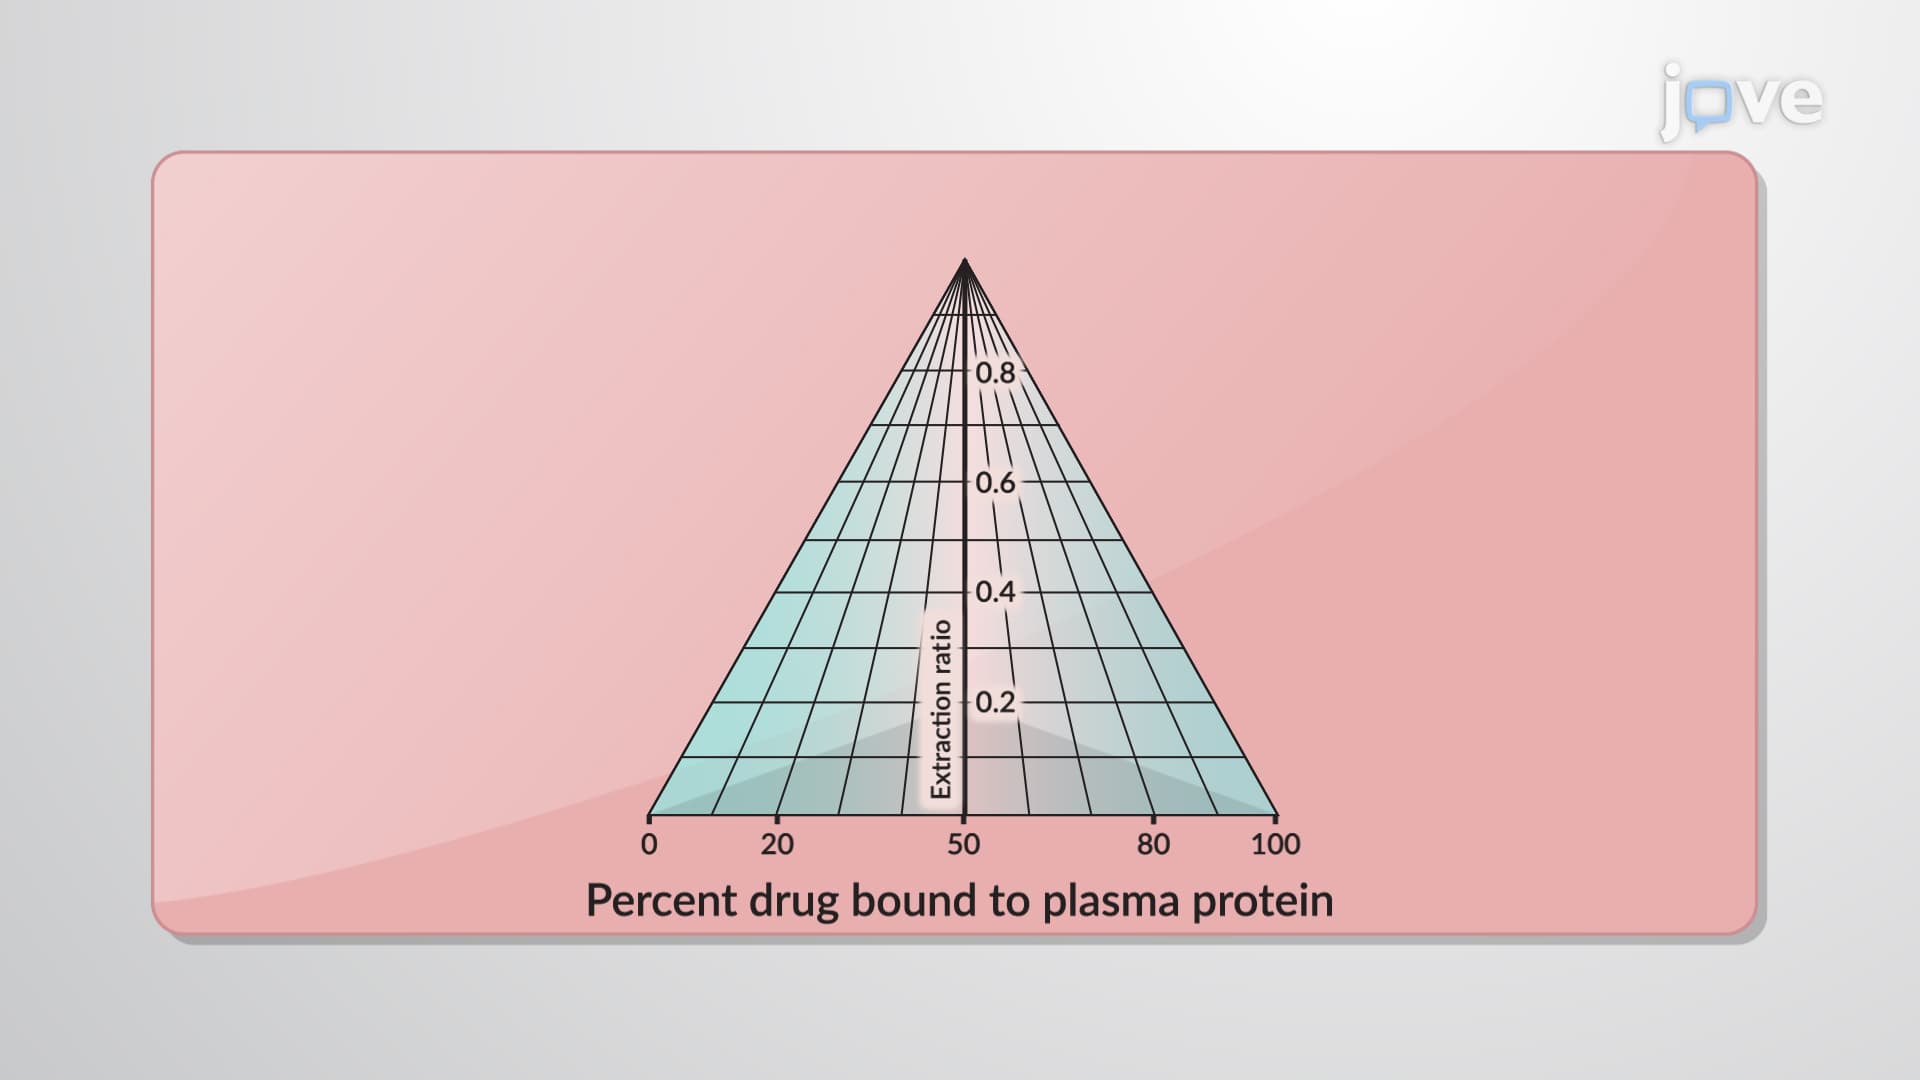
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...
 01:29
Drug Distribution: Plasma Protein Binding
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods