Related Experiment Video
Updated: Jul 10, 2026

Reaction Kinetics and Combustion Dynamics of I4O9 and Aluminum Mixtures
Published on: November 7, 2016
Thermochemistry, bond energies, and internal rotor potentials of dimethyl tetraoxide
Gabriel da Silva1, Joseph W Bozzelli
1Department of Chemistry and Environmental Science, New Jersey Institute of Technology, Newark, NJ 07102, USA.

Abstract:
Thermochemical properties of dimethyl tetraoxide (CH(3)OOOOCH(3)), the dimer of the methylperoxy radical, are studied using ab initio and density functional theory methods. Methylperoxy radicals are known to be important intermediates in the tropospheric ozone cycle, and the self-reaction of methylperoxy radicals, which is thought to proceed via dimethyl tetraoxide, leads to significant chain radical termination in this process. Dimethyl tetraoxide has five internal rotors, three of them unique; the potential energy profiles are calculated for these rotors, as well as for those in the CH(3)OO, CH(3)OOO, and CH(3)OOOO radicals. The dimethyl tetraoxide internal rotor profiles show barriers to rotation of 2-8 kcal mol(-1). Using B3LYP/6-31(d) geometries, frequencies, internal rotor potentials, and moments of inertia, we determine entropy and heat capacity values for dimethyl tetraoxide and its radicals. Isodesmic work reactions with the G3B3 and CBS-APNO methods are used; we calculate this enthalpy as -9.8 kcal mol(-1). Bond dissociation energies (BDEs) are calculated for all C-O and O-O bonds in dimethyl tetraoxide, again with the G3B3 and CBS-APNO theoretical methods, and we suggest the following BDEs: 46.0 kcal mol(-1) for CH(3)-OOOOCH(3), 20.0 kcal mol(-1) for CH(3)O-OOOCH(3), and 13.9 kcal mol(-1) for CH(3)OO-OOCH(3). From the BDE calculations and the isodesmic enthalpy of formation for dimethyl tetraoxide, we suggest enthalpies of 2.1, 5.8, and 1.4 kcal mol(-1) for the CH(3)OO, CH(3)OOO, and CH(3)OOOO radicals, respectively. We evaluate the suitability of 10 different density functional theory (DFT) methods for calculating thermochemical properties of dimethyl tetraoxide and its radicals with the 6-31G(d) and 6-311++G(3df,3pd) basis sets, using a variety of work reaction schemes. Overall, the best-performed DFT methods of those tested were TPSSh, BMK, and B1B95. Significant improvements in accuracy were made by moving from atomization to isodesmic work reactions, with most DFT methods yielding errors of less than 2 kcal mol(-1) with the 6-311++G(3df,3pd) basis set for isodesmic calculations on the dimethyl tetraoxide enthalpy. These isodesmic calculations were basis set consistent, with a considerable reduction in error found by using the 6-311++G(3df,3pd) basis set over the 6-31G(d) basis set. This was not the case, however, for atomization and bond dissociation work reactions, where the two basis sets returned similar results. Improved group additivity terms for the O-O-O moiety (O/O2 central atom group) are also determined.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:11
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 01:27
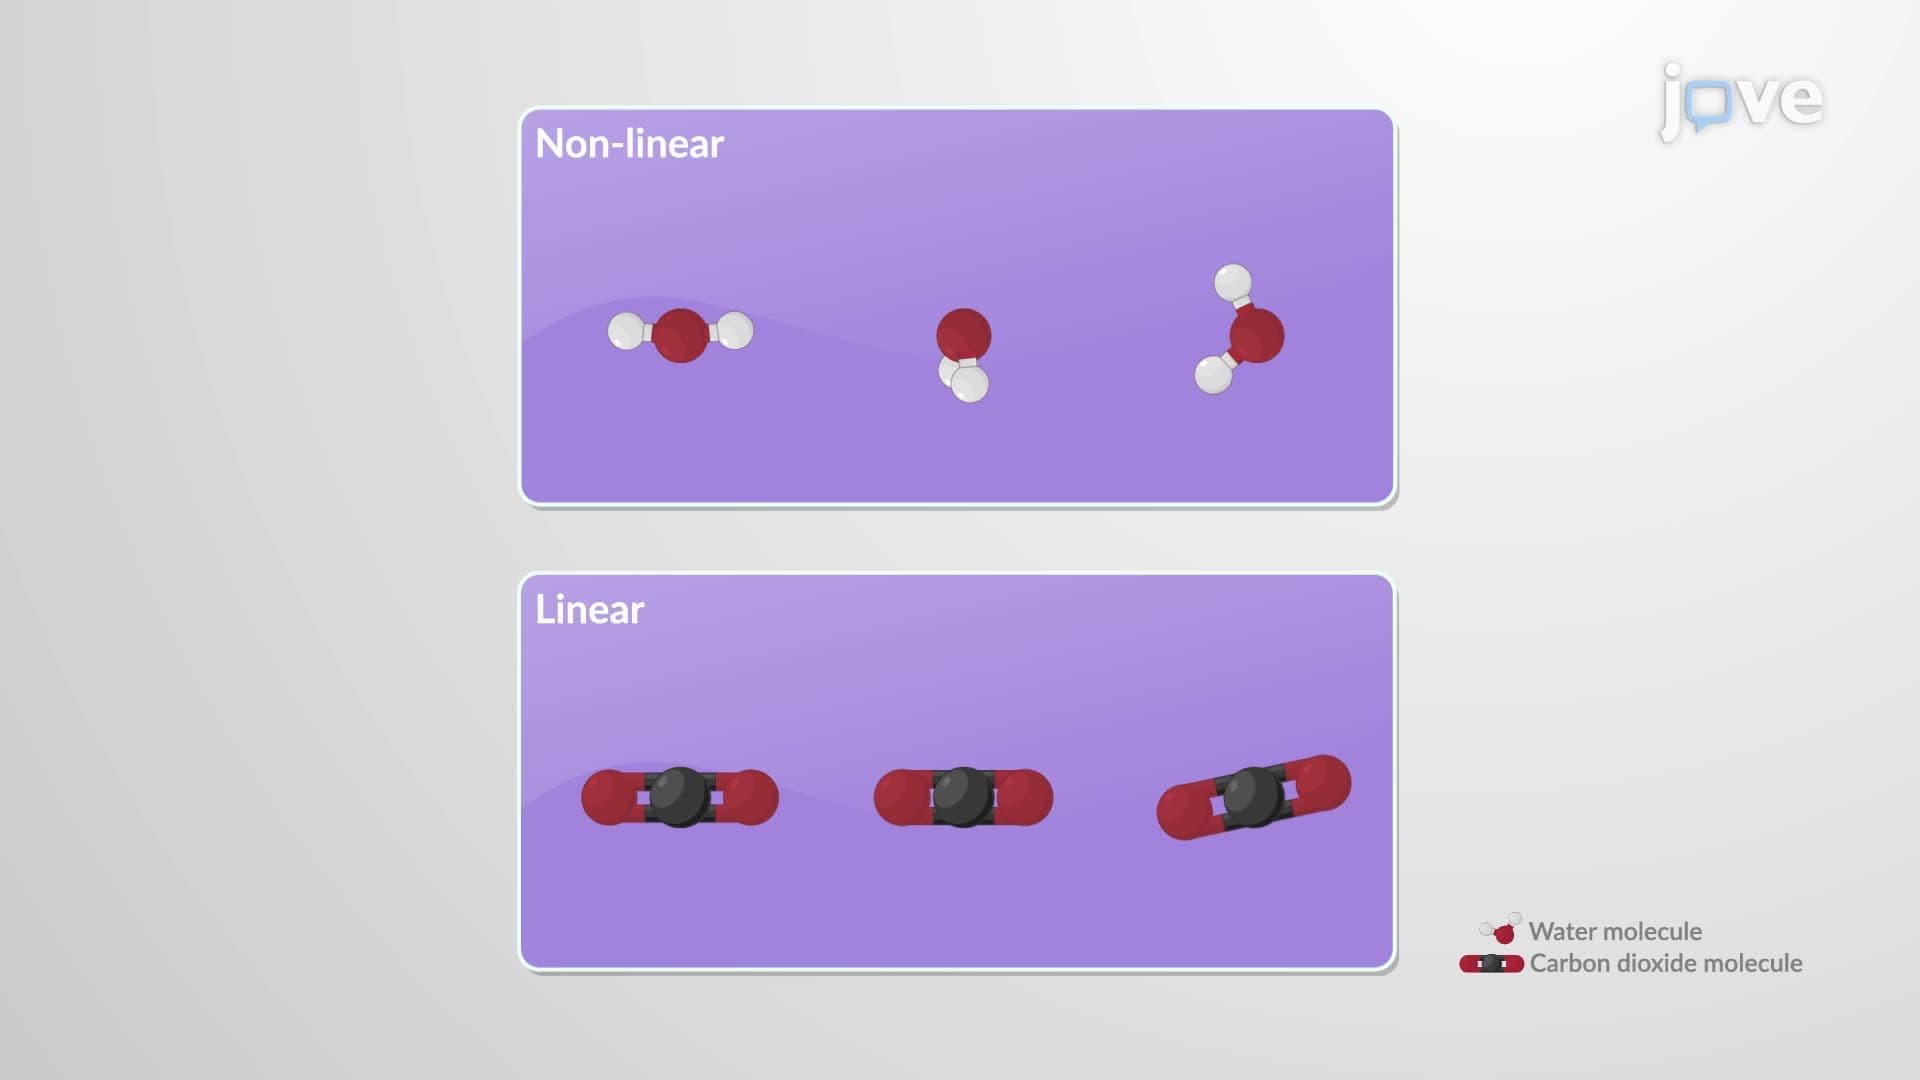
The Molecular Nature of Internal Energy
 02:49
Bond Energies and Bond Lengths
 02:13
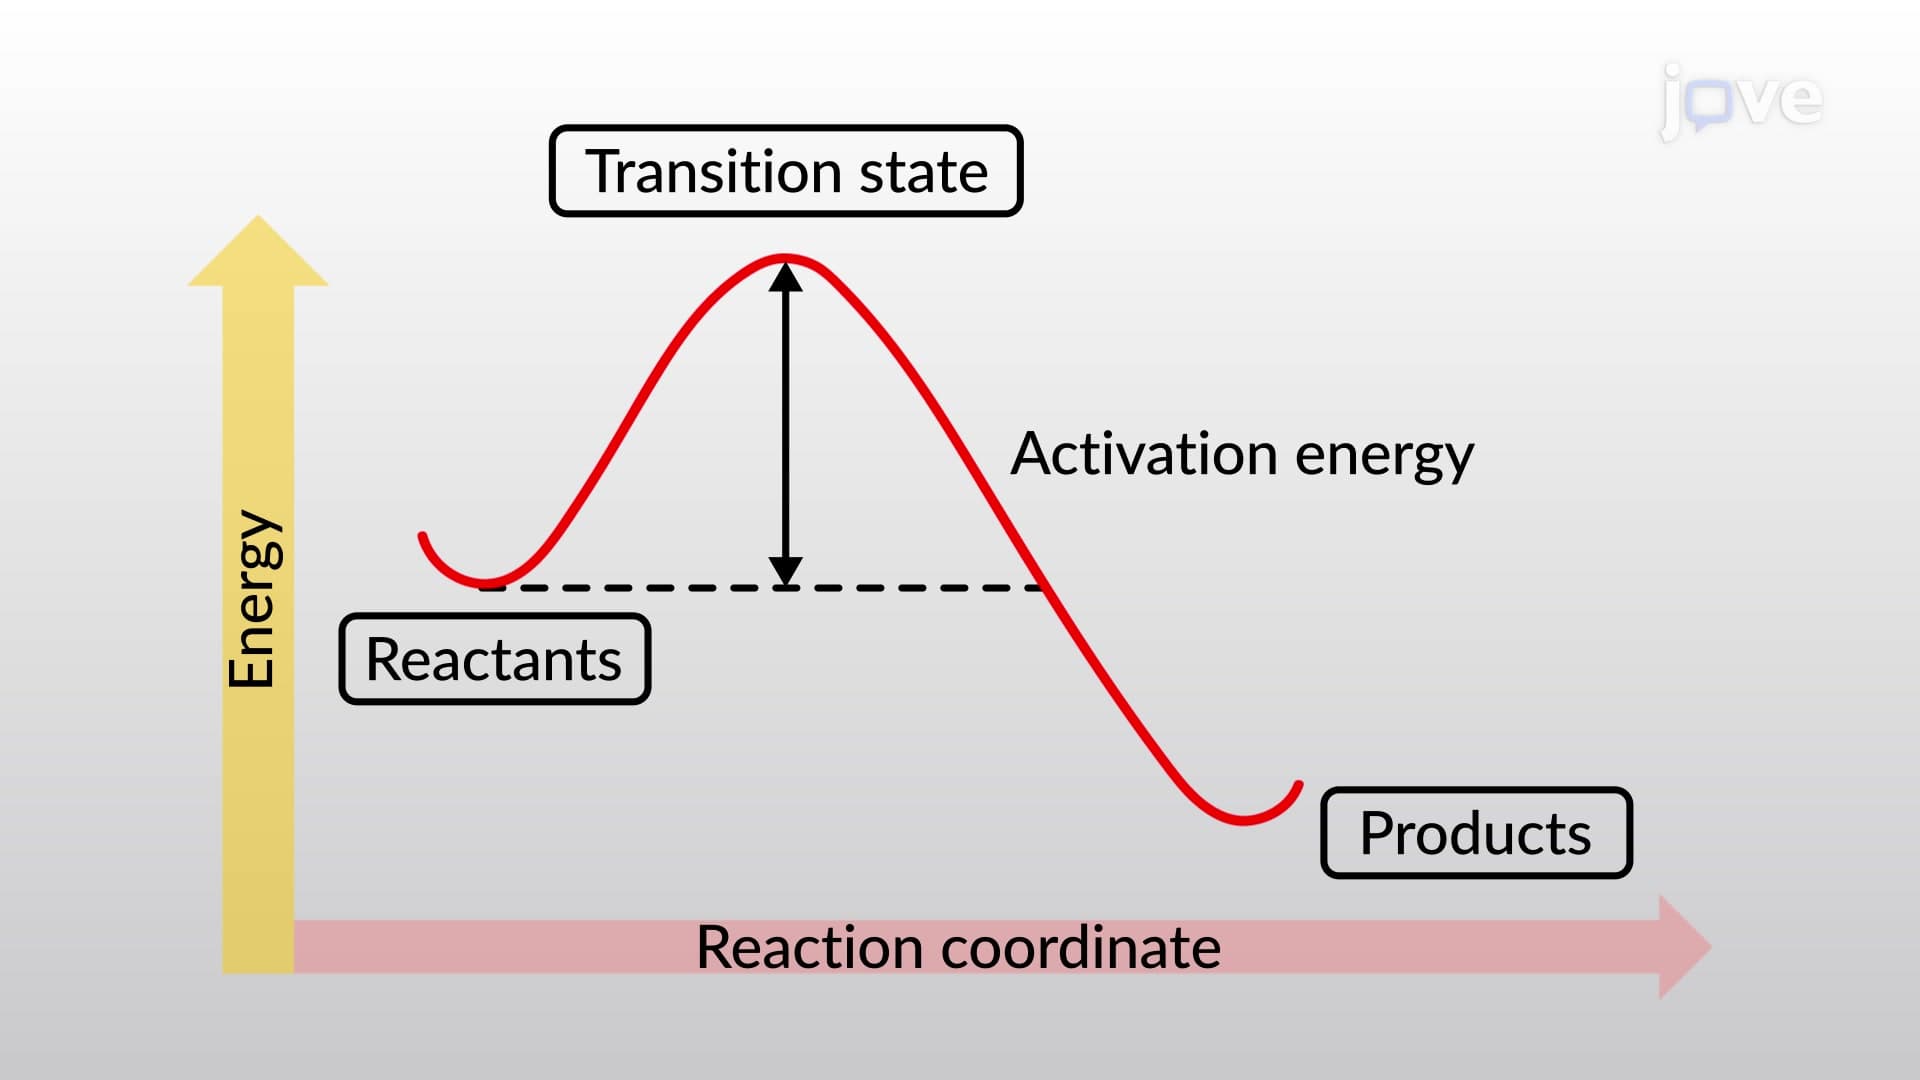
Bond Dissociation Energy and Activation Energy
 00:54
Radical Formation: Homolysis
 01:16
Cycloaddition Reactions: MO Requirements for Thermal Activation
 03:51
Molecular Orbital Theory II