Related Experiment Video
Updated: Jul 10, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Method for efficient computation of the density of states in water-explicit biopolymer simulations on a lattice
Bryan A Patel1, Pablo G Debenedetti, Frank H Stillinger
1Department of Chemical Engineering, and Department of Chemistry, Princeton University, Princeton, New Jersey 08544, USA.

Abstract:
We present a method for fast computation of the density of states of binary systems. The contributions of each of the components to the density of states can be separated based on the conditional independence of the individual components' degrees of freedom. The conditions establishing independence are the degrees of freedom of the interfacial region between the two components. The separate contributions of the components to the density of states can then be calculated using the Wang-Landau algorithm [Wang, F.; Landau, D. P. Phys. Rev. Lett. 2001, 86, 2050]. We apply this method to a 2D lattice model of a hydrophobic homopolymer in water that exhibits protein-like cold, pressure, and thermal unfolding. The separate computation of the protein and water density of states contributions is faster and more accurate than the combined simulation of both components and allows for the investigation of larger systems.
Related Concept Videos
 01:19
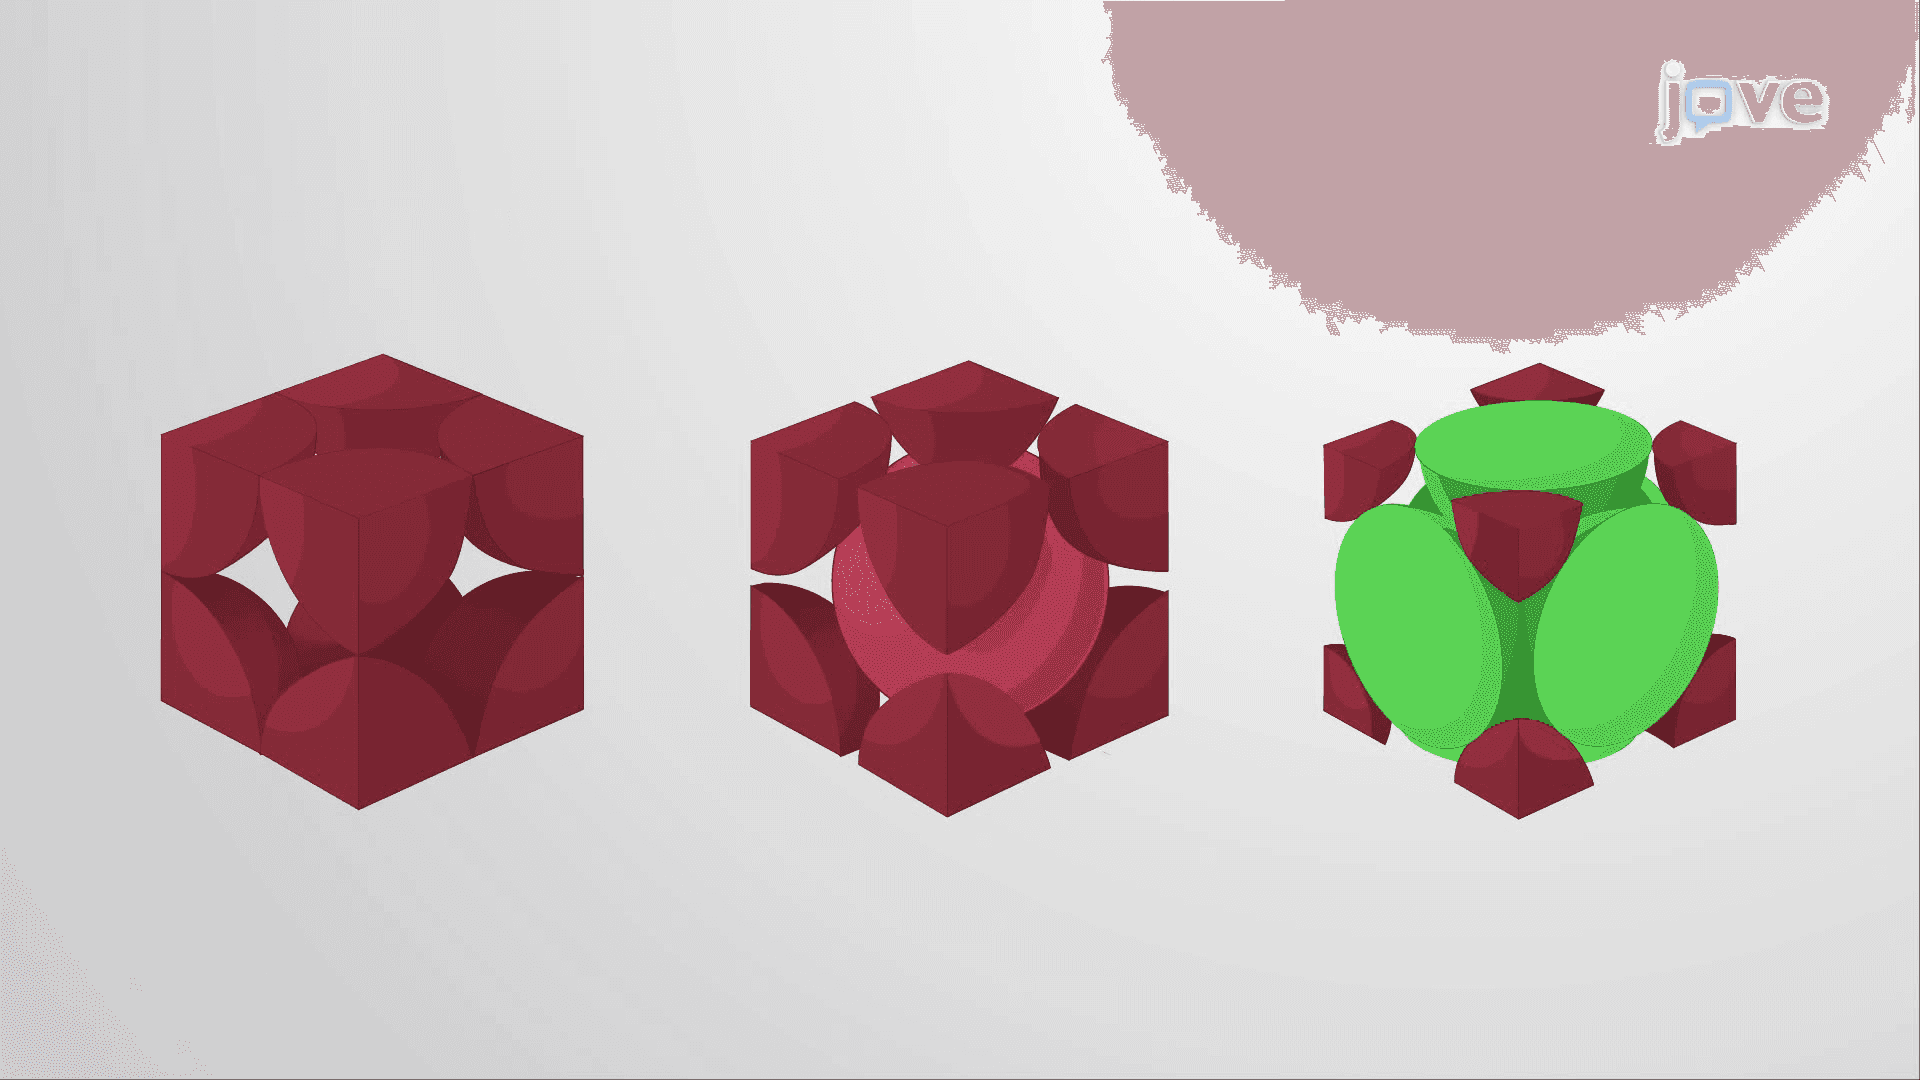
Crystal Density
 02:54
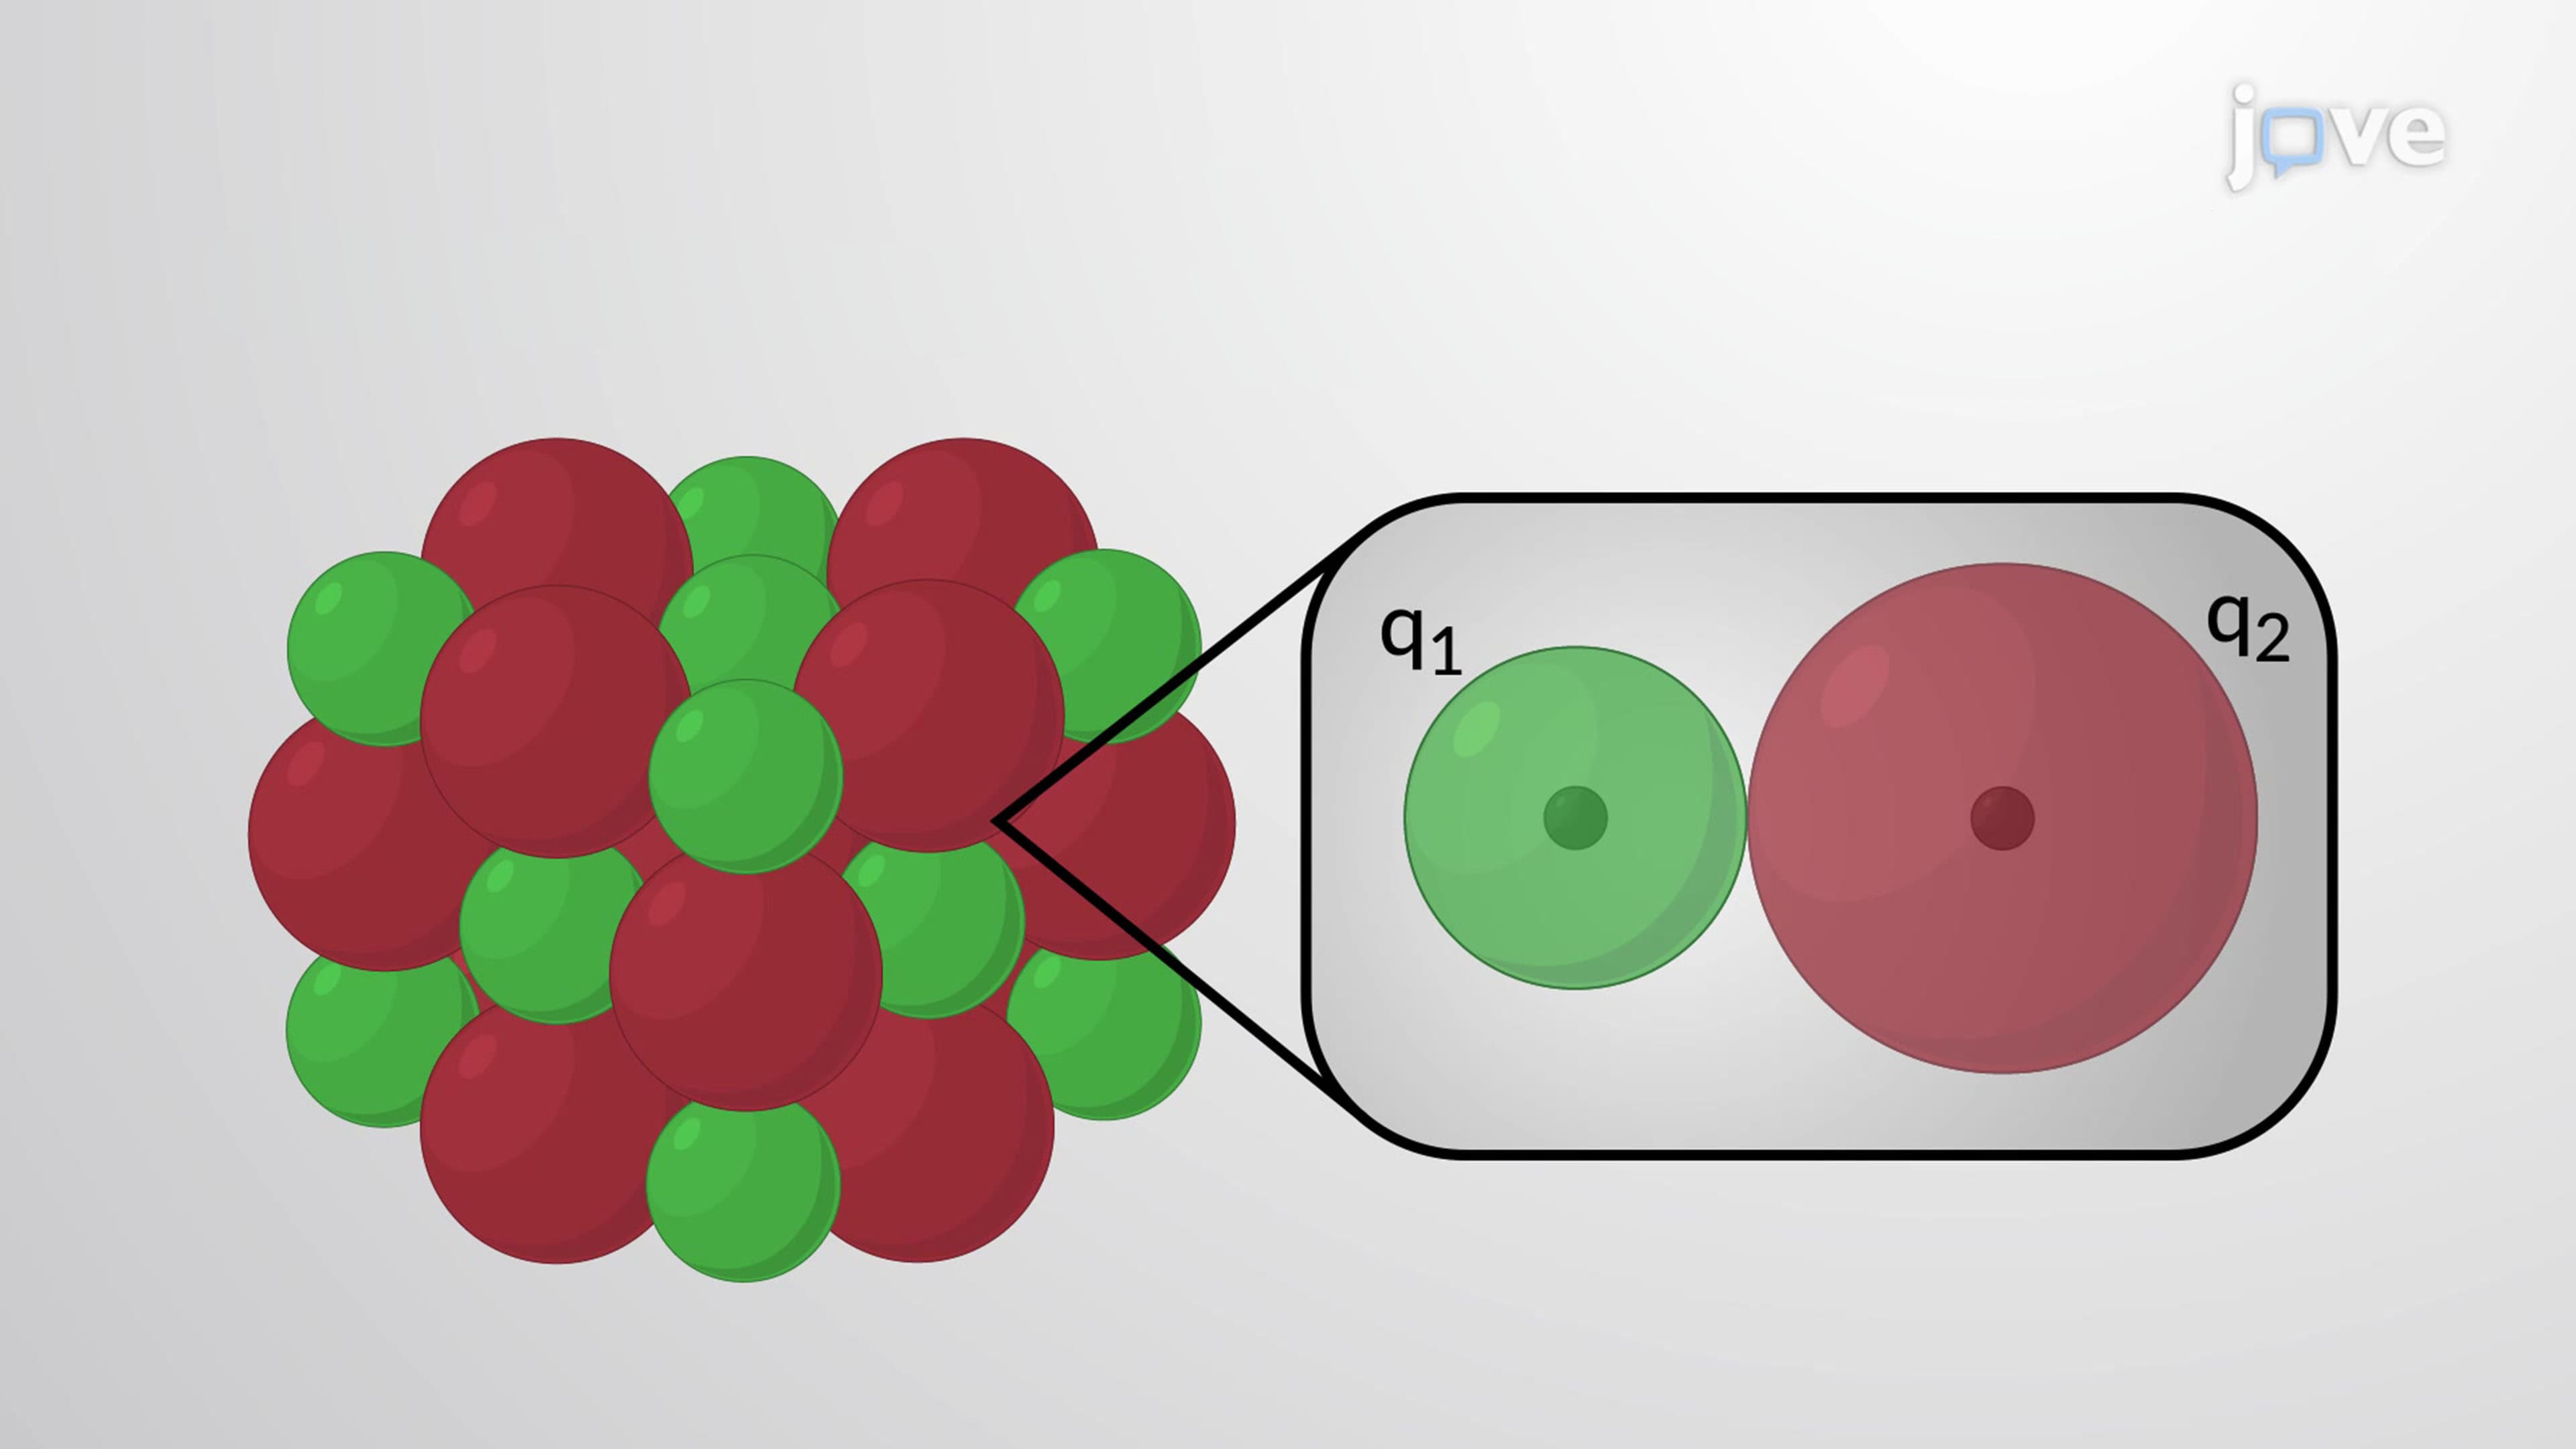
Trends in Lattice Energy: Ion Size and Charge
 01:28
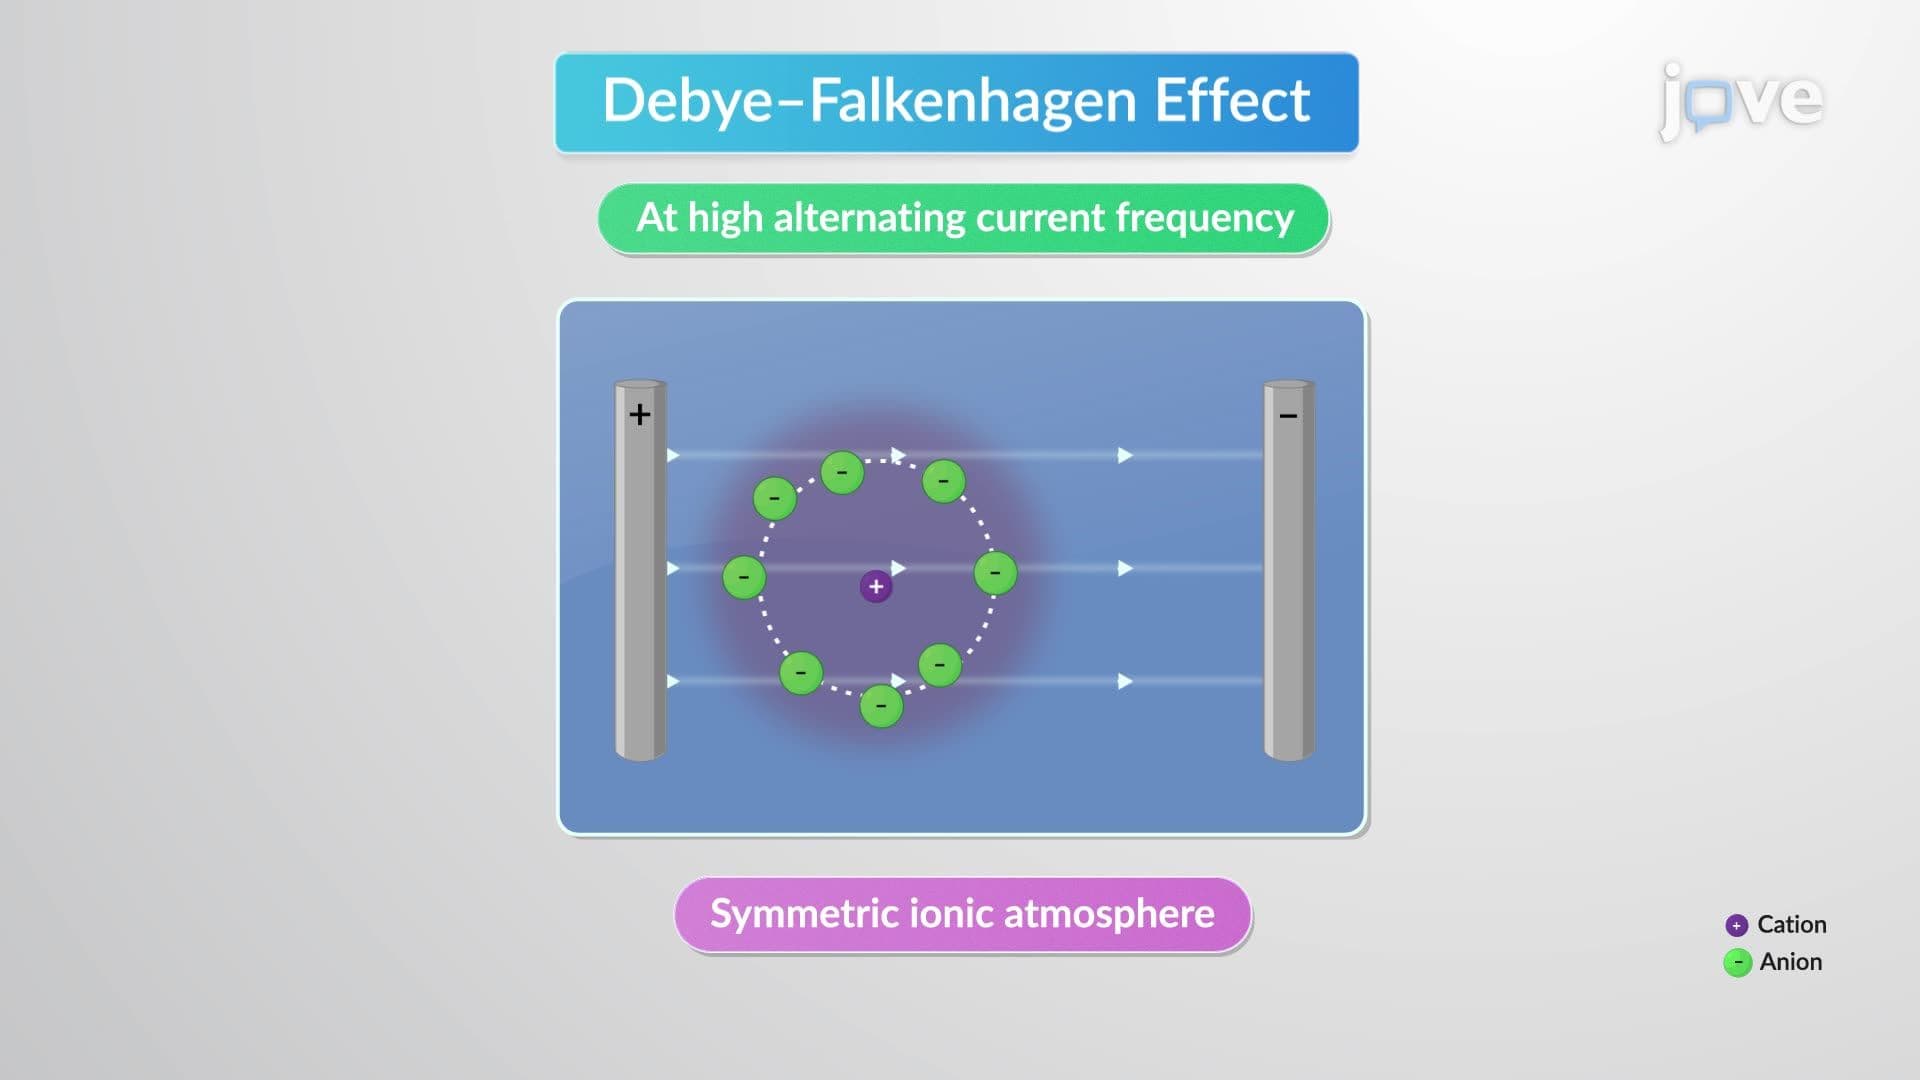
Debye–Huckel–Onsager Conductance Equation
 01:27
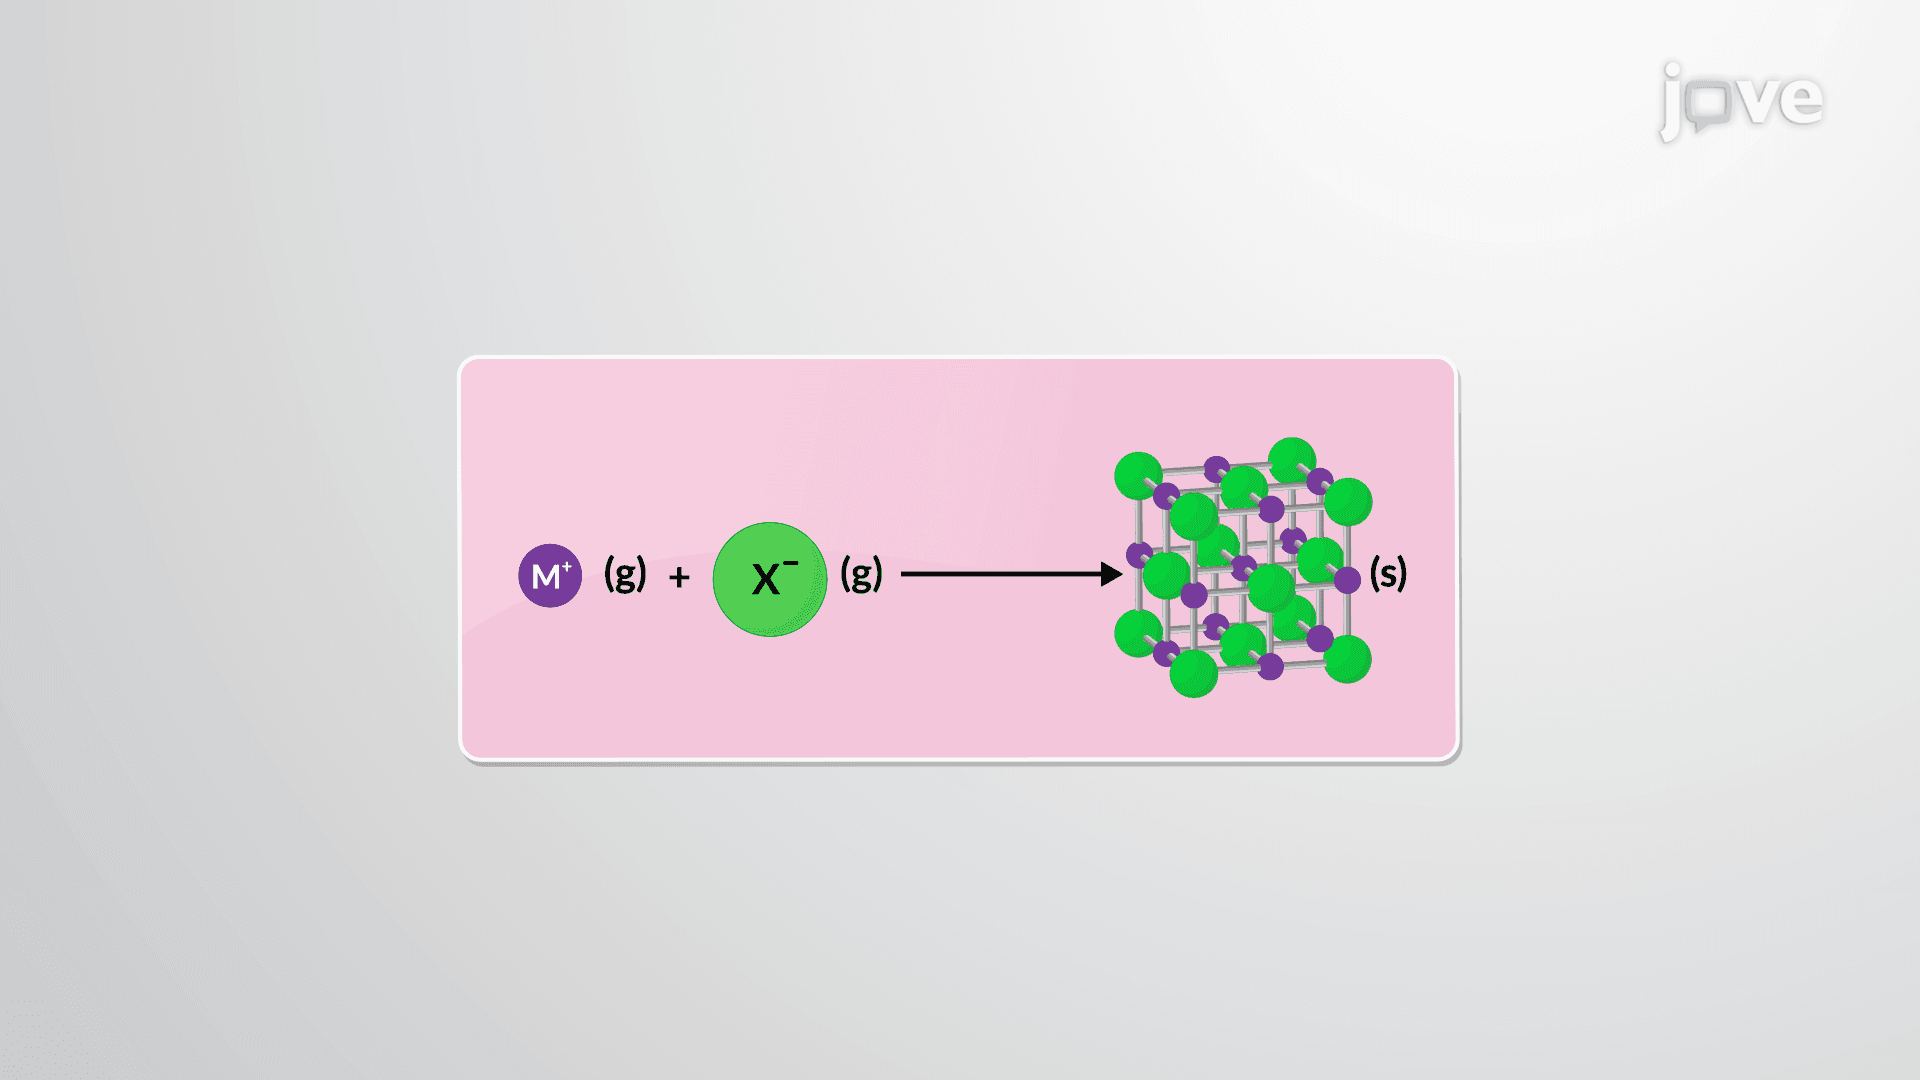
Lattice Energies of Ionic Crystals
 01:08
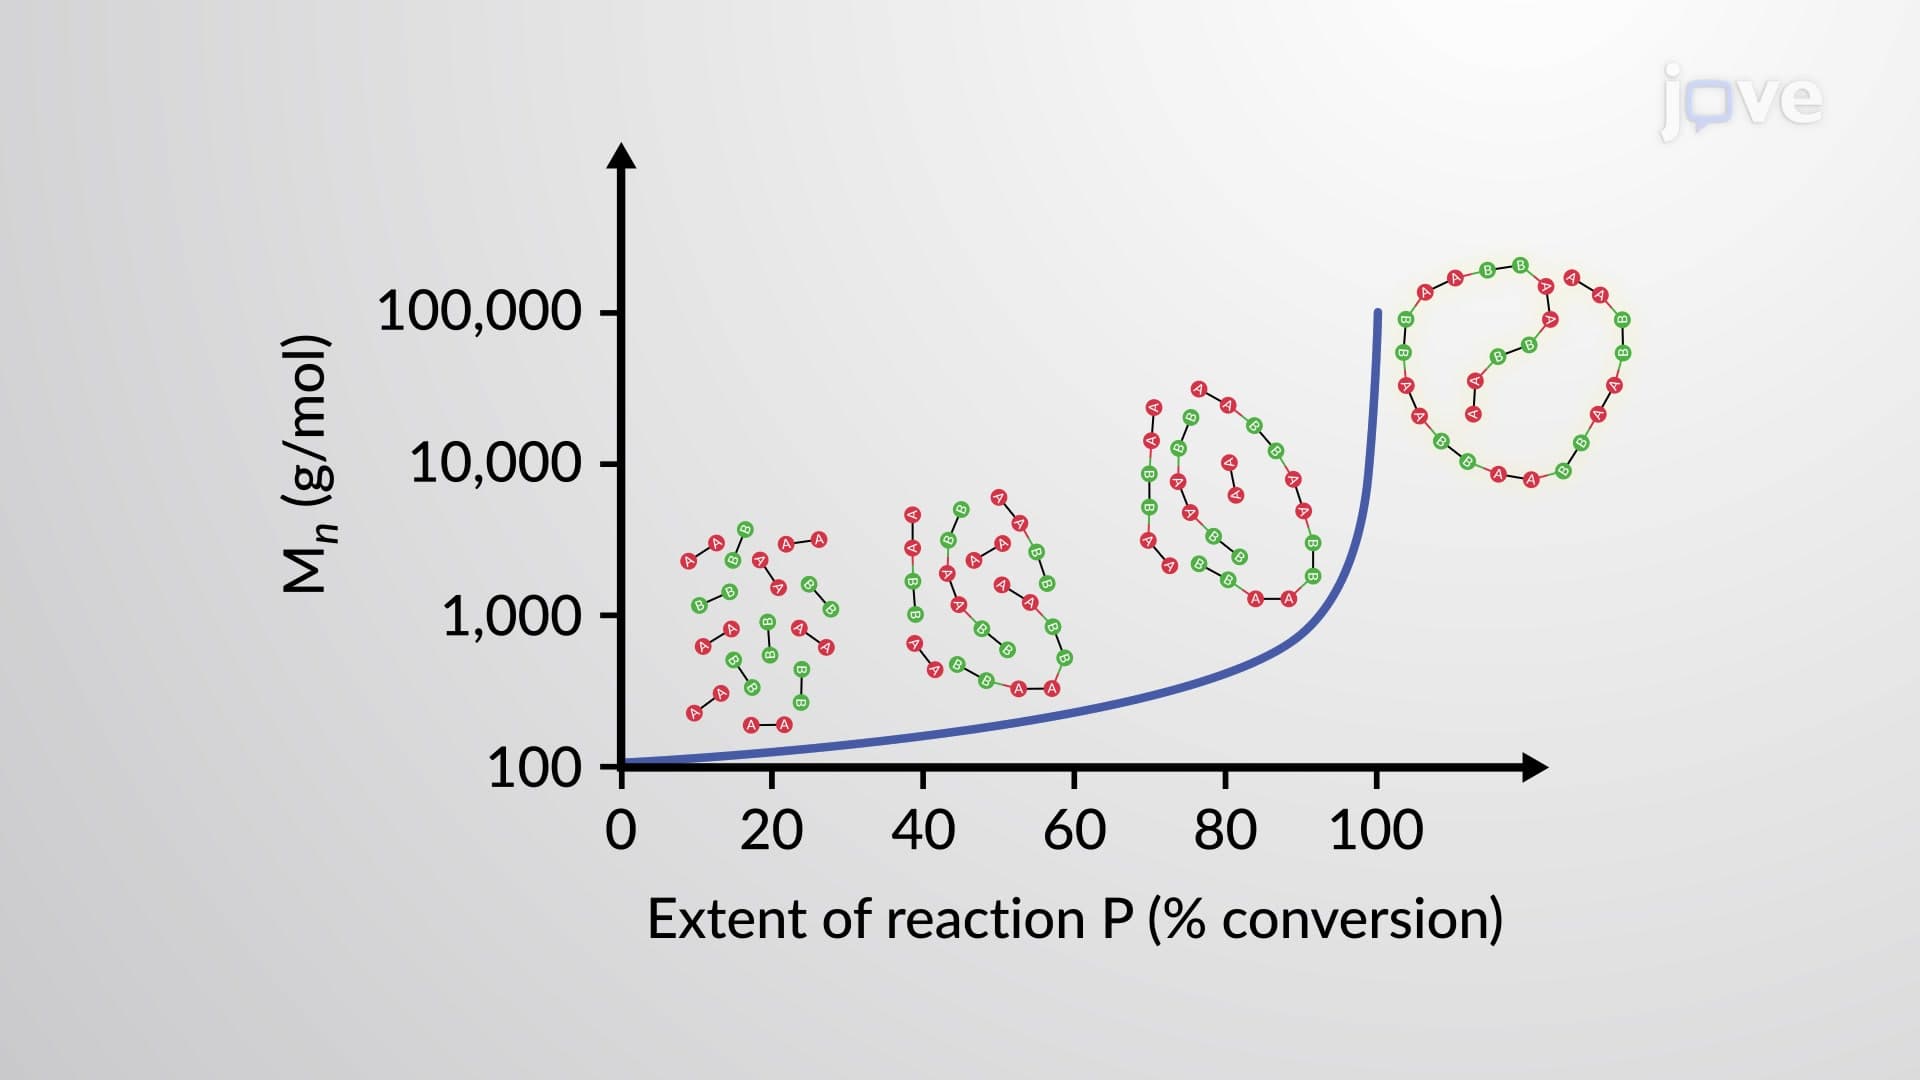
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:10
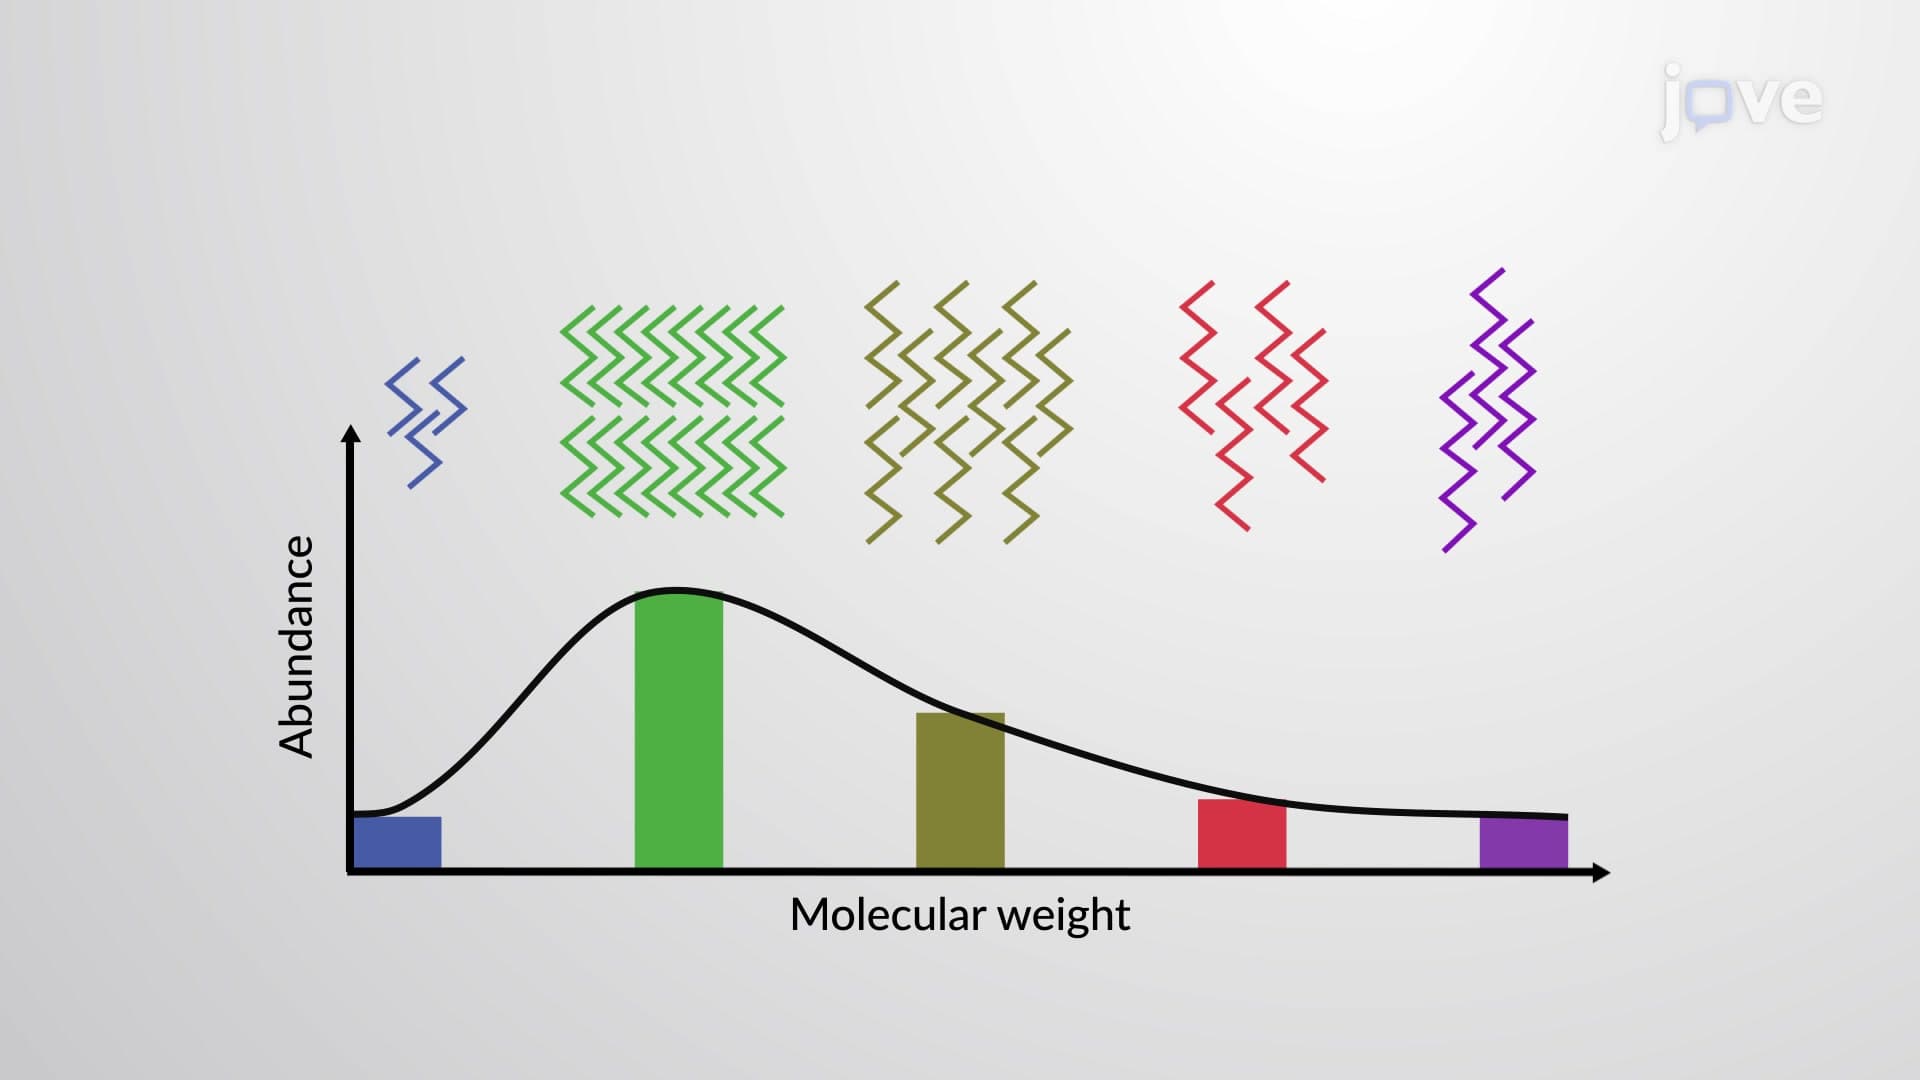
Polymers: Molecular Weight Distribution