Related Experiment Video
Updated: Jul 9, 2026
Measuring Glucose Uptake in Drosophila Models of TDP-43 Proteinopathy
Published on: August 3, 2021
Glucose-6-phosphate dehydrogenase Buenos Aires: a novel de novo missense mutation associated with severe enzyme
Angelo Minucci1, Paola Concolino, Francesca Vendittelli
1Laboratory of Clinical Molecular Biology Institute of Biochemistry and Clinical Biochemistry, Catholic University of Rome, Italy.

Objective:
: Glucose 6-phosphate dehydrogenase (G6PD) catalyzes the first committed steps in the pentose phosphate pathway: the generation of NADPH by this enzyme is essential for protection against oxidative stress. The human enzyme is in a dimer<-->tetramer equilibrium and its stability depends on NADP(+) concentration. Herein, we report a case of a symptomatic baby affected by severe deficiency of G6PD activity due to a novel de novo genetic mutation (g1465C>T) in the thirteenth exon of its gene.
Methods:
: Clinical, biochemical and genetic evaluations of the affected baby and his mother were performed.
Results:
: We found the g1465C>T novel mutation, in the thirteenth exon of G6PD gene (named "G6PD Buenos Aires variant"). This g1465C>T mutation produce a P489S substitution at protein level. The P489S mutation was absent in his mother, suggesting that G6PD Buenos Aires resulted from a de novo mutation.
Conclusions:
: The absence of mosaicism in the baby's DNA (from saliva and blood samples) suggests that a de novo mutation event may occur in the very early stages in embryogenesis or in the mother's germ cell lines.
More Related Videos
 09:34
Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
 13:35
Structural Biology and Analytical Chemistry Approaches for Characterizing C-Glycoside Metabolic Enzymes in Human Gut Microbiota
Published on: May 23, 2025
Related Concept Videos
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:20
Inborn Errors of Metabolism
 01:02
Glucose Homeostasis: Regulation of Blood Glucose
During fasting, when blood glucose levels are low, the pancreas secretes glucagon. it...
 01:21
Huntington Disease l: Introduction
 02:59
Animal Mitochondrial Genetics
 01:21
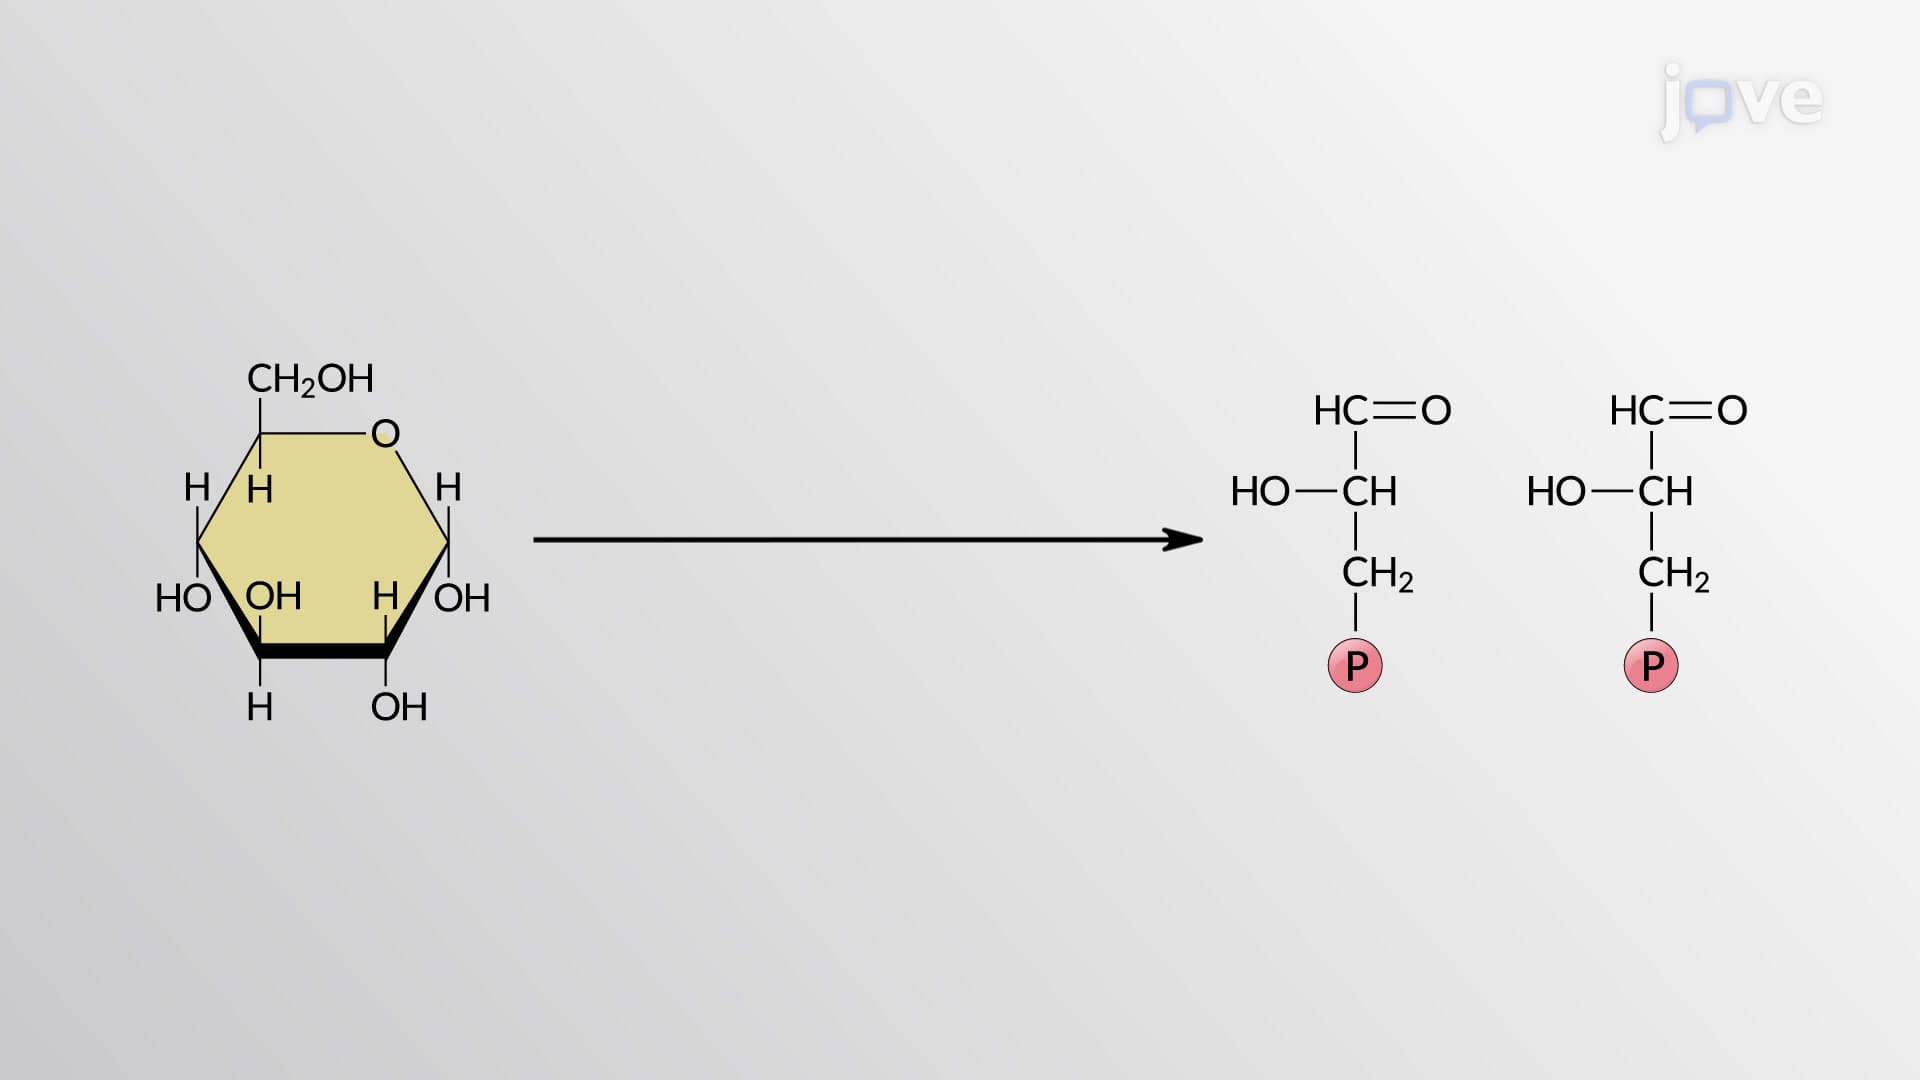
Glycolysis: Preparatory Phase