Related Experiment Video
Updated: Jul 9, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
CONTRAST: a discriminative, phylogeny-free approach to multiple informant de novo gene prediction
Samuel S Gross1, Chuong B Do, Marina Sirota
1Computer Science Department, Stanford University, Stanford, CA, USA. ssgross@cs.stanford.edu

Abstract:
We describe CONTRAST, a gene predictor which directly incorporates information from multiple alignments rather than employing phylogenetic models. This is accomplished through the use of discriminative machine learning techniques, including a novel training algorithm. We use a two-stage approach, in which a set of binary classifiers designed to recognize coding region boundaries is combined with a global model of gene structure. CONTRAST predicts exact coding region structures for 65% more human genes than the previous state-of-the-art method, misses 46% fewer exons and displays comparable gains in specificity.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 03:21
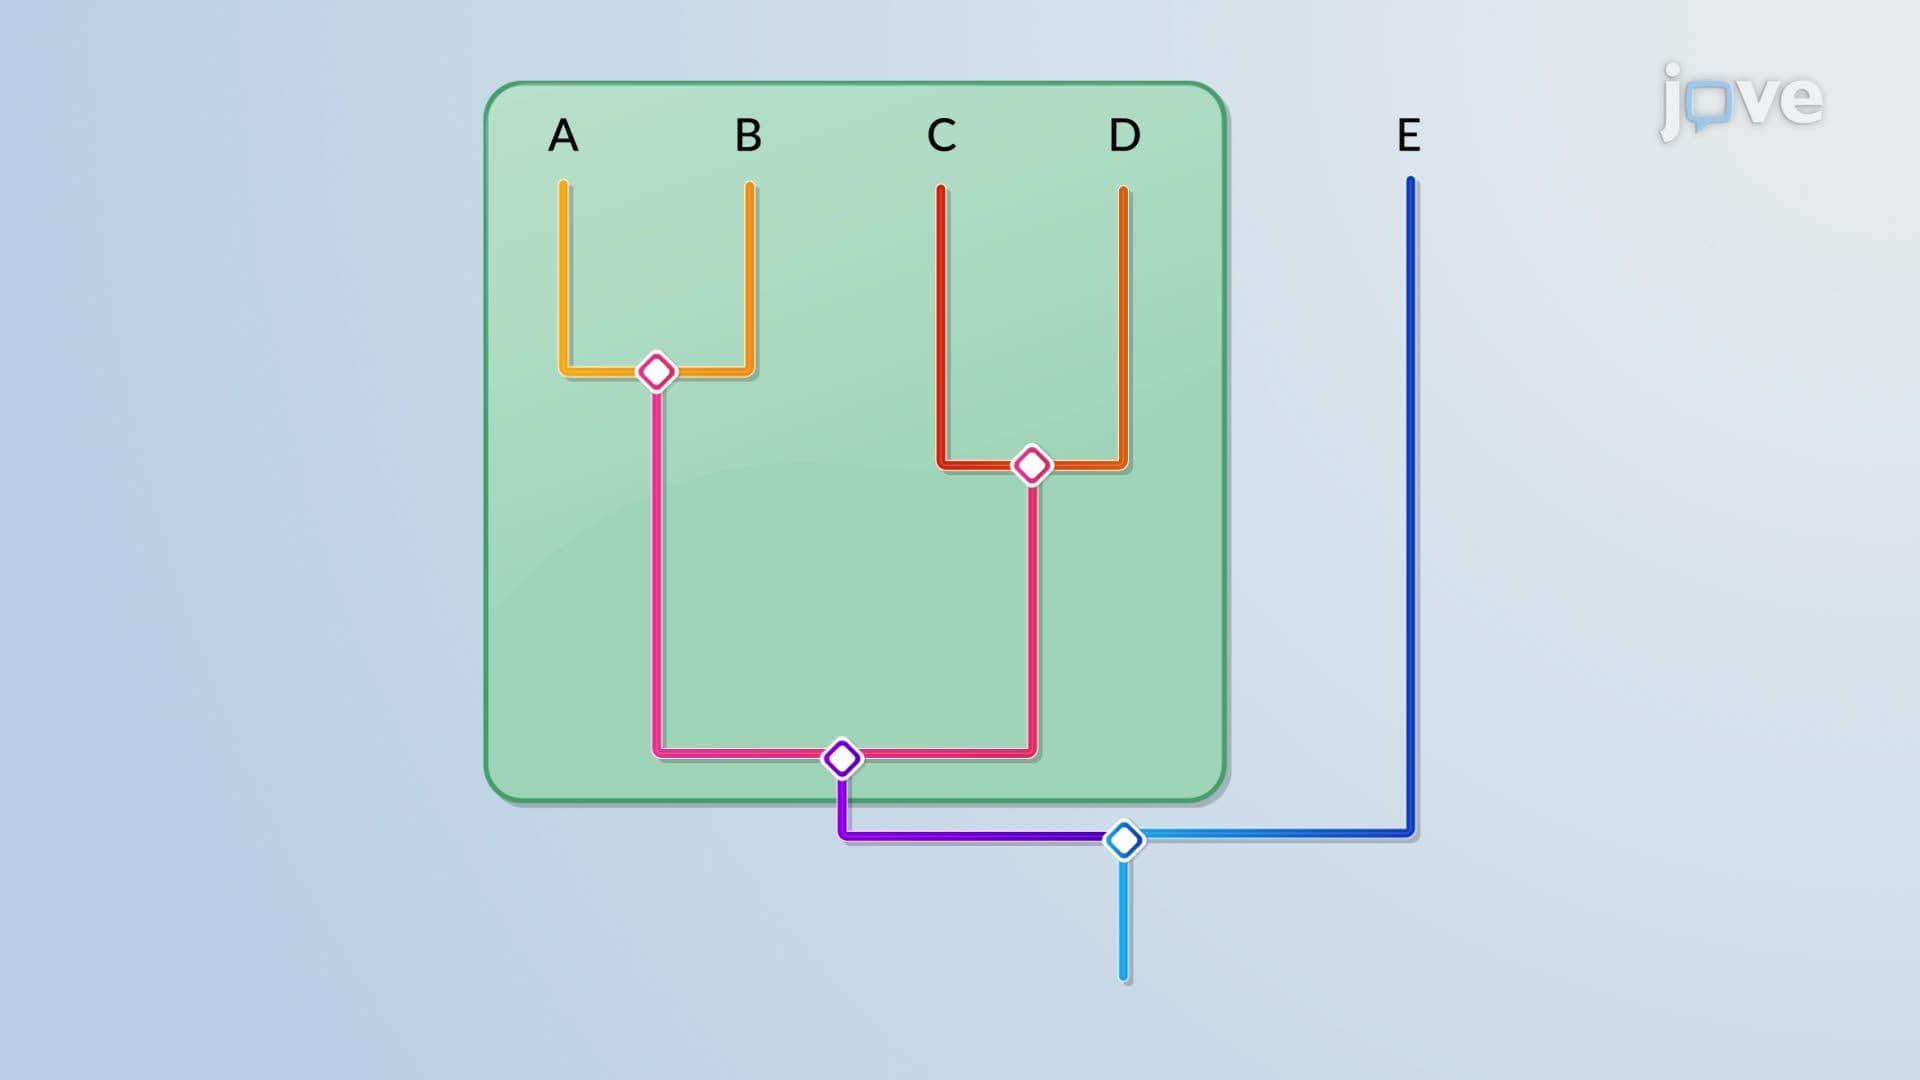
Phylogenetic Trees
03:21Phylogenetic Trees
 02:37
Gene Duplication and Divergence
The duplicated copies of the gene are called Paralogs. Paralogs with similar sequences and functions form a gene family. Across several species, a large number of gene families are characterized.