Related Experiment Video
Updated: Jul 8, 2026

Modeling Charcot-Marie-Tooth Disease In Vitro by Transfecting Mouse Primary Motoneurons
Published on: January 7, 2019
POLG1 mutations manifesting as autosomal recessive axonal Charcot-Marie-Tooth disease
Timothy Harrower1, Joanna D Stewart, Gavin Hudson
1Department of Neurology, Essex Center for Neurological Sciences, Queen's Hospital, Romford, England.

Background:
Although a molecular diagnosis is possible in most patients having Charcot-Marie-Tooth disease (CMT), recessively inherited and axonal neuropathies still present a diagnostic challenge.
Objective:
To determine the cause of axonal CMT type 2 in 3 siblings.
Design:
Case report.
Setting:
Academic research.
Participants:
Three siblings who subsequently developed profound cerebellar ataxia.
Main Outcome Measures:
Muscle biopsy specimen molecular genetic analysis of the POLG1 (polymerase gamma-1) gene, as well as screening of control subjects for POLG1 sequence variants.
Results:
Cytochrome c oxidase deficient fibers and multiple deletions of mitochondrial DNA were detected in skeletal muscle. Three compound heterozygous substitutions were detected in POLG1.
Conclusion:
Even in the absence of classic features of mitochondrial disease, POLG1 should be considered in patients having axonal CMT that may be associated with tremor or ataxia.
More Related Videos
 06:35
In Vivo Electrophysiological Measurement of Compound Muscle Action Potential from the Forelimbs in Mouse Models of Motor Neuron Degeneration
Published on: June 15, 2018
 06:49
A Quick Phenotypic Neurological Scoring System for Evaluating Disease Progression in the SOD1-G93A Mouse Model of ALS
Published on: October 6, 2015
Related Concept Videos
 01:35
Pedigree Analysis
 01:33
Pleiotropy
 01:21
Huntington Disease l: Introduction
 01:24
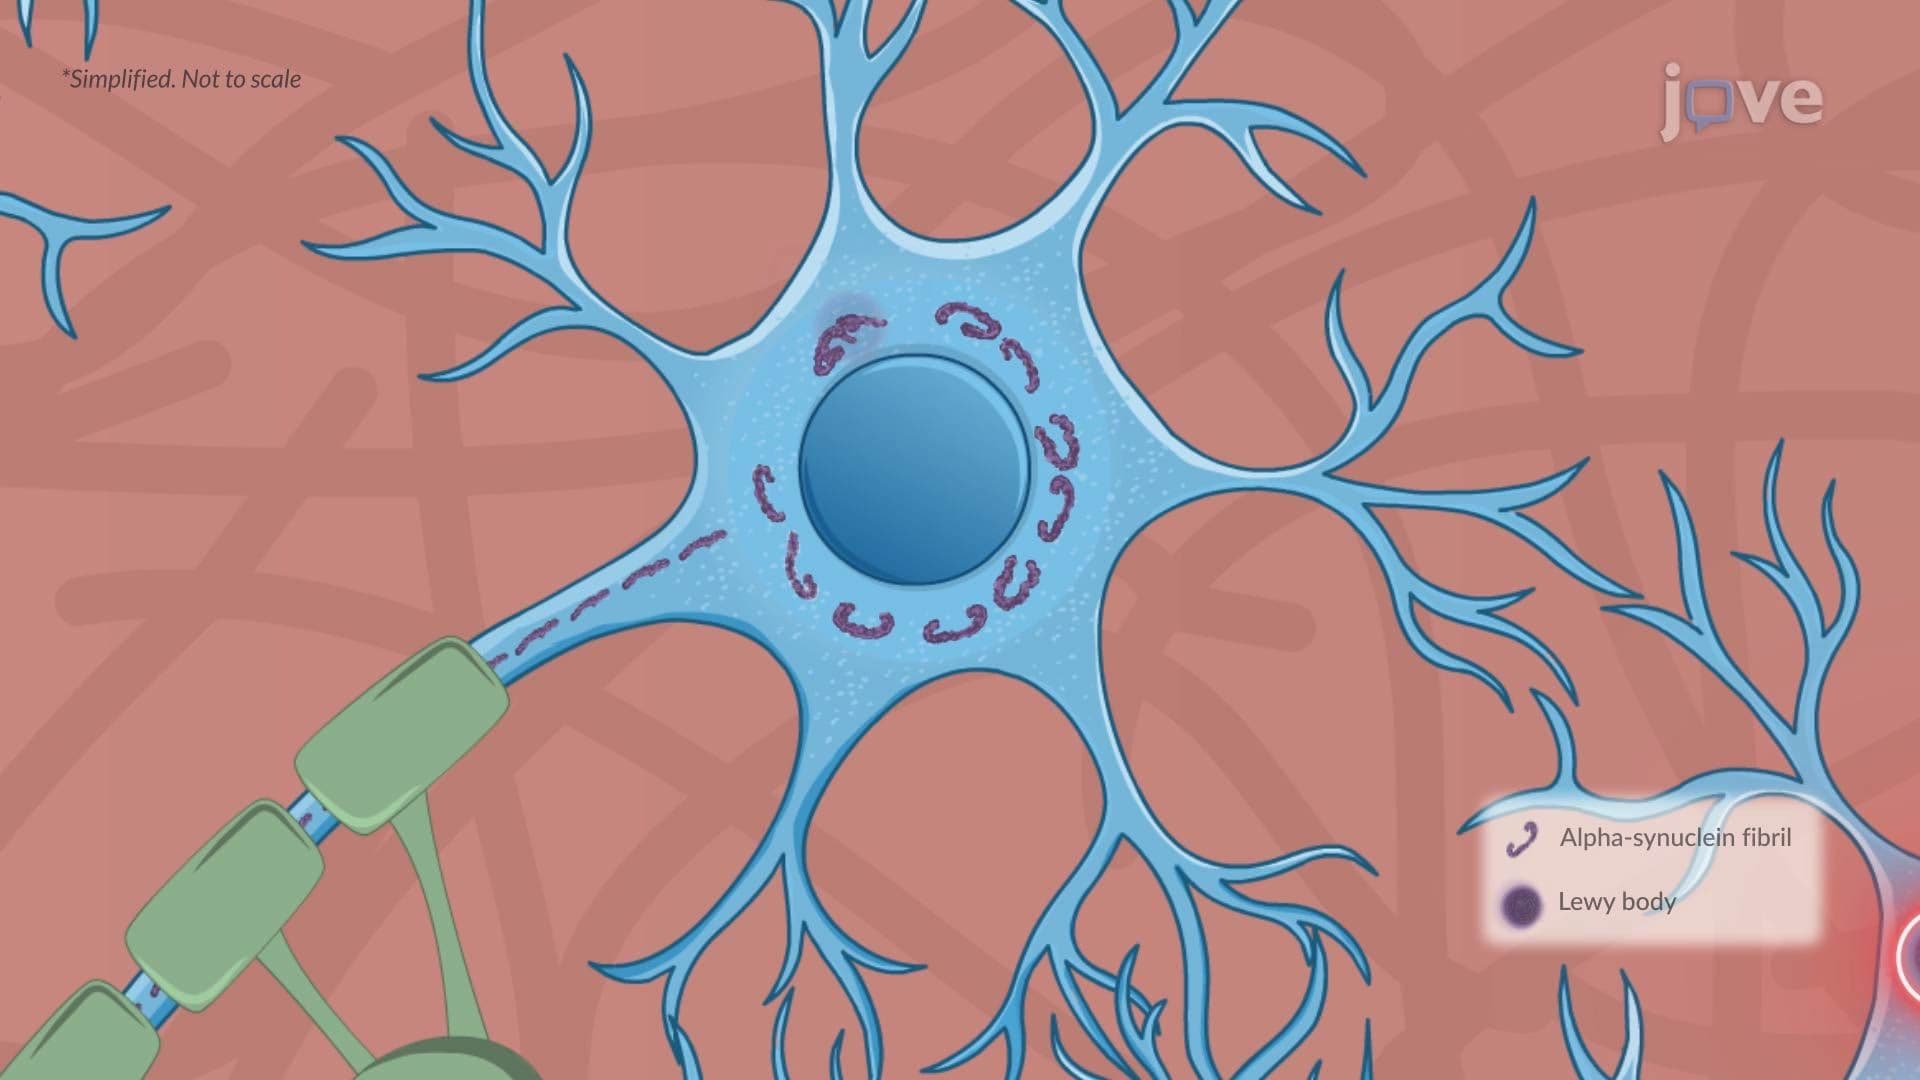
Parkinson Disease ll: Pathophysiology
 01:20
Inborn Errors of Metabolism
 01:43
Sex-linked Disorders