Related Experiment Video
Updated: Jul 8, 2026
Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Celecoxib is a CYP1A2 inhibitor in vitro but not in vivo
Marjo J Karjalainen1, Pertti J Neuvonen, Janne T Backman
1Department of Clinical Pharmacology, University of Helsinki, Helsinki, Finland.

Background And Objective:
We recently discovered that rofecoxib is a potent mechanism-based inhibitor of CYP1A2. The effect of the widely used cyclo-oxygenase-2 selective non-steroidal anti-inflammatory drug celecoxib on CYP1A2 activity has not been reported.
Methods:
The effect of celecoxib on CYP1A2 activity (phenacetin O-deethylation) was first studied in vitro using human liver microsomes. This was followed by a randomized, placebo-controlled, cross-over study in which 12 healthy volunteers were given celecoxib (200 mg twice daily) or placebo for 4 days. On day 3, a caffeine test was performed. On day 4, the subjects ingested 2 mg tizanidine. Plasma samples for the measurement of the concentrations of tizanidine, its metabolites and celecoxib were collected up to 24 h post-administration. Pharmacodynamic variables (e.g. blood pressure, subjective drowsiness and drug effect) were recorded up to 12 h post-adm.
Results:
Celecoxib was found to be a moderately potent competitive inhibitor of CYP1A2 in vitro with a K(i) (inhibitor constant) of 25.4 microM. However, in vivo, celecoxib did not affect the caffeine test, or the peak concentration, time to peak concentration, area under the concentration-time curve or half-life of tizanidine. The pharmacodynamic variables of tizanidine also remained unchanged.
Conclusions:
Unlike rofecoxib, celecoxib does not clinically to significantly inhibit CYP1A2. The lack of significant in vivo inhibition of CYP1A2 can be correctly predicted on the basis of in vitro K(i) data and the free peripheral or portal plasma concentration of celecoxib.
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:20
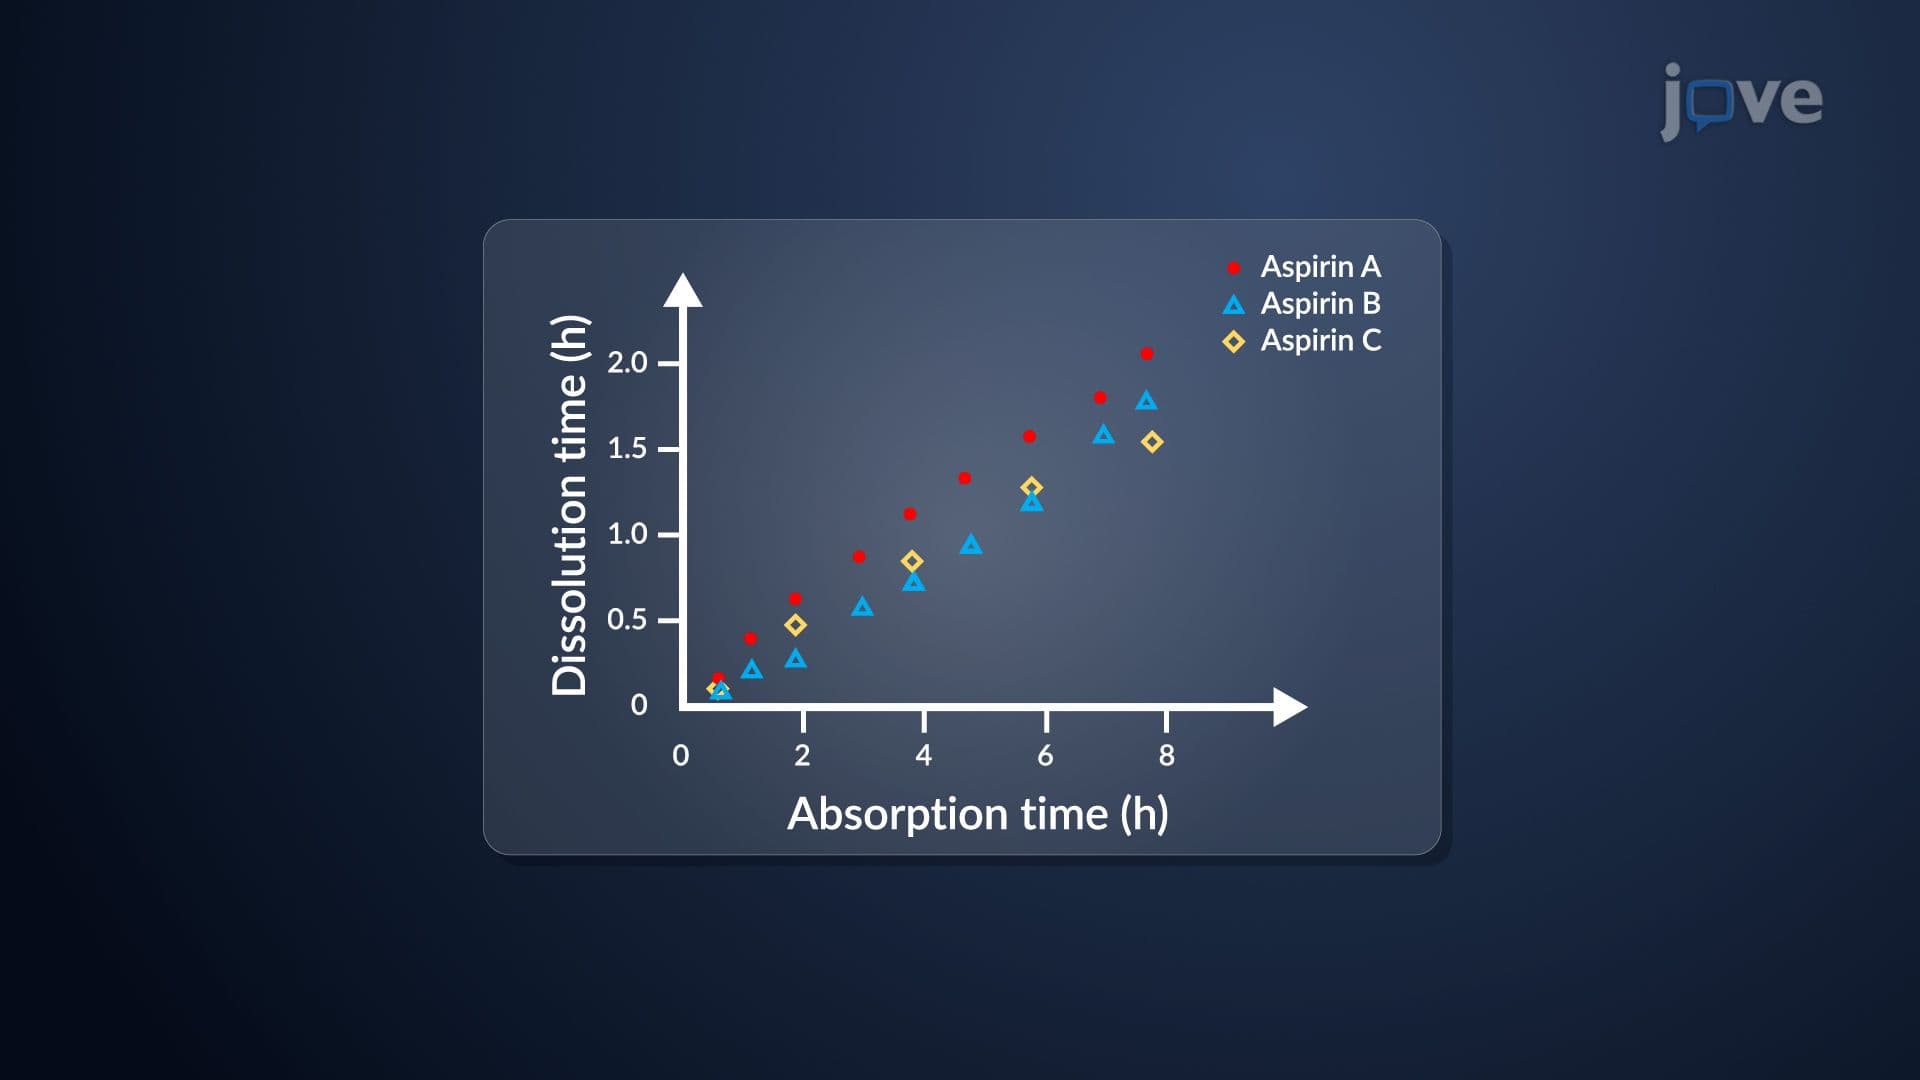
Drug Product Performance: In Vitro–In Vivo Correlation
 01:16
Methods for Studying Drug Absorption: In vitro
The diffusion cell method uses a two-compartment cell, including a donor compartment with the drug solution, which simulates the environment where the drug is applied, and a receptor compartment with a buffer solution, which simulates the environment...
 01:17
Hepatic Drug Excretion: Enterohepatic Cycling
Post-release drugs and metabolites can be reabsorbed into the body from the intestine. For conjugated metabolites like glucuronides, reabsorption requires enzymatic hydrolysis by intestinal microflora. This...
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:16
Hepatic Drug Excretion: Influencing Factors