Related Experiment Video
Updated: Jul 8, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quantum mechanics/molecular mechanics minimum free-energy path for accurate reaction energetics in solution and
Hao Hu1, Zhenyu Lu, Jerry M Parks
1Department of Chemistry, Duke University, Durham, North Carolina 27708, USA.

Abstract:
To accurately determine the reaction path and its energetics for enzymatic and solution-phase reactions, we present a sequential sampling and optimization approach that greatly enhances the efficiency of the ab initio quantum mechanics/molecular mechanics minimum free-energy path (QM/MM-MFEP) method. In the QM/MM-MFEP method, the thermodynamics of a complex reaction system is described by the potential of mean force (PMF) surface of the quantum mechanical (QM) subsystem with a small number of degrees of freedom, somewhat like describing a reaction process in the gas phase. The main computational cost of the QM/MM-MFEP method comes from the statistical sampling of conformations of the molecular mechanical (MM) subsystem required for the calculation of the QM PMF and its gradient. In our new sequential sampling and optimization approach, we aim to reduce the amount of MM sampling while still retaining the accuracy of the results by first carrying out MM phase-space sampling and then optimizing the QM subsystem in the fixed-size ensemble of MM conformations. The resulting QM optimized structures are then used to obtain more accurate sampling of the MM subsystem. This process of sequential MM sampling and QM optimization is iterated until convergence. The use of a fixed-size, finite MM conformational ensemble enables the precise evaluation of the QM potential of mean force and its gradient within the ensemble, thus circumventing the challenges associated with statistical averaging and significantly speeding up the convergence of the optimization process. To further improve the accuracy of the QM/MM-MFEP method, the reaction path potential method developed by Lu and Yang [Z. Lu and W. Yang, J. Chem. Phys. 121, 89 (2004)] is employed to describe the QM/MM electrostatic interactions in an approximate yet accurate way with a computational cost that is comparable to classical MM simulations. The new method was successfully applied to two example reaction processes, the classical SN2 reaction of Cl-+CH3Cl in solution and the second proton transfer step of the reaction catalyzed by the enzyme 4-oxalocrotonate tautomerase. The activation free energies calculated with this new sequential sampling and optimization approach to the QM/MM-MFEP method agree well with results from other simulation approaches such as the umbrella sampling technique with direct QM/MM dynamics sampling, demonstrating the accuracy of the iterative QM/MM-MFEP method.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:19
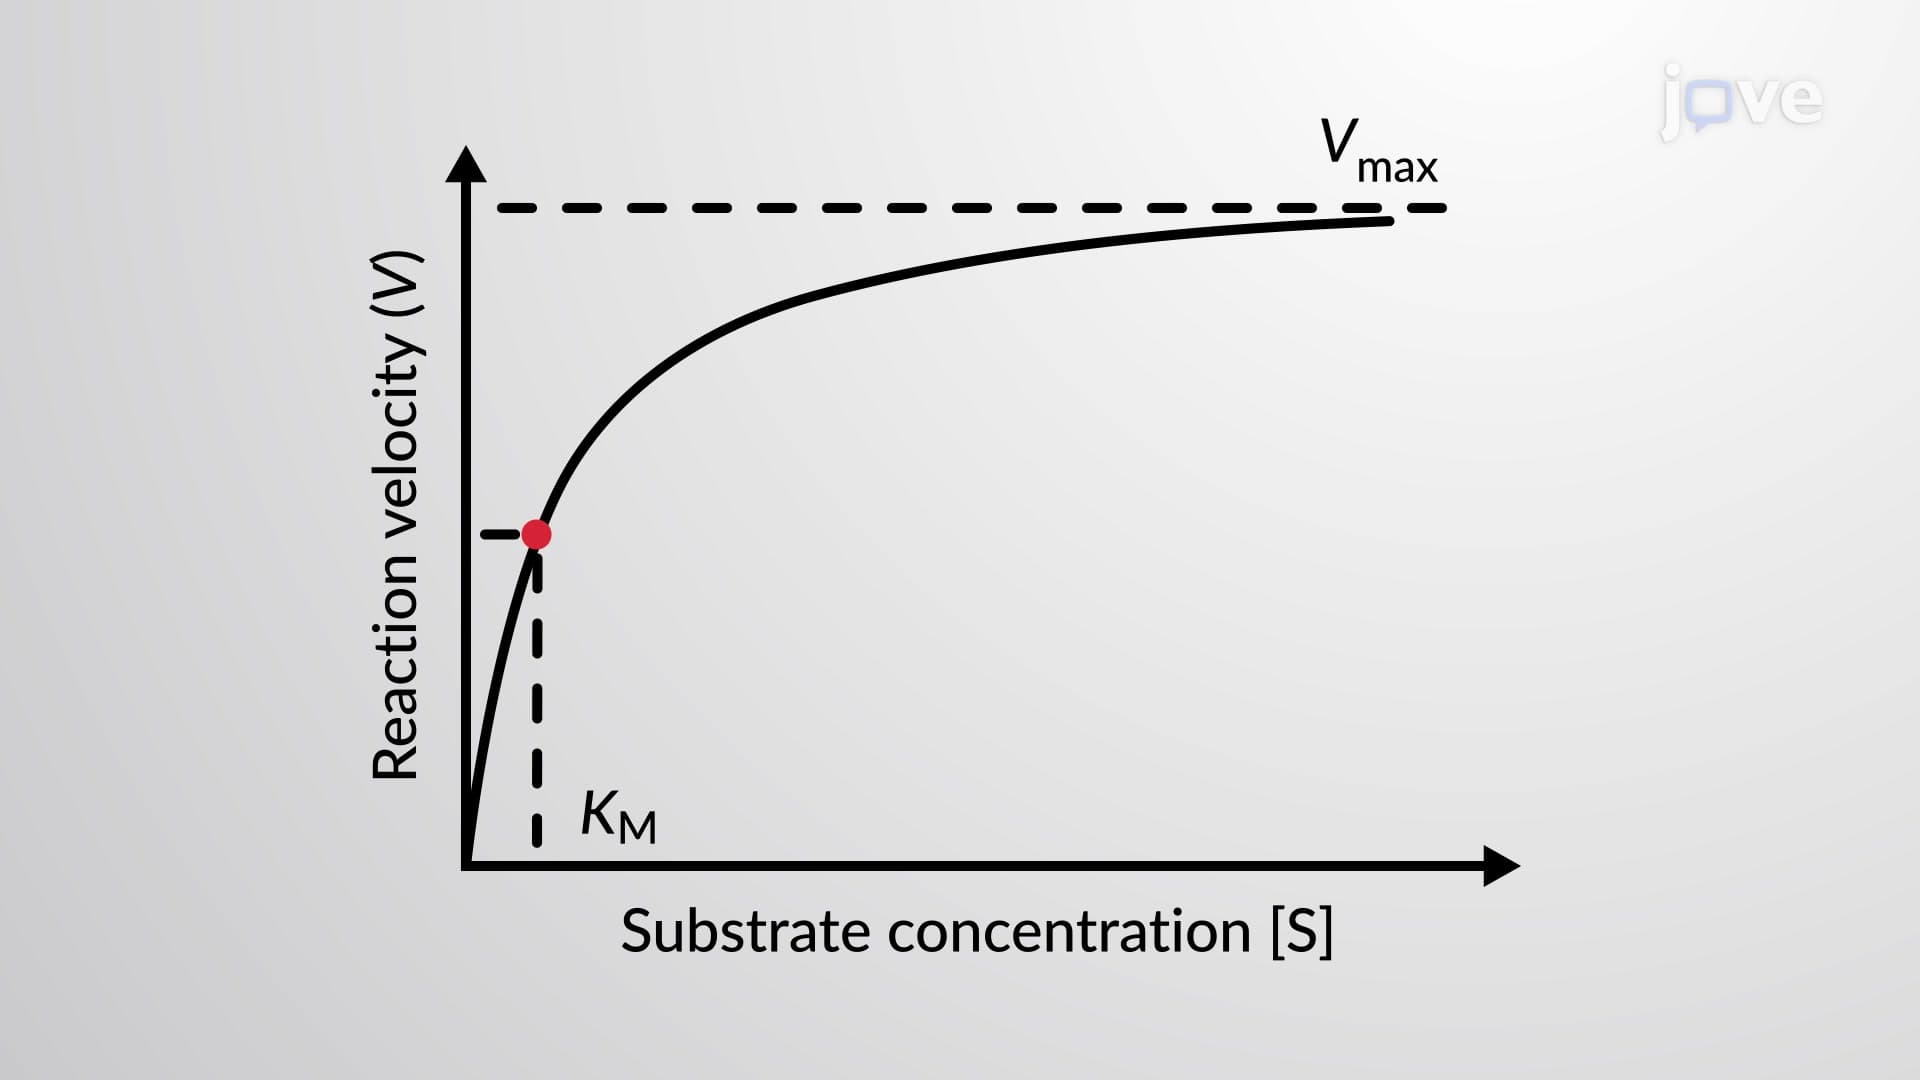
Introduction to Enzyme Kinetics
The experimenter can then plot the initial reaction rate or velocity (Vo) of a given trial against the substrate concentration ([S]) to obtain a graph of the reaction properties. For many enzymatic reactions involving a...
 01:26
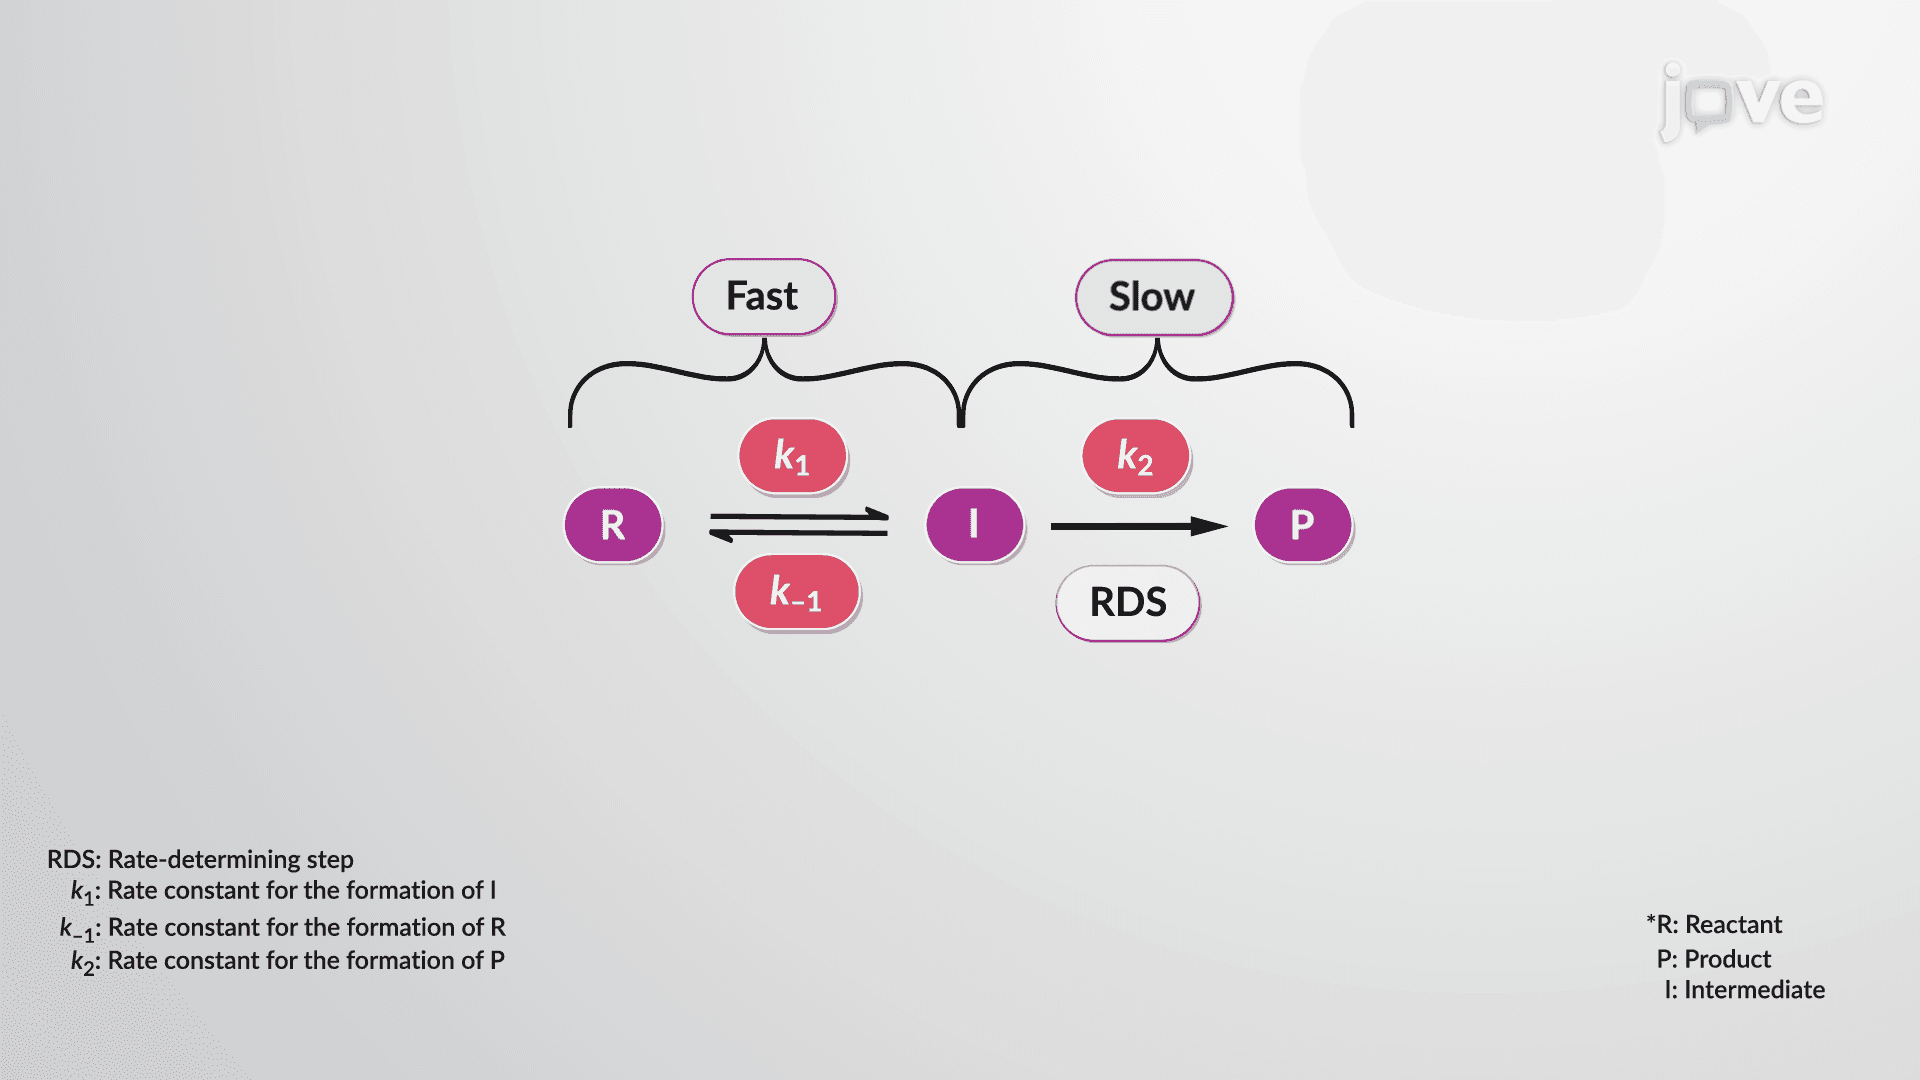
Reaction Mechanisms: The Steady-State Approximation
 03:25
Free Energy Changes for Nonstandard States
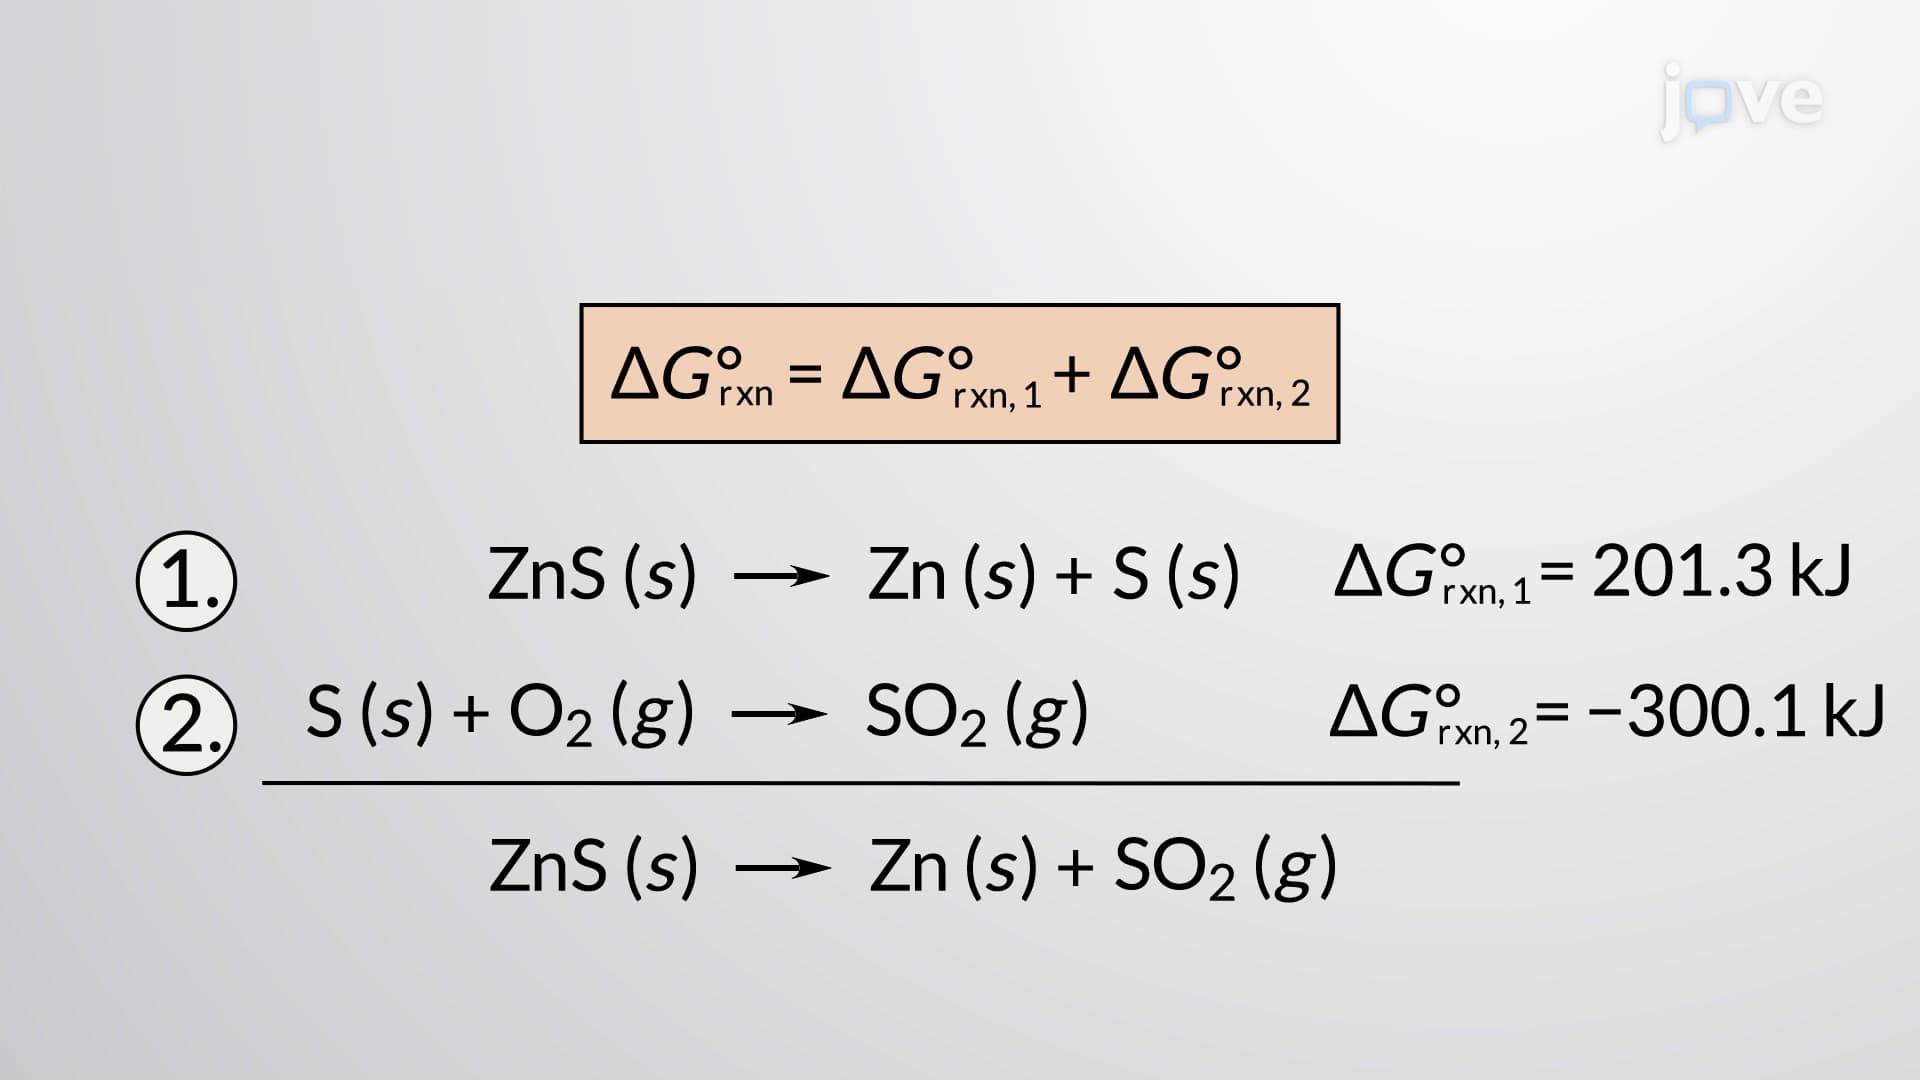 02:49
Calculating Standard Free Energy Changes
 02:34
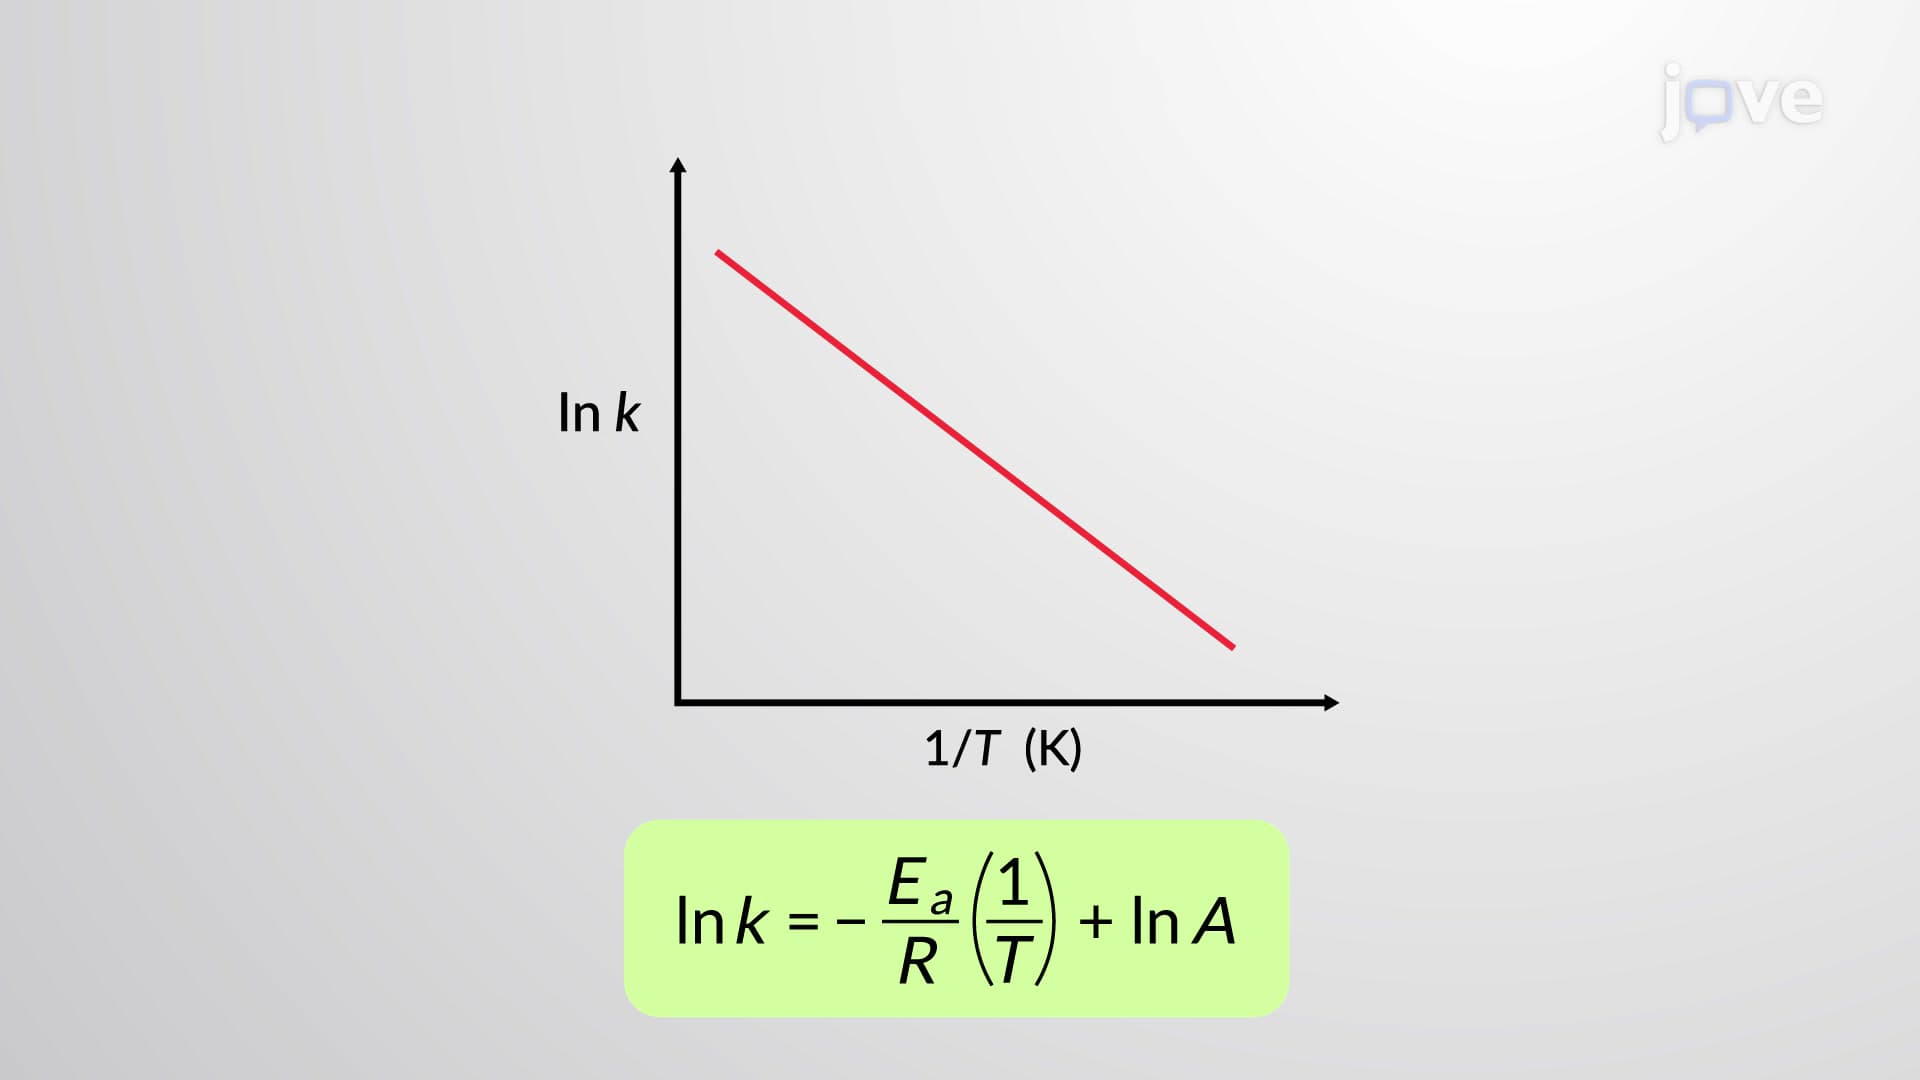
Arrhenius Plots
The Arrhenius equation can be used to...
 01:13
Introduction to Mechanisms of Enzyme Catalysis