Related Experiment Video
Updated: Jul 7, 2026

Prion Safety Laboratory Swipe Test
Published on: February 14, 2025
MM1-type sporadic Creutzfeldt-Jakob disease with unusually prolonged disease duration presenting with
Akira Hoshino1, Yasushi Iwasaki, Masayuki Izumi
1Department of Neurology, Aichi Medical University School of Medicine, Aichi, Japan.

Abstract:
We report an autopsy case of MM1-type sporadic Creutzfeldt-Jakob disease (sCJD) with an unusually prolonged disease duration of 58 months. The initial symptom was progressive mental disorder followed by advanced cognitive impairment. Clinical progression was generally slow; myoclonus appeared at 17 months and periodic sharp-wave complexes on electroencephalogram at 21 months. A state of akinetic mutism occurred 29 months after the onset of symptoms. MRI showed gradually progressive cerebral atrophy. Neuropathologic examination showed widespread severe brain involvement. In the cerebral neocortex, widespread severe tissue rarefaction, hypertrophic astrocytosis and neuron loss (so-called status spongiosus) were observed. The cerebral white matter showed diffuse myelin pallor with intense hypertrophic astrocytosis, numerous foamy macrophages and emperipolesis, indicating panencephalopathic-type sCJD pathology. The brainstem was relatively preserved from sCJD pathology, with the exception of the pontine nucleus and pyramidal tract. This may explain the prolonged disease duration without respiratory insufficiency until the terminal stage. Immunohistochemistry for prion protein (PrP) showed widespread synaptic-type PrP deposits in the cerebral neocortex, hippocampus and thalamus. The striatum and cerebellar cortex showed faint synaptic-type PrP deposition with some areas of small plaque-like PrP deposition. Sparse PrP deposition was also observed in the brainstem. Analysis of the PrP gene showed no mutation but methionine homozygosity at polymorphic codon 129. Western blot analysis of protease-resistant PrP indicated type 1 PrP. To our knowledge, this is the longest reported disease duration of MM1-type sCJD.
Insights
This case study details an MM1-type sporadic Creutzfeldt-Jakob disease (sCJD) patient with a record 58-month duration. The slow progression and preserved brainstem highlight unique neuropathological findings in this prion disease.
Area of Science:
- Neurology
- Neuroscience
- Pathology
Background:
- Sporadic Creutzfeldt-Jakob disease (sCJD) is a fatal neurodegenerative prion disease.
- MM1-type sCJD is the most common subtype, typically presenting with a shorter disease duration.
- Unusually prolonged disease courses offer insights into disease mechanisms and variability.
Observation:
- An autopsy case of MM1-type sCJD with a 58-month disease duration, significantly longer than typical.
- Initial symptoms included progressive mental disorder and cognitive decline, with myoclonus appearing late at 17 months.
- Electroencephalogram showed periodic sharp-wave complexes at 21 months, and akinetic mutism at 29 months.
- MRI revealed progressive cerebral atrophy.
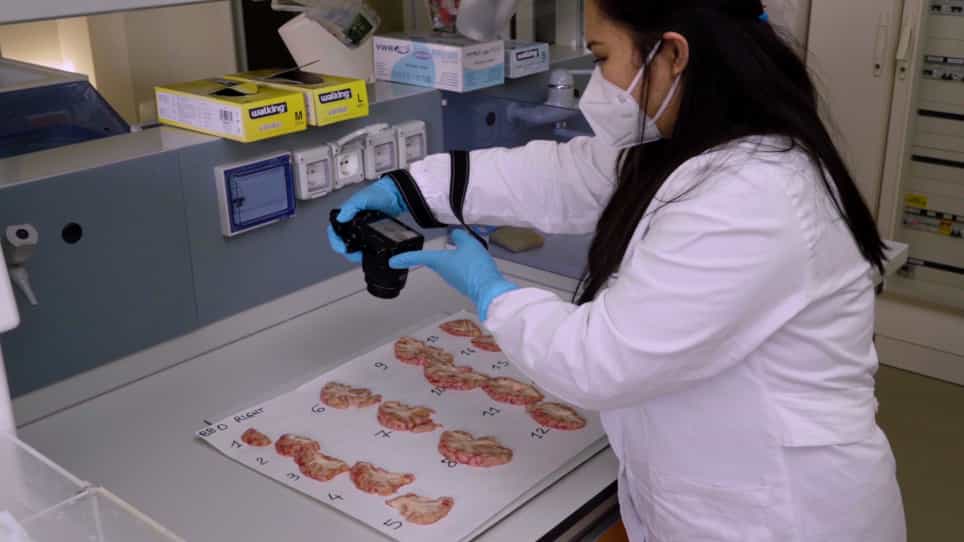
- Neuropathology demonstrated widespread spongiform changes, astrocytosis, and neuron loss in the cerebral cortex, indicative of panencephalopathic-type sCJD.
- The brainstem was relatively spared, potentially contributing to the prolonged survival.
Findings:
- The case represents the longest reported disease duration for MM1-type sCJD.
- Prion protein (PrP) immunohistochemistry revealed widespread synaptic-type deposits in the cortex, hippocampus, and thalamus.
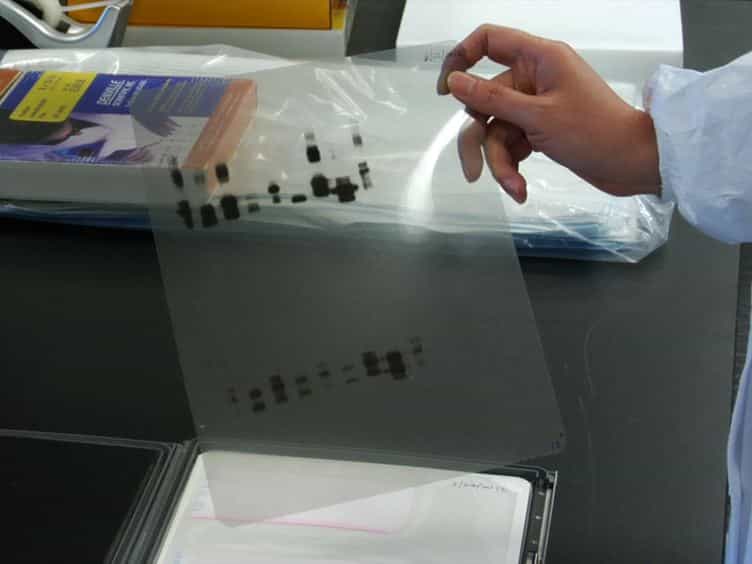
- Genetic analysis confirmed methionine homozygosity at codon 129 (MM) and Western blot identified type 1 PrP.
- The neuropathological findings, particularly the panencephalopathic changes and relative brainstem preservation, correlate with the prolonged clinical course.
Implications:
- This case expands the known spectrum of clinical and pathological variability in MM1-type sCJD.
- Understanding factors contributing to prolonged disease duration is crucial for disease management and future therapeutic strategies.
- Further research into the neuropathological correlates of extended survival in prion diseases is warranted.
Related Concept Videos
 01:19
Encephalitis l: Introduction
 01:26
Encephalitis ll: Pathophysiology
 01:23
Alzheimer Disease ll: Pathophysiology
 01:24
Parkinson Disease ll: Pathophysiology
 01:21
Huntington Disease l: Introduction
 01:29
Alzheimer Disease l: Introduction