

Frontier orbital consistent quantum capping potential (FOC-QCP) for bulky ligand of transition metal complexes
Yu-ya Ohnishi1, Yoshihide Nakao, Hirofumi Sato
1Department of Molecular Engineering, Graduate School of Engineering, Kyoto University, Nishikyo-ku, Kyoto 615-8510, Japan.

Abstract:
Chemically reasonable models of PR3 (R = Me, Et, iPr, and tBu) were constructed to apply the post Hartree-Fock method to large transition metal complexes. In this model, R is replaced by the H atom including the frontier orbital consistent quantum capping potential (FOC-QCP) which reproduces the frontier orbital energy of PR3. The steric effect is incorporated by the new procedure named steric repulsion correction (SRC). To examine the performance of this FOC-QCP method with the SRC, the activation barriers and reaction energies of the reductive elimination reactions of C2H6 and H2 from M(R1)2(PR2(3))2 (M = Ni, Pd, or Pt; R1 = Me for R2 = Me, Et, or iPr, or R1 = H for R2 = tBu) were evaluated with the DFT[B3PW91], MP4(SDQ), and CCSD(T) methods. The FOC-QCP method reproduced well the DFT[B3PW91]- and MP4(SDQ)-calculated energy changes of the real complexes with PMe3. For more bulky phosphine, the SRC is important to present correct energy change, in which the MP2 method presents reliable steric repulsion correction like the CCSD(T) method because the systems calculated in the SRC do not include a transition metal element. The monomerization energy of [RhCl(PiPr3)2]2 and the coordination energies of CO, H2, N2, and C2H4 with [RhCl(PiPr3)2]2 were theoretically calculated by the CCSD(T) method combined with the FOC-QCP and the SRC. The CCSD(T)-calculated energies agree well with the experimental ones, indicating the excellent performance of the combination of the FOC-QCP with the SRC. On the other hand, the DFT[B3PW91]-calculated energies of the real complexes considerably deviate from the experimental ones.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:09
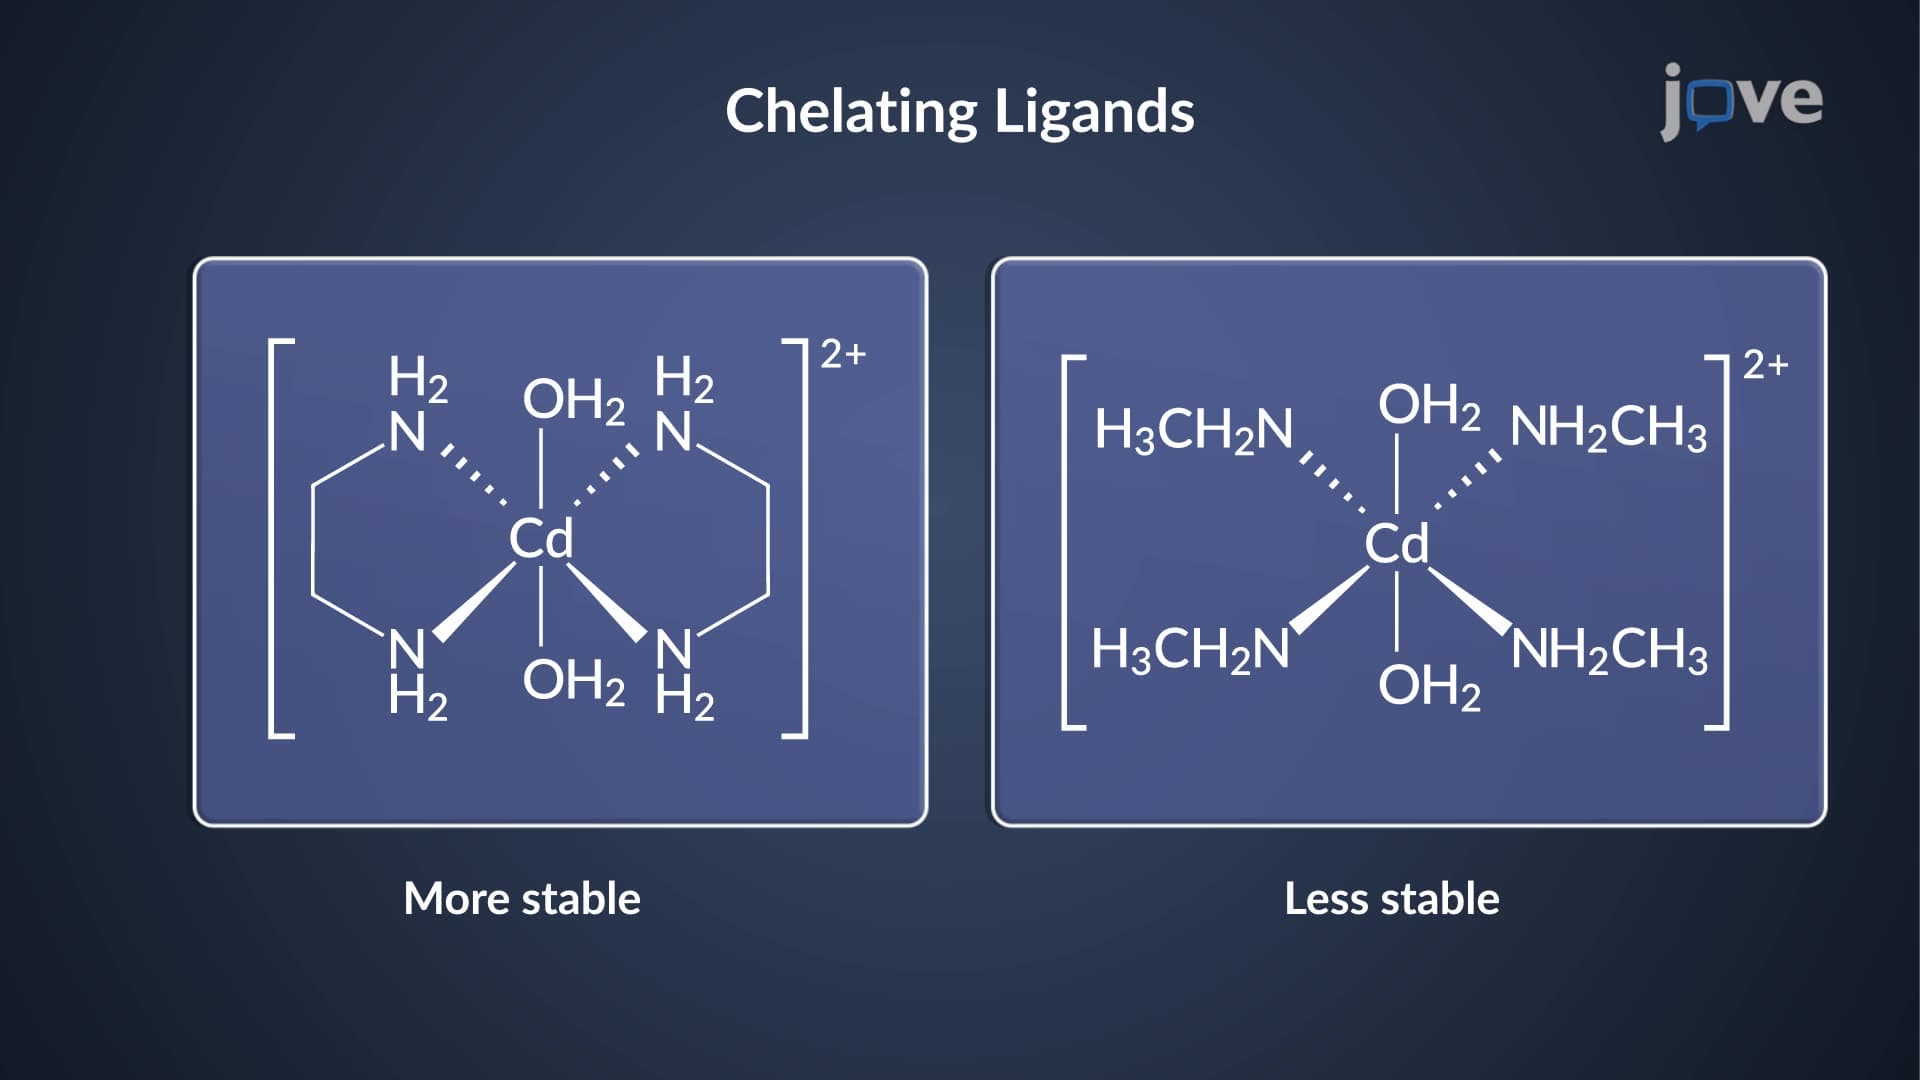
Complexation Equilibria: Factors Influencing Stability of Complexes
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.