Flexible ligand docking to multiple receptor conformations: a practical alternative
Related Experiment Video
Updated: Jul 7, 2026
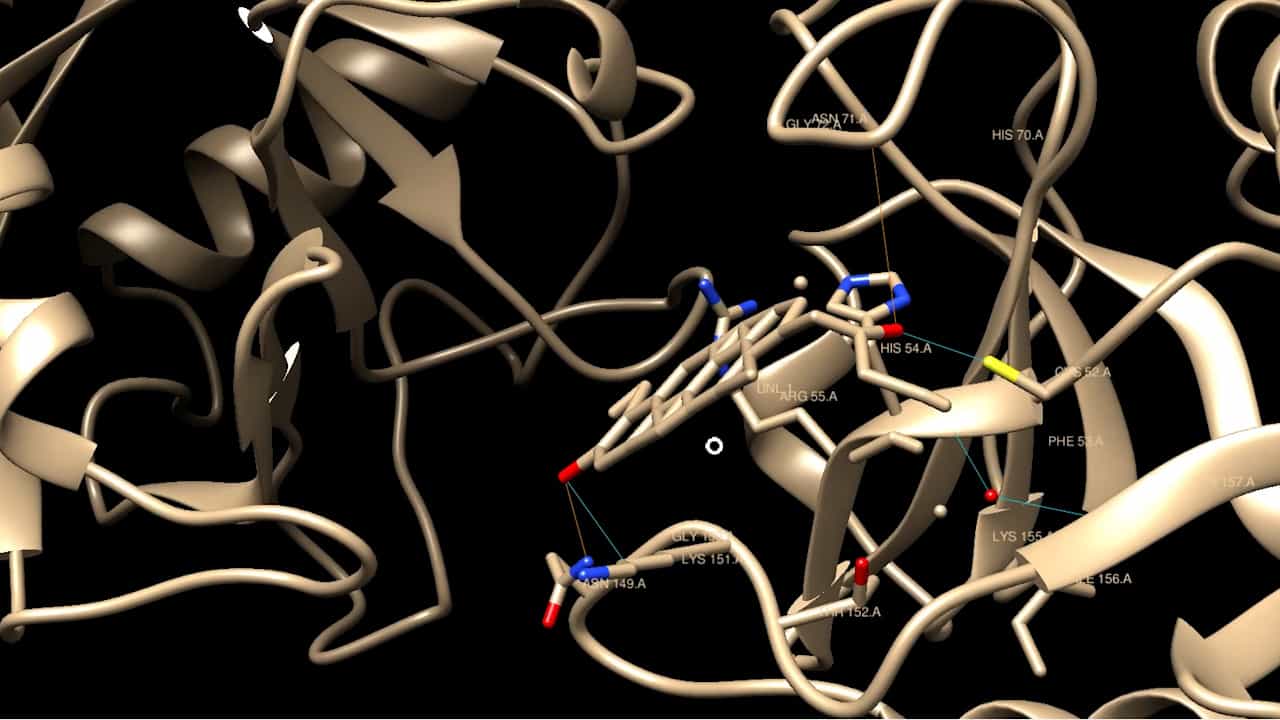
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
1Molsoft, 3366 N. Torrey Pines Court, Suite 300, CA 92037, United States. max@molsoft.com

Abstract:
State of the art docking algorithms predict an incorrect binding pose for about 50-70% of all ligands when only a single fixed receptor conformation is considered. In many more cases, lack of receptor flexibility results in meaningless ligand binding scores, even when the correct pose is obtained. Incorporating conformational rearrangements of the receptor binding pocket into predictions of both ligand binding pose and binding score is crucial for improving structure-based drug design and virtual ligand screening methodologies. However, direct modeling of protein binding site flexibility remains challenging because of the large conformational space that must be sampled, and difficulties remain in constructing a suitably accurate energy function. Here we show that using multiple fixed receptor conformations, either experimentally determined by crystallography or NMR, or computationally generated, is a practical shortcut that may improve docking calculations. In several cases, such an approach has led to experimentally validated predictions.

Related Concept Videos
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:58
Cooperative Allosteric Transitions
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...