Related Experiment Video
Updated: Jul 7, 2026
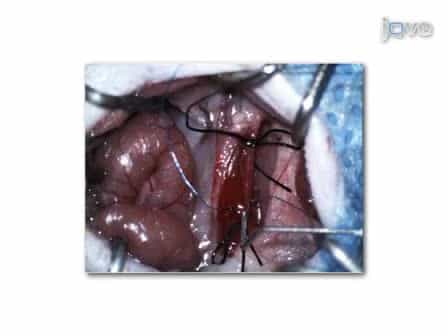
Creation of Murine Experimental Abdominal Aortic Aneurysms with Elastase
Published on: July 23, 2009
Matrix metalloproteinase 9 activity leads to elastin breakdown in an animal model of Kawasaki disease
Andrew C Lau1, Trang T Duong, Shinya Ito
1Hospital for Sick Children, and University of Toronto, Toronto, Ontario, Canada.

Insights
Matrix metalloproteinase 9 (MMP-9) plays a key role in coronary artery damage and aneurysm formation in a Kawasaki disease (KD) model. Inhibiting MMP-9 significantly reduces elastin breakdown, offering potential therapeutic targets for KD.
Area of Science:
- Cardiovascular Research
- Immunology
- Pathology
Background:
- Kawasaki disease (KD) is a critical pediatric vasculitis affecting coronary arteries, potentially leading to aneurysms.
- Studying KD mechanisms is challenging due to the unavailability of human cardiac tissue.
- Experimental models are essential for investigating KD-induced coronary artery damage.
Purpose of the Study:
- To investigate the role of matrix metalloproteinase 9 (MMP-9) in coronary artery inflammation and vascular damage within an experimental KD model.
- To determine if MMP-9 is essential for the development of coronary artery pathology in KD.
Main Methods:
- Induction of coronary arteritis in C57BL/6 mice using Lactobacillus casei cell wall extract.
- Assessment of MMP-9 expression, activity, and localization via immunoblotting, confocal microscopy, and zymography.
- Utilizing MMP-9-deficient mice to evaluate the necessity of MMP-9 in disease progression and vascular damage.
Main Results:
- Coronary arteritis induced elastin breakdown and aneurysm formation, independent of smooth muscle cell apoptosis.
- Elevated MMP-9 protein and enzymatic activity were observed following disease induction.
- Tumor necrosis factor alpha (TNFα) upregulated MMP-9, primarily produced by vascular smooth muscle cells.
- MMP-9 deficiency significantly reduced elastin breakdown, though inflammation persisted.
Conclusions:
- TNFα upregulates MMP-9, a crucial protease for extracellular matrix degradation, driving coronary artery damage in this KD model.
- Targeting MMP-9 presents a potential therapeutic strategy to mitigate coronary artery damage and improve outcomes in children with KD.
Objective:
Kawasaki disease (KD) is a multisystem vasculitis leading to damage in the coronary circulation and aneurysm formation. Because cardiac tissue from affected children is not available, investigation of the mechanisms responsible for coronary artery damage in KD requires use of a disease model. The present study was undertaken to examine, in an experimental model, the role of matrix metalloproteinase 9 (MMP-9) on coronary artery inflammation and vascular damage.
Methods:
C57BL/6 mice were injected with Lactobacillus casei cell wall extract to induce coronary arteritis. Hearts were isolated and assayed for MMP-9 protein expression and enzymatic activity by immunoblotting or confocal microscopy and zymography, respectively. MMP-9-deficient mice were used to examine the necessity of MMP-9 for disease development.
Results:
Localized inflammation at the coronary artery led to elastin breakdown and aneurysm formation. This occurred in the absence of smooth muscle cell apoptosis. Following disease induction, there was an increase in the amount and enzymatic activity of MMP-9, an elastolytic protease. MMP-9 was up-regulated by tumor necrosis factor alpha (TNFalpha) and produced primarily by vascular smooth muscle cells. In MMP-9-deficient animals, vascular inflammation continued to develop, but the incidence of elastin breakdown was significantly reduced. Elastin breakdown in the coronary artery was virtually eliminated by ablation of MMP-9.
Conclusion:
These findings show that TNFalpha up-regulates expression of MMP-9, an important proteinase responsible for extracellular matrix breakdown, leading to coronary artery damage in this model of KD. These results have important implications regarding treatments for improving coronary outcome in affected children.
