Related Experiment Video
Updated: Jul 6, 2026


In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
Enzyme replacement therapy with idursulfase in patients with mucopolysaccharidosis type II
1Royal Manchester Children's Hospital, Manchester, UK. ed.wraith@cmmc.nhs.uk

Unlabelled:
Mucopolysaccharidosis type II (MPS II; Hunter syndrome) is a rare X-linked recessive disease caused by deficiency of the lysosomal enzyme iduronate-2-sulphatase. This leads to the progressive accumulation of glycosaminoglycans in tissues throughout the body. As a result, patients may suffer from severe airway obstruction, skeletal deformities, cardiomyopathy and, in severely affected patients, there may be progressive neurological decline. Despite the early onset of signs and symptoms, diagnosis is often delayed. Until recently, treatment for MPS II has been largely palliative; however, enzyme replacement therapy with recombinant iduronate-2-sulphatase, which is produced in a human cell line and targets the underlying cause of the disease, has now been approved.
Conclusion:
This short review provides an overview of the natural history of MPS II and current experience of enzyme replacement therapy with idursulfase.
Related Concept Videos
 01:14
Drug Metabolism: Phase II Reactions
 01:19
Phase II Reactions: Sulfation and Conjugation with α-Amino Acids
 01:24
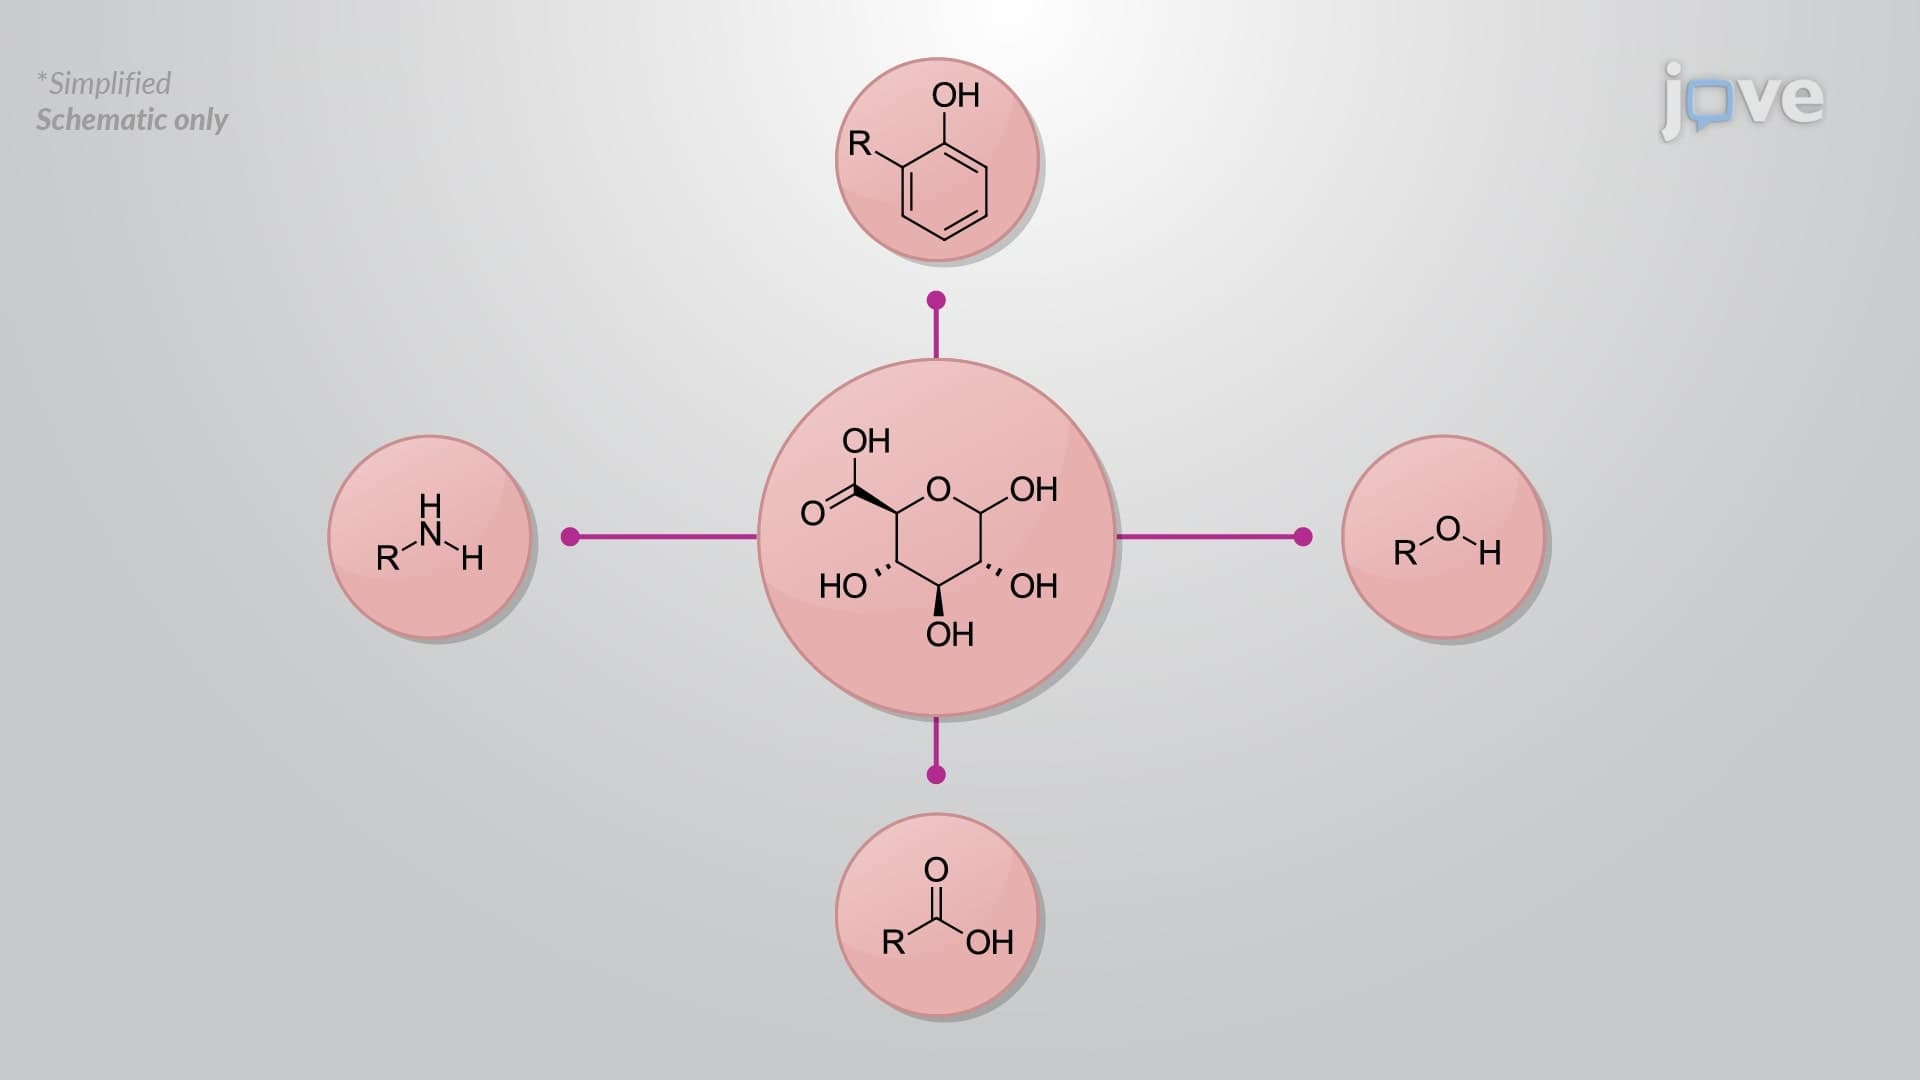
Phase II Reactions: Glucuronidation
 01:22
Lysosomal Hydrolases
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...