Related Experiment Video
Updated: Jul 6, 2026
10:28

A Semi-High-Throughput Adaptation of the NADH-Coupled ATPase Assay for Screening Small Molecule Inhibitors
Published on: August 17, 2019
[Improvement of precision in pharmacological studies using the non-linear regression method: estimation of enzyme
Akiko Yamazaki1, Eisuke Kume, Sumiko Tanaka
1hamada@ms.kagu.tus.ac.jp
Nihon Yakurigaku Zasshi. Folia Pharmacologica Japonica
|March 18, 2008

Abstract
No abstract available in PubMed .
Related Concept Videos
 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems
The empirical approach to drug therapy optimization relies on correlating pharmacological response with administered dosage. Such an approach can be costly, time-consuming, and often yields poor correlation due to variables like formulation factors and drug elimination characteristics. A more precise approach correlates response with plasma drug concentration or the amount of drug in the body, rather than dosage. This is achieved through pharmacokinetic-pharmacodynamic (PK/PD) modeling, which...
 01:19
Nonlinear Pharmacokinetics: Overview
Nonlinear or dose-dependent pharmacokinetics is a phenomenon that occurs when the pharmacokinetic parameters of certain drugs deviate from linear pharmacokinetics at higher doses. These drugs do not follow the expected first-order kinetics, where the rate of drug elimination is directly proportional to the drug concentration. Instead, they exhibit a nonlinear relationship, which can be attributed to several factors.
Nonlinearity can arise due to the saturation of plasma protein-binding or...
Nonlinearity can arise due to the saturation of plasma protein-binding or...
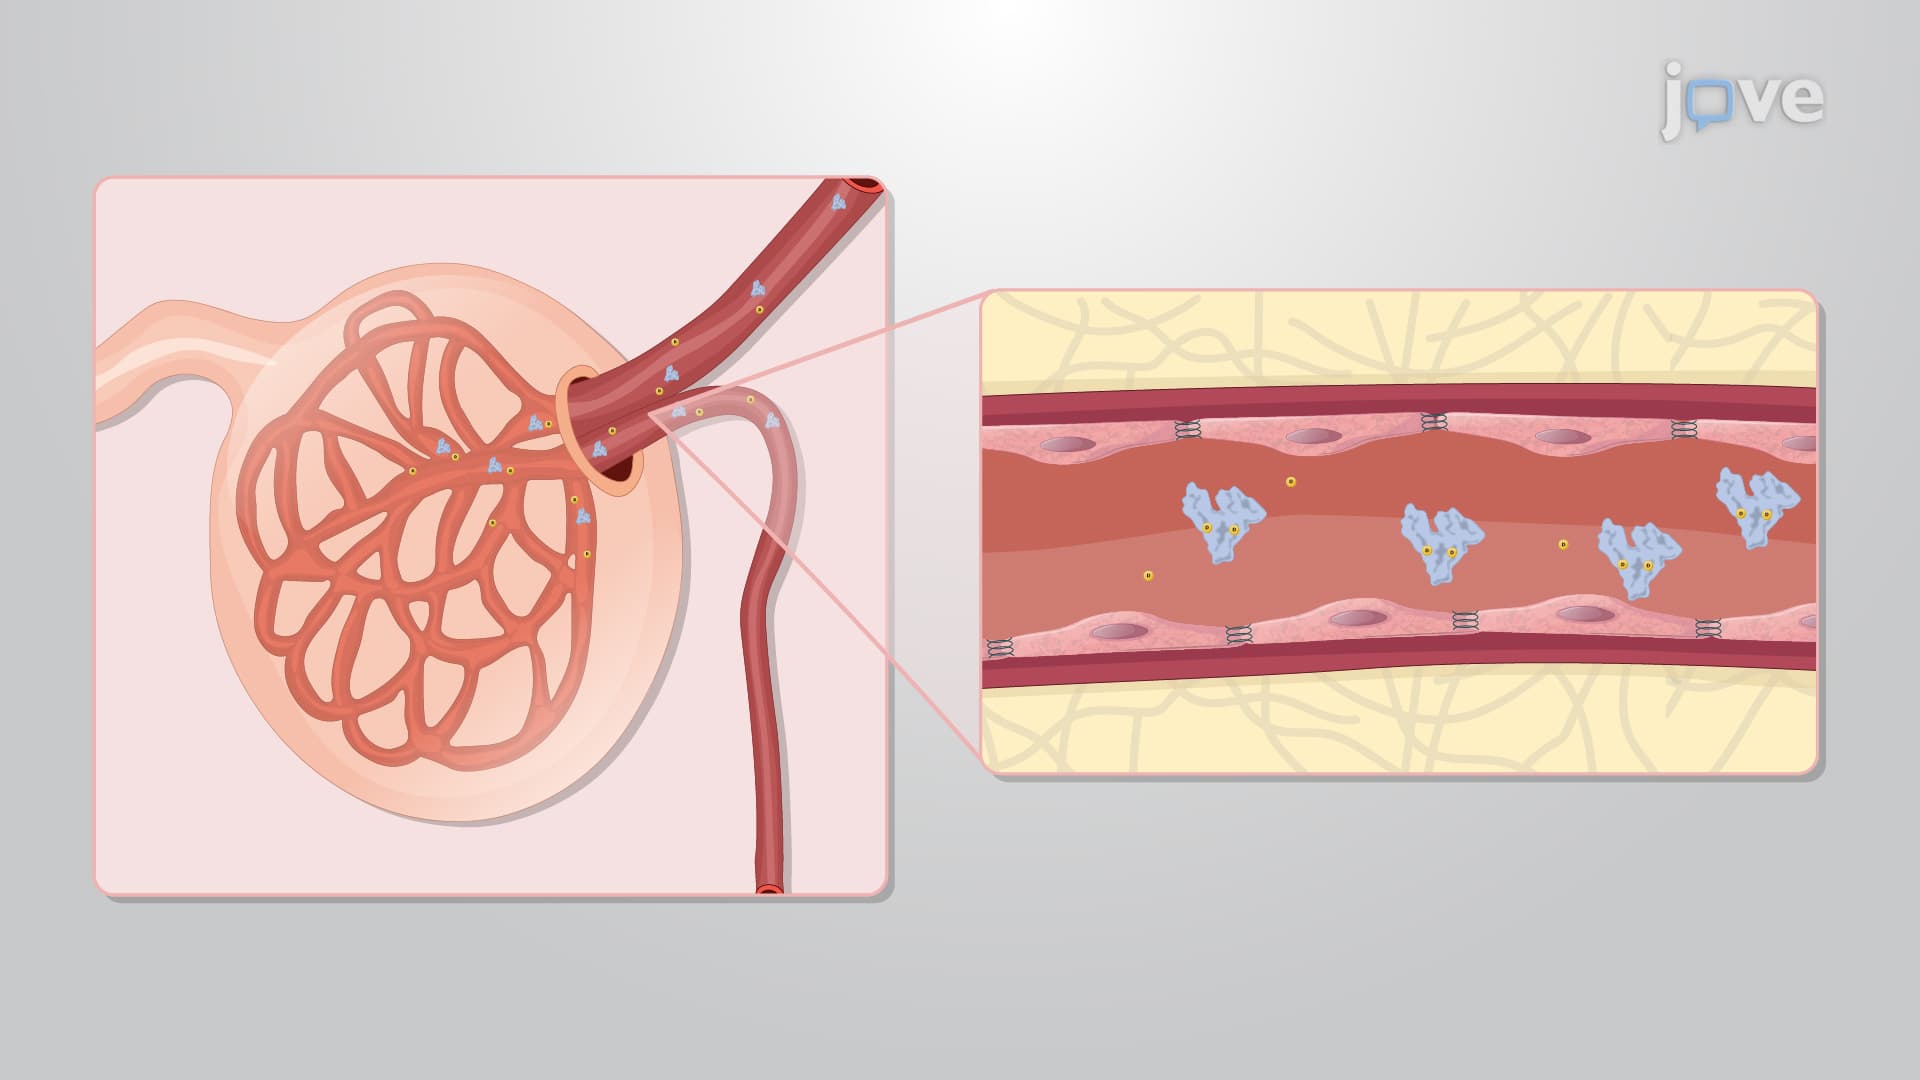 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 00:59
Model-Independent Approaches for Pharmacokinetic Data: Noncompartmental Analysis
Noncompartmental analyses offer an alternative method for describing drug pharmacokinetics without relying on a specific compartmental model. In this approach, the drug's pharmacokinetics are assumed to be linear, with the terminal phase log-linear. This assumption allows for simplified analysis and interpretation of the drug's behavior in the body.
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This relationship...
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This relationship...
 01:22
Nonlinear Pharmacokinetics: Causes of Nonlinearity
Nonlinearity in drug pharmacokinetics is caused by various factors influencing how a drug is absorbed, distributed, metabolized, and excreted. Understanding these nonlinear processes is crucial for predicting drug behavior in the body and optimizing drug dosing regimens.
Nonlinear drug absorption can occur when the process is rate-limited by solubility, carrier-mediated transport systems, or saturation of the presystemic gut wall or hepatic metabolism. For instance, high doses of riboflavin...
Nonlinear drug absorption can occur when the process is rate-limited by solubility, carrier-mediated transport systems, or saturation of the presystemic gut wall or hepatic metabolism. For instance, high doses of riboflavin...
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
The linear concentration–effect model, underpinned by the principle that pharmacological effect (E) is directly proportional to plasma drug concentration (C), emerges as a pivotal simplification of the Emax model for conditions where C is significantly less than EC50. This model portrays a linear trajectory of the concentration–effect relationship when drug levels are markedly below the EC50 threshold.Despite its inherent assumption of continuous effect augmentation with increasing drug...