Related Experiment Video
Updated: Jul 6, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Computation of binding free energy with molecular dynamics and grand canonical Monte Carlo simulations
1Biosciences Division, Argonne National Laboratory, Argonne, IL 60439, USA.

Abstract:
The binding of a ligand to a receptor is often associated with the displacement of a number of bound water molecules. When the binding site is exposed to the bulk region, this process may be sampled adequately by standard unbiased molecular dynamics trajectories. However, when the binding site is deeply buried and the exchange of water molecules with the bulk region may be difficult to sample, the convergence and accuracy in free energy perturbation (FEP) calculations can be severely compromised. These problems are further compounded when a reduced system including only the region surrounding the binding site is simulated. To address these issues, we couple molecular dynamics (MD) with grand canonical Monte Carlo (GCMC) simulations to allow the number of water to fluctuate during an alchemical FEP calculation. The atoms in a spherical inner region around the binding pocket are treated explicitly while the influence of the outer region is approximated using the generalized solvent boundary potential (GSBP). At each step during thermodynamic integration, the number of water in the inner region is equilibrated with GCMC and energy data generated with MD is collected. Free energy calculations on camphor binding to a deeply buried pocket in cytochrome P450cam, which causes about seven water molecules to be expelled, are used to test the method. It concluded that solvation free energy calculations with the GCMC/MD method can greatly improve the accuracy of the computed binding free energy compared to simulations with fixed number of water.

Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
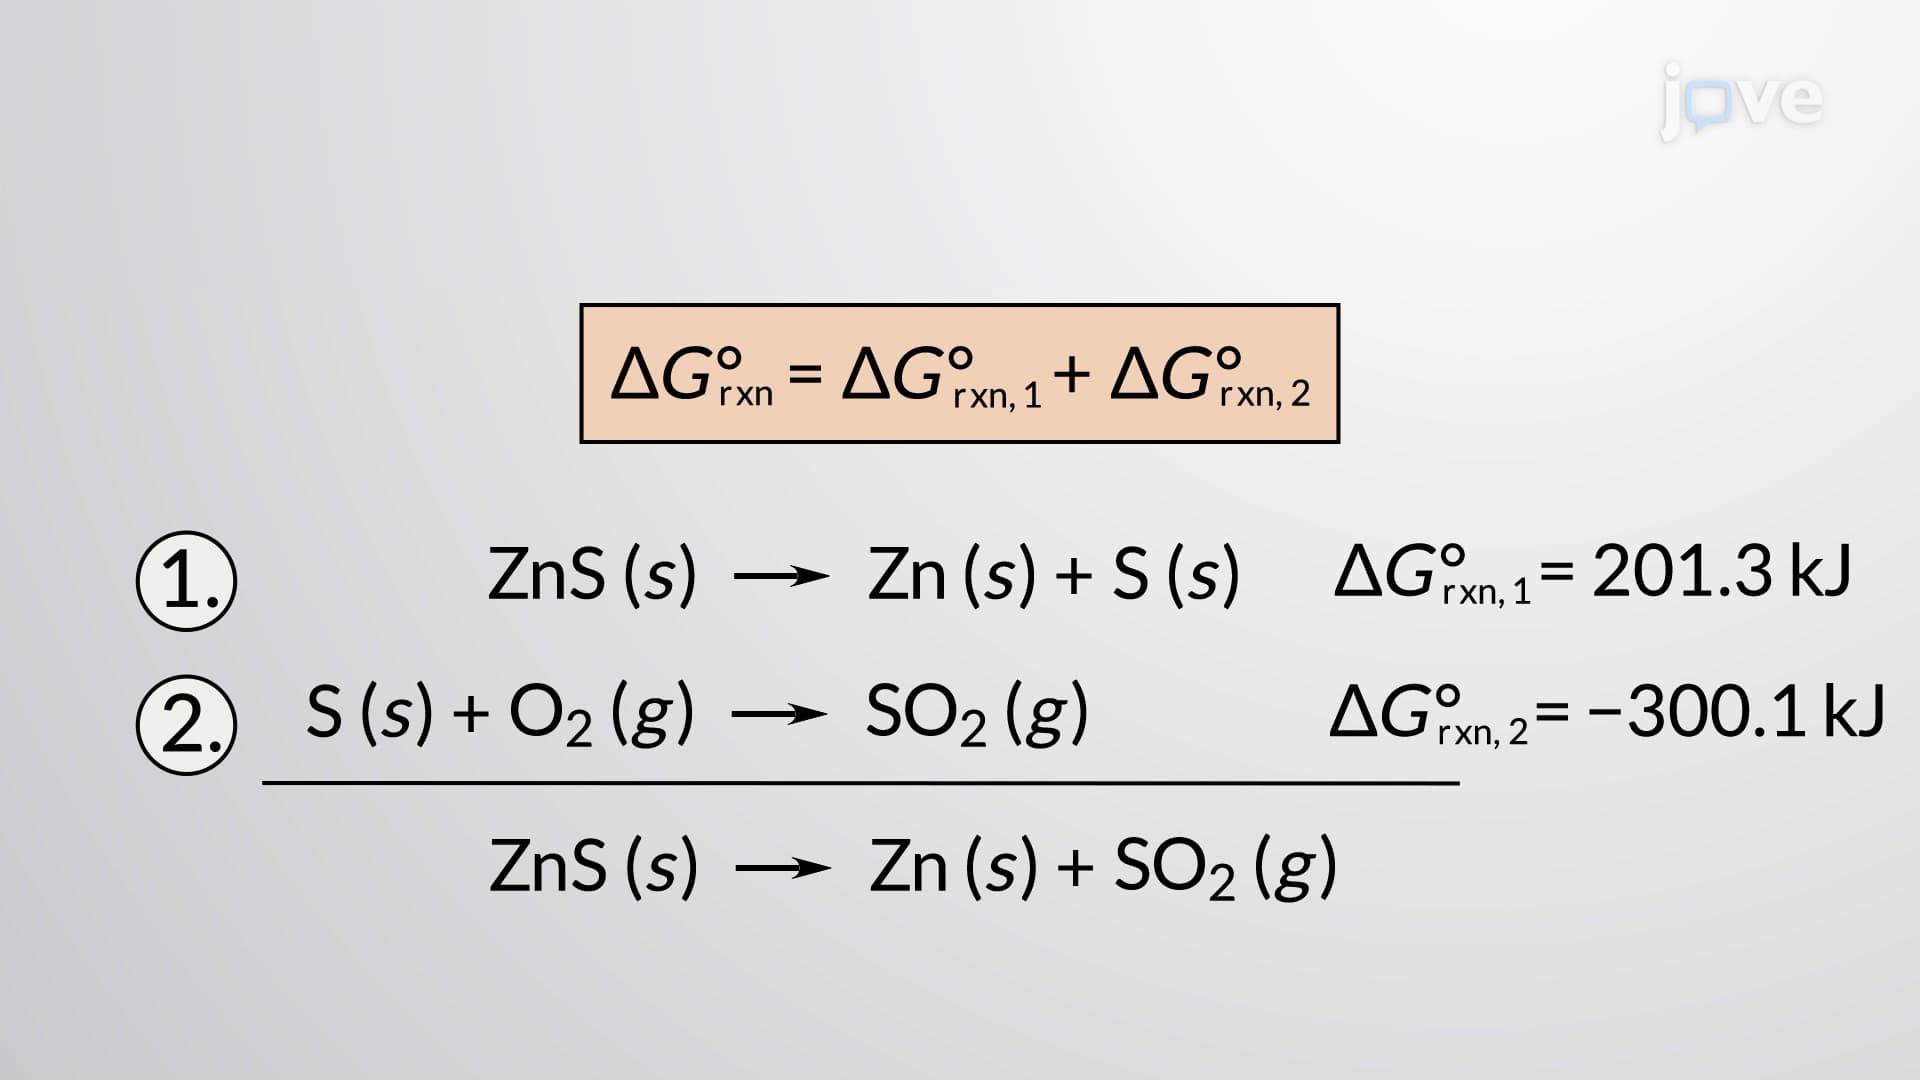 02:49
Calculating Standard Free Energy Changes
 02:39
Gibbs Free Energy
 02:23
Gibbs Free Energy and Thermodynamic Favorability
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...