Ab initio protein structure prediction with force field parameters derived from water-phase quantum chemical calculation
Related Experiment Video
Updated: Jul 6, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Daisuke Katagiri1, Hideyoshi Fuji, Saburo Neya
1Graduate School of Pharmaceutical Sciences, Chiba University, Chiba 263-8522, Japan.

Abstract:
Molecular dynamics (MD) simulations are extensively used in the study of the structures and functions of proteins. Ab initio protein structure prediction is one of the most important subjects in computational biology, and many trials have been performed using MD simulation so far. Since the results of MD simulations largely depend on the force field, reliable force field parameters are indispensable for the success of MD simulation. In this work, we have modified atom charges in a standard force field on the basis of water-phase quantum chemical calculations. The modified force field turned out appropriate for ab initio protein structure prediction by the MD simulation with the generalized Born method. Detailed analysis was performed in terms of the conformational stability of amino acid residues, the stability of secondary structure of proteins, and the accuracy for prediction of protein tertiary structure, comparing the modified force field with a standard one. The energy balance between alpha-helix and beta-sheet structures was significantly improved by the modification of charge parameters.


Related Concept Videos
 01:24
Protein Organization
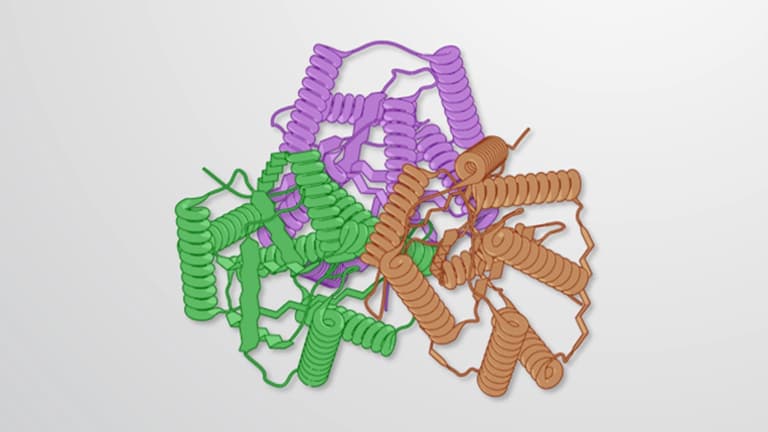
The primary structure of a protein is its amino acid sequence.
 01:13
Protein Organization
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:27
Predicting Molecular Geometry