Related Experiment Video
Updated: Jul 6, 2026
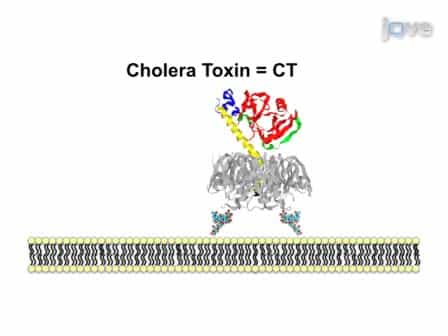
Detection of Toxin Translocation into the Host Cytosol by Surface Plasmon Resonance
Published on: January 3, 2012
Multivalent drug design and inhibition of cholera toxin by specific and transient protein-ligand interactions
Jiyun Liu1, Darren Begley1, Daniel D Mitchell1
1Department of Chemistry, University of Washington, Seattle, WA 98195, USABiomolecular Structure Center, University of Washington, Seattle, WA 98195, USADepartment of Biochemistry, University of Washington, Seattle, WA 98195, USA.

Abstract:
Multivalent inhibitors of the cholera toxin B pentamer are potential therapeutic drugs for treating cholera and serve as models for demonstrating multivalent ligand effects through a structure-based approach. A crucial yet often overlooked aspect of multivalent drug design is the length, rigidity and chemical composition of the linker used to connect multiple binding moieties. To specifically study the role of chemical linkers in multivalent ligand design, we have synthesized a series of compounds with one and two binding motifs connected by several different linkers. These compounds have affinity for and potency against the cholera toxin B pentamer despite the fact that none can simultaneously bind two toxin receptor sites. Results from saturation transfer difference NMR reveal transient, non-specific interactions between the cholera toxin and linker groups contribute significantly to overall binding affinity of monovalent compounds. However, the same random protein-ligand interactions do not appear to affect binding of bivalent molecules. Moreover, the binding affinities and potencies of these 'non-spanning' bivalent ligands appear to be wholly independent of linker length. Our detailed analysis identifies multiple effects that account for the improved inhibitory potencies of bivalent ligands and suggest approaches to further improve the activity of this class of compounds.
More Related Videos
 09:39
Determination of Tolerable Fatty Acids and Cholera Toxin Concentrations Using Human Intestinal Epithelial Cells and BALB/c Mouse Macrophages
Published on: May 30, 2013
 10:17
Creating Highly Specific Chemically Induced Protein Dimerization Systems by Stepwise Phage Selection of a Combinatorial Single-Domain Antibody Library
Published on: January 14, 2020
Related Concept Videos
 01:25
Cholera
 01:25
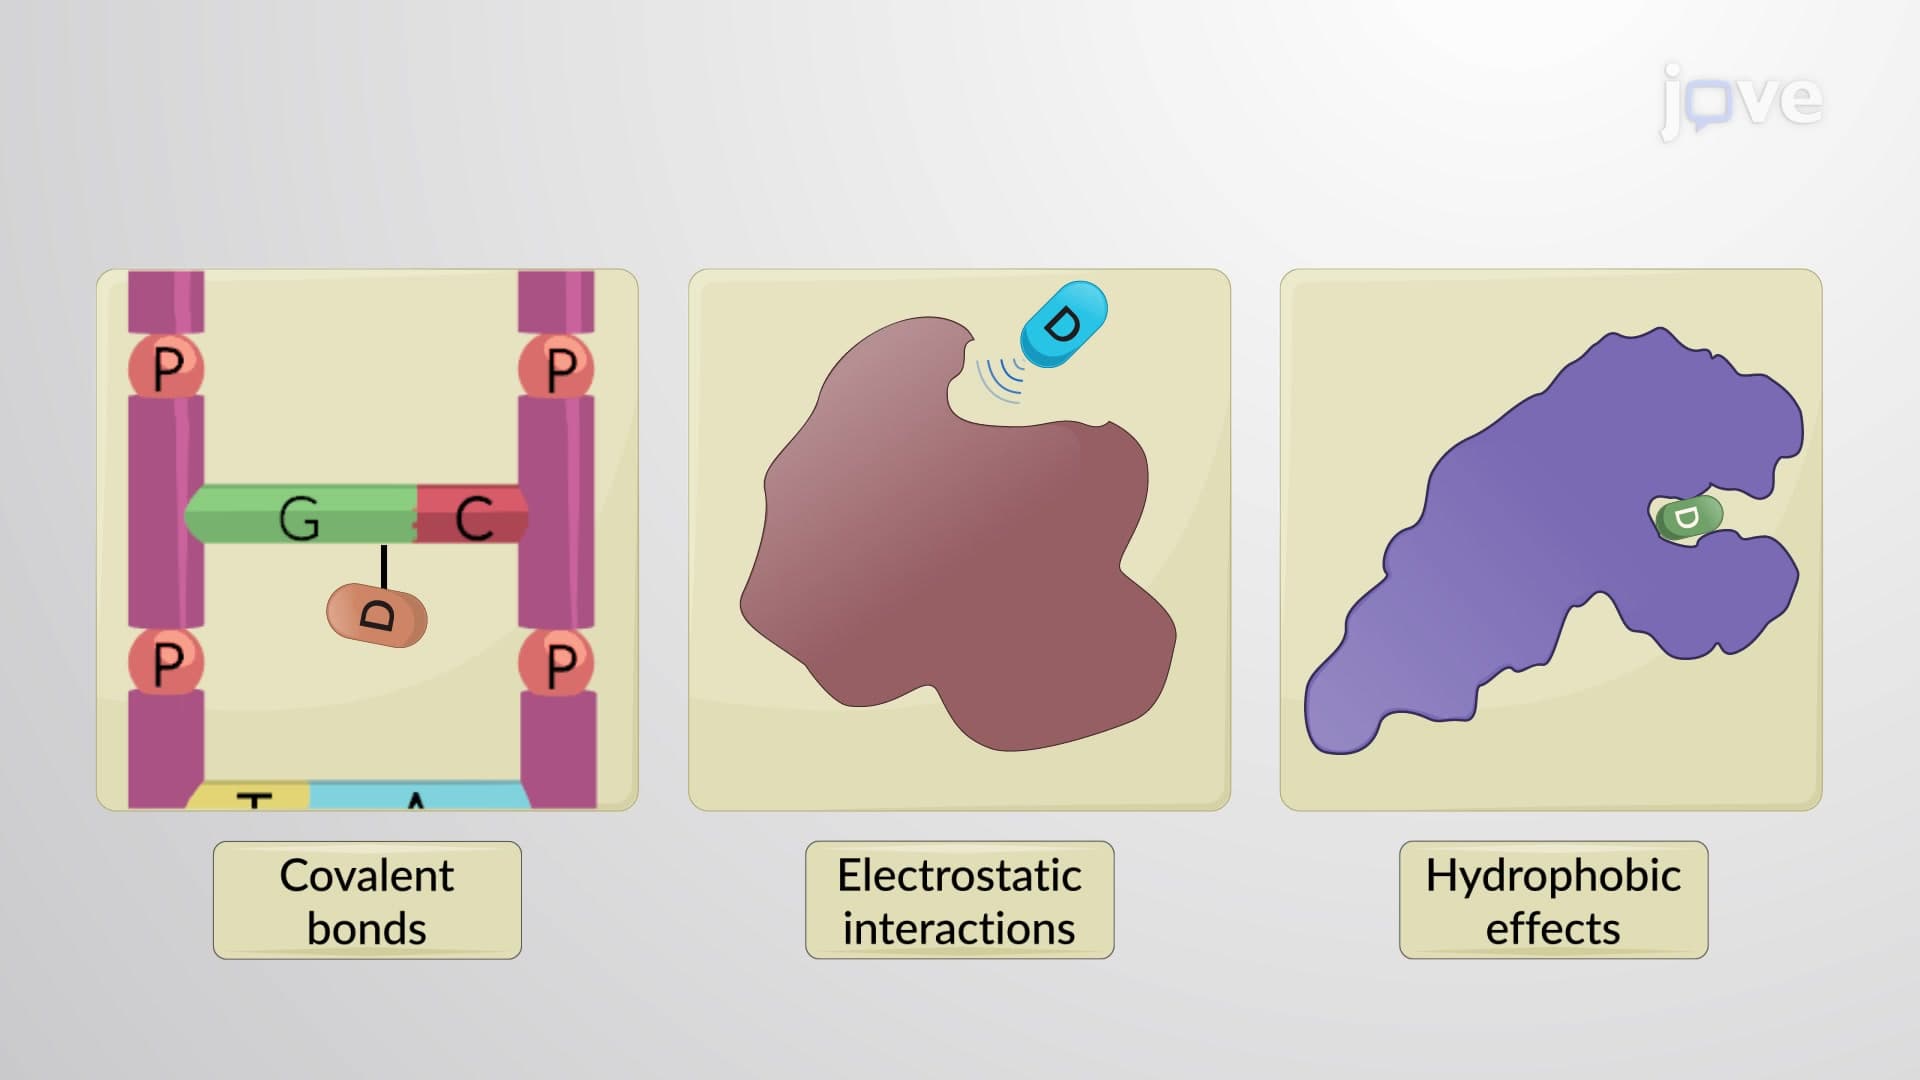
Drug-Receptor Bonds
In...
 01:58
Cooperative Allosteric Transitions
 01:29
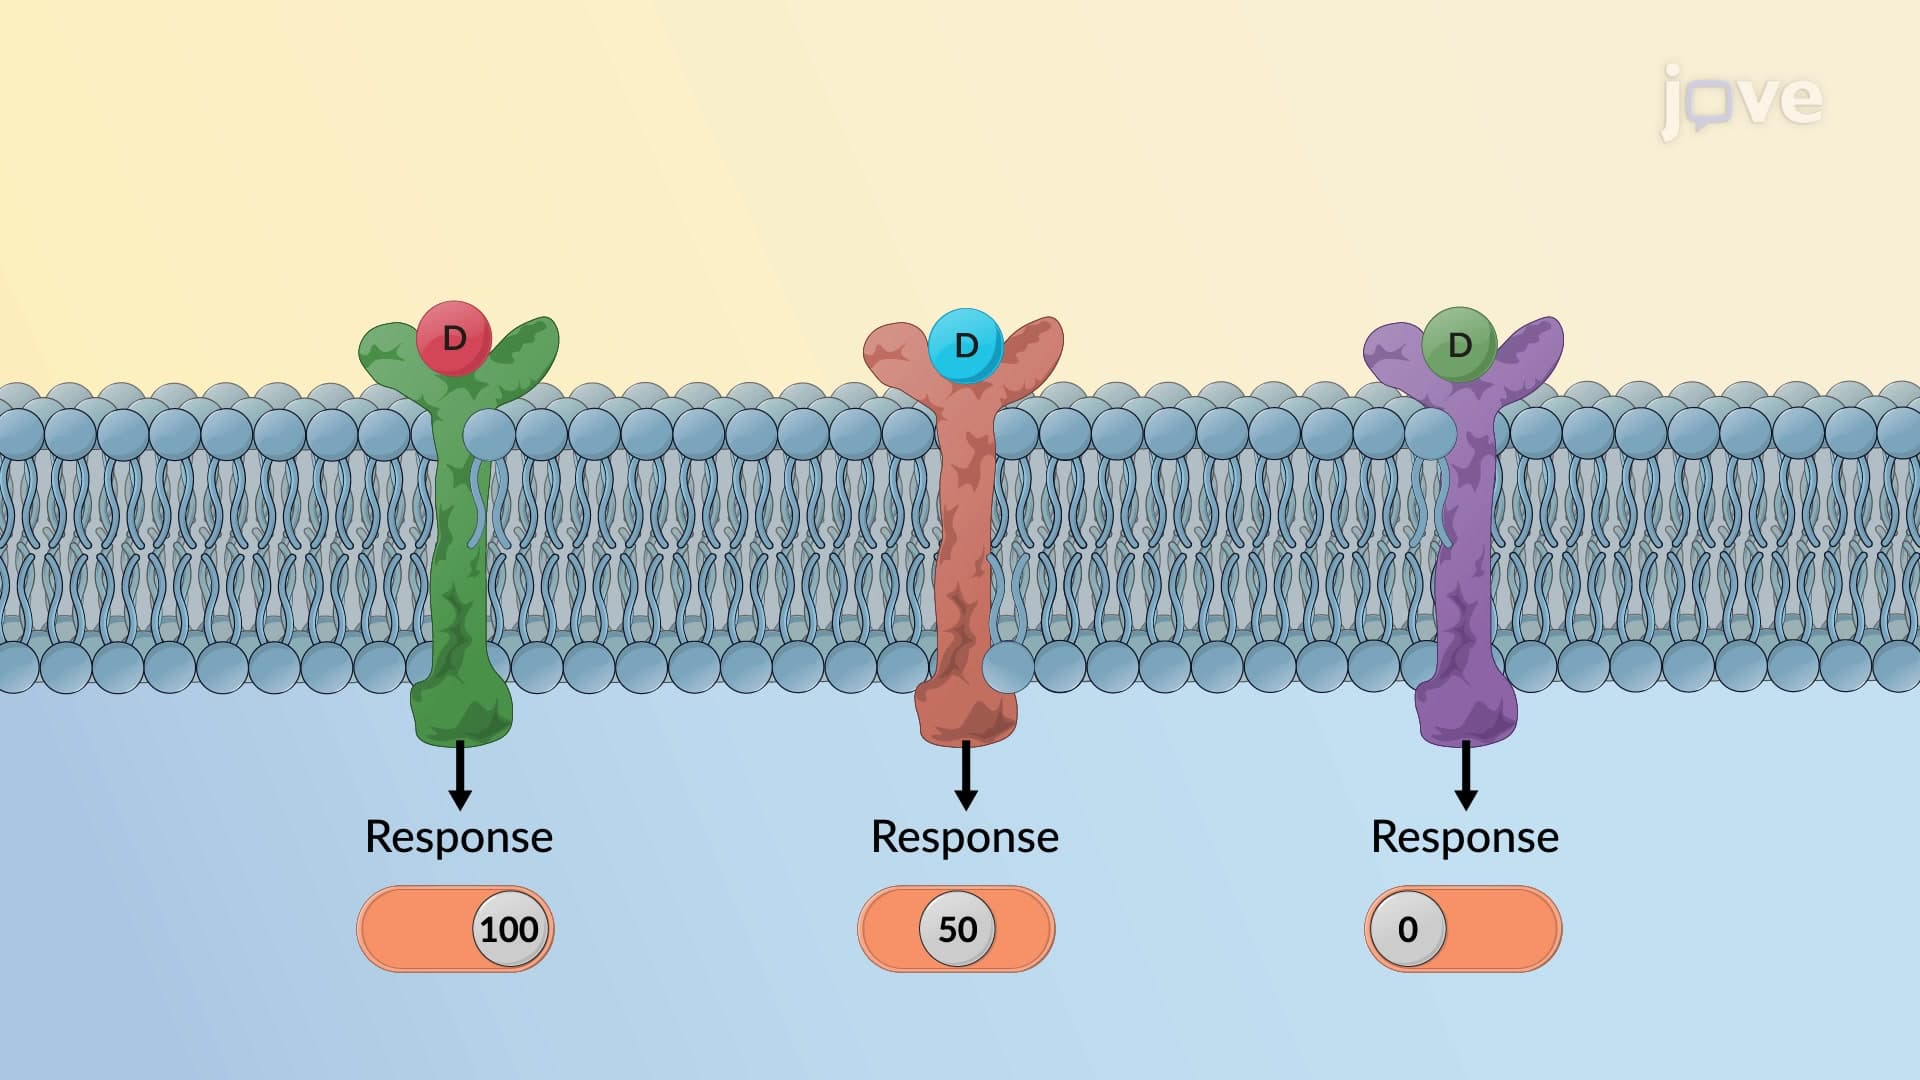
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:19
Complexation Equilibria: The Chelate Effect