PPARs as therapeutic targets for correction of inborn mitochondrial fatty acid oxidation disorders
1CNRS UPR 9078, Université Paris Descartes, Faculté Necker-Enfants Malades, Paris, France.

Abstract:
Enzyme defects in the mitochondrial fatty acid oxidation (FAO) are a large family of inherited metabolic disease well characterized clinically and genetically, but for which pharmacological strategies remain limited. It is now well established that regulation of genes involved in mitochondrial FAO is under control of the PPAR (peroxisome proliferator activated receptor) signalling pathway, and this led us to test a possible pharmacological correction of FAO disorders by fibrates and other PPAR activators. This review presents the basic data supporting our initial hypothesis, summarizes the results obtained in cells from patients with CPT II (carnitine palmitoyltransferase II) or VLCAD (very long-chain acyl-CoA dehydrogenase) deficiency, and discusses the perspectives and limits of this approach for therapy of these disorders.
Insights
Pharmacological activation of PPAR signaling may offer a new therapeutic approach for inherited metabolic disorders affecting mitochondrial fatty acid oxidation (FAO). This strategy shows promise for conditions like CPT II and VLCAD deficiencies.
Area of Science:
- Biochemistry
- Genetics
- Metabolic Disorders
Background:
- Enzyme defects in mitochondrial fatty acid oxidation (FAO) represent a significant group of inherited metabolic diseases.
- Current pharmacological treatments for FAO disorders are limited, despite extensive clinical and genetic characterization.
Purpose of the Study:
- To investigate the potential of pharmacological correction for FAO disorders using PPAR activators.
- To explore the therapeutic efficacy of fibrates and other PPAR agonists in treating FAO deficiencies.
Main Methods:
- Review of existing data on PPAR signaling pathway regulation of mitochondrial FAO genes.
- Testing PPAR activators in cellular models derived from patients with CPT II and VLCAD deficiencies.
Main Results:
- Demonstrated that PPAR signaling pathway regulates genes involved in mitochondrial FAO.
- Observed positive effects of PPAR activators in cellular models of CPT II and VLCAD deficiencies.
Conclusions:
- PPAR activators represent a promising pharmacological strategy for treating FAO disorders.
- Further research is needed to fully understand the perspectives and limitations of this therapeutic approach.

Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:20
Inborn Errors of Metabolism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:48
ATP Synthase: Mechanism
 01:19
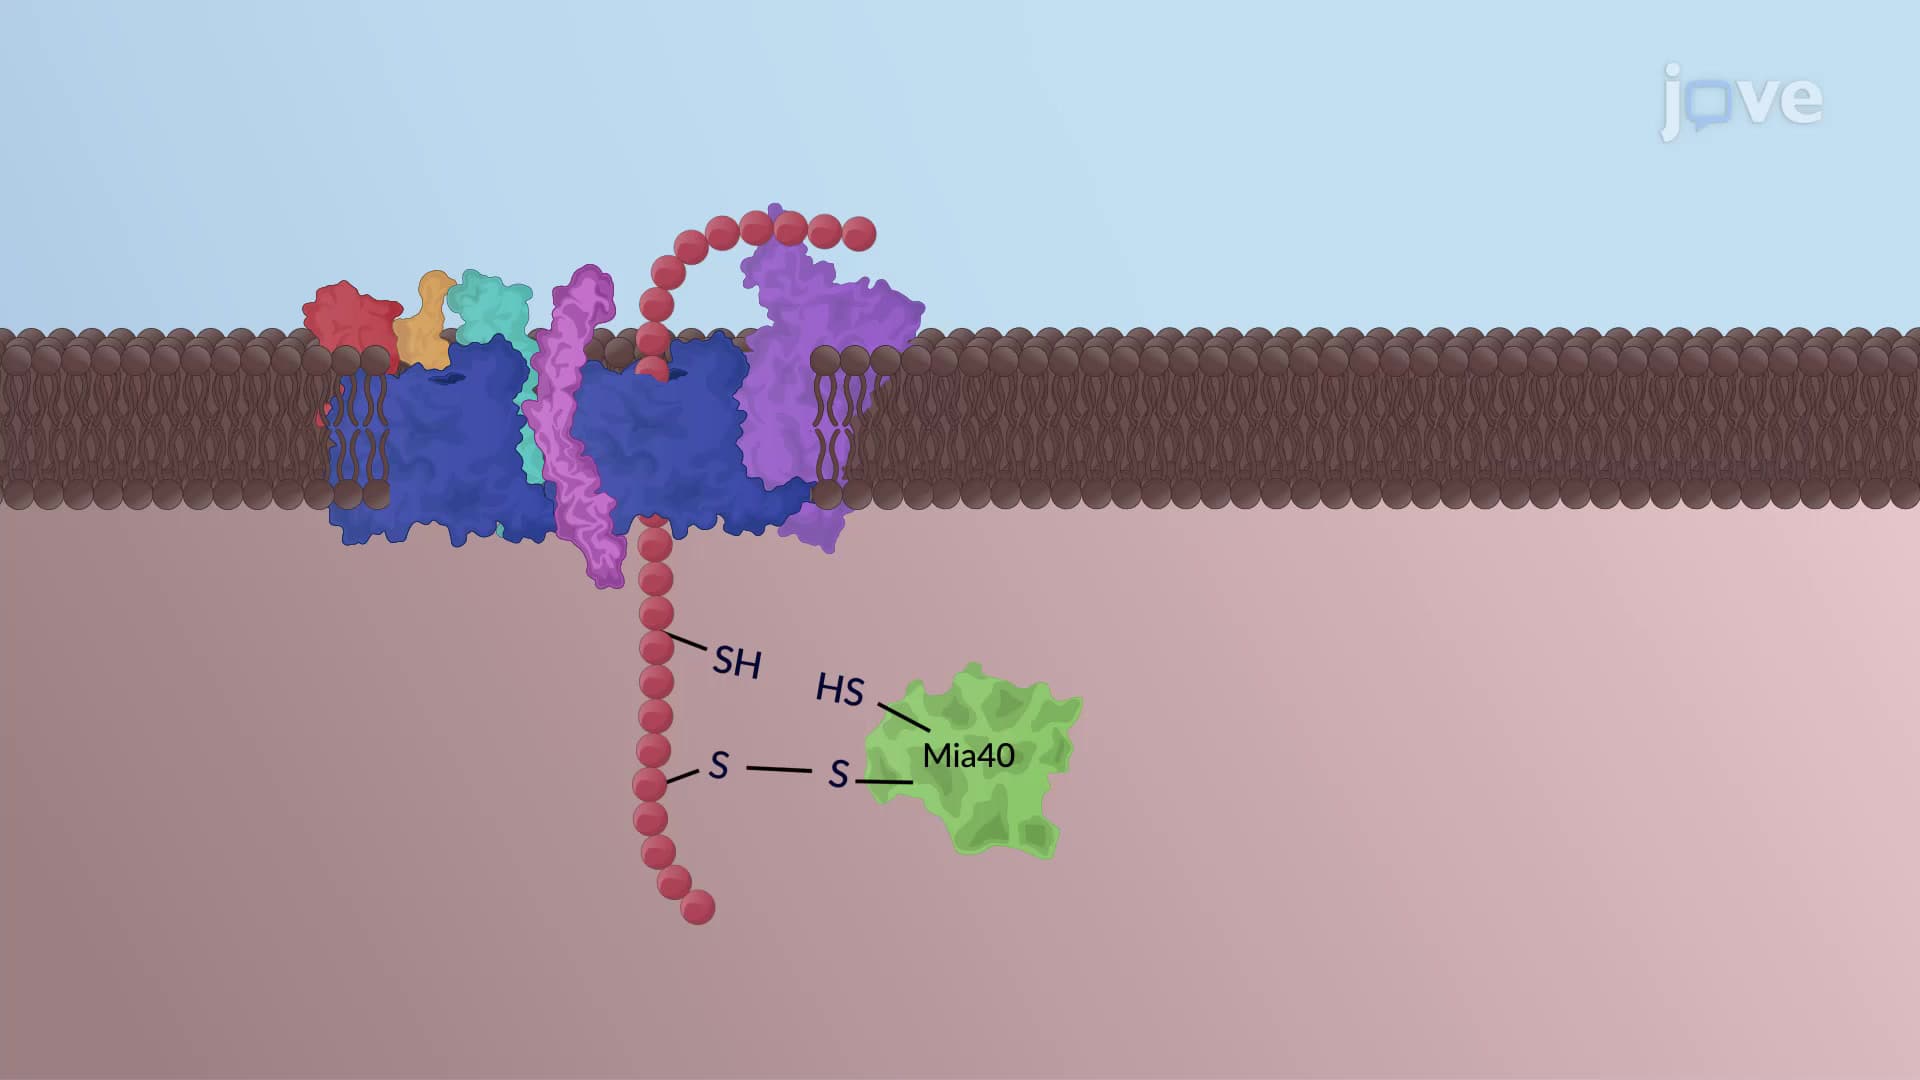
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:12
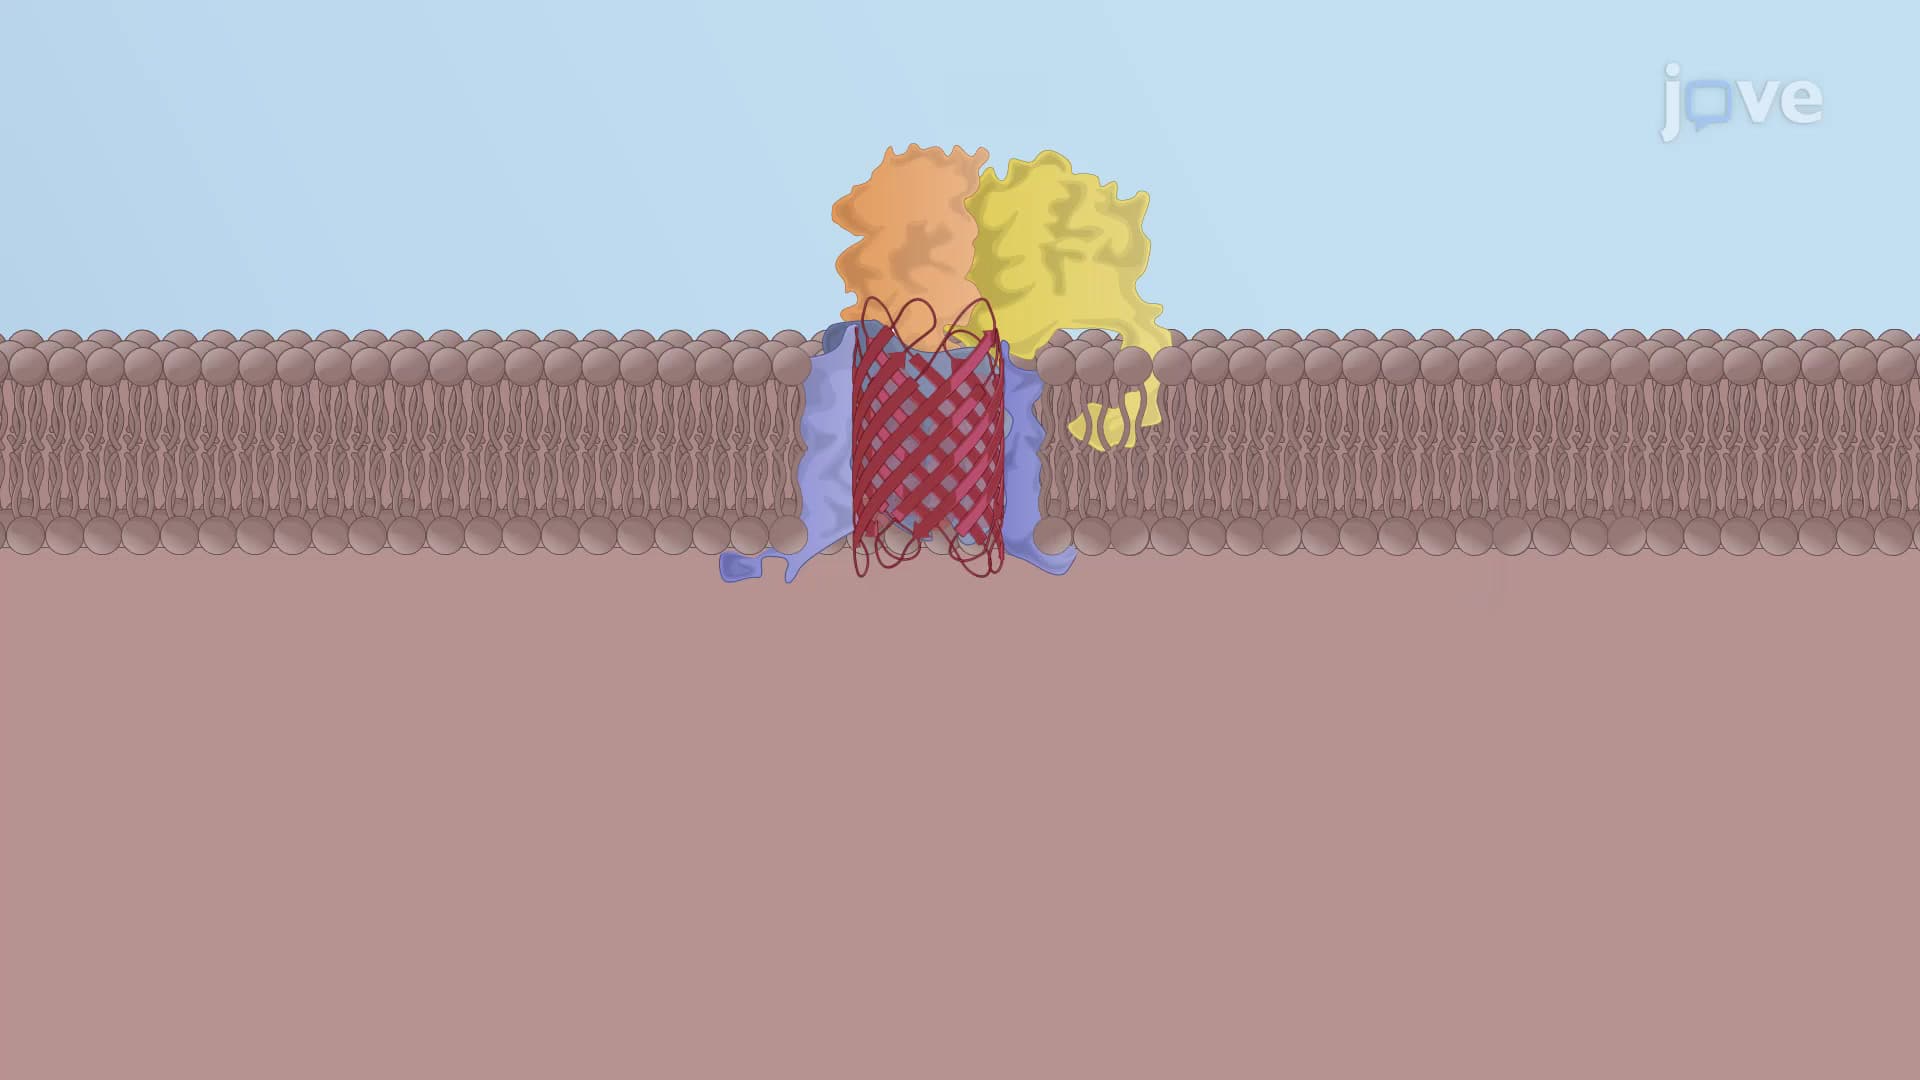
Porin Insertion in the Outer Mitochondrial Membrane
Three models describe the assembly of porins by the SAM complex and their insertion into the outer membrane. Model 1 suggests that porins are assembled outside the SAM channel as the...