In vitro metabolic stability of moisture-sensitive rabeprazole in human liver microsomes and its modulation by pharmaceutical excipients
Related Experiment Video
Updated: Jul 6, 2026
An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Shan Ren1, Mi-Jin Park, Aera Kim
1National Research Laboratory for Bioavailability Control, College of Pharmacy, Kangwon National University, Chuncheon 200-701, Korea.

Abstract:
A reliable method to assess in vitro metabolic stability of rabeprazole and its modulation by Generally Recognized As Safe (GRAS)-listed pharmaceutical excipients was established in human liver microsomes. The metabolic stability of rabeprazole decreased as a function of incubation time, resulting in the formation of thioether rabeprazole via nonenzymatic degradation and enzymatic metabolism. Buffer type was also a determining factor for the degree of both nonenzymatic degradation and enzymatic metabolism. The net extent of enzymatic drug metabolism, obtained by calculating the difference in drug degradation between a microsome-present reaction system and a microsome-free solution, was about 9.20 +/- 0.67% in phosphate buffer and 2.27 +/- 1.76% in Tris buffer, respectively. Rabeprazole exhibited first-order kinetics in microsome-free solution but showed non-linear kinetics in the microsome-present reaction system. The maximal velocity, Vmax, in phosphate buffer was 5.07 microg mL(-1) h(-1) and the Michaelis-Menten constant, Km, was 10.39 microg mL(-1) by computer-fitting to the classical Michaelis-Menten equation for pattern of time-dependent change in the substrate concentration. The intact drug and its thioether form were well resolved and successfully identified by HPLC chromatography and liquid chromatography mass spectroscopy (LC/MS). The metabolic stability of rabeprazole was also modulated by the presence of pharmaceutical excipients. Among the five pharmaceutical excipients tested, poloxamer 188 and Gelucire 44/14 had potentially inhibitory effects on rabeprazole metabolism in human liver microsomes (p < 0.05). A greater understanding of metabolic stability and its modulation by pharmaceutical excipients would be useful for optimizing the bioavailability of rabeprazole at the early formulation stages.
Related Concept Videos
 01:30
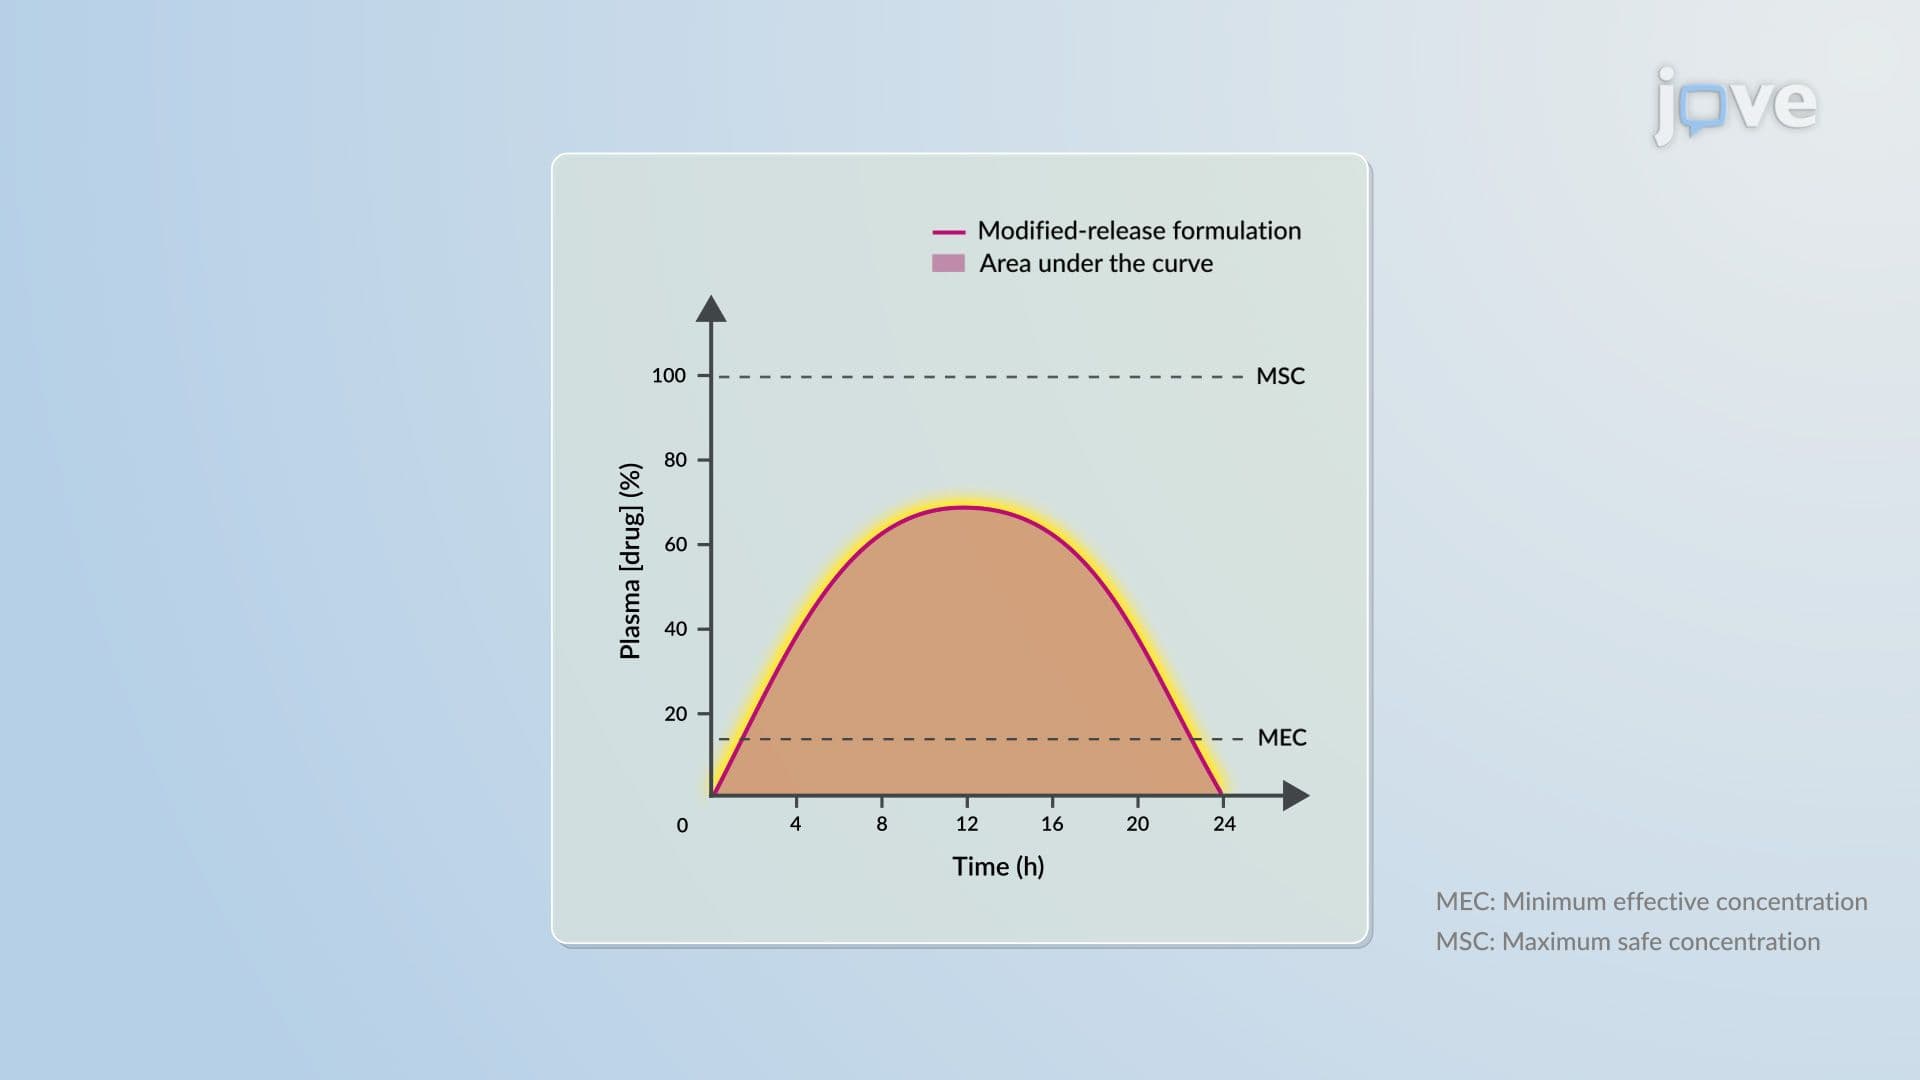
Modified-Release Drug Delivery Systems: Bioavailability
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
 01:05
Bioavailability Enhancement: Drug Stability Enhancement and GI Retention
 01:16
Drug Product Stability
 01:28
Factors Influencing Drug Absorption: Pharmaceutical Parameters
 01:16
Bioavailability Enhancement: Drug Solubility Enhancement