Related Experiment Video
Updated: Jul 5, 2026

Treating SCA1 Mice with Water-Soluble Compounds to Non-Specifically Boost Mitochondrial Function
Published on: January 22, 2017
Spinocerebellar ataxia 17 (SCA17) and Huntington's disease-like 4 (HDL4)
Giovanni Stevanin1, Alexis Brice
1INSERM U679, Groupe Pitié-Salpetrière, 47 Boulevard de l'Hopital, 75651 Paris Cedex 13, France. stevanin@ccr.jussieu.fr

Abstract:
Spinocerebellar ataxia 17 (SCA17) or Huntington's disease-like-4 is a neurodegenerative disease caused by the expansion above 44 units of a CAG/CAA repeat in the coding region of the TATA box binding protein (TBP) gene leading to an abnormal expansion of a polyglutamine stretch in the corresponding protein. Alleles with 43 and 44 repeats have been identified in sporadic cases and their pathogenicity remains uncertain. Furthermore, incomplete penetrance of pathological alleles with up to 49 repeats has been suggested. The imperfect nature of the repeat makes intergenerational instability extremely rare and de novo mutations are most likely the result of partial duplications. This is one of the rarer forms of autosomal dominant cerebellar ataxia but the associated phenotype is often severe, involving various systems (cerebral cortex, striatum, and cerebellum), with extremely variable age at onset (range: 3-75 years) and clinical presentation. This gene is thought to account for a small proportion of patients with a Huntington's disease-like phenotype and cerebellar signs. Parkinson's disease-like, Creutzfeldt-Jakob disease-like and Alzheimer disease-like phenotypes have also been described with small SCA17 expansions. The abnormal protein is expressed at the same level as its normal counterpart and forms neuronal intranuclear inclusions containing other proteins involved in protein folding or degradation. The increase in the size of the glutamine stretch enhances transcription in vitro, probably leading to transcription deregulation. Interestingly, the TBP protein mutated in SCA17 is recruited in the inclusions of other polyglutaminopathies, suggesting its involvement in the transcription down-regulation observed in these diseases.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:23
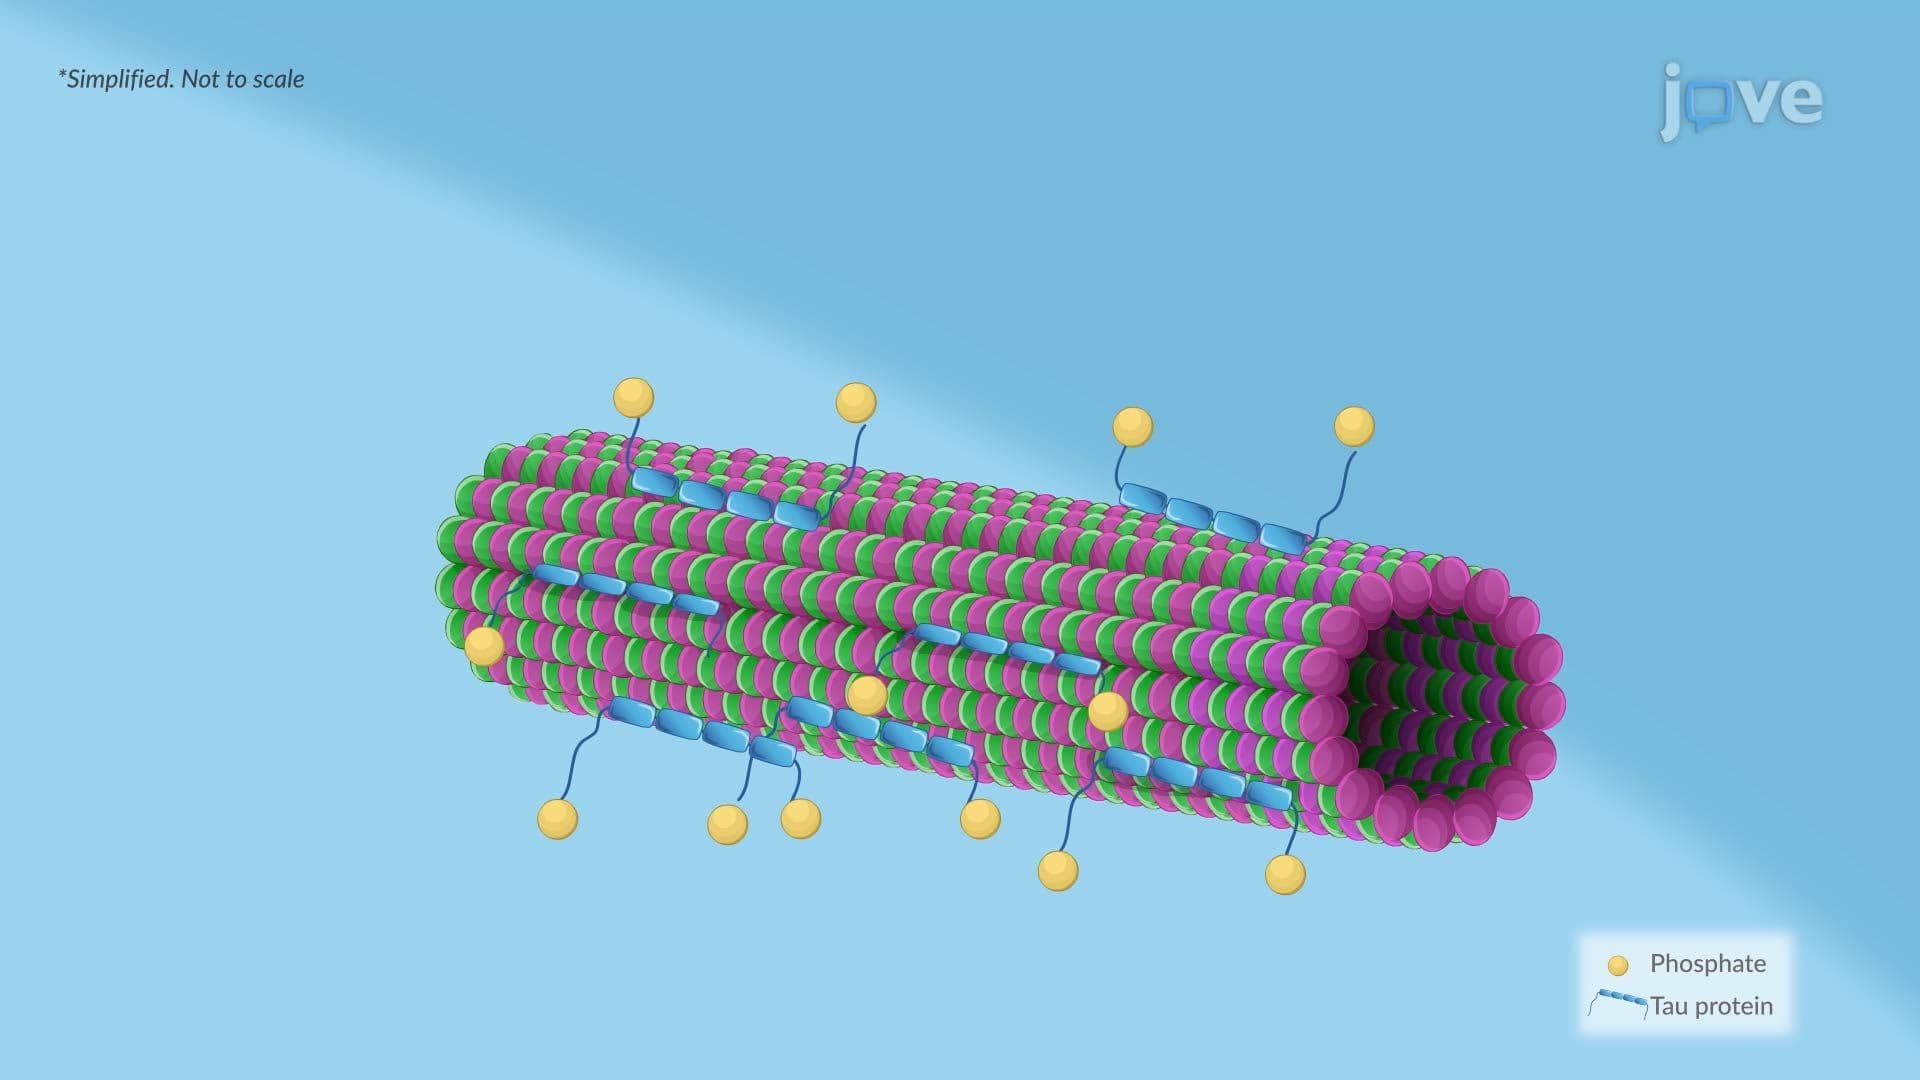
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:24
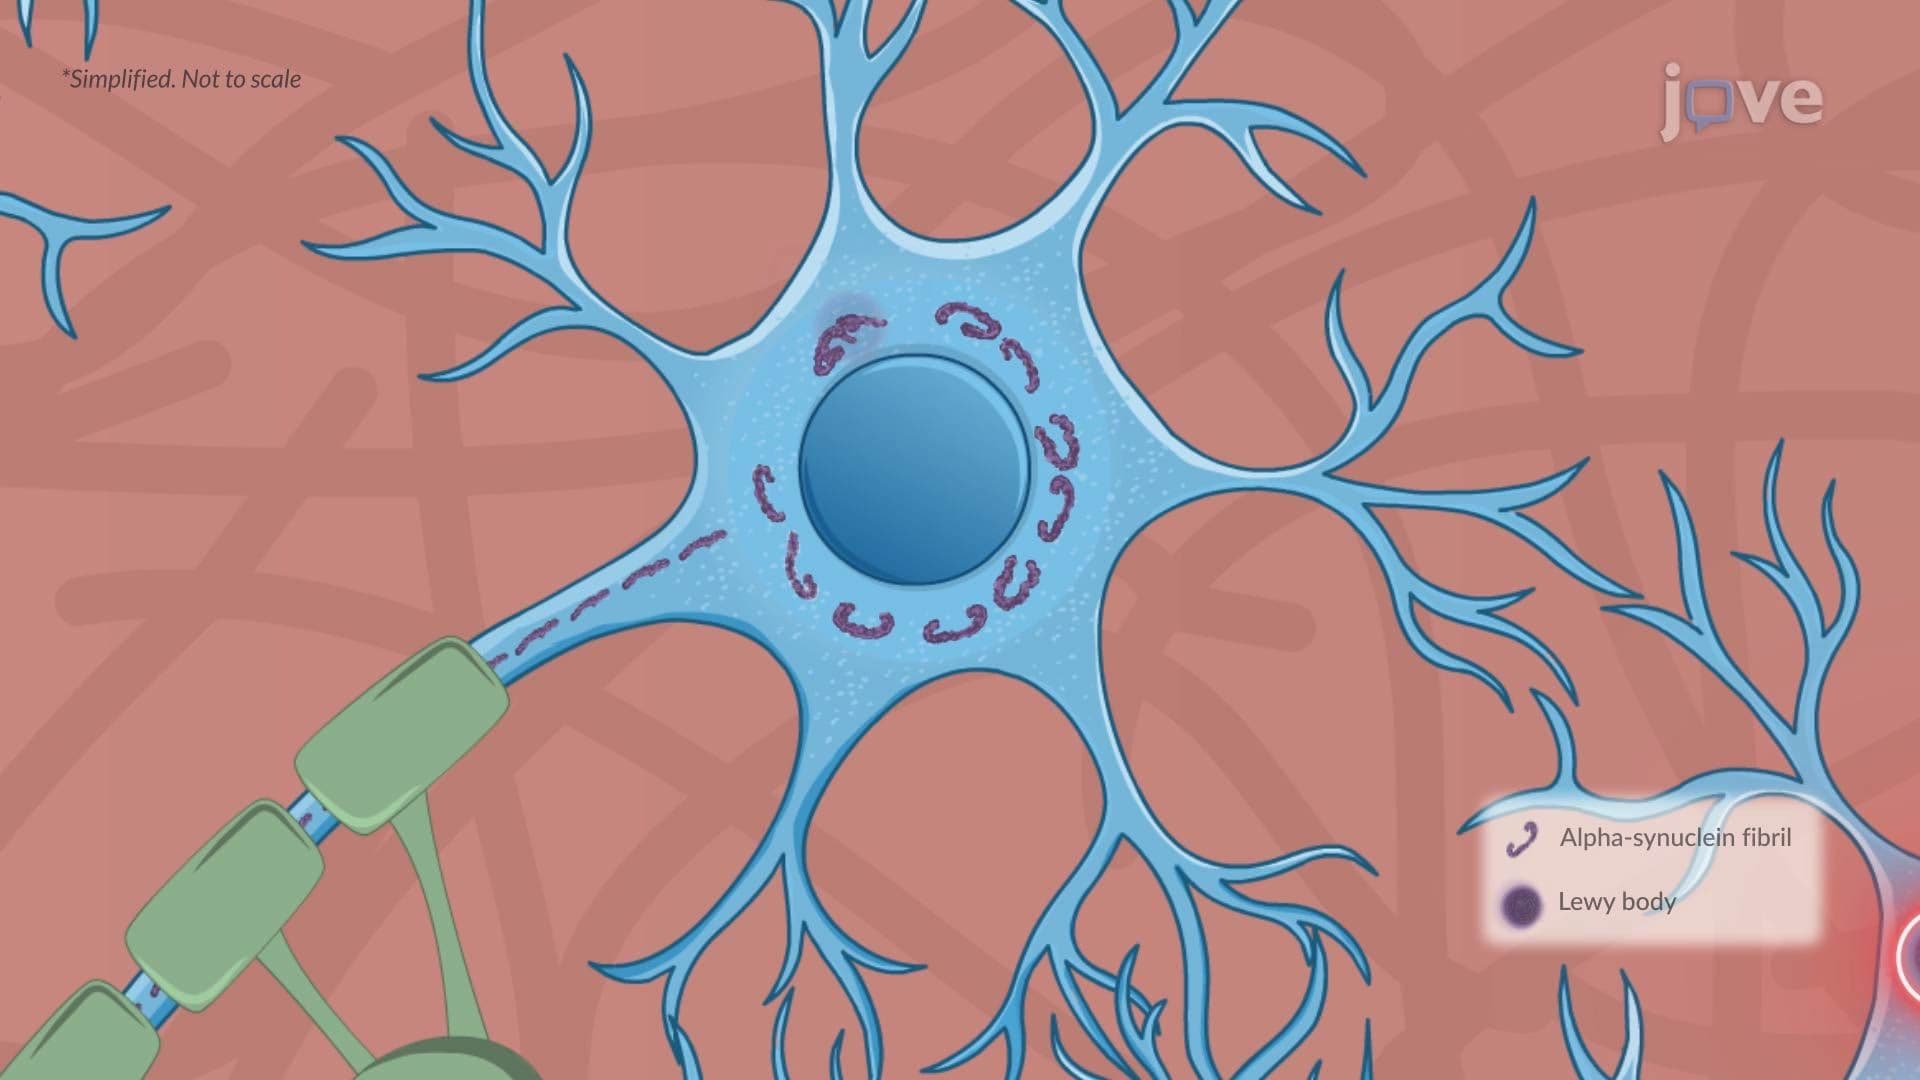
Parkinson Disease ll: Pathophysiology
 01:11
Genetic Lingo