Related Experiment Video
Updated: Jul 5, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Monte Carlo free energy calculations using electronic structure methods
Daniel R Matusek1, Sébastien Osborne, Alain St-Amant
1Department of Chemistry, University of Ottawa, 10 Marie Curie, Ottawa, ON, Canada.

Abstract:
The molecular mechanics-based importance sampling function (MMBIF) algorithm [R. Iftimie, D. Salahub, D. Wei, and J. Schofield, J. Chem. Phys. 113, 4852 (2000)] is extended to incorporate semiempirical electronic structure methods in the secondary Markov chain, creating a fully quantum mechanical Monte Carlo sampling method for simulations of reactive chemical systems which, unlike the MMBIF algorithm, does not require the generation of a system-specific force field. The algorithm is applied to calculating the potential of mean force for the isomerization reaction of HCN using thermodynamic integration. Constraints are implemented in the sampling using a modification of the SHAKE algorithm, including that of a fixed, arbitrary reaction coordinate. Simulation results show that sampling efficiency with the semiempirical secondary potential is often comparable in quality to force fields constructed using the methods suggested in the original MMBIF work. The semiempirical based importance sampling method presented here is a useful alternative to MMBIF sampling as it can be applied to systems for which no suitable MM force field can be constructed.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
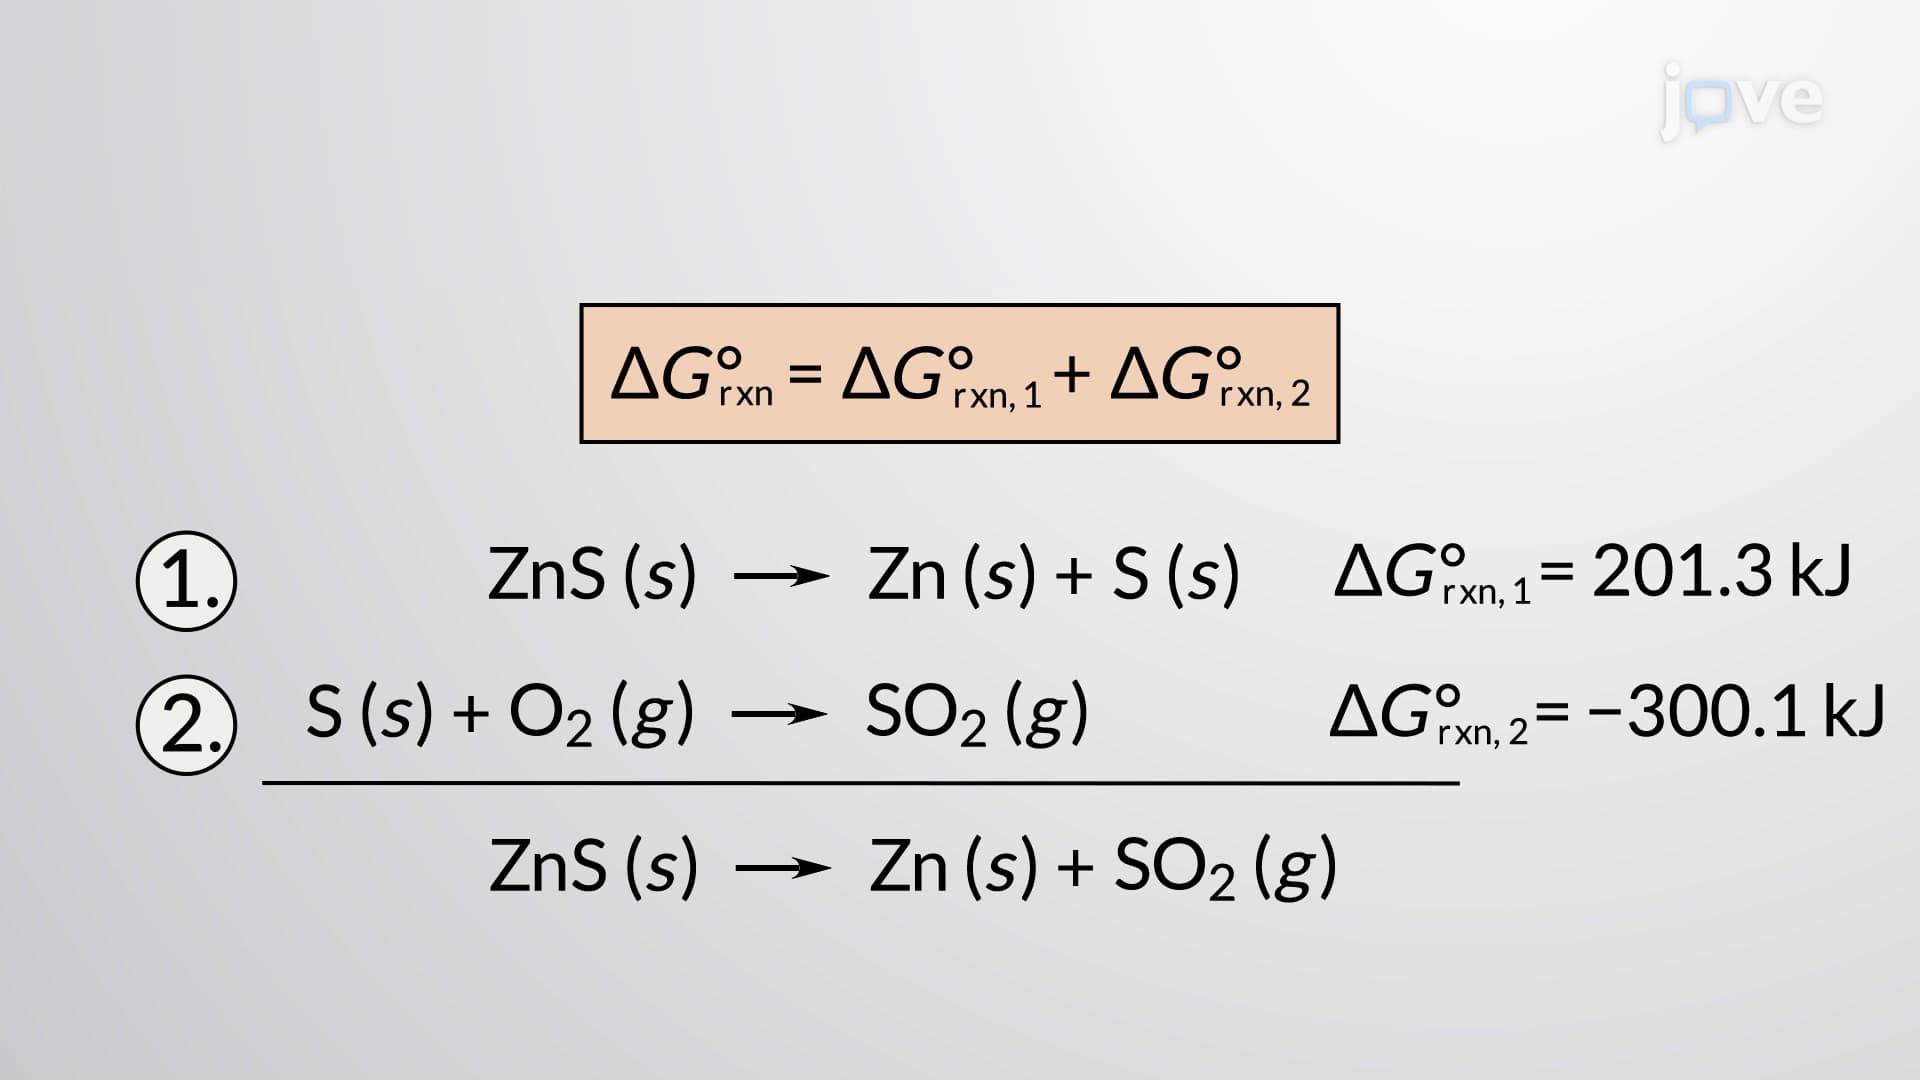 02:49
Calculating Standard Free Energy Changes
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 03:25
Free Energy Changes for Nonstandard States
 01:26
Thermodynamic Potentials
 01:27
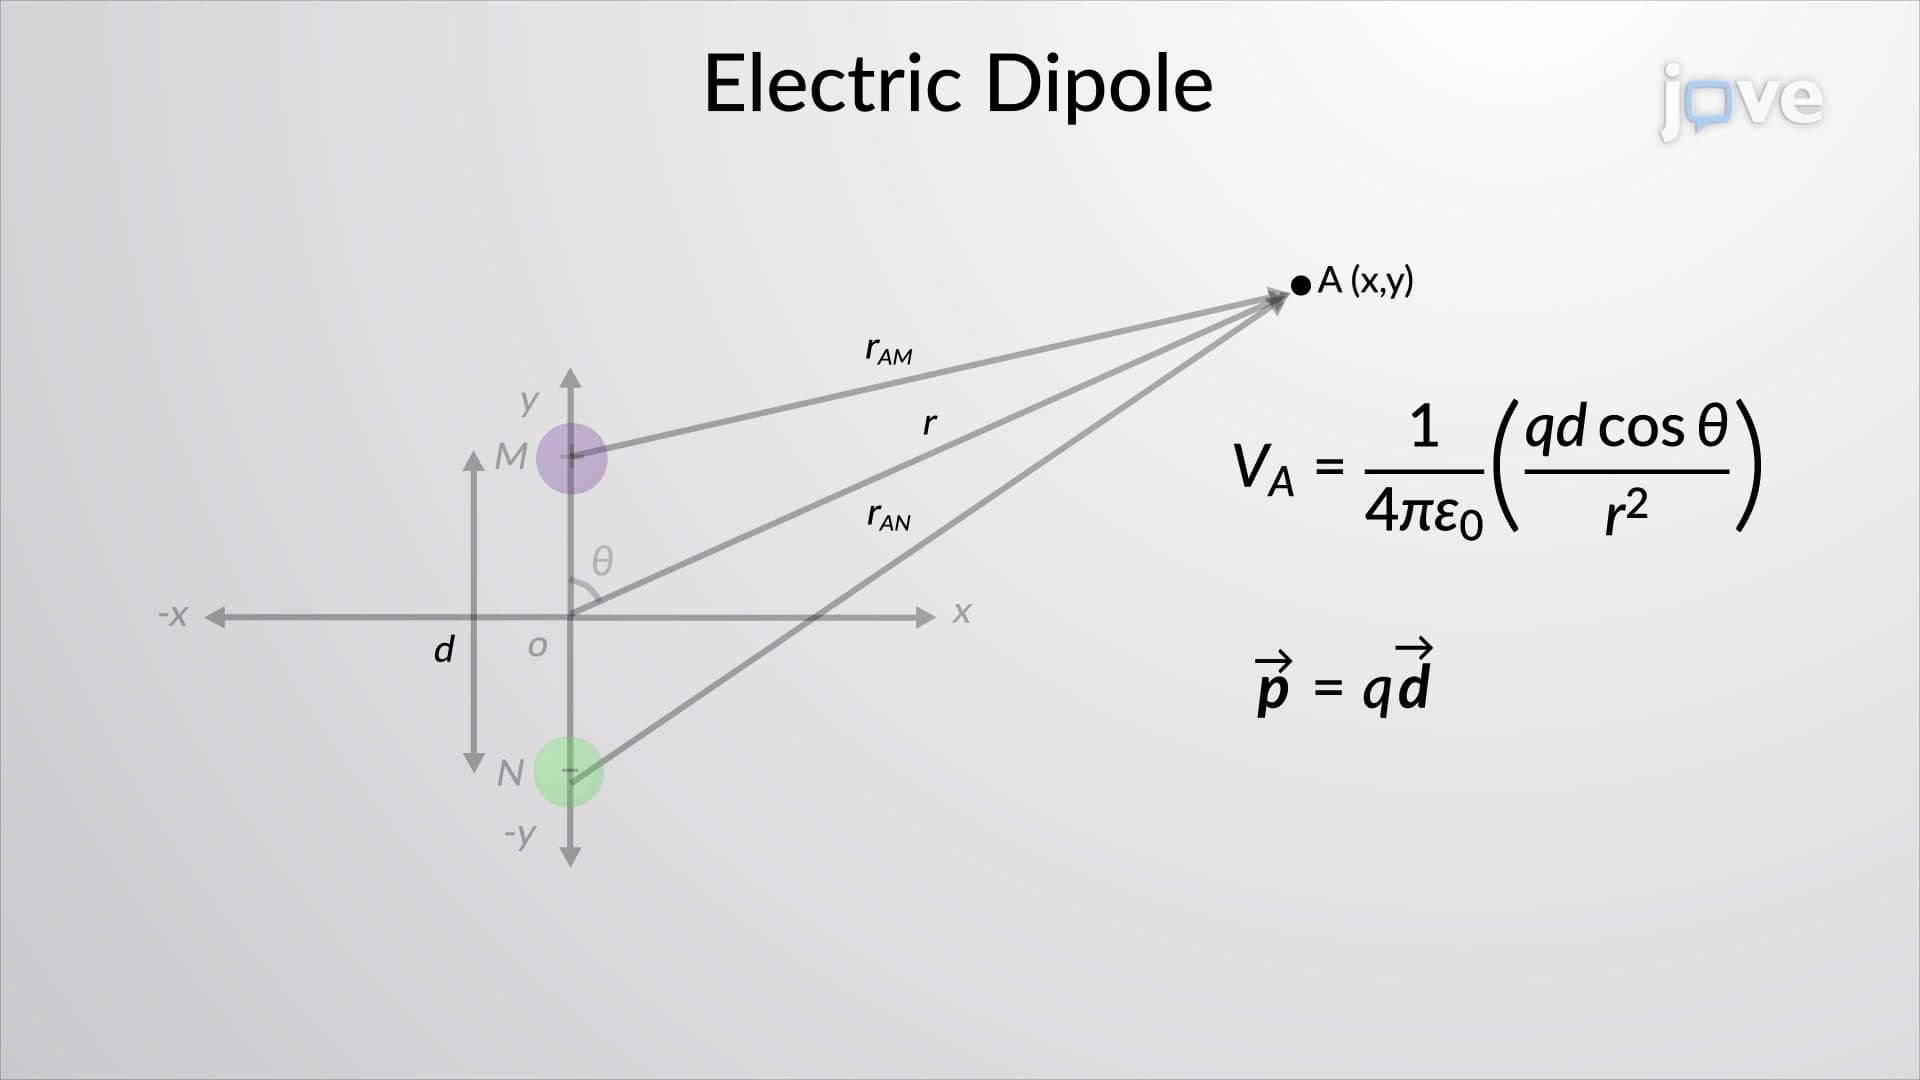
Calculations of Electric Potential II
Consider a...
 01:27
The Molecular Nature of Internal Energy