Related Experiment Video
Updated: Jul 5, 2026

A Fast and Reliable Pipeline for Bacterial Transcriptome Analysis Case study: Serine-dependent Gene Regulation in Streptococcus pneumoniae
Published on: April 25, 2015
Large-scale computational and statistical analyses of high transcription potentialities in 32 prokaryotic genomes
Christine Sinoquet1, Sylvain Demey, Frédérique Braun
1Computer Science Institute of Nantes-Atlantic (Lina), U.M.R. C.N.R.S. 6241, University of Nantes, 2 rue de la Houssinière, BP 92208, 44322 Nantes Cedex, France. christine.sinoquet@univ-nantes.fr

Abstract:
This article compares 32 bacterial genomes with respect to their high transcription potentialities. The sigma70 promoter has been widely studied for Escherichia coli model and a consensus is known. Since transcriptional regulations are known to compensate for promoter weakness (i.e. when the promoter similarity with regard to the consensus is rather low), predicting functional promoters is a hard task. Instead, the research work presented here comes within the scope of investigating potentially high ORF expression, in relation with three criteria: (i) high similarity to the sigma70 consensus (namely, the consensus variant appropriate for each genome), (ii) transcription strength reinforcement through a supplementary binding site--the upstream promoter (UP) element--and (iii) enhancement through an optimal Shine-Dalgarno (SD) sequence. We show that in the AT-rich Firmicutes' genomes, frequencies of potentially strong sigma70-like promoters are exceptionally high. Besides, though they contain a low number of strong promoters (SPs), some genomes may show a high proportion of promoters harbouring an UP element. Putative SPs of lesser quality are more frequently associated with an UP element than putative strong promoters of better quality. A meaningful difference is statistically ascertained when comparing bacterial genomes with similarly AT-rich genomes generated at random; the difference is the highest for Firmicutes. Comparing some Firmicutes genomes with similarly AT-rich Proteobacteria genomes, we confirm the Firmicutes specificity. We show that this specificity is neither explained by AT-bias nor genome size bias; neither does it originate in the abundance of optimal SD sequences, a typical and significant feature of Firmicutes more thoroughly analysed in our study.
More Related Videos
 12:54
Real-time Analysis of Transcription Factor Binding, Transcription, Translation, and Turnover to Display Global Events During Cellular Activation
Published on: March 7, 2018
 08:03
Heuristic Mining of Hierarchical Genotypes and Accessory Genome Loci in Bacterial Populations
Published on: December 7, 2021
Related Concept Videos
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique helps...
 01:29
Modern Molecular Taxonomy
 01:28
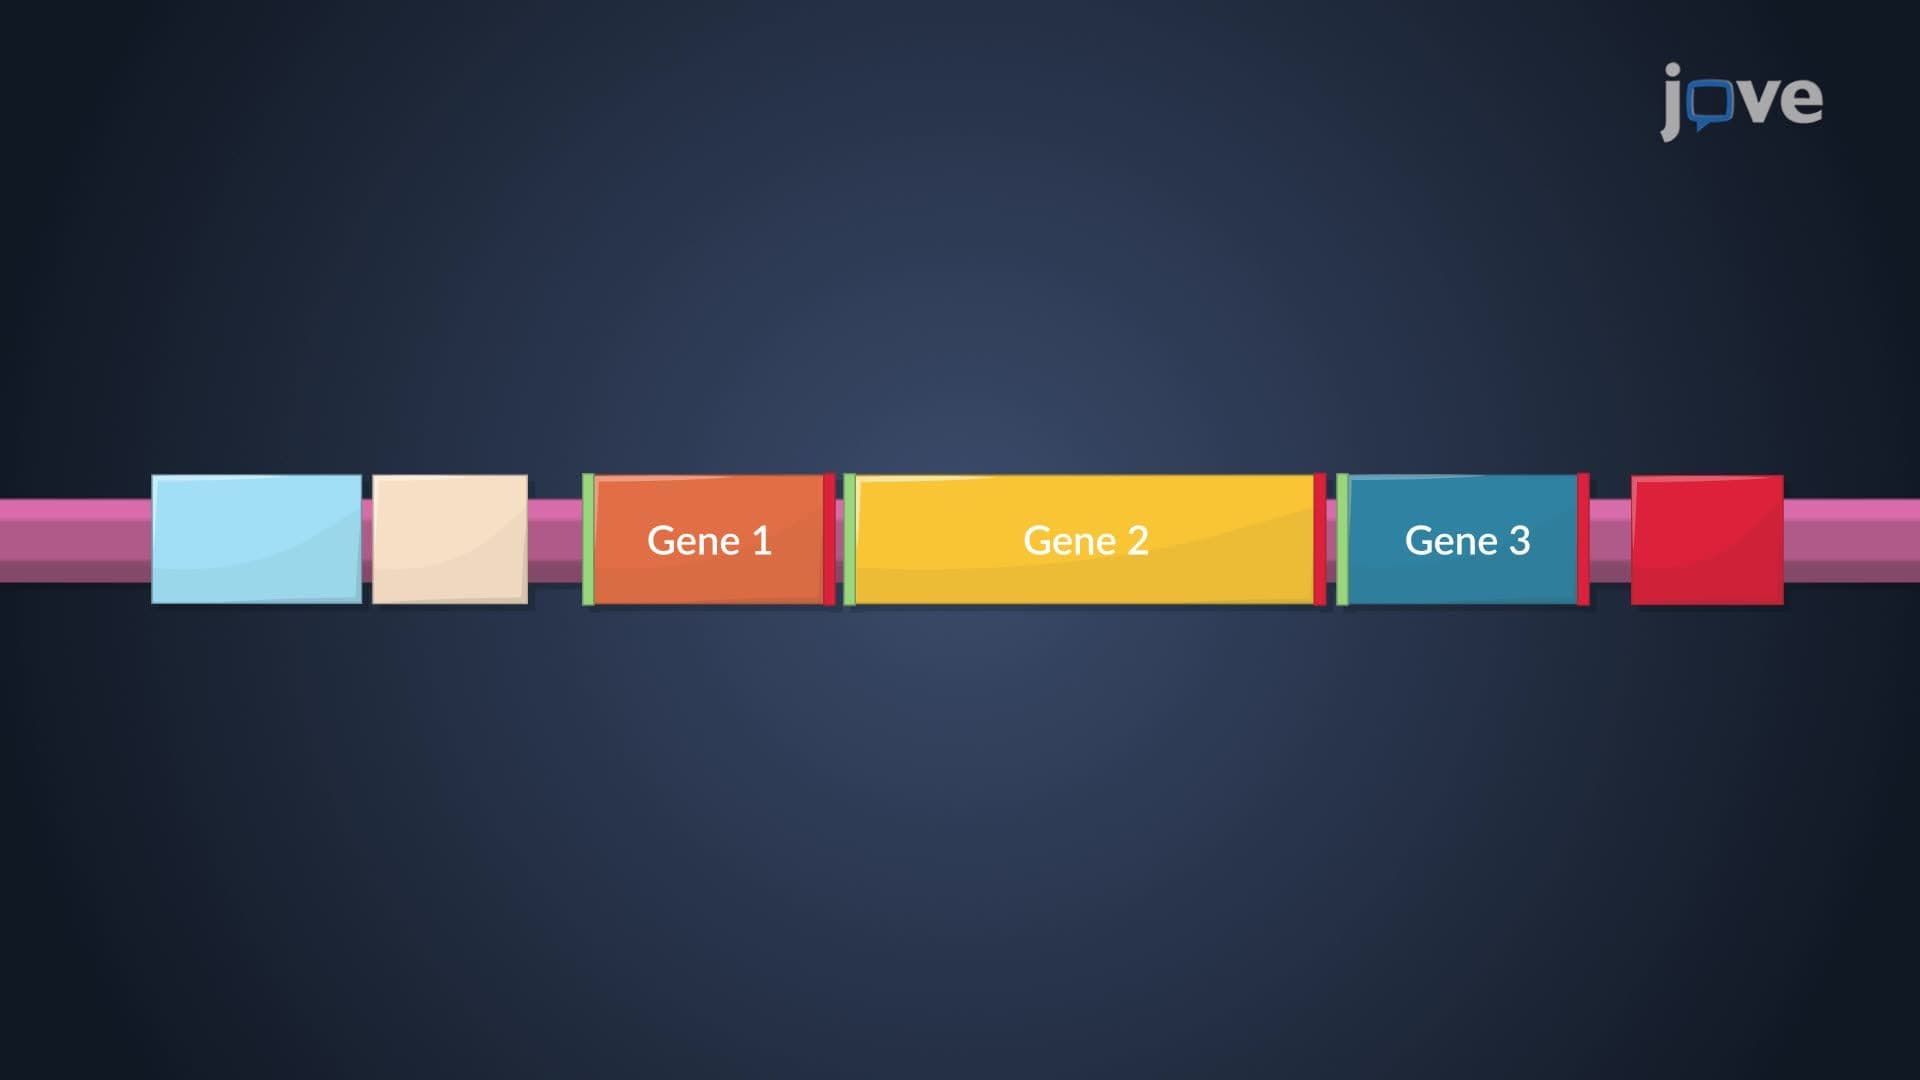
Prokaryotic Gene Structure and Organization
 01:28
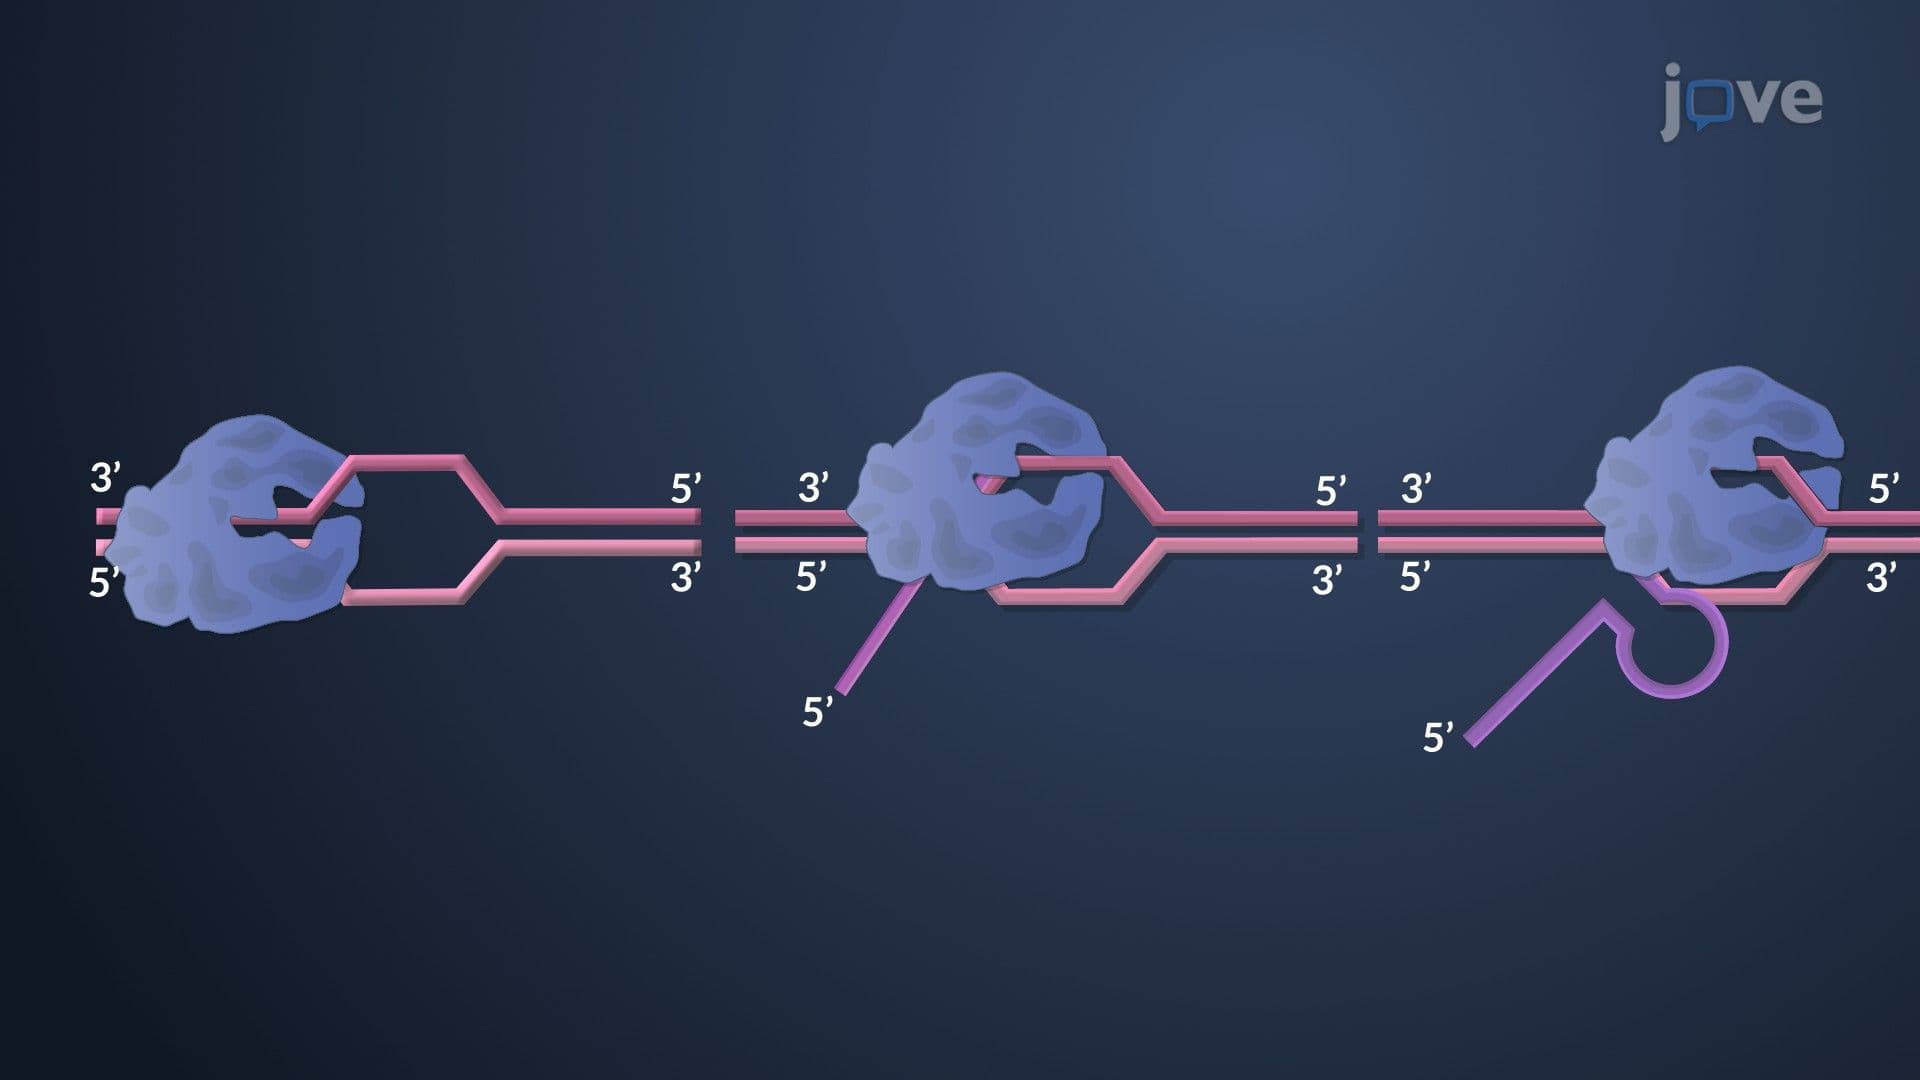
Transcription in Prokaryotes