Related Experiment Video
Updated: Jul 5, 2026
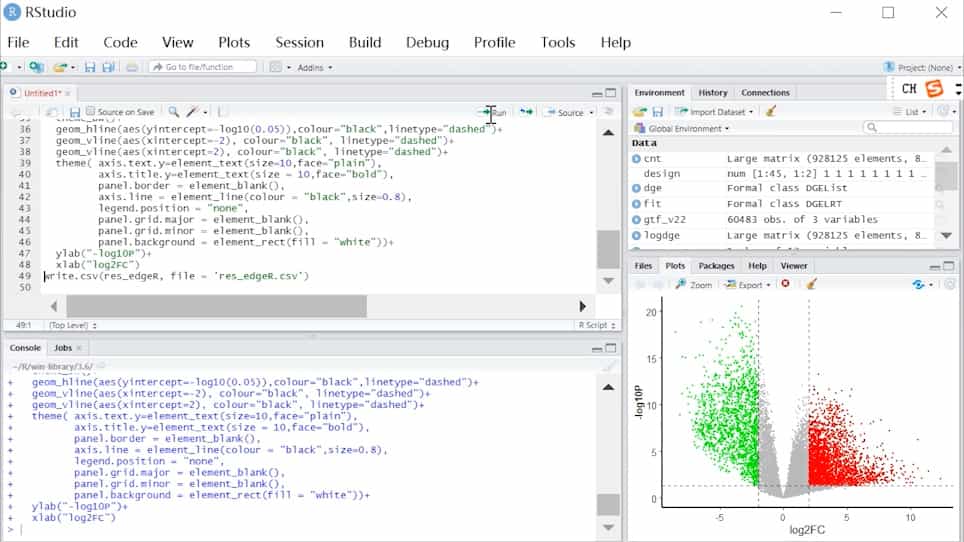

Three Differential Expression Analysis Methods for RNA Sequencing: limma, EdgeR, DESeq2
Published on: September 18, 2021
An efficient method to identify differentially expressed genes in microarray experiments
Huaizhen Qin1, Tao Feng, Scott A Harding
1Department of Mathematical Sciences, Biotechnology Research Center, School of Forest Resources and Environmental Science, Michigan Technological University, Houghton, MI 49931, USA.

Motivation:
Microarray experiments typically analyze thousands to tens of thousands of genes from small numbers of biological replicates. The fact that genes are normally expressed in functionally relevant patterns suggests that gene-expression data can be stratified and clustered into relatively homogenous groups. Cluster-wise dimensionality reduction should make it feasible to improve screening power while minimizing information loss.
Results:
We propose a powerful and computationally simple method for finding differentially expressed genes in small microarray experiments. The method incorporates a novel stratification-based tight clustering algorithm, principal component analysis and information pooling. Comprehensive simulations show that our method is substantially more powerful than the popular SAM and eBayes approaches. We applied the method to three real microarray datasets: one from a Populus nitrogen stress experiment with 3 biological replicates; and two from public microarray datasets of human cancers with 10 to 40 biological replicates. In all three analyses, our method proved more robust than the popular alternatives for identification of differentially expressed genes.
Availability:
The C++ code to implement the proposed method is available upon request for academic use.