
Fragmentation-based QM/MM simulations: length dependence of chain dynamics and hydrogen bonding of polyethylene oxide
1School of Chemistry and Chemical Engineering, Institute of Theoretical and Computational Chemistry, Key Laboratory of Mesoscopic Chemistry of MOE, Nanjing University, Nanjing, P. R. China.

Abstract:
Molecular fragmentation quantum mechanics (QM) calculations have been combined with molecular mechanics (MM) to construct the fragmentation QM/MM method for simulations of dilute solutions of macromolecules. We adopt the electrostatics embedding QM/MM model, where the low-cost generalized energy-based fragmentation calculations are employed for the QM part. Conformation energy calculations, geometry optimizations, and Born-Oppenheimer molecular dynamics simulations of poly(ethylene oxide), PEO(n) (n = 6-20), and polyethylene, PE(n) ( n = 9-30), in aqueous solution have been performed within the framework of both fragmentation and conventional QM/MM methods. The intermolecular hydrogen bonding and chain configurations obtained from the fragmentation QM/MM simulations are consistent with the conventional QM/MM method. The length dependence of chain conformations and dynamics of PEO and PE oligomers in aqueous solutions is also investigated through the fragmentation QM/MM molecular dynamics simulations.
More Related Videos
Related Concept Videos
 01:17
Radical Chain-Growth Polymerization: Chain Branching
 02:35
Free-Radical Chain Reaction and Polymerization of Alkenes
 01:27
Determination of Molar Masses of Polymers II
 01:08
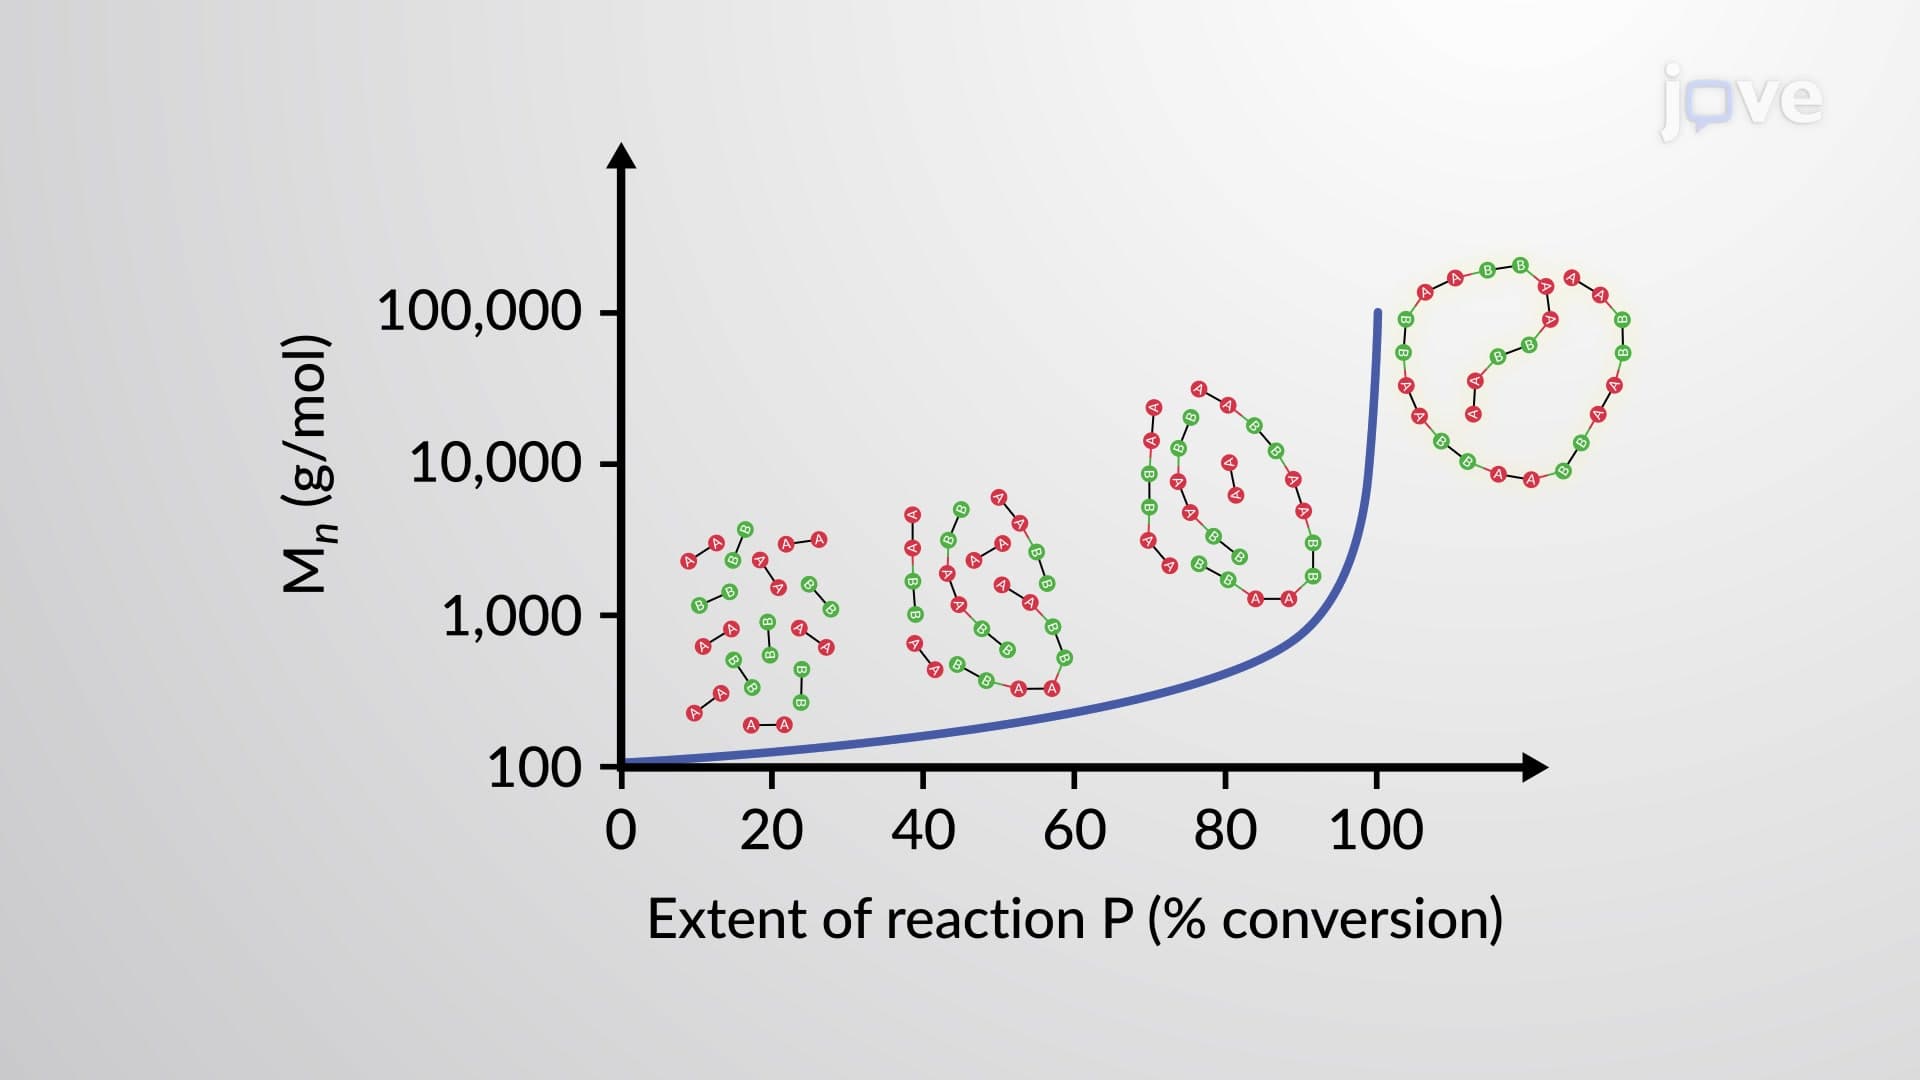
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:18
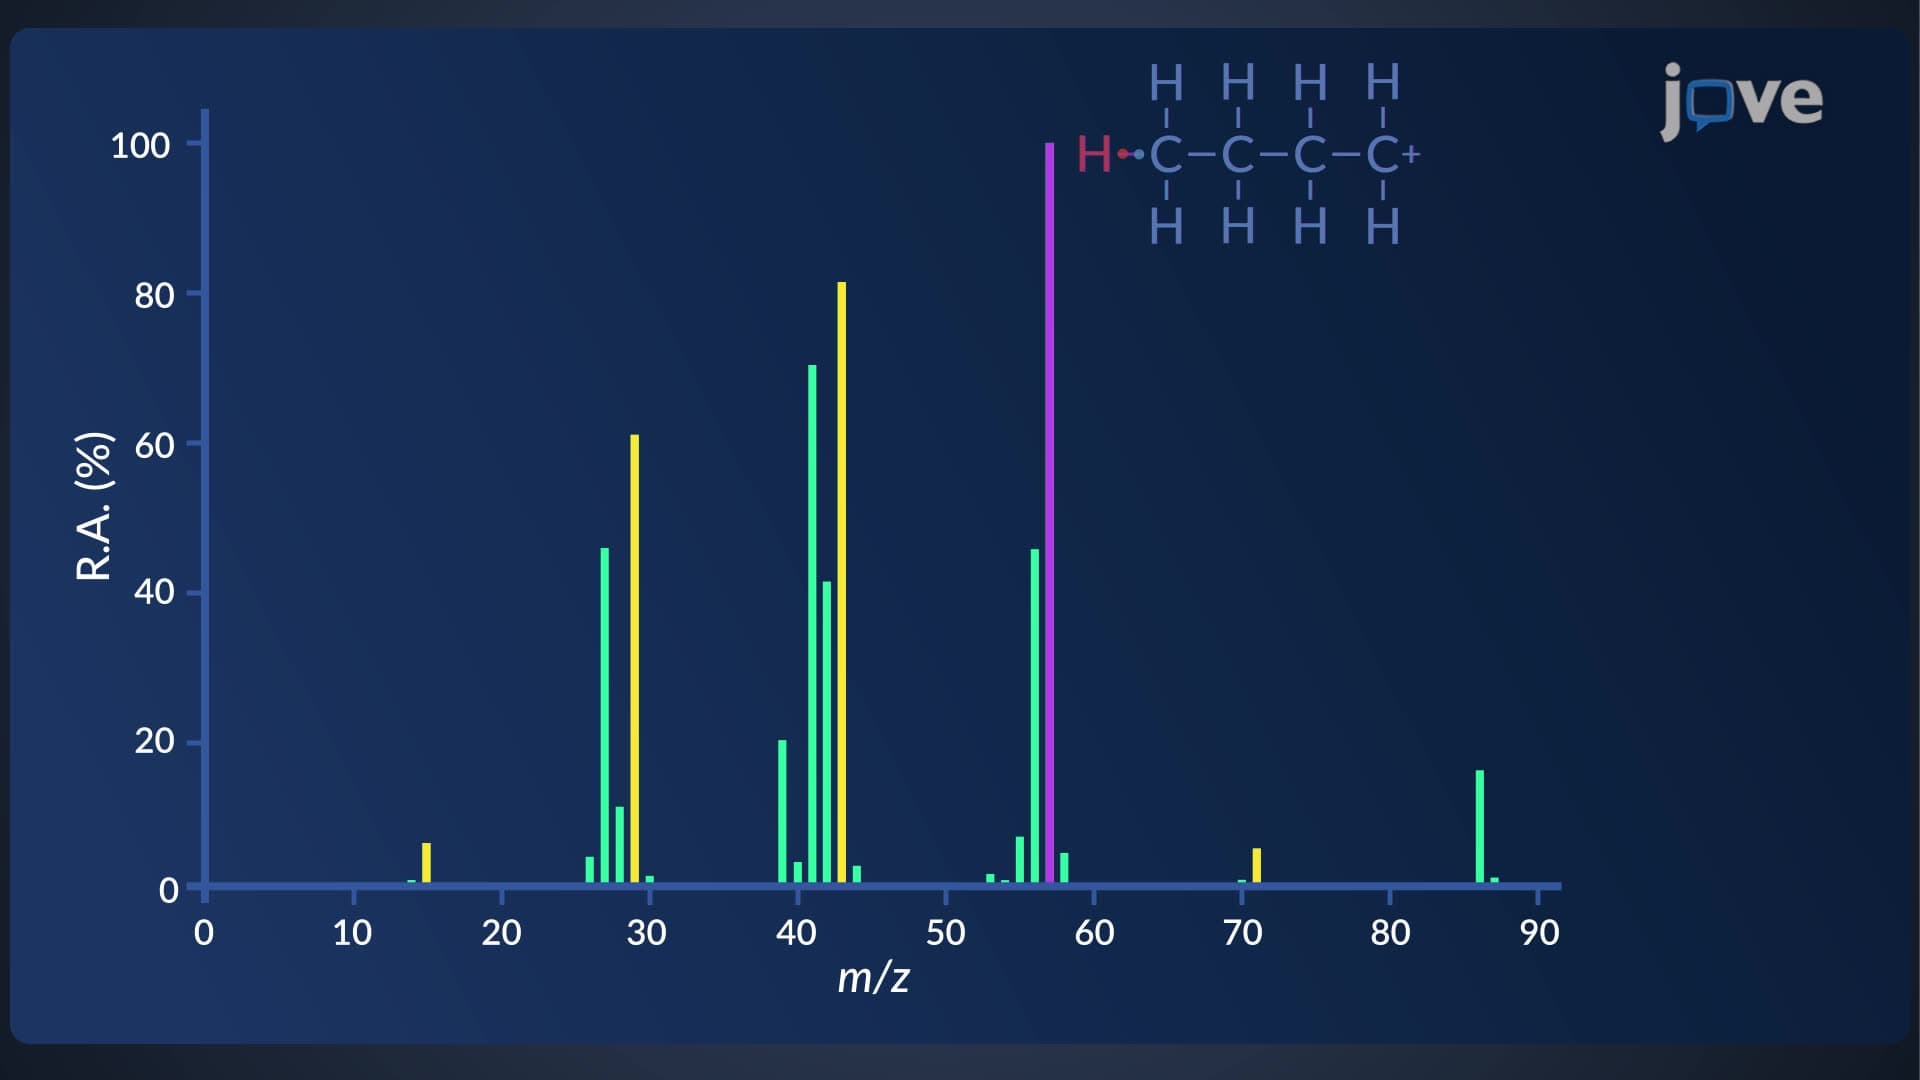
Mass Spectrometry: Long-Chain Alkane Fragmentation
 01:24
Determination of Molar Masses of Polymers I