Related Experiment Video
Updated: Jul 5, 2026

Visualizing the DNA Damage Response in Purkinje Cells Using Cerebellar Organotypic Cultures
Published on: December 27, 2024
Pyruvate dehydrogenase deficiency presenting as intermittent isolated acute ataxia
F-G Debray1, M Lambert, R Gagne
1Division of Medical Genetics, Department of Pediatrics, CHU Sainte-Justine, Université de Montreal, Montreal, Québec, Canada.

Insights
Pyruvate dehydrogenase (PDH) deficiency can present with unusual intermittent neurological symptoms, even with normal lactate levels. Early recognition is crucial for managing this rare genetic disorder.
Area of Science:
- Biochemistry
- Genetics
- Neurology
Background:
- Pyruvate dehydrogenase (PDH) deficiency is a rare metabolic disorder affecting cellular energy production.
- Unusual and intermittent clinical presentations can delay diagnosis.
Observation:
- Two siblings with a novel PDHA1 gene mutation (G585C) exhibited recurrent isolated ataxia in infancy.
- Symptoms initially resolved but progressed to severe encephalopathy, with death in their twenties.
- Normal blood and CSF lactate levels were observed, and no thiamine responsiveness was noted.
Findings:
- A new mutation (G585C) in the PDHA1 gene was identified as the cause of PDH deficiency.
- The mutation, though in the thiamine pyrophosphate binding domain, did not result in thiamine responsiveness.
- The study highlights a spectrum of intermittent neurological symptoms associated with PDH deficiency.
Implications:
- PDH deficiency should be considered in the differential diagnosis of unexplained intermittent neurological symptoms.
- The long-term prognosis for PDH deficiency remains uncertain and requires further investigation.
- Normal CSF lactate levels do not exclude the diagnosis of PDH deficiency.
Objective:
The aim of this study is to report and emphasize unusual presentations of pyruvate dehydrogenase (PDH) deficiency (OMIM 312170).
Methods:
PDH activity and PDHA1 gene were studied in two siblings presenting with intermittent ataxia in childhood. Similar presentations in reported PDH-deficient patients were searched for using the Medline database.
Results:
Both patients had PDH deficiency caused by a new mutation (G585C) in the PDHA1 gene, which is predicted to replace a highly conserved glycine at codon 195 by alanine. Although this mutation lies within the thiamine pyrophosphate binding domain, there was no thiamine responsiveness IN VIVO. The patients presented recurrent episodes of acute isolated ataxia in infancy. Both had normal blood and CSF lactate levels. Although symptoms initially resolved between episodes during the first decade, both patients subsequently worsened and developed progressive and severe encephalopathy, leading to death in their twenties. The spectrum of intermittent presentations in PDH deficiency includes episodic ataxia, intermittent peripheral weakness, recurrent dystonia and extrapyramidal movement disorders.
Conclusions:
PDH deficiency should be considered in patients with unexplained intermittent and recurrent acute neurological symptoms. Long-term prognosis and outcome remain uncertain. PDH deficiency can occur even with normal CSF lactate concentration.
More Related Videos
 09:40
Phosphorus-31 Magnetic Resonance Spectroscopy: A Tool for Measuring In Vivo Mitochondrial Oxidative Phosphorylation Capacity in Human Skeletal Muscle
Published on: January 19, 2017
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase (COX/SDH) Double-labeling Histochemistry
Published on: November 23, 2011
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:29
Hepatic Encephalopathy
 01:21
Huntington Disease l: Introduction
 01:15
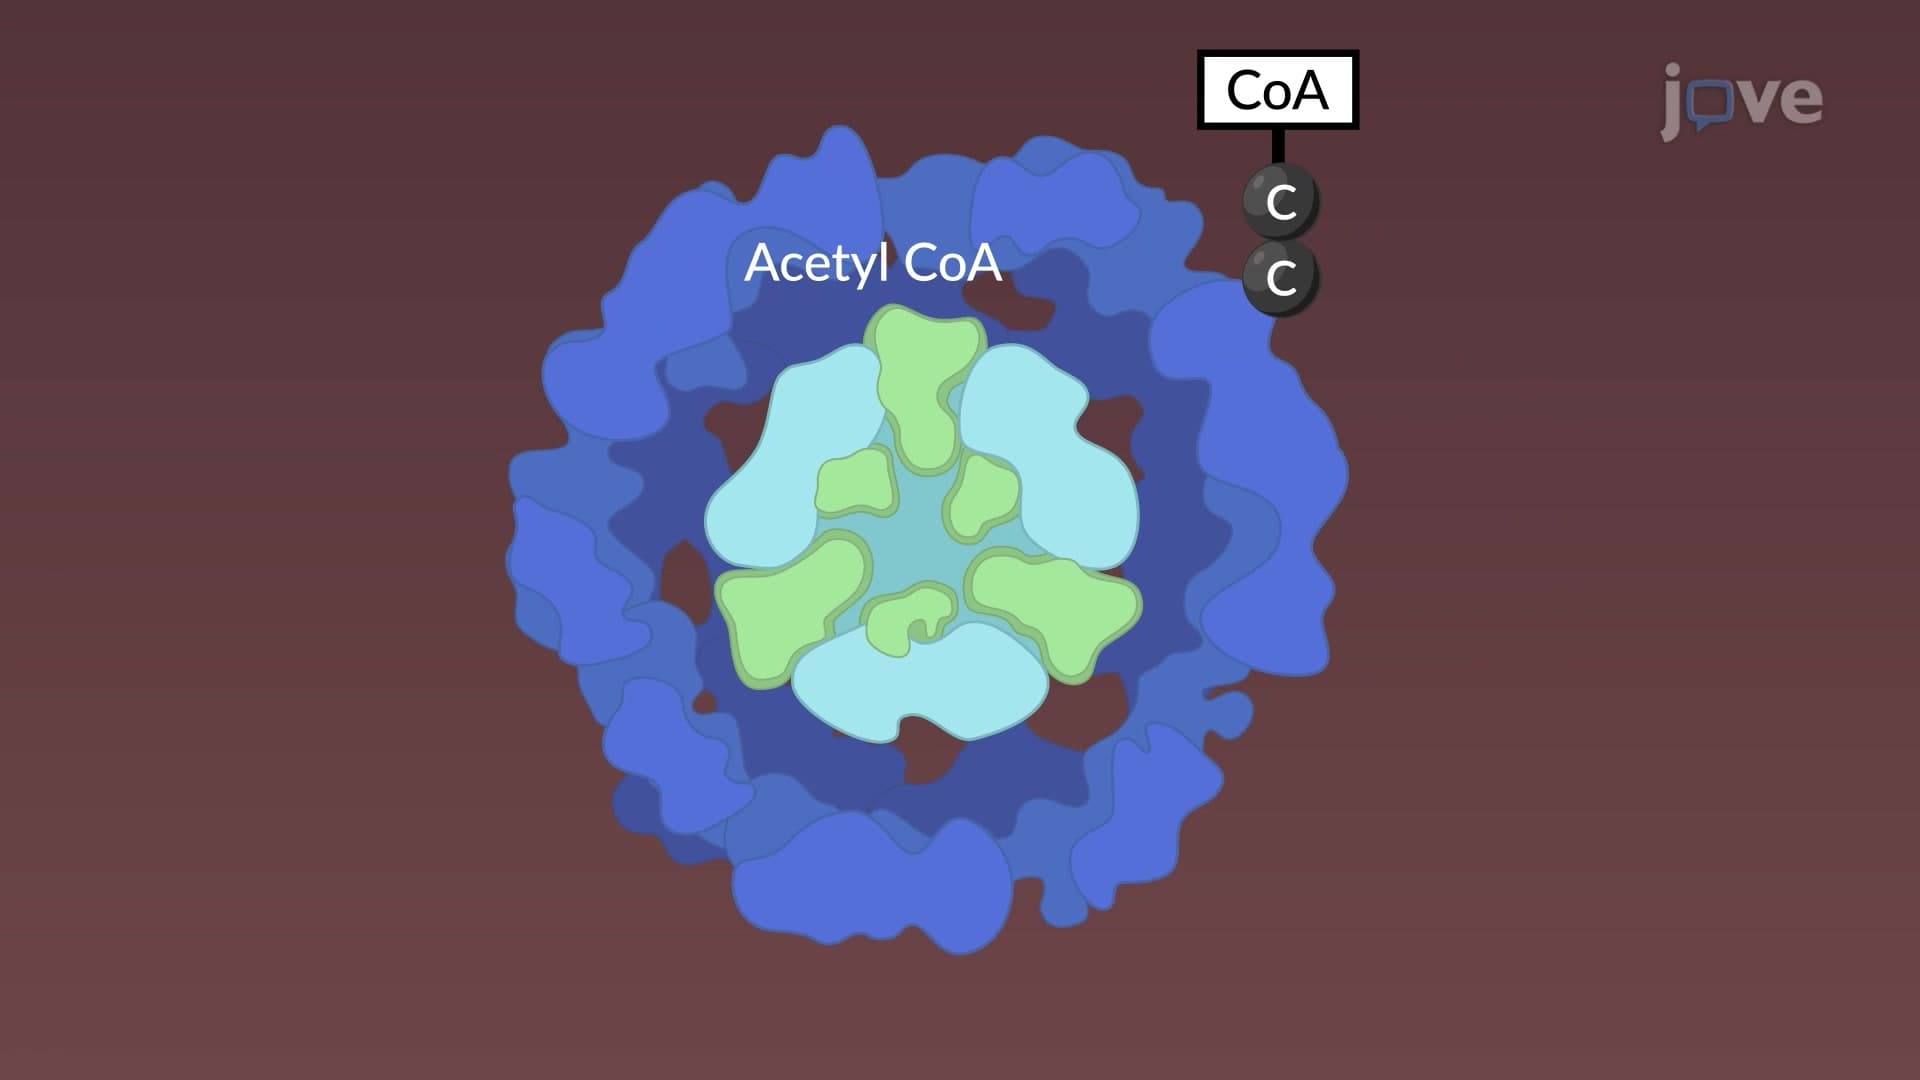
Pyruvate Oxidation
First, the enzyme pyruvate dehydrogenase removes the carboxyl group from pyruvate and releases it as carbon dioxide. The stripped molecule is then oxidized and releases electrons, which are then picked up by NAD+...
 01:24
Parkinson Disease ll: Pathophysiology
 01:24
Parkinson Disease l: Introduction