Brain adenosine A2A receptor occupancy by a novel A1/A2A receptor antagonist, ASP5854, in rhesus monkeys: relationship to anticataleptic effect
Related Experiment Video
Updated: Jul 4, 2026
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Takuma Mihara1, Akihiro Noda, Hiroshi Arai
1Pharmacology Research Laboratories, Astellas Pharma Inc., Tsukuba, Ibaraki, Japan. takuma.mihara@jp.astellas.com

Unlabelled:
The purpose of the present study was to measure adenosine A(2A) receptor (A(2A)R) occupancy in the brain by a novel adenosine A(1)/A(2A) antagonist, 5-[5-amino-3-(4fluorophenyl)pyrazin-2-yl]-1-isopropylpyridine-2(1H)-one (ASP5854), and to determine the degree of receptor occupancy necessary to inhibit haloperidol-induced catalepsy in rhesus monkeys.
Methods:
A(2A)R occupancy by ASP5854 (0.001-0.1 mg/kg) was examined in the striatum using an A(2A)R-specific radiotracer, (11)C-SCH442416, and PET in conscious rhesus monkeys. A(2A)R occupancy was monitored after a single intravenous administration of ASP5854 in 3 animals, and a dynamic PET scan was performed at 1, 4, and 8 h after an intravenous bolus injection of the tracer for approximately 740 MBq. Catalepsy was induced by haloperidol (0.03 mg/kg, intramuscularly) and examined for incidence and duration.
Results:
ASP5854 dose-dependently increased A(2A)R occupancy in the striatum and showed long-lasting occupancy even after the reduction of plasma concentration. Haloperidol induced severe catalepsy at 40 min after intramuscular injection. The incidence and duration of cataleptic posture were dose-dependently reduced by ASP5854 at 1 h after oral administration, and the minimum ED(50) value was 0.1 mg/kg. Administration of a dose of 0.1 mg/kg yielded a plasma concentration of 97 +/- 16.3 ng/mL, which corresponded to 85%-90% of A(2A)R occupancy.
Conclusion:
These results showed that ASP5854 antagonized A(2A)R in the striatum, and the dissociation from A(2A)R was relatively slow. In addition, more than 85% A(2A)R occupancy by ASP5854 resulted in an inhibition of haloperidol-induced catalepsy. Thus, such a pharmacodynamic study directly demonstrates both the kinetics of a drug in the brain and the relationship between dose-dependent receptor occupancy and plasma level.
Related Concept Videos
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:17
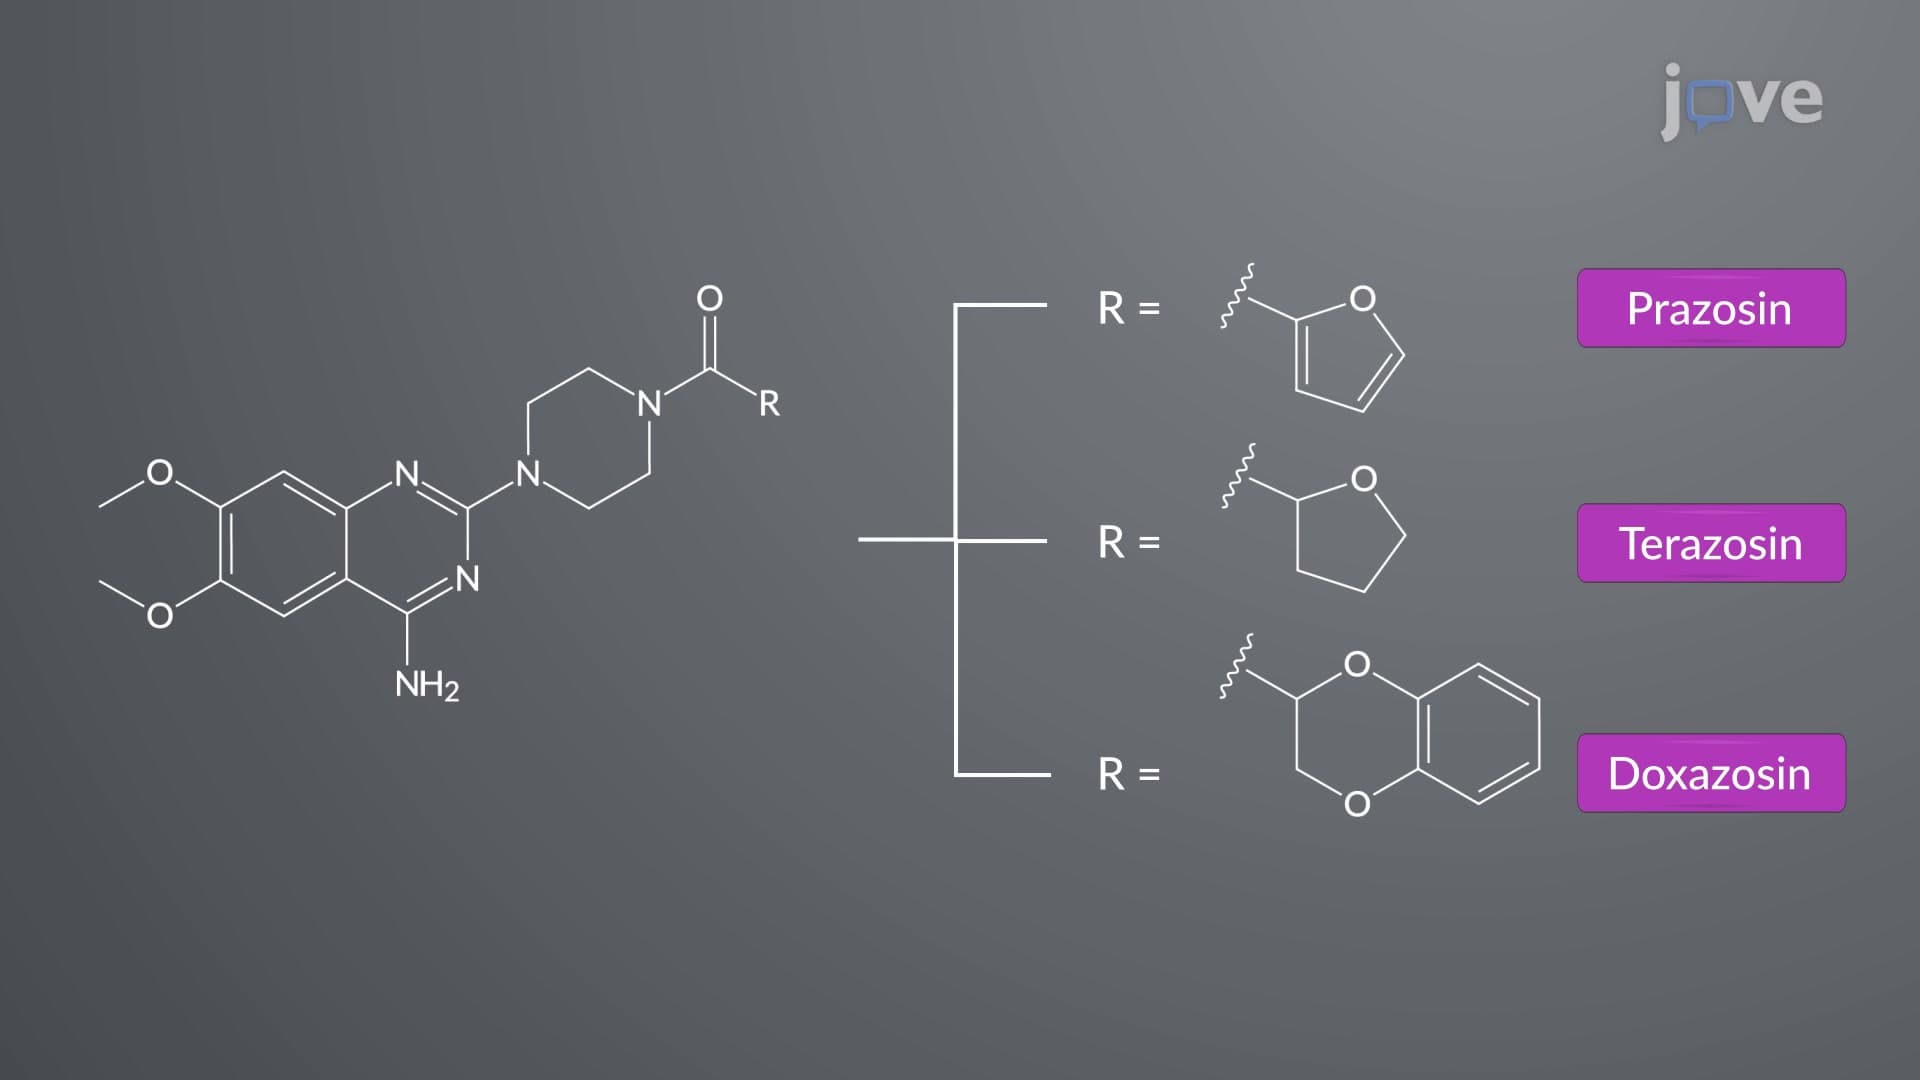
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:21
Drugs Affecting Neurotransmitter Release or Uptake
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:25
Adrenergic Agonists: Indirect-Acting Agents
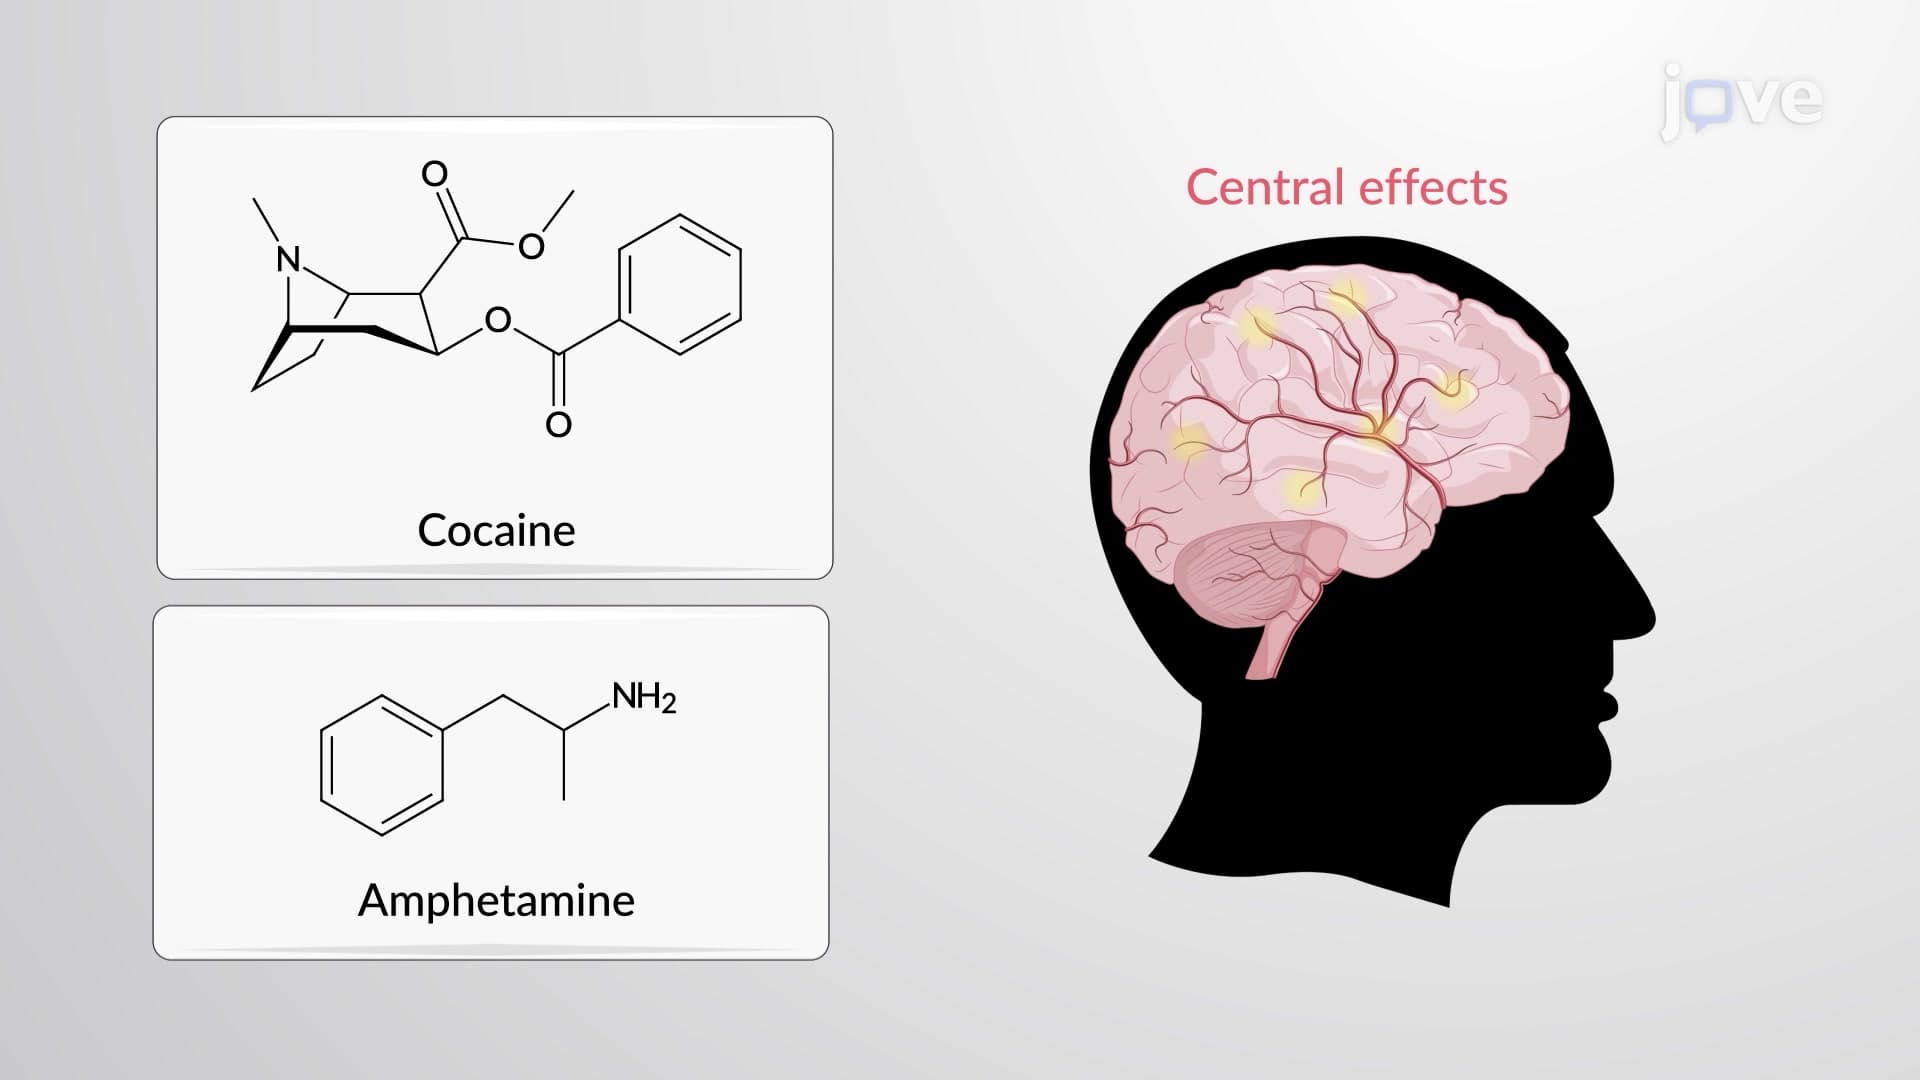
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...