Related Experiment Video
Updated: Jul 4, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
A first-principles study of molecular oxygen dissociation at an electrode surface: a comparison of potential
Sally A Wasileski1, Michael J Janik
1Department of Chemistry, University of North Carolina at Asheville, One University Heights CPO #2310, Asheville, NC 28804, USA. swasiles@unca.edu

Abstract:
Influences of coadsorbed sodium and water, aqueous solvent, and electrode potential on the kinetics of O(2) dissociation over Pt(111) are systematically investigated using density functional theory models of vacuum and electrochemical interfaces. Na coadsorption alters the electronic states of Pt to stabilize the reactant (O(2)*), transition, and product (2O*) states by facilitating electron donation to oxygen, causing a more exothermic reaction energy (-0.84 eV for Na and O(2), -0.81 eV for isolated O(2)) and a decrease in dissociation barrier (0.39 eV for Na and O(2), 0.57 eV for isolated O(2)). Solvation decreases the reaction energy (-0.67 eV) due to enhanced hydrogen bond stabilization of O(2)* compared to 2O*. The influence of Na is less pronounced at the solvated interface (barrier decreases by only 0.11 eV) because H(2)O screens Na charge-donation. In the electrochemical model system, the dissociation energy becomes more exothermic and the barrier decreases toward more positive potentials. Potential-dependent behavior results from changes in interfacial dipole moment and polarizability between O(2)*, the dissociation transition state, and 2O*; each are influenced by changes in adsorption and hydrogen bonding. Coadsorption of Na in the solvated system dampens the dipole moment change between O(2)* and 2O* and significantly increases the polarizability at the dissociation transition state and for 2O*; the combination causes little change in the reaction energy but reduces the activation barrier by 0.08 eV at 0 V versus NHE. The potential-dependent behavior contrasts that determined at a constant surface charge or from an applied electric field, illustrating the importance of considering the electrochemical potential at the fully-solvated interface in determining reaction energetics, even for non-redox reactions.
More Related Videos
 08:31
Anaerobic Protein Purification and Kinetic Analysis via Oxygen Electrode for Studying DesB Dioxygenase Activity and Inhibition
Published on: October 3, 2018
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Related Concept Videos
 01:30
Processes at Electrodes
 01:30
The Electrical Double Layer
 01:06
Interfacial Electrochemical Methods: Overview
 03:02
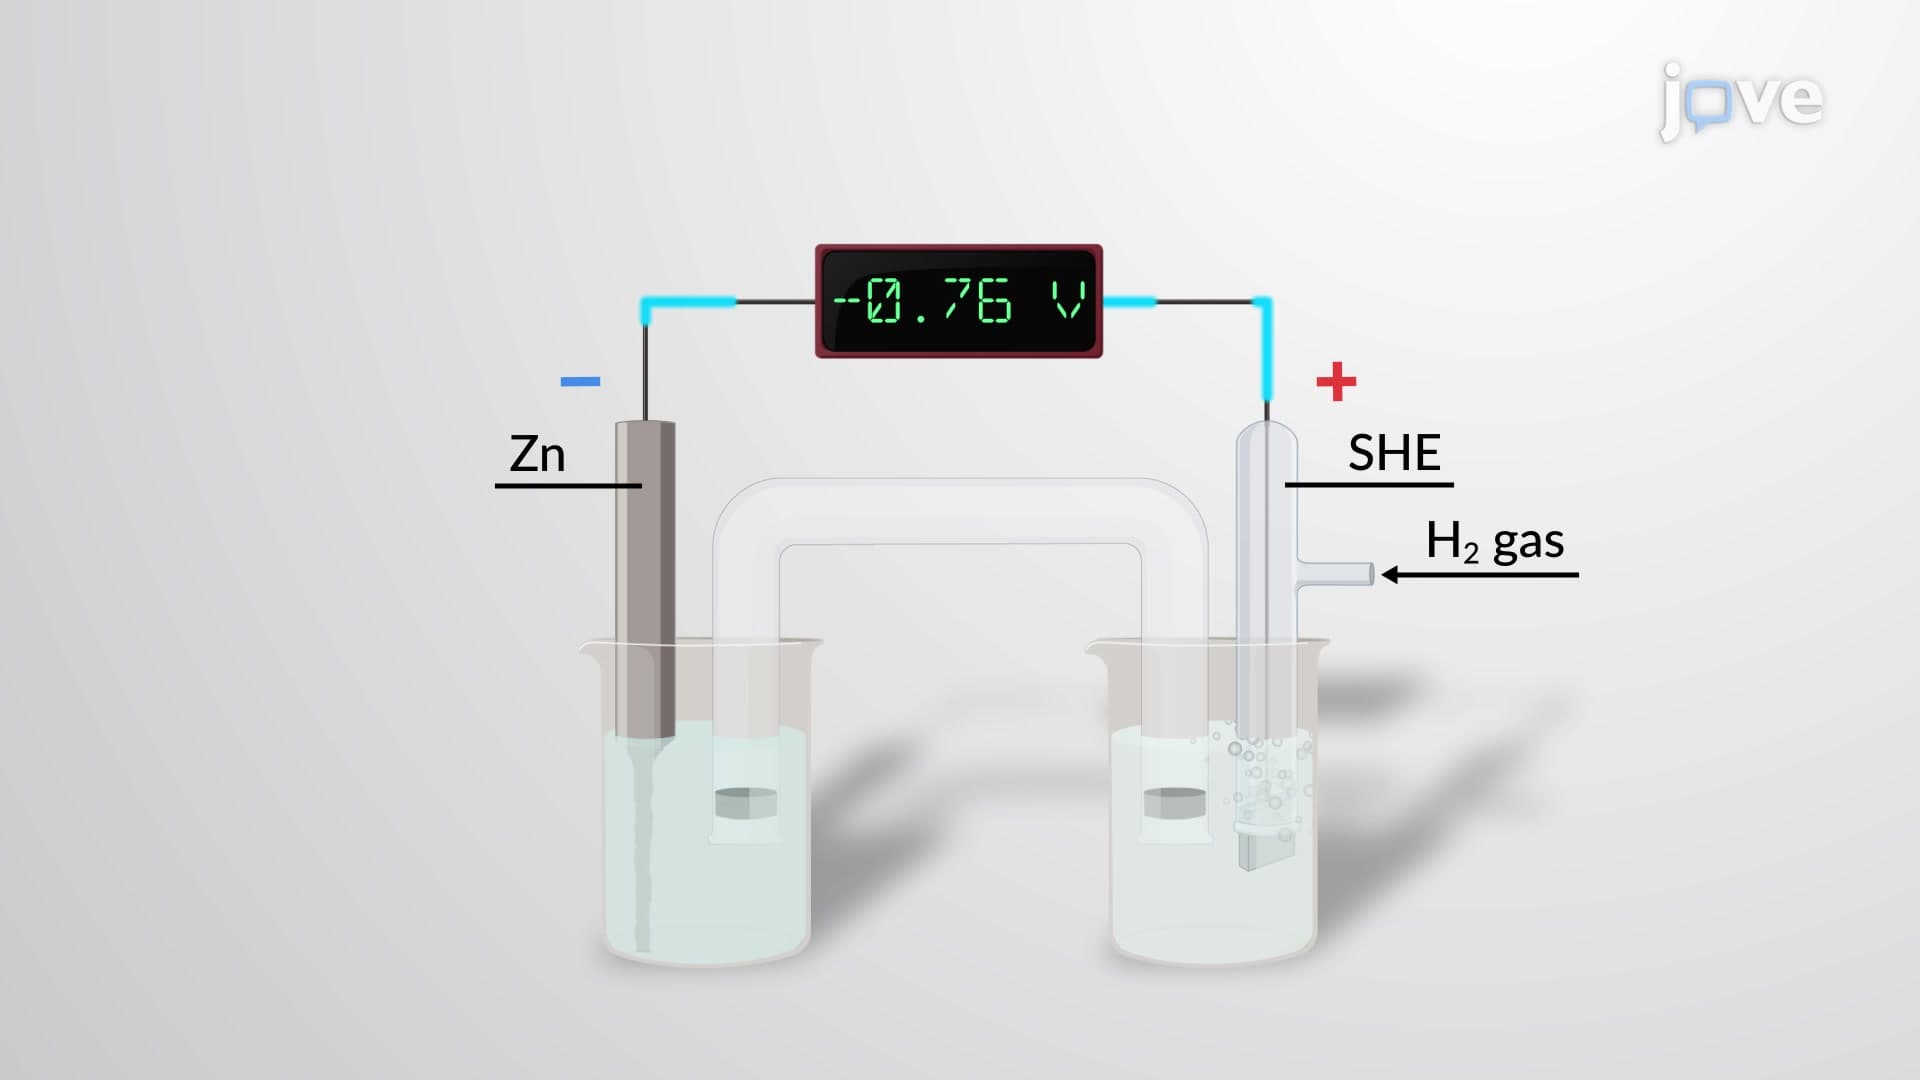
Standard Electrode Potentials
 01:21
Voltammetry: Factors Affecting Measurements
 01:23
Redox Equilibria: Overview