Related Experiment Video
Updated: Jul 4, 2026
Enzymatic Synthesis of Epoxidized Metabolites of Docosahexaenoic, Eicosapentaenoic, and Arachidonic Acids
Published on: June 28, 2019
Stereochemistry of eudesmane cation formation during catalysis by aristolochene synthase from Penicillium roqueforti
David J Miller1, Jiali Gao, Donald G Truhlar
1School of Chemistry, Main Building, Cardiff University, Park Place, Cardiff, UK CF10 3AT.

Abstract:
The aristolochene synthase catalysed cyclisation of farnesyl diphosphate (1) has been postulated to proceed through (S)-germacrene A (3). However, the active site acid that reprotonates this neutral intermediate has so far proved difficult to identify and, based on high level ab initio molecular orbital and density functional theory calculations, a proton transfer mechanism has recently been proposed, in which proton transfer from C12 of germacryl cation to the C6,C7-double bond of germacryl cation (2) proceeds either directly or via a tightly bound water molecule. In this work, the stereochemistry of the elimination and protonation reactions was investigated by the analysis of the reaction products from incubation of 1 and of [12,12,12,13,13,13-(2)H(6)]-farnesyl diphosphate (15) with aristolochene synthase from Penicillium roqueforti (PR-AS) in H(2)O and D(2)O. The results reveal proton loss from C12 during the reaction and incorporation of another proton from the solvent. Incubation of with PR-AS in D(2)O led to the production of (6R)-[6-(2)H] aristolochene, indicating that protonation occurs from the face of the 10-membered germacrene ring opposite the isopropylidene group. Hence these results firmly exclude proton transfer from C12 to C6 of germacryl cation. We propose here Lys 206 as the general acid/base during PR-AS catalysis. This residue is part of a conserved network of hydrogen bonds, along which protons could be delivered from the solvent to the active site.
Related Concept Videos
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 02:43
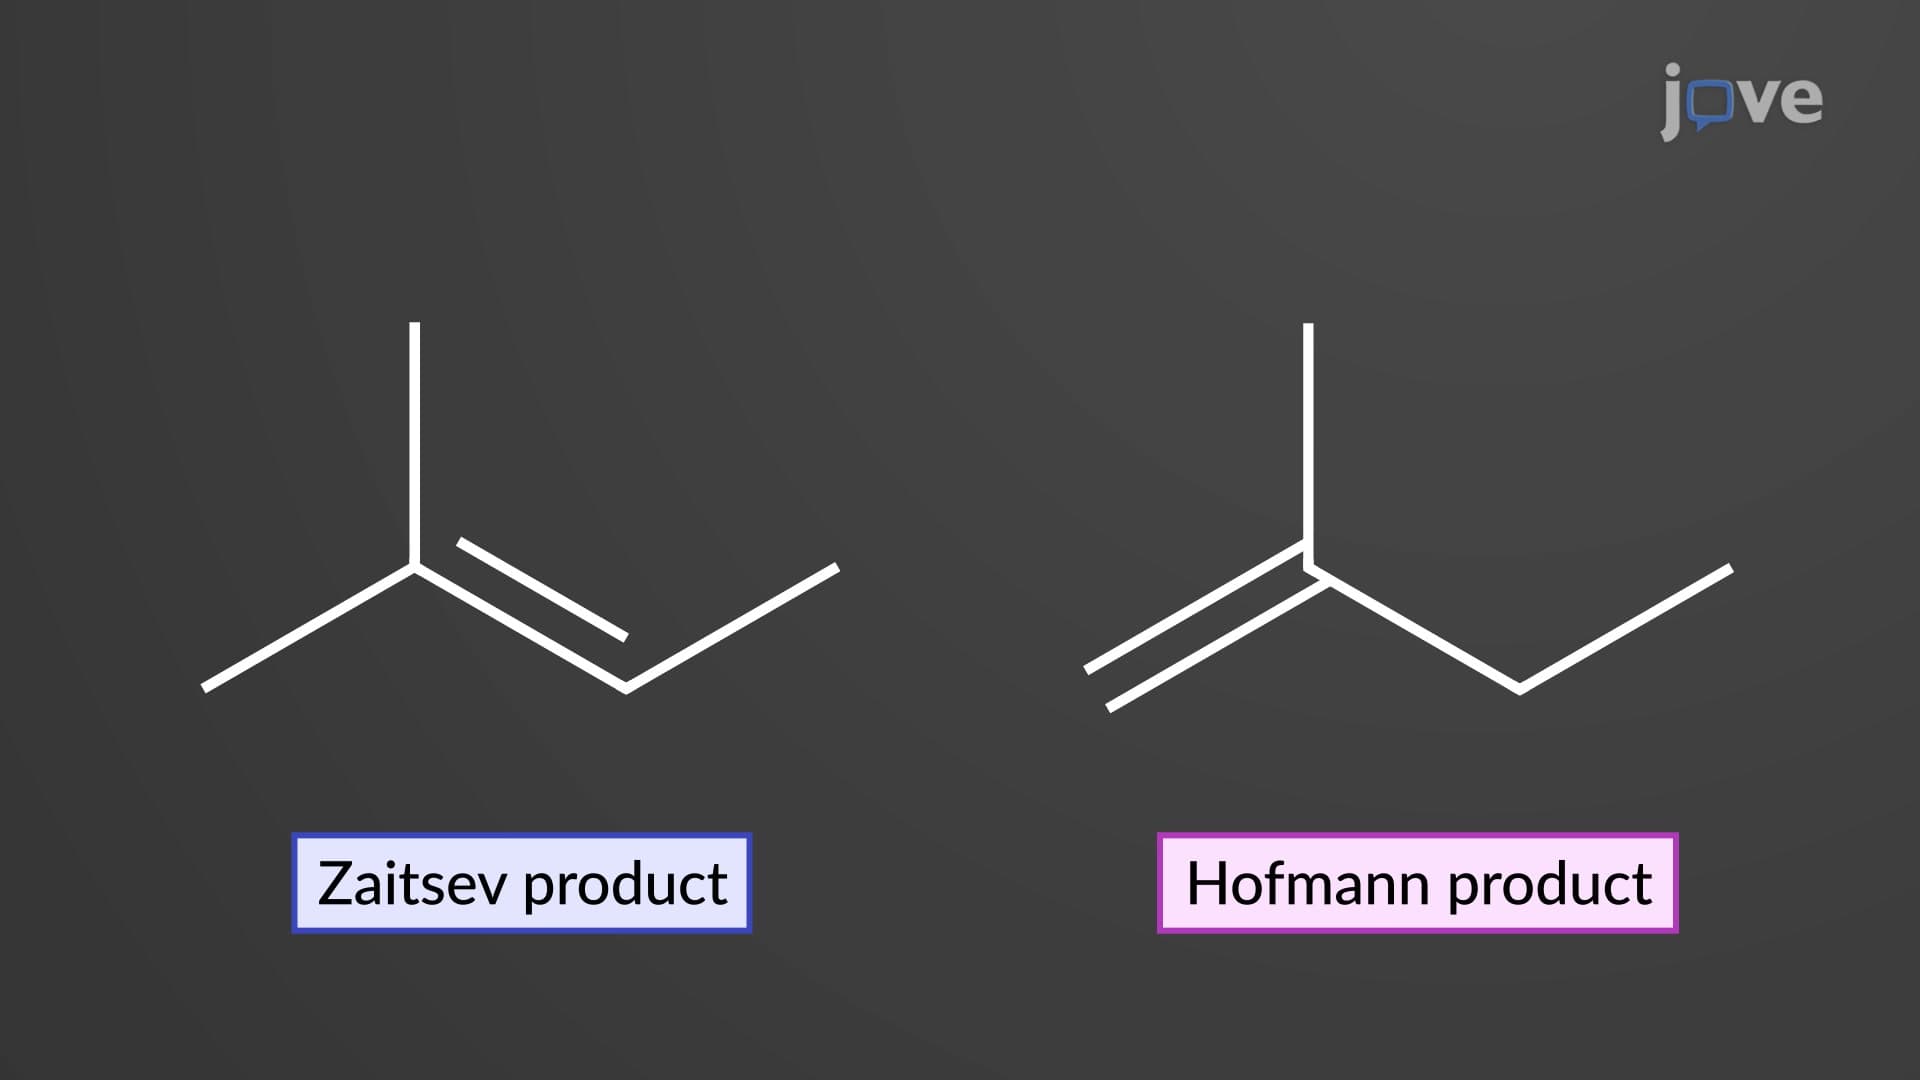
E1 Reaction: Stereochemistry and Regiochemistry
 01:28
Diels–Alder Reaction Forming Cyclic Products: Stereochemistry
 02:44
Oxidation of Alkenes: Syn Dihydroxylation with Osmium Tetraoxide