Related Experiment Video
Updated: Jul 4, 2026
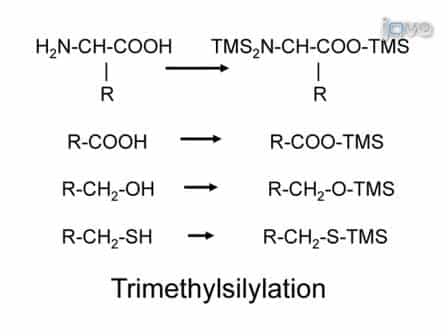

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Phenylketonuria: an inborn error of phenylalanine metabolism
Robin A Williams1, Cyril D S Mamotte, John R Burnett
1Department of Core Clinical Pathology & Biochemistry, PathWest Laboratory Medicine, Royal Perth Hospital, Perth, WA 6847, Australia. robin.williams@health.wa.gov.au

Abstract:
Phenylketonuria (PKU) is an autosomal recessive inborn error of phenylalanine (Phe) metabolism resulting from deficiency of phenylalanine hydroxylase (PAH). Most forms of PKU and hyperphenylalaninaemia (HPA) are caused by mutations in the PAH gene on chromosome 12q23.2. Untreated PKU is associated with an abnormal phenotype which includes growth failure, poor skin pigmentation, microcephaly, seizures, global developmental delay and severe intellectual impairment. However, since the introduction of newborn screening programs and with early dietary intervention, children born with PKU can now expect to lead relatively normal lives. A better understanding of the biochemistry, genetics and molecular basis of PKU, as well as the need for improved treatment options, has led to the development of new therapeutic strategies.
More Related Videos
 10:38
Simultaneous Quantification of Selected Kynurenines Analyzed by Liquid Chromatography-Mass Spectrometry in Medium Collected from Cancer Cell Cultures
Published on: May 9, 2020
 09:54
Lentiviral Vector-mediated Gene Therapy of Hepatocytes Ex Vivo for Autologous Transplantation in Swine
Published on: November 4, 2018
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:24
Pharmacokinetics in Pediatric Patients: Drug Metabolism
 01:23
Urea Cycle
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:29
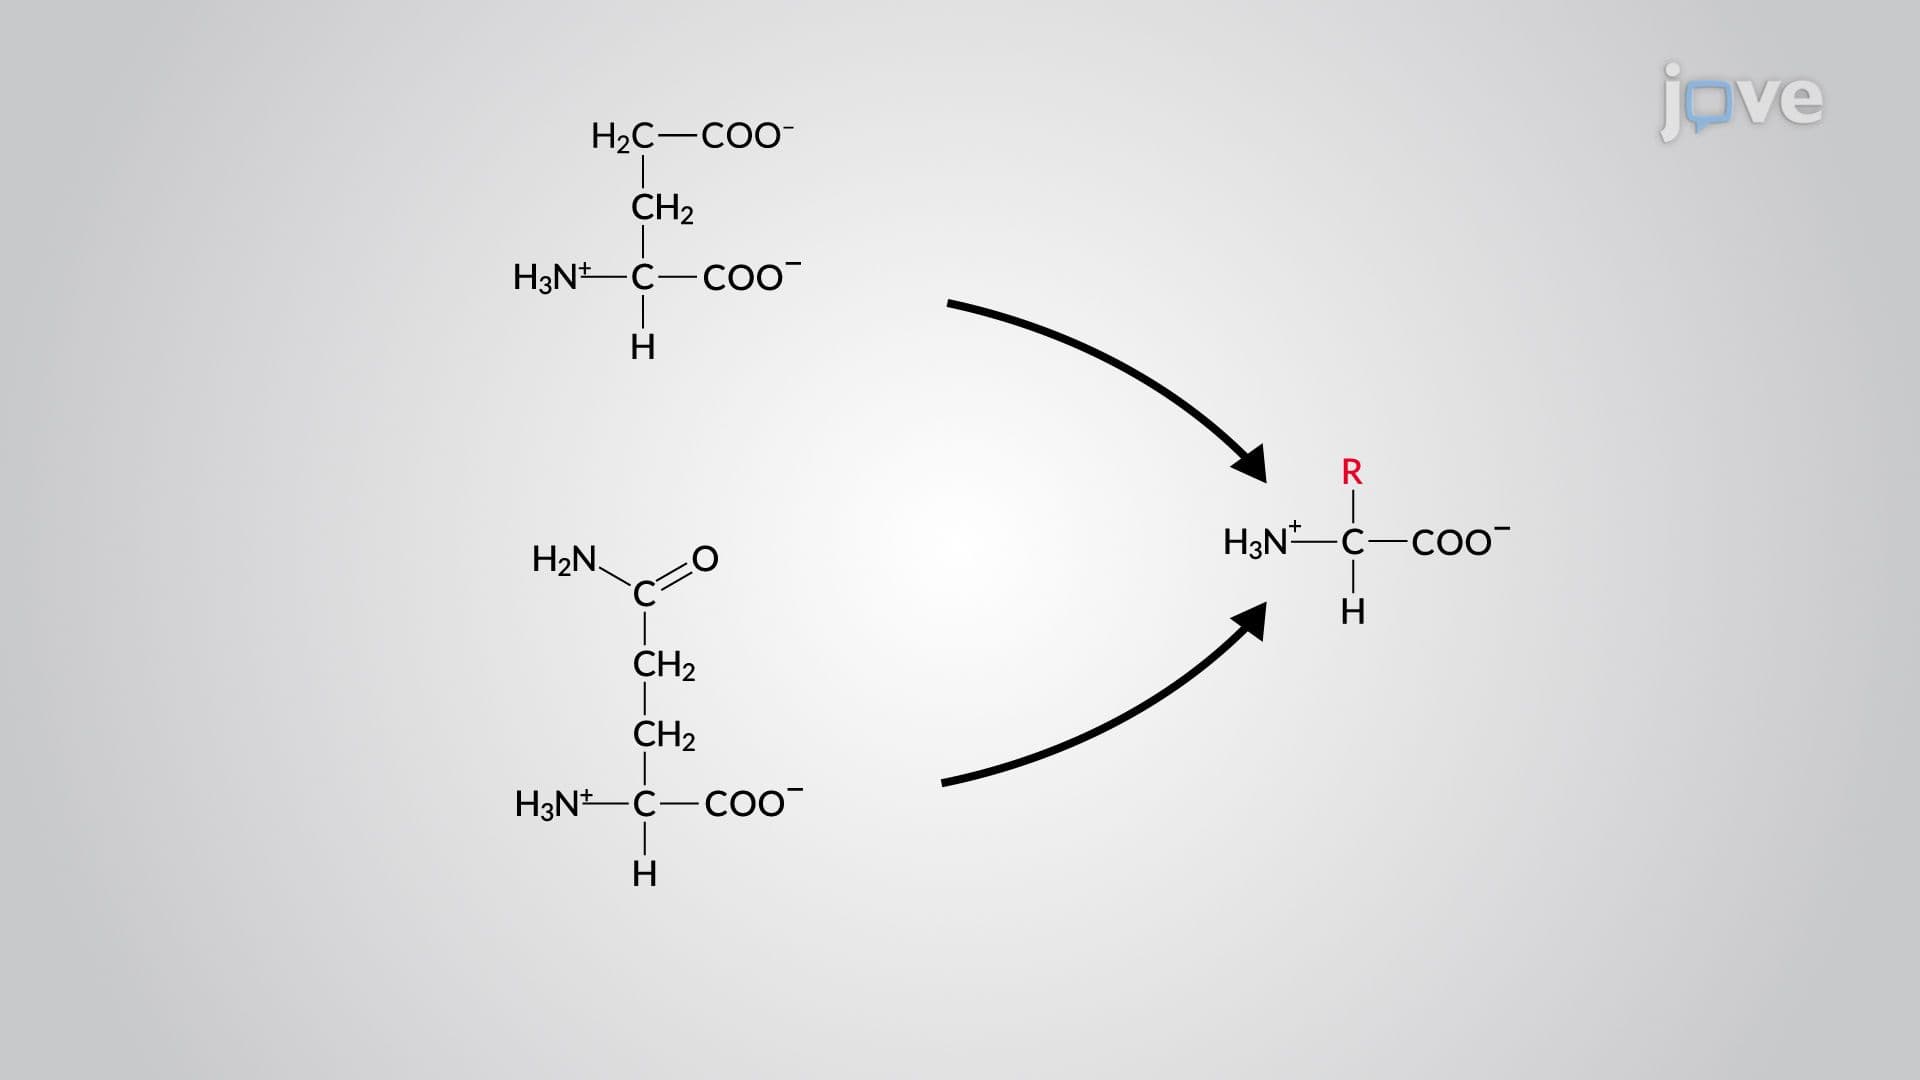
Amino Acid Biosynthetic Pathways