Related Experiment Video
Updated: Jul 4, 2026
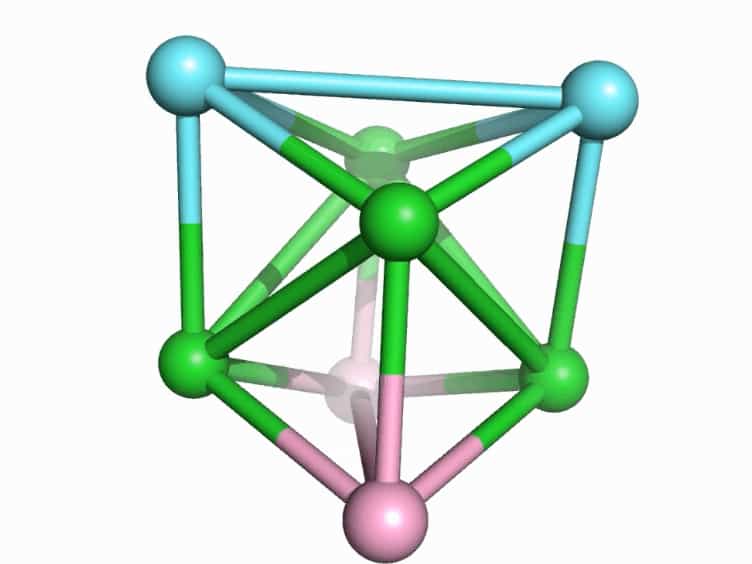

Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Molecular structures and energetics of the (TiO2)n (n = 1-4) clusters and their anions
1Chemistry Department, The University of Alabama, Shelby Hall, Box 870336, Tuscaloosa, Alabama 35487-0336, USA.

Abstract:
The (TiO2)n clusters and their anions for n = 1-4 have been studied with coupled cluster theory [CCSD(T)] and density functional theory (DFT). For n > 1, numerous conformations are located for both the neutral and anionic clusters, and their relative energies are calculated at both the DFT and CCSD(T) levels. The CCSD(T) energies are extrapolated to the complete basis set limit for the monomer and dimer and calculated up to the triple-zeta level for the trimer and tetramer. The adiabatic and vertical electron detachment energies of the anionic clusters to the ground and first excited states of the neutral clusters are calculated at both levels and compared with the experimental results. The comparison allows for the definitive assignment of the ground-state structures of the anionic clusters. Anions of the dimer and tetramer are found to have very closely lying conformations within 2 kcal/mol at the CCSD(T) level, whereas that of the trimer does not. In addition, accurate clustering energies and heats of formation are calculated for the neutral clusters and compared with the available experimental data. Estimates of the titanium-oxygen bond energies show that they are stronger than the group VIB transition metal-oxygen bonds except for tungsten. The atomization energies of these clusters display much stronger basis set dependence than the clustering energies. This allows the calculation of more accurate heats of formation for larger clusters on the basis of calculated clustering energies.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:11
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:27
Predicting Molecular Geometry
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:48
Ionic Bonding and Electron Transfer