Ab-initio simulations of materials using VASP: Density-functional theory and beyond
Related Experiment Video
Updated: Jul 3, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
1Faculty of Physics and Center for Computational Materials Science, Universität Wien, Sensengasse 8, A-1090 Wien, Austria. juergen.hafner@univie.ac.at

Abstract:
During the past decade, computer simulations based on a quantum-mechanical description of the interactions between electrons and between electrons and atomic nuclei have developed an increasingly important impact on solid-state physics and chemistry and on materials science-promoting not only a deeper understanding, but also the possibility to contribute significantly to materials design for future technologies. This development is based on two important columns: (i) The improved description of electronic many-body effects within density-functional theory (DFT) and the upcoming post-DFT methods. (ii) The implementation of the new functionals and many-body techniques within highly efficient, stable, and versatile computer codes, which allow to exploit the potential of modern computer architectures. In this review, I discuss the implementation of various DFT functionals [local-density approximation (LDA), generalized gradient approximation (GGA), meta-GGA, hybrid functional mixing DFT, and exact (Hartree-Fock) exchange] and post-DFT approaches [DFT + U for strong electronic correlations in narrow bands, many-body perturbation theory (GW) for quasiparticle spectra, dynamical correlation effects via the adiabatic-connection fluctuation-dissipation theorem (AC-FDT)] in the Vienna ab initio simulation package VASP. VASP is a plane-wave all-electron code using the projector-augmented wave method to describe the electron-core interaction. The code uses fast iterative techniques for the diagonalization of the DFT Hamiltonian and allows to perform total-energy calculations and structural optimizations for systems with thousands of atoms and ab initio molecular dynamics simulations for ensembles with a few hundred atoms extending over several tens of ps. Applications in many different areas (structure and phase stability, mechanical and dynamical properties, liquids, glasses and quasicrystals, magnetism and magnetic nanostructures, semiconductors and insulators, surfaces, interfaces and thin films, chemical reactions, and catalysis) are reviewed.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 01:10
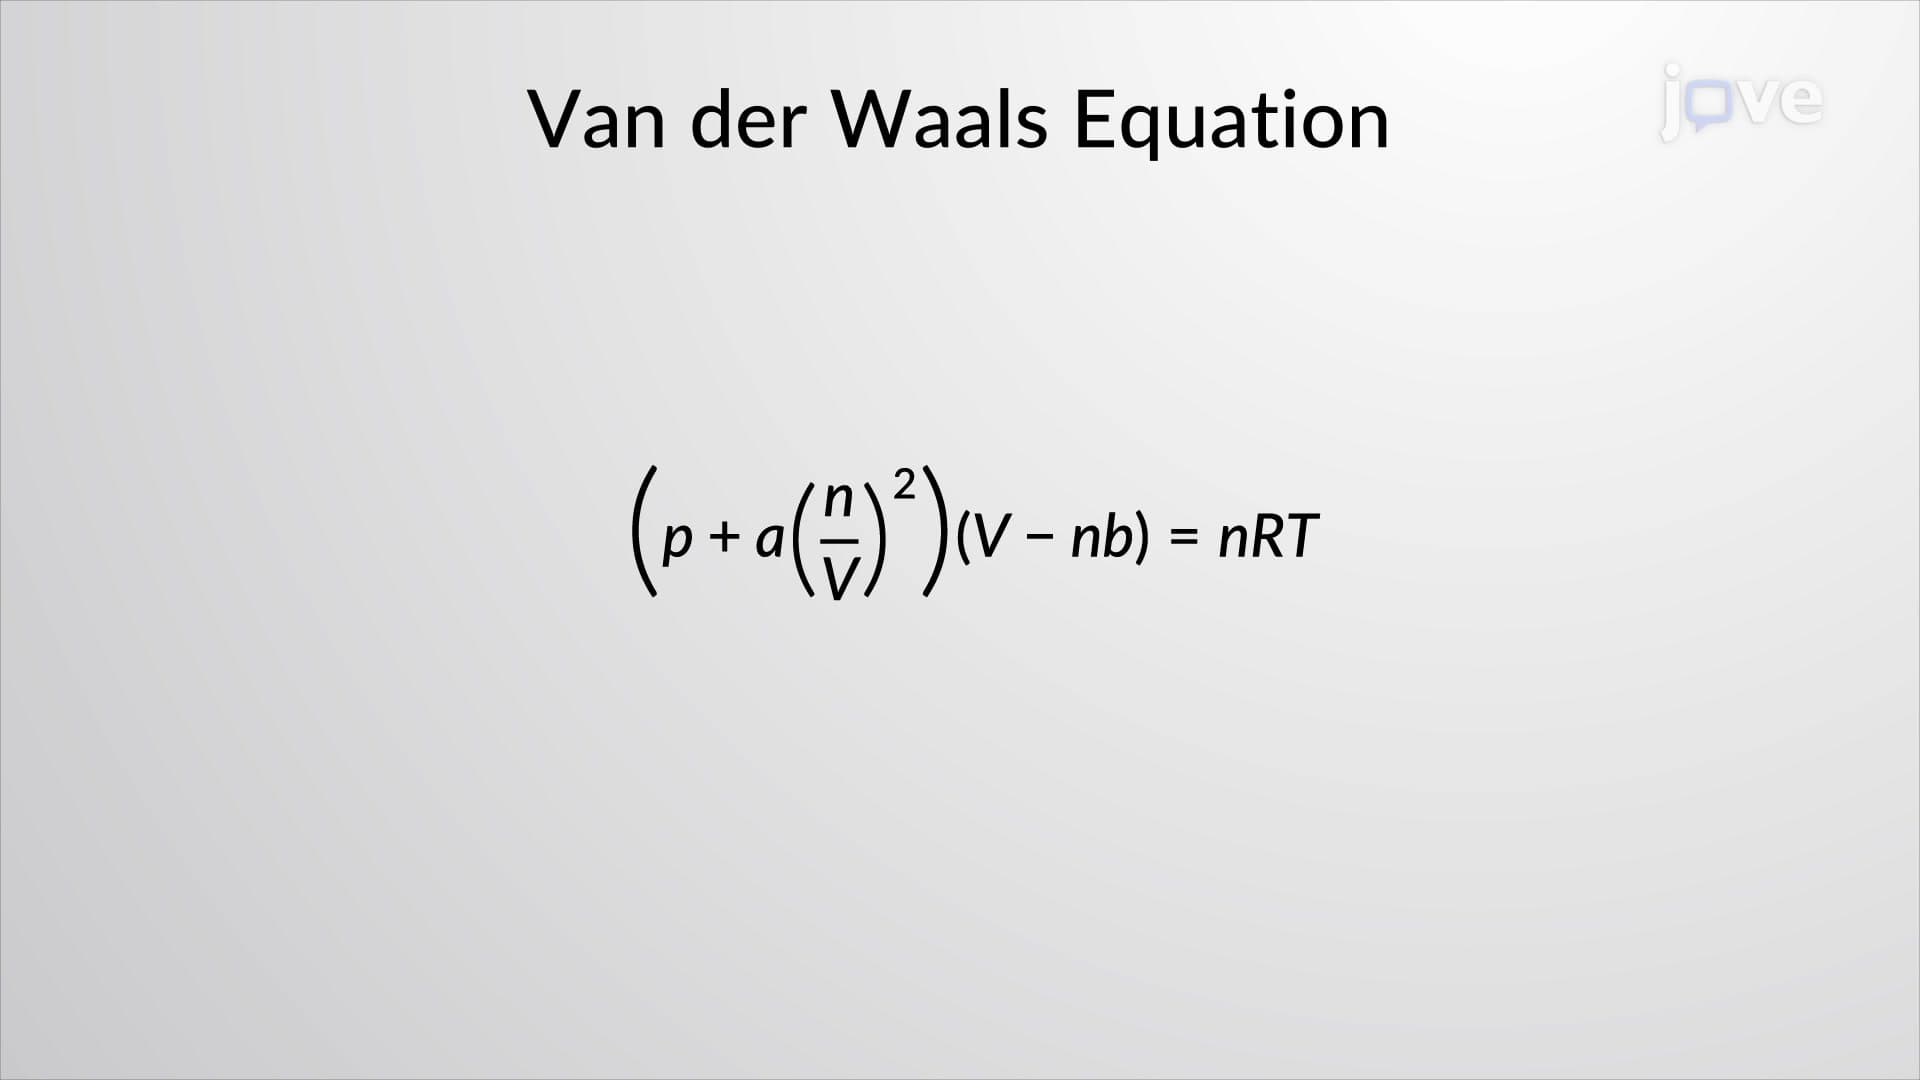
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:35
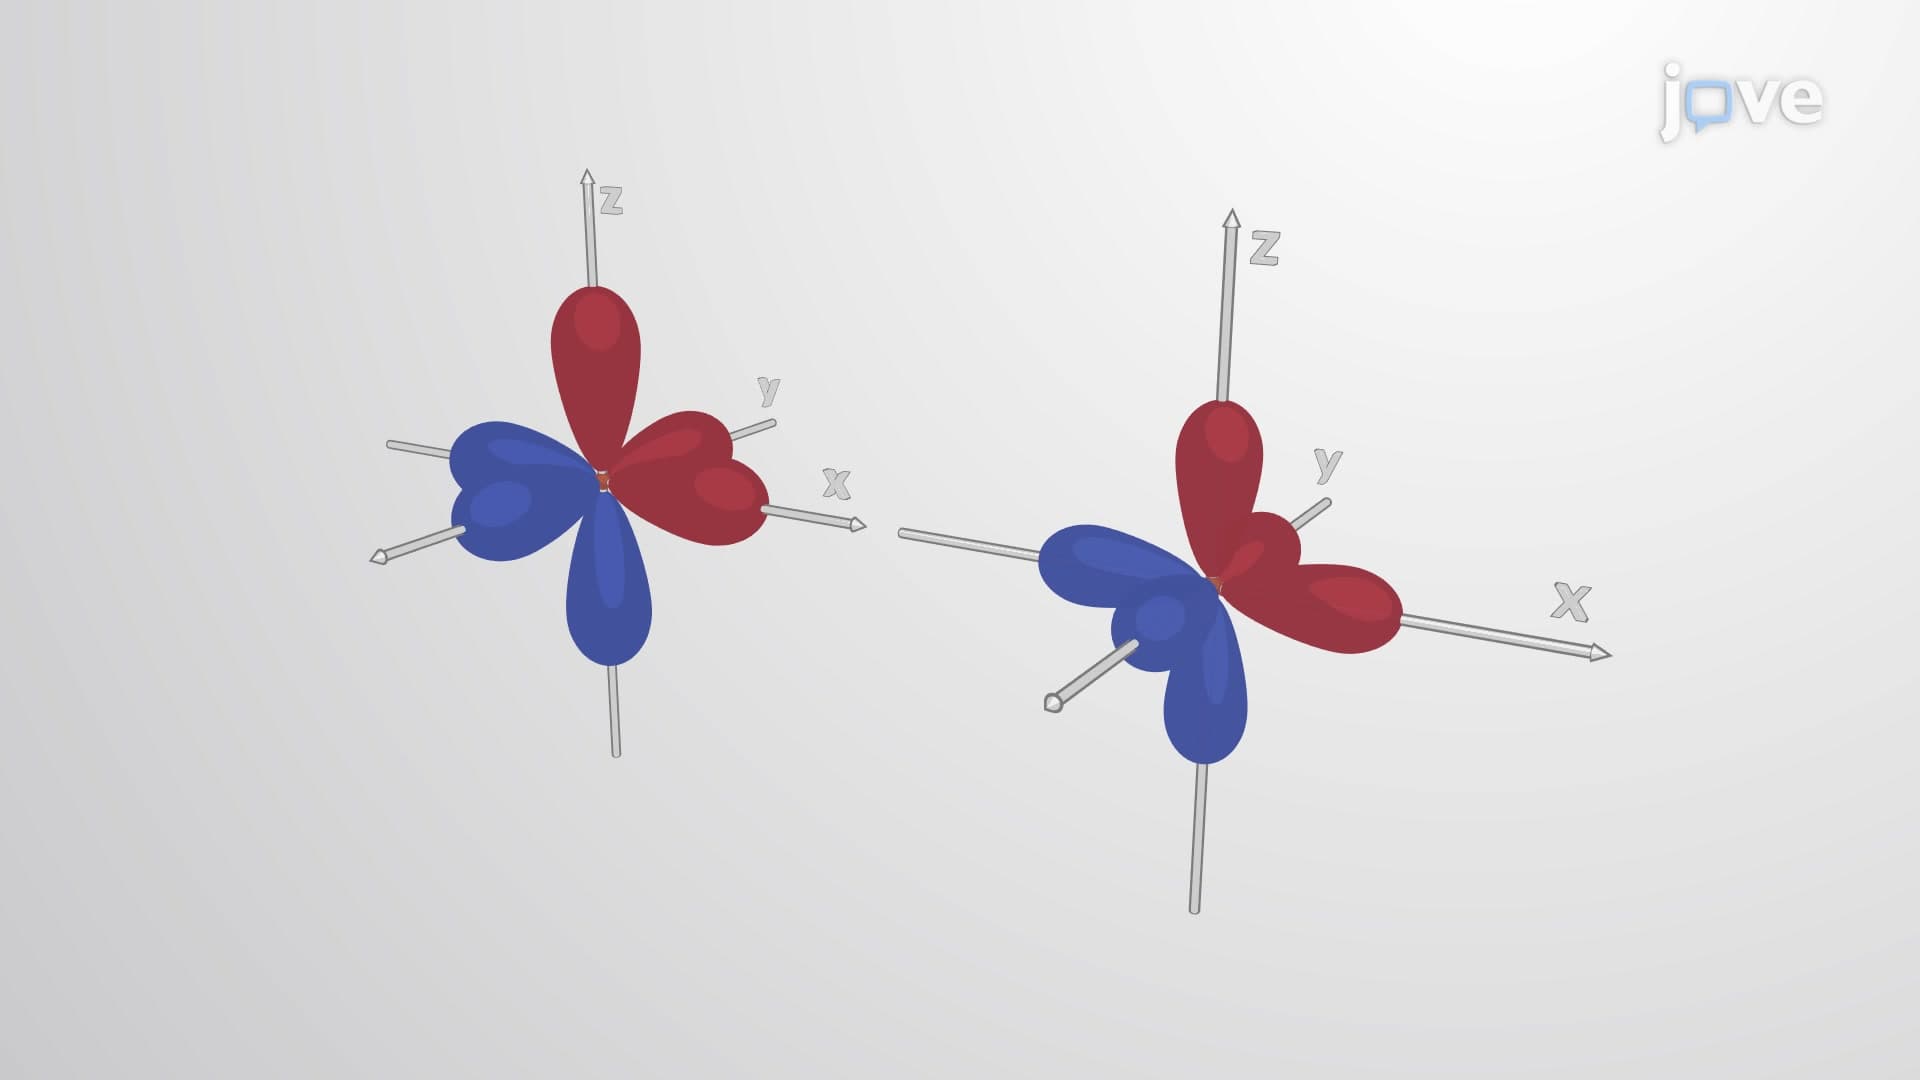
Molecular Orbital Theory I
 02:38
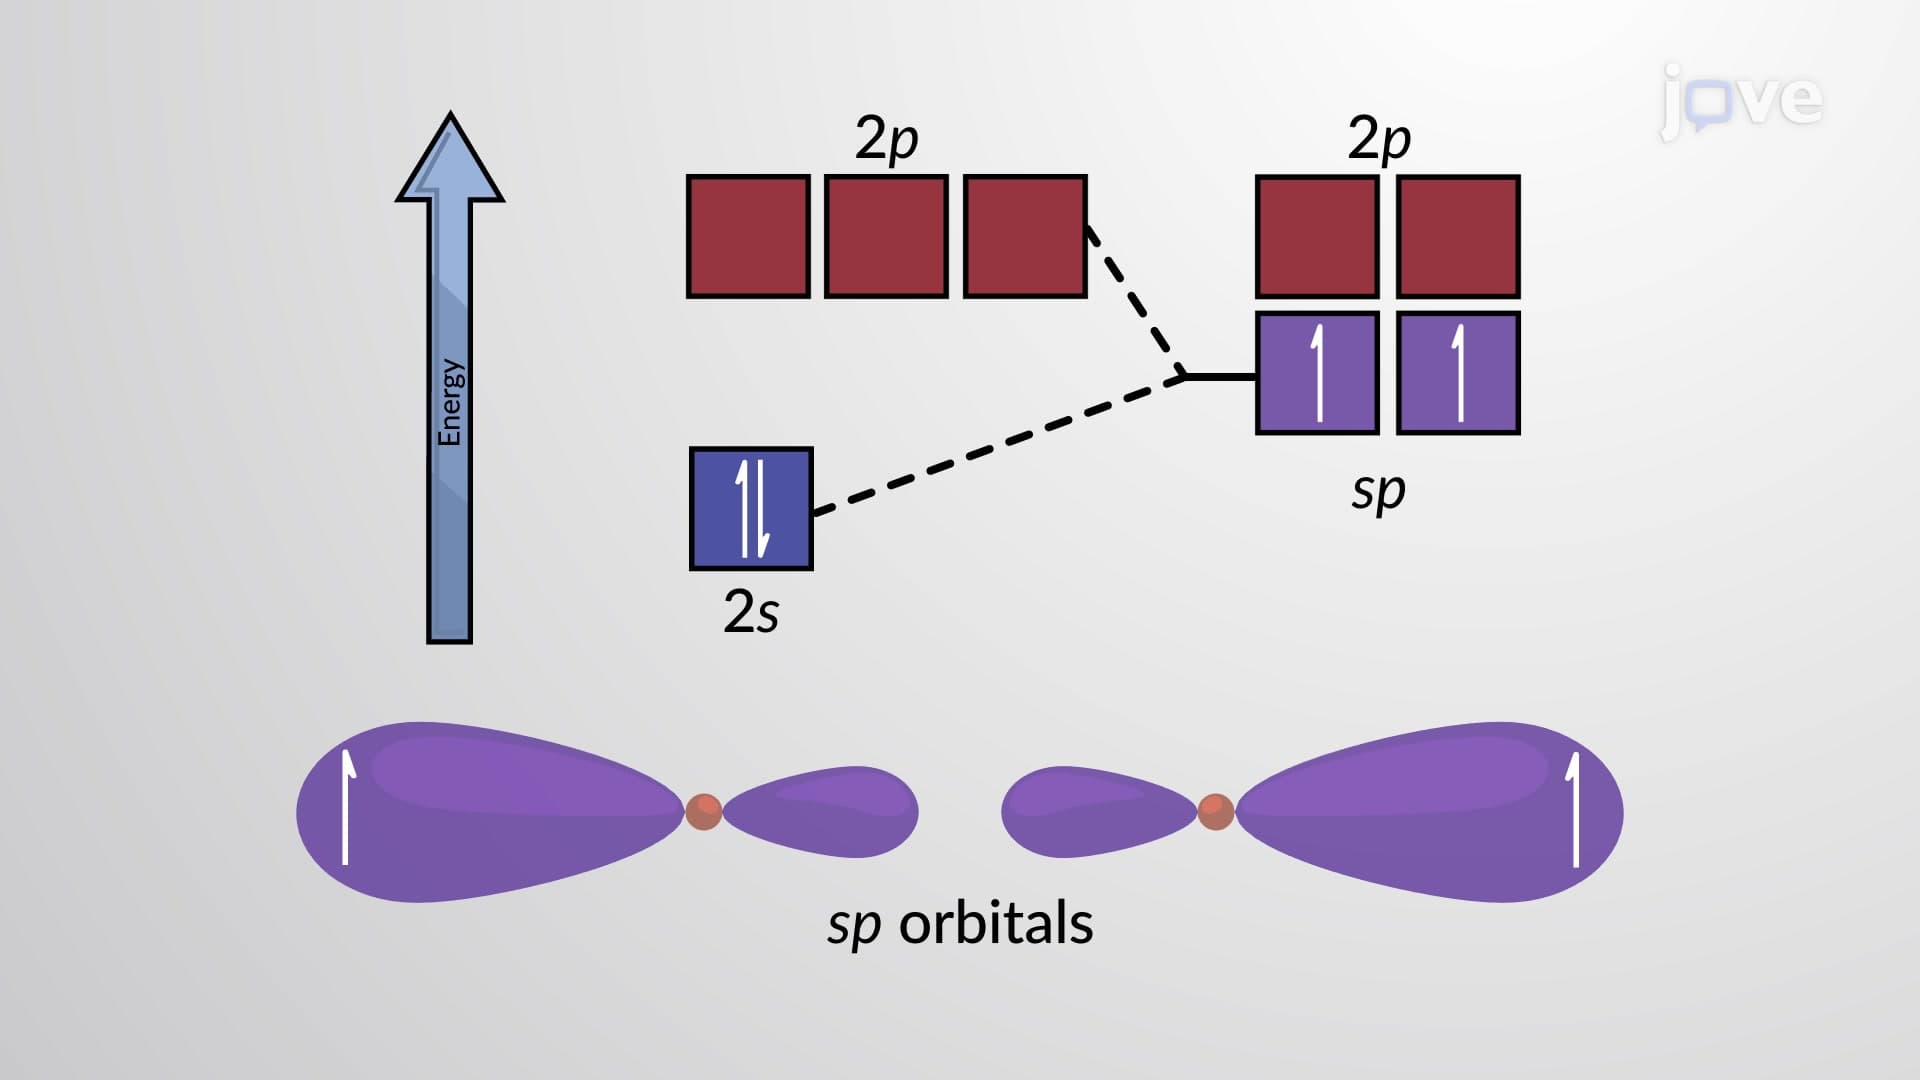
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:24
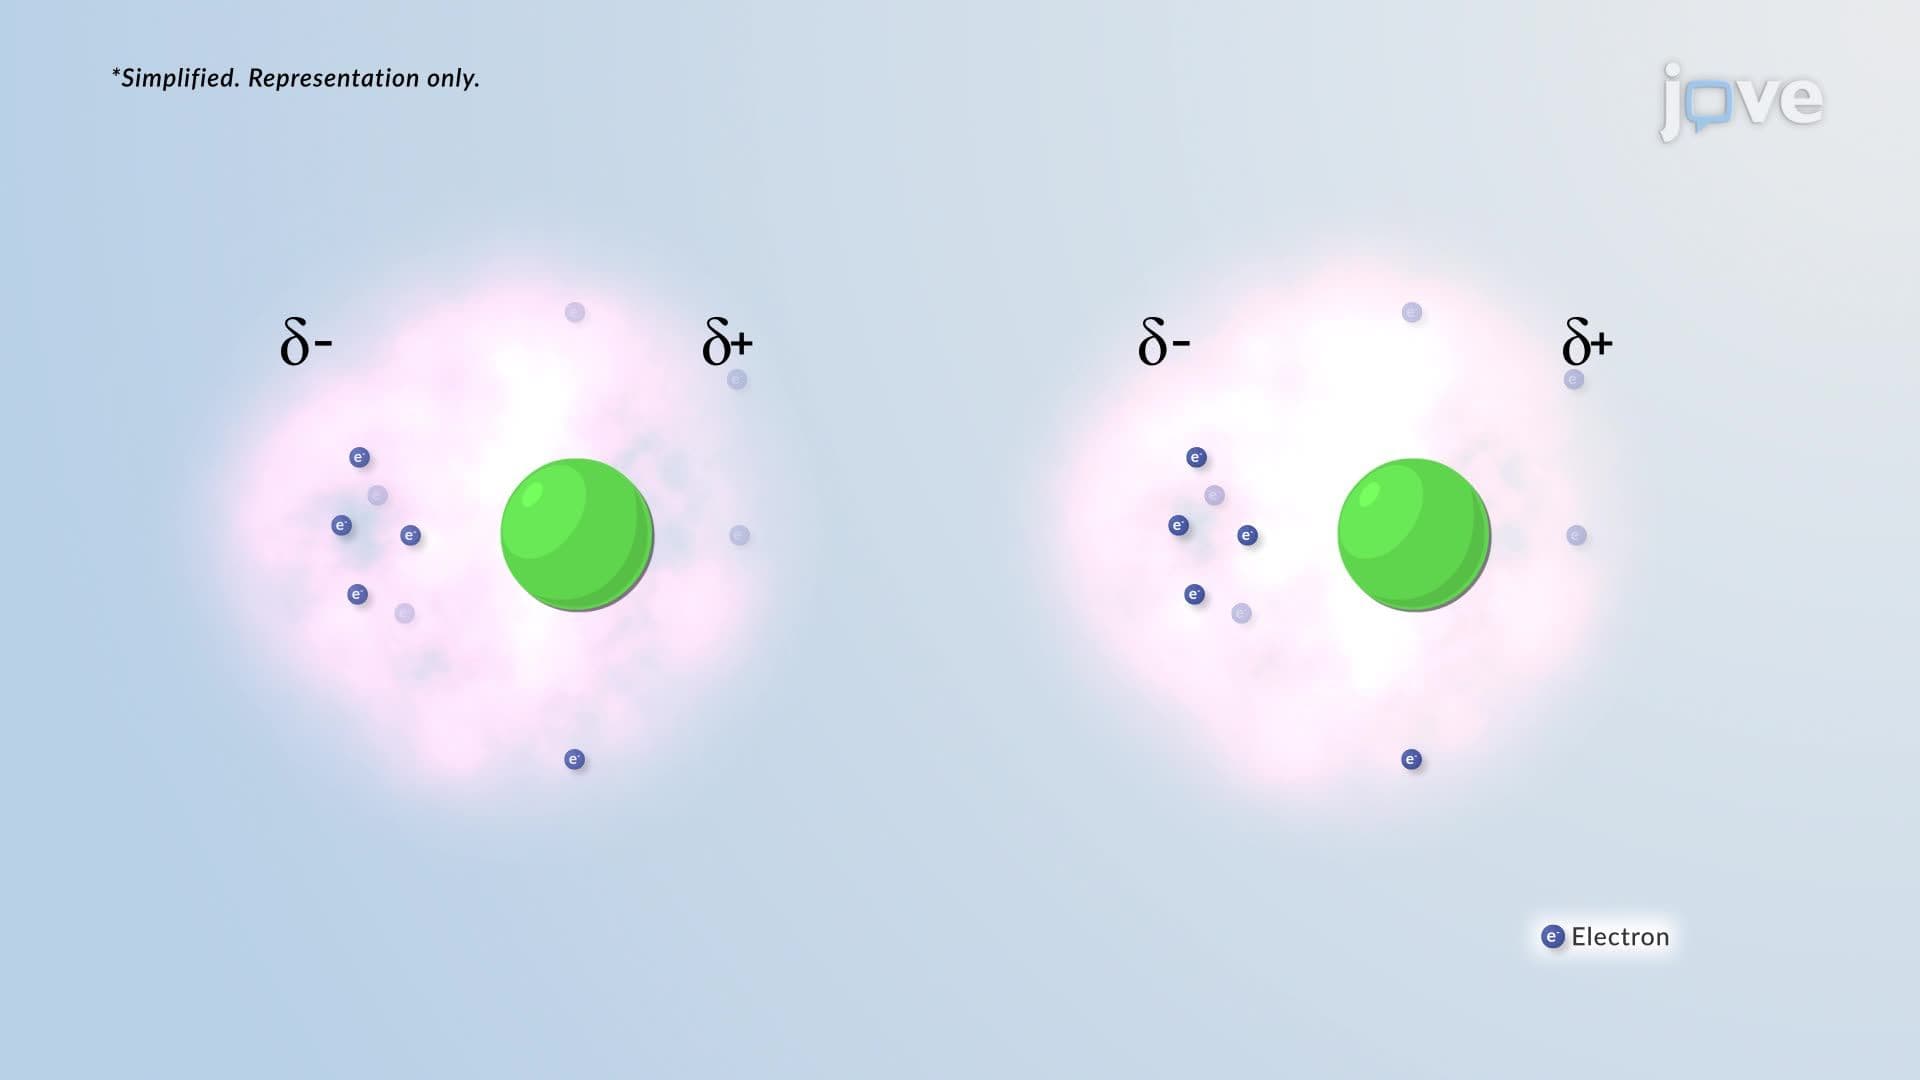
Van der Waals Interactions
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...